GNPTG-Mucolipidosis
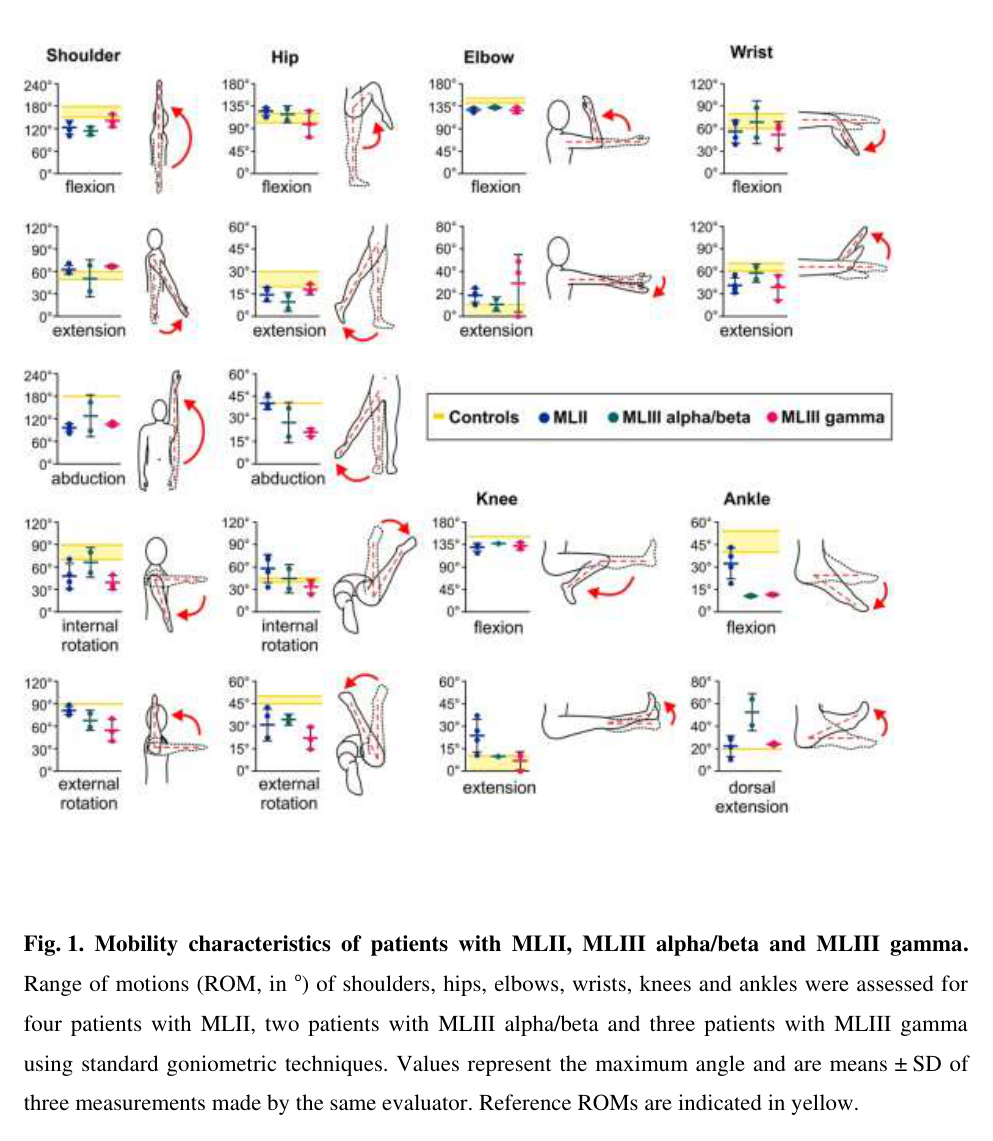
GNPTG-mucolipidosis (mucolipidosis III gamma) is the attenuated form of GlcNAc-1-phosphotransferase deficiency, caused by biallelic variants in GNPTG encoding the gamma subunit of the enzyme that initiates mannose-6-phosphate (M6P) tagging of soluble lysosomal hydrolases. Loss of M6P tagging missorts and hypersecretes lysosomal enzymes, leaving lysosomes functionally deficient and causing slowly progressive substrate accumulation. It is a slowly progressive disorder mainly affecting skeletal, joint, and connective tissues, generally milder than GNPTAB-related mucolipidosis, with survival into adulthood and relatively preserved cognition.
Ask OpenScientist
Ask a research question about GNPTG-Mucolipidosis. OpenScientist will conduct autonomous deep research using the Disorder Mechanisms Knowledge Base and PubMed literature (typically 10-30 minutes).
Do not include personal health information in your question. Questions and results are cached in your browser's local storage.
Classifications
Mappings
Definitions
1Show evidence (1 reference)
Inheritance
1Show evidence (1 reference)
Discussions and Knowledge Gaps
1- Both lines show chondroitin sulfate accumulation and matrix change at the deeper endpoints while both remain unremarkable on histology alone, which would make depth of assay the explanation and the cartilage phenotype a property of gamma-subunit loss rather than of one line.
- The Hamburg line reproduces the 2020 findings and the Lexicon line remains unaffected at the same endpoints and ages, which would make the line difference the explanation and leave the cartilage phenotype unestablished for gamma-subunit loss in general.
- Residual beta-glucuronidase activity in cartilage tracks inversely with cartilage involvement across the lines, supporting a graded-activity threshold above which the tissue is spared.
- Residual beta-glucuronidase activity is comparable in the two lines despite their differing cartilage findings, which would eliminate the 2009 authors' account and leave line provenance, assay depth or animal age carrying the discrepancy.
Show evidence (3 references)
Pathophysiology
2Show evidence (2 references)
Show evidence (1 reference)
Pathograph
Phenotypes
9Cardiovascular 2
Show evidence (1 reference)
Show evidence (1 reference)
Musculoskeletal 2
Show evidence (1 reference)
Show evidence (1 reference)
Nervous System 3
Show evidence (1 reference)
Show evidence (1 reference)
Show evidence (1 reference)
Respiratory 1
Show evidence (1 reference)
Constitutional 1
Show evidence (1 reference)
Genetic Associations
1Show evidence (1 reference)
Medical Actions
4Show evidence (1 reference)
Show evidence (1 reference)
Show evidence (1 reference)
Show evidence (1 reference)
Diagnosis
2Show evidence (1 reference)
Show evidence (1 reference)
Differential Diagnoses
1Conditions with similar clinical presentations that must be differentiated from GNPTG-Mucolipidosis:
- GNPTAB defects with severe infantile-onset disease and death in early childhood, versus the attenuated GNPTG form.
Show evidence (1 reference)
Animal Models
2Show evidence (1 reference)
Show evidence (1 reference)
Show evidence (1 reference)
Show evidence (1 reference)
Show evidence (1 reference)
Notes
Show evidence (1 reference)
Notes
Source YAML
click to showname: GNPTG-Mucolipidosis
creation_date: "2026-06-13T00:00:00Z"
description: >-
GNPTG-mucolipidosis (mucolipidosis III gamma) is the attenuated form of
GlcNAc-1-phosphotransferase deficiency, caused by biallelic variants in GNPTG encoding the
gamma subunit of the enzyme that initiates mannose-6-phosphate (M6P) tagging of soluble
lysosomal hydrolases. Loss of M6P tagging missorts and hypersecretes lysosomal enzymes,
leaving lysosomes functionally deficient and causing slowly progressive substrate
accumulation. It is a slowly progressive disorder mainly affecting skeletal, joint, and
connective tissues, generally milder than GNPTAB-related mucolipidosis, with survival into
adulthood and relatively preserved cognition.
category: Mendelian
disease_term:
preferred_term: GNPTG-mucolipidosis
term:
id: MONDO:0009652
label: GNPTG-mucolipidosis
classifications:
isds_skeletal_category:
- classification_value: lysosomal_storage_with_skeletal_involvement
notes: >-
ISDS Nosology and Classification of Genetic Skeletal Disorders, 2019
revision (Mortier et al., PMID:31633310), Table 1 group 27 "Lysosomal
storage diseases with skeletal involvement (dysostosis multiplex group)";
listed as "Mucolipidosis III (pseudo-Hurler polydystrophy), gamma type".
mappings:
mondo_mappings:
- term:
id: MONDO:0009652
label: GNPTG-mucolipidosis
mapping_predicate: skos:exactMatch
mapping_source: MONDO
mapping_justification: Primary MONDO disease identifier for this mucolipidosis III gamma entry.
references:
- reference: PMID:20301784
title: "Mucolipidosis III Gamma."
tags:
- GeneReviews
synonyms:
- Mucolipidosis III gamma
- Mucolipidosis type IIIC
- Pseudo-Hurler polydystrophy, gamma
- GNPTG deficiency
parents:
- Lysosomal Storage Disorder
pathophysiology:
- name: GlcNAc-1-Phosphotransferase Gamma-Subunit Deficiency and M6P Targeting Failure
conforms_to: "lysosomal_substrate_accumulation#Lysosomal Hydrolase or Cofactor Deficiency"
biological_scale: MOLECULAR
description: >-
Biallelic GNPTG variants impair the gamma subunit of GlcNAc-1-phosphotransferase,
reducing mannose-6-phosphate tagging of soluble acid hydrolases. Untagged enzymes are
missorted and hypersecreted into plasma rather than delivered to the lysosome, leaving
the lysosome functionally deficient in multiple hydrolases.
gene:
preferred_term: GNPTG
term:
id: hgnc:23026
label: GNPTG
biological_processes:
- preferred_term: protein targeting to lysosome
term:
id: GO:0006622
label: protein targeting to lysosome
modifier: DECREASED
cell_types:
- preferred_term: fibroblast
term:
id: CL:0000057
label: fibroblast
evidence:
- reference: PMID:34172897
reference_title: "Mucolipidosis type II and type III: a systematic review of 843 published cases."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "Mucolipidosis (ML) II, MLIII alpha/beta, and MLIII gamma are rare\nautosomal recessive lysosomal storage disorders."
explanation: "Mucolipidosis III gamma is a lysosomal storage disorder in the GlcNAc-1-phosphotransferase spectrum."
- reference: PMID:30882951
reference_title: "The lysosomal storage disorders mucolipidosis type II, type III alpha/beta, and type III gamma: Update on GNPTAB and GNPTG mutations."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "MLIII alpha/beta and MLIII gamma are\nattenuated forms of the disease"
explanation: "GNPTG-related ML III gamma is an attenuated form within the GNPTAB/GNPTG disease spectrum."
downstream:
- target: Lysosomal Substrate Accumulation in Connective Tissue
description: Functional lysosomal hydrolase deficiency leads to substrate accumulation.
- name: Lysosomal Substrate Accumulation in Connective Tissue
conforms_to: "lysosomal_substrate_accumulation#Lysosomal Substrate Accumulation"
biological_scale: TISSUE
description: >-
Functional deficiency of multiple lysosomal hydrolases leads to slowly progressive
accumulation of undegraded substrates, predominantly affecting skeletal, joint, and
connective tissues.
biological_processes:
- preferred_term: lysosomal transport
term:
id: GO:0007041
label: lysosomal transport
modifier: ABNORMAL
cell_types:
- preferred_term: fibroblast
term:
id: CL:0000057
label: fibroblast
cellular_components:
- preferred_term: lysosome
term:
id: GO:0005764
label: lysosome
evidence:
- reference: PMID:20301784
reference_title: "Mucolipidosis III Gamma."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "Mucolipidosis III gamma (ML IIIγ) is a slowly progressive inborn error of metabolism mainly affecting skeletal, joint, and connective tissues."
explanation: "Substrate accumulation predominantly affects skeletal, joint, and connective tissues."
downstream:
- target: Skeletal dysplasia
description: Connective-tissue and skeletal lysosomal storage produces progressive skeletal dysplasia.
- target: Limitation of joint mobility
description: Joint and connective-tissue involvement restricts mobility.
- target: Chronic pain
description: Progressive skeletal and joint disease produces chronic pain and functional impairment.
- target: Cardiac valve disease
description: Connective-tissue storage affects cardiac valves, producing valvular thickening and insufficiency.
- target: Carpal tunnel syndrome
description: Connective-tissue thickening can compress the median nerve in the carpal tunnel.
phenotypes:
- name: Skeletal dysplasia
description: Progressive skeletal dysplasia (dysostosis) affecting the skeleton.
phenotype_term:
preferred_term: Skeletal dysplasia
term:
id: HP:0002652
label: Skeletal dysplasia
evidence:
- reference: PMID:20301784
reference_title: "Mucolipidosis III Gamma."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "Mucolipidosis III gamma (ML IIIγ) is a slowly progressive inborn error of metabolism mainly affecting skeletal, joint, and connective tissues."
explanation: "Skeletal involvement is a major feature."
- name: Limitation of joint mobility
description: Progressive joint stiffness and restriction of movement.
phenotype_term:
preferred_term: Limitation of joint mobility
term:
id: HP:0001376
label: Limitation of joint mobility
evidence:
- reference: PMID:20301784
reference_title: "Mucolipidosis III Gamma."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "mainly affecting skeletal, joint, and connective tissues"
explanation: "Joint involvement with progressive restriction is characteristic."
- name: Chronic pain
description: Significant morbidity from chronic pain with severe functional impairment.
phenotype_term:
preferred_term: Chronic pain
term:
id: HP:0012532
label: Chronic pain
evidence:
- reference: PMID:20301784
reference_title: "Mucolipidosis III Gamma."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "severe functional impairment and significant morbidity from chronic\npain"
explanation: Chronic pain is a major source of morbidity in ML III gamma.
- name: Cardiac valve disease
description: Thickening and insufficiency of the mitral and aortic valves, usually mild.
phenotype_term:
preferred_term: Mitral regurgitation
term:
id: HP:0001653
label: Mitral regurgitation
evidence:
- reference: PMID:20301784
reference_title: "Mucolipidosis III Gamma."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "thickening and insufficiency of the mitral and aortic valves"
explanation: Cardiac valve thickening and insufficiency occur in ML III gamma.
- name: Carpal tunnel syndrome
description: Carpal tunnel syndrome may require surgical release.
phenotype_term:
preferred_term: Constrictive median neuropathy
term:
id: HP:0012185
label: Constrictive median neuropathy
evidence:
- reference: PMID:20301784
reference_title: "Mucolipidosis III Gamma."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "carpal tunnel and, rarely, tarsal tunnel syndrome may\nrequire surgica"
explanation: Carpal tunnel syndrome is a recognized complication requiring surgical management.
- name: Restrictive lung disease
description: >
Restrictive lung disease arising from thoracic (chest wall/skeletal)
involvement; a rarely clinically significant cardiorespiratory
complication.
phenotype_term:
preferred_term: Restrictive ventilatory defect
term:
id: HP:0002091
label: Restrictive ventilatory defect
evidence:
- reference: PMID:20301784
reference_title: "Mucolipidosis III Gamma."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "restrictive lung disease from thoracic"
explanation: GeneReviews reports restrictive lung disease from thoracic involvement as a cardiorespiratory complication of ML III gamma.
- name: Aortic valve insufficiency
description: >
Thickening and insufficiency (regurgitation) of the aortic valve;
accompanies the mitral valve involvement and is usually mild.
phenotype_term:
preferred_term: Aortic regurgitation
term:
id: HP:0001659
label: Aortic regurgitation
evidence:
- reference: PMID:20301784
reference_title: "Mucolipidosis III Gamma."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "thickening and insufficiency of the mitral and aortic valves"
explanation: GeneReviews reports thickening and insufficiency of the aortic (as well as mitral) valve in ML III gamma.
- name: Motor delay
description: >
Delayed acquisition of motor milestones secondary to progressive
joint disease.
phenotype_term:
preferred_term: Motor delay
term:
id: HP:0001270
label: Motor delay
evidence:
- reference: PMID:20301784
reference_title: "Mucolipidosis III Gamma."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "Motor milestones can be affected due to joint"
explanation: GeneReviews attributes affected motor milestones to joint disease in ML III gamma.
- name: Cognitive impairment
description: >
Mild cognitive impairment reported in a small minority (probably <5%)
of affected individuals.
phenotype_term:
preferred_term: Cognitive impairment
term:
id: HP:0100543
label: Cognitive impairment
frequency: VERY_RARE
evidence:
- reference: PMID:20301784
reference_title: "Mucolipidosis III Gamma."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "A few (probably <5%) affected individuals display mild cognitive"
explanation: GeneReviews reports mild cognitive impairment in a minority (<5%) of individuals with ML III gamma.
inheritance:
- name: Autosomal recessive
inheritance_term:
preferred_term: Autosomal recessive inheritance
term:
id: HP:0000007
label: Autosomal recessive inheritance
evidence:
- reference: PMID:34172897
reference_title: "Mucolipidosis type II and type III: a systematic review of 843 published cases."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "rare\nautosomal recessive lysosomal storage disorders"
explanation: "Mucolipidosis III gamma is autosomal recessive."
genetic:
- name: GNPTG
association: Biallelic GNPTG variants impairing the GlcNAc-1-phosphotransferase gamma subunit
relationship_type: CAUSATIVE
variant_origin: GERMLINE
gene_term:
preferred_term: GNPTG
term:
id: hgnc:23026
label: GNPTG
evidence:
- reference: PMID:30882951
reference_title: "The lysosomal storage disorders mucolipidosis type II, type III alpha/beta, and type III gamma: Update on GNPTAB and GNPTG mutations."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "GNPTAB and GNPTG encode the α/β-precursor and the\nγ-subunit of N-acetylglucosamine (GlcNAc)-1-phosphotransferase, respectively"
explanation: "GNPTG encodes the gamma subunit of GlcNAc-1-phosphotransferase; its biallelic loss causes ML III gamma."
diagnosis:
- name: Plasma lysosomal enzyme activity and GNPTG sequencing
diagnosis_term:
preferred_term: clinical laboratory procedure
term:
id: NCIT:C25294
label: Laboratory Procedure
description: >-
Markedly elevated activities of multiple lysosomal enzymes in plasma/serum (from
hypersecretion) with reduced intracellular activity suggests a mannose-6-phosphate
targeting defect; diagnosis is confirmed by GNPTG sequencing.
markers: Elevated plasma lysosomal enzyme activities with reduced intracellular activity.
evidence:
- reference: PMID:20301784
reference_title: "Mucolipidosis III Gamma."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "The diagnosis of ML IIIγ is established in a proband with\nsuggestive clinical and radiographic findings and biallelic pathogenic variants\nin GNPTG identified by molecular genetic testing"
explanation: "Diagnosis combines suggestive clinical/radiographic findings with biallelic GNPTG variants on molecular testing."
- name: GNPTG molecular genetic testing
diagnosis_term:
preferred_term: genetic testing
term:
id: NCIT:C15709
label: Genetic Testing
description: Confirmatory biallelic GNPTG sequencing.
evidence:
- reference: PMID:20301784
reference_title: "Mucolipidosis III Gamma."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "biallelic pathogenic variants\nin GNPTG identified by molecular genetic testing"
explanation: "Biallelic GNPTG variants on molecular genetic testing provide diagnostic confirmation."
differential_diagnoses:
- name: Mucolipidosis type II
description: >-
The severe end of the GlcNAc-1-phosphotransferase spectrum (GNPTAB), with infantile onset
and early death.
disease_term:
preferred_term: mucolipidosis type II
term:
id: MONDO:0009650
label: mucolipidosis type II
distinguishing_features:
- GNPTAB defects with severe infantile-onset disease and death in early childhood, versus the attenuated GNPTG form.
evidence:
- reference: PMID:34172897
reference_title: "Mucolipidosis type II and type III: a systematic review of 843 published cases."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "Mucolipidosis (ML) II, MLIII alpha/beta, and MLIII gamma are rare\nautosomal recessive lysosomal storage disorders."
explanation: "ML II and ML III gamma are distinct severities within the same phosphotransferase spectrum."
treatments:
- name: Supportive Care
description: >-
No disease-modifying therapy is established; management is supportive, with orthopedic
and physiotherapeutic care for the musculoskeletal disease.
treatment_term:
preferred_term: Supportive Care
term:
id: NCIT:C15747
label: Supportive Care
evidence:
- reference: PMID:20301784
reference_title: "Mucolipidosis III Gamma."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "Braces (especially of the hands) at night may\nimprove daily functions"
explanation: Supportive measures such as night braces improve daily function in ML III gamma.
- name: Bisphosphonate Therapy
description: >-
Bisphosphonate treatment is used in individuals with significant skeletal disease and
bone pain.
treatment_term:
preferred_term: Pharmacotherapy
term:
id: NCIT:C15986
label: Pharmacotherapy
evidence:
- reference: PMID:20301784
reference_title: "Mucolipidosis III Gamma."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "Bisphosphonate treatment in individuals with significant\nskeletal dis"
explanation: Bisphosphonates are used for the significant skeletal disease and bone pain.
- name: Joint Replacement Surgery
description: >-
Joint replacement (especially hip and knee) relieves pain and improves function in older
adolescents and adults.
treatment_term:
preferred_term: joint replacement
term:
id: NCIT:C131579
label: Joint Replacement
evidence:
- reference: PMID:20301784
reference_title: "Mucolipidosis III Gamma."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "joint replacement has\nbeen successful in relieving hip pain and knee"
explanation: Joint replacement successfully relieves hip and knee pain in ML III gamma.
- name: Pain Management
description: >-
As skeletal disease intensifies, management focuses on pain relief, including for
chronic pain.
treatment_term:
preferred_term: supportive care
term:
id: NCIT:C15747
label: Supportive Care
evidence:
- reference: PMID:20301784
reference_title: "Mucolipidosis III Gamma."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "management\nfocuses on pain relief"
explanation: Pain relief is a central component of ML III gamma management.
definitions:
- name: Clinical case definition of mucolipidosis III gamma
definition_type: CASE_DEFINITION
description: >-
Mucolipidosis III gamma is the attenuated GlcNAc-1-phosphotransferase deficiency caused by
biallelic GNPTG variants, defined by a mannose-6-phosphate targeting defect with slowly
progressive skeletal, joint, and connective-tissue disease.
scope: Disease-level case definition for GNPTG-mucolipidosis (ML III gamma).
evidence:
- reference: PMID:20301784
reference_title: "Mucolipidosis III Gamma."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "Mucolipidosis III gamma (ML IIIγ) is a slowly progressive inborn error of metabolism mainly affecting skeletal, joint, and connective tissues."
explanation: "Anchors the case definition in the GNPTG-related connective-tissue mucolipidosis."
animal_models:
- name: Gnptg-null mouse, Lexicon line (Gnptg tm1Lex)
species: Mouse
genotype: Gnptg<tm1Lex>/Gnptg<tm1Lex>
alleles:
- Gnptg<tm1Lex>
genes:
- preferred_term: GNPTG
term:
id: hgnc:23026
label: GNPTG
publication: PMID:19261645
description: >-
Knockout of the GlcNAc-1-phosphotransferase gamma subunit, studied alongside
the alpha/beta-subunit knockout that models mucolipidosis II. The pairing is
what makes this model informative: losing the gamma subunit causes only a
partial loss of mannose-6-phosphate tagging, where losing alpha/beta abolishes
it, and the severity of storage disease in the two lines tracks the residual
enzyme activity. That graded relationship is the mouse counterpart of the
milder human ML III gamma against ML II.
notes: >-
MGI records Gnptg<tm1Lex> (MGI:3757947, Lexicon Genetics) with PMID:17652091
as its originating reference and annotates that paper to this genotype, so
the phosphorylation measurement below is allele-confirmed. MGI carries no
annotation for PMID:19261645; that paper is assigned to this line on author
and allele-series continuity with PMID:17652091 rather than on an MGI record,
and the assignment should be treated as such. A separate gamma-null line from
a different group is curated below, and the two disagree about cartilage.
modeled_mechanisms:
- target: GlcNAc-1-Phosphotransferase Gamma-Subunit Deficiency and M6P Targeting
Failure
relationship: RECAPITULATES
fidelity: HIGH
model_scale: ORGANISM
description: >-
Germline loss of the orthologous subunit, with the partial rather than
complete tagging defect that distinguishes gamma-subunit disease from
alpha/beta-subunit disease.
limitations: >-
A complete gamma-subunit null models the severe end of the GNPTG allelic
spectrum, whereas most human ML III gamma alleles retain some subunit
function.
readouts:
- name: Mannose 6-phosphorylation of acid hydrolases
target: GlcNAc-1-Phosphotransferase Gamma-Subunit Deficiency and M6P Targeting
Failure
direction: DECREASED
interpretation: >-
Partial, not abolished. The same study measured complete loss in the
alpha/beta knockout, so this readout is the quantitative difference
between ML III gamma and ML II rather than a shared defect.
evidence:
- reference: PMID:17652091
reference_title: 'Murine UDP-GlcNAc:lysosomal enzyme N-acetylglucosamine-1-phosphotransferase
lacking the gamma-subunit retains substantial activity toward acid hydrolases.'
supports: SUPPORT
evidence_source: MODEL_ORGANISM
snippet: disruption of the alpha/beta gene completely abolishes phosphorylation
of high mannose oligosaccharides on acid hydrolases whereas knock-out of
the gamma gene results in only a partial loss of phosphorylation
explanation: Measures the residual phosphotransferase activity in this genotype
against the alpha/beta null assayed in the same study.
- name: Serum lysosomal enzyme levels
target: GlcNAc-1-Phosphotransferase Gamma-Subunit Deficiency and M6P Targeting
Failure
direction: INCREASED
interpretation: >-
Enzyme amount rising in the wrong compartment because the tagging that
routes hydrolases to the lysosome is impaired. It must not be read as
preserved lysosomal function.
evidence:
- reference: PMID:19261645
reference_title: Comparative pathology of murine mucolipidosis types II and
IIIC.
supports: SUPPORT
evidence_source: MODEL_ORGANISM
snippet: We found that both lines of mutant mice had elevated levels of serum
lysosomal enzymes and cytoplasmic alterations in secretory cells of several
exocrine glands; however, lesions in gamma-subunit deficient (Gnptg(-/-))
mice were milder and more restricted in distribution than in alpha/beta-subunit
deficient (Gnptab(-/-)) mice.
explanation: Reports the raised circulating hydrolases and the milder, more
restricted lesion distribution relative to the alpha/beta null.
evidence:
- reference: PMID:19261645
reference_title: Comparative pathology of murine mucolipidosis types II and IIIC.
supports: SUPPORT
evidence_source: MODEL_ORGANISM
snippet: The milder disease in the gamma-subunit deficient mice correlated with
residual synthesis of the mannose 6-phosphate recognition marker.
explanation: Ties the milder phenotype directly to residual M6P tagging, which
is the mechanism this node names.
- target: Lysosomal Substrate Accumulation in Connective Tissue
relationship: PARTIALLY_RECAPITULATES
fidelity: LOW
model_scale: ORGANISM
description: >-
Storage develops, but not in the tissue compartment that defines the human
disease.
limitations: >-
Neither this line nor the alpha/beta line develops the cytoplasmic vacuolar
inclusions in fibrocytes and mesenchymal cells - the I-cells - that
characterise the skeletal and connective-tissue disease in human ML II and
ML III. The predominant murine lesion is in exocrine secretory epithelium
instead, which is not where the human disease is felt. This line is also
reported to lack cartilage defects outright, which a second gamma-null line
contradicts; see the cartilage controversy recorded in this entry.
Storage-burden or connective-tissue endpoints should not be transferred from
this model to patients.
readouts:
- name: Cytoplasmic vacuolar inclusions in fibrocytes and mesenchymal cells (I-cells)
target: Lysosomal Substrate Accumulation in Connective Tissue
direction: UNCHANGED
interpretation: >-
A negative result, and the reason this link is PARTIALLY_RECAPITULATES at
LOW fidelity. The lesion that names the human disease does not form in
either mouse line.
evidence:
- reference: PMID:19261645
reference_title: Comparative pathology of murine mucolipidosis types II and
IIIC.
supports: SUPPORT
evidence_source: MODEL_ORGANISM
snippet: Of significance, neither strain of mutant mice developed cytoplasmic
vacuolar inclusions in fibrocytes or mesenchymal cells (I-cells), the characteristic
lesion associated with the prominent skeletal and connective tissue abnormalities
in humans with MLII and MLIII.
explanation: Documents the absent I-cell lesion in both knockout lines.
evidence:
- reference: PMID:19261645
reference_title: Comparative pathology of murine mucolipidosis types II and IIIC.
supports: SUPPORT
evidence_source: MODEL_ORGANISM
snippet: We conclude that mice lacking either alpha/beta or gamma subunits displayed
clinical and pathologic features that differed substantially from those reported
in humans having mutations in orthologous genes.
explanation: The authors' own bound on how far either mouse line stands in for
the human connective-tissue disease.
- name: Gnptg-null mouse, Hamburg line (Gnptg ko)
species: Mouse
genotype: Gnptg ko, homozygous
background: C57BL/6
genes:
- preferred_term: GNPTG
term:
id: hgnc:23026
label: GNPTG
publication: PMID:33023972
description: >-
A second, independently derived gamma-subunit null line, curated separately
from the Lexicon line above because it is a different mouse. The reporting
paper's Animals section states that its Gnptg ko (C57Bl/6) mice "have been
described previously (Kollmann et al., 2012, 2013; Di Lorenzo et al., 2018)",
a provenance distinct from the Lexicon allele. It is the line in which the
cartilage and joint phenotype was found.
notes: >-
The allele is not named in the paper's abstract, and the full text is not in
the local reference cache, so the line is recorded as the authors write it
rather than by MGI allele symbol. MGI lists Gnptg<tm1d(KOMP)Wtsi>
(MGI:6270380) with the synonym Gnptg<ko> and Di Lorenzo 2018 (PMID:29773673)
as its reference, which is the plausible MGI identity for this line, but the
match is not asserted here because it has not been confirmed against the
paper.
modeled_mechanisms:
- target: Lysosomal Substrate Accumulation in Connective Tissue
relationship: PARTIALLY_RECAPITULATES
fidelity: MODERATE
model_scale: ORGANISM
description: >-
Substrate accumulates inside chondrocytes and the cartilage matrix is
altered, which is the connective-tissue storage this node names.
limitations: >-
Chondrocytes and tendon were assayed; the I-cell inclusions in fibrocytes
and mesenchymal cells that define the human lesion were not reported here
and were absent in the Lexicon line. The Lexicon line is reported to lack
cartilage defects altogether, so this readout is contradicted by the other
gamma-null model in this entry.
readouts:
- name: Chondrocyte chondroitin sulfate content and differentiation
target: Lysosomal Substrate Accumulation in Connective Tissue
direction: INCREASED
interpretation: >-
Substrate accumulating inside the cell type that builds cartilage, with
differentiation impaired alongside it.
evidence:
- reference: PMID:33023972
reference_title: Imbalanced cellular metabolism compromises cartilage homeostasis
and joint function in a mouse model of mucolipidosis type III gamma.
supports: SUPPORT
evidence_source: MODEL_ORGANISM
snippet: missorting of a number of lysosomal enzymes is associated with intracellular
accumulation of chondroitin sulfate in Gnptgko chondrocytes and their impaired
differentiation, as well as with altered microstructure of the cartilage
extracellular matrix (ECM)
explanation: Reports the substrate accumulation and matrix change in cartilage
in this line.
- target: Chronic pain
relationship: PARTIALLY_RECAPITULATES
fidelity: LOW
model_scale: ORGANISM
description: >-
A functional joint deficit that the authors attribute to fatigue or pain at
the joint, which is the closest available model correlate of the chronic
joint pain reported by patients.
limitations: >-
Rotarod performance under fatigue is a composite motor readout, not a pain
measurement. The authors offer fatigue or pain as alternative explanations
and do not separate them, and no nociceptive assay was performed, so this
link rests on the authors' interpretation rather than on a direct measure of
pain.
readouts:
- name: Limb motor performance on rotarod under fatigue
target: Chronic pain
direction: DECREASED
interpretation: >-
Reduced endurance on the rotarod under fatigue conditions, taken as a
whole-animal functional measure of how well the joint bears sustained
load.
evidence:
- reference: PMID:33023972
reference_title: Imbalanced cellular metabolism compromises cartilage homeostasis
and joint function in a mouse model of mucolipidosis type III gamma.
supports: SUPPORT
evidence_source: MODEL_ORGANISM
snippet: This idea was supported by a deficit of the limb motor function in
Gnptgko mice challenged on a rotarod under fatigue-associated conditions,
suggesting that the impaired motor performance of Gnptgko mice was caused
by fatigue and/or pain at the joint.
explanation: Reports the motor deficit and the authors' attribution of it to
fatigue or joint pain.
evidence:
- reference: PMID:33023972
reference_title: Imbalanced cellular metabolism compromises cartilage homeostasis
and joint function in a mouse model of mucolipidosis type III gamma.
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: Patients with MLIII gamma typically present with osteoarthritis and
joint stiffness, suggesting cartilage involvement.
explanation: States the human joint phenotype this model is being used to represent.
discussions:
- discussion_id: gnptg_mouse_cartilage_conflict
kind: CONTROVERSY
status: OPEN
prompt: Do Gnptg-null mice have a cartilage phenotype?
attaches_to:
- animal_models#Gnptg-null mouse, Lexicon line (Gnptg tm1Lex)
- animal_models#Gnptg-null mouse, Hamburg line (Gnptg ko)
- pathophysiology#Lysosomal Substrate Accumulation in Connective Tissue
rationale: >-
Two reports of gamma-subunit null mice reach opposite conclusions. The 2009
comparative pathology study reports that the gamma-null mice "lacked cartilage
defects", and uses that absence, with the absent retinal degeneration, to argue
the line is milder than the alpha/beta knockout. The 2020 study reports
intracellular chondroitin sulfate accumulation in Gnptg knockout chondrocytes,
impaired chondrocyte differentiation, and altered cartilage extracellular
matrix microstructure. Four candidate explanations are on the table and none
has been tested against the others. First, the two papers do not use the same
mouse: the 2020 Animals section attributes its line to Kollmann and Di Lorenzo,
a different provenance from the Lexicon allele carrying the 2009 result, so
line and background differences are a live explanation rather than a
formality. Second, depth of assay differs - the earlier study read histology
and gross phenotype, the later one measured matrix ultrastructure, substrate
content and chondrocyte differentiation, so a defect invisible at one
resolution may be plain at another. Third, the 2009 authors offer their own
mechanistic account, attributing the absent chondrocyte and retinal lesions to
residual beta-glucuronidase activity, which is a residual-activity explanation
of the same kind that organises the rest of this entry. Fourth, the animals may
not be of comparable age: the 2020 work reports on aged joints and tendon,
while the 2009 abstract does not state its timepoints. Recorded so the entry
does not silently adopt one report over the other.
evidence:
- reference: PMID:19261645
reference_title: Comparative pathology of murine mucolipidosis types II and IIIC.
supports: REFUTE
evidence_source: MODEL_ORGANISM
snippet: In contrast to mice deficient in the alpha/beta subunits, the mice lacking
the gamma subunits were of normal size, lacked cartilage defects, and did not
develop retinal degeneration.
explanation: The report that the Lexicon line has no cartilage defect.
- reference: PMID:19261645
reference_title: Comparative pathology of murine mucolipidosis types II and IIIC.
supports: REFUTE
evidence_source: MODEL_ORGANISM
snippet: The absence of retinal and chondrocyte lesions in Gnptg(-/-) mice might
be attributed to residual beta-glucuronidase activity.
explanation: The sentence presupposes the absent chondrocyte lesions in order
to explain them, so it asserts the absence as well as accounting for it. Its
distinctive content is that account - residual beta-glucuronidase activity -
which the rationale carries as one of the four candidate explanations.
- reference: PMID:33023972
reference_title: Imbalanced cellular metabolism compromises cartilage homeostasis
and joint function in a mouse model of mucolipidosis type III gamma.
supports: SUPPORT
evidence_source: MODEL_ORGANISM
snippet: missorting of a number of lysosomal enzymes is associated with intracellular
accumulation of chondroitin sulfate in Gnptgko chondrocytes and their impaired
differentiation, as well as with altered microstructure of the cartilage extracellular
matrix (ECM)
explanation: The report that a gamma-null line does have a cartilage phenotype.
proposed_experiments:
- experiment_id: gnptg_cartilage_head_to_head_matched_age
name: Both gamma-null lines assayed side by side at matched ages with the deeper
cartilage endpoints
description: >-
Take the Lexicon line and the Hamburg line into one experiment and apply the
2020 endpoints to both at the same ages: chondrocyte chondroitin sulfate
content, chondrocyte differentiation markers, and cartilage extracellular
matrix ultrastructure, alongside the histology and gross skeletal assessment
the 2009 study used. This is the one design that separates the first two
candidate explanations from each other, because a line difference and a
depth-of-assay difference make opposite predictions once line and assay are
varied independently.
would_support:
- pathophysiology#Lysosomal Substrate Accumulation in Connective Tissue
supporting_outcome:
- >-
Both lines show chondroitin sulfate accumulation and matrix change at the
deeper endpoints while both remain unremarkable on histology alone, which
would make depth of assay the explanation and the cartilage phenotype a
property of gamma-subunit loss rather than of one line.
would_refute:
- pathophysiology#Lysosomal Substrate Accumulation in Connective Tissue
refuting_outcome:
- >-
The Hamburg line reproduces the 2020 findings and the Lexicon line remains
unaffected at the same endpoints and ages, which would make the line
difference the explanation and leave the cartilage phenotype unestablished
for gamma-subunit loss in general.
- experiment_id: gnptg_residual_beta_glucuronidase_in_cartilage
name: Residual beta-glucuronidase activity in cartilage of both gamma-null lines
description: >-
Measure beta-glucuronidase activity in chondrocytes and cartilage of both
lines against wild-type and against the alpha/beta knockout, to test the 2009
authors' own account of the absence they report. The claim is currently an
untested hypothesis offered in a discussion sentence, and it is the candidate
explanation most directly checkable, since the residual activity it invokes
is a quantity rather than a difference in husbandry or method.
supporting_outcome:
- >-
Residual beta-glucuronidase activity in cartilage tracks inversely with
cartilage involvement across the lines, supporting a graded-activity
threshold above which the tissue is spared.
refuting_outcome:
- >-
Residual beta-glucuronidase activity is comparable in the two lines despite
their differing cartilage findings, which would eliminate the 2009 authors'
account and leave line provenance, assay depth or animal age carrying the
discrepancy.
References & Deep Research
References
1Deep Research
1Deep research results are used as seeds for research; they do not undergo the same validation as the main records and may contain errors. How we use deep research.
Question: You are an expert researcher providing comprehensive, well-cited information.
Provide detailed information focusing on: 1. Key concepts and definitions with current understanding 2. Recent developments and latest research (prioritize 2023-2024 sources) 3. Current applications and real-world implementations 4. Expert opinions and analysis from authoritative sources 5. Relevant statistics and data from recent studies
Format as a comprehensive research report with proper citations. Include URLs and publication dates where available. Always prioritize recent, authoritative sources and provide specific citations for all major claims.
Disease Characteristics Research Template
Target Disease
- Disease Name: GNPTG-Mucolipidosis
- MONDO ID: (if available)
- Category: Mendelian
Research Objectives
Please provide a comprehensive research report on GNPTG-Mucolipidosis covering all of the disease characteristics listed below. This report will be used to populate a disease knowledge base entry. Be thorough and cite primary literature (PMID preferred) for all claims.
For each section, suggested databases/resources are listed. These are the first places you should search for information on each topic.
1. Disease Information
Search first: OMIM, Orphanet, ICD-10/ICD-11, MeSH, PubMed
- What is the disease? Provide a concise overview.
- What are the key identifiers? (OMIM, Orphanet, ICD-10/ICD-11, MeSH, Mondo)
- What are the common synonyms and alternative names?
- Is the information derived from individual patients (e.g., EHR) or aggregated disease-level resources?
2. Etiology
- Disease Causal Factors: What are the primary causes? (genetic, environmental, infectious, mechanistic)
- Risk Factors:
Search first: PubMed, Cochrane Library, UpToDate, clinical guidelines, ClinVar, ClinGen, GWAS Catalog, PheGenI, CTD, CDC, WHO, epidemiological databases
- Genetic risk factors (causal variants, susceptibility loci, modifier genes)
- Environmental risk factors (toxins, lifestyle, occupational exposures, age, sex, family history)
- Protective Factors:
Search first: PubMed, Cochrane Library, clinical trial databases, GWAS Catalog, gnomAD, WHO, CDC, nutrition databases
- Genetic protective factors (protective variants, modifier alleles)
- Environmental protective factors (diet, lifestyle, exposures that reduce risk)
- Gene-Environment Interactions: How do genetic and environmental factors interact to influence disease?
Search first: CTD, PubMed, PheGenI, GxE databases
3. Phenotypes
Search first: HPO (Human Phenotype Ontology), OMIM, Orphanet, PubMed, clinicaltrials.gov, MedDRA, SNOMED CT, DECIPHER, LOINC
For each phenotype, provide: - Phenotype type: symptoms, clinical signs, physical manifestations, behavioral changes, or laboratory abnormalities
For symptoms/signs: HPO, OMIM, Orphanet, PubMed For behavioral changes: HPO, DSM, RDoC (Research Domain Criteria), PubMed For laboratory abnormalities: LOINC, SNOMED CT, LabTests Online, PubMed - Phenotype characteristics: Search first: OMIM, Orphanet, HPO, PubMed - Age of symptom onset (neonatal, childhood, adult-onset, late-onset) - Symptom severity (mild, moderate, severe, variable) - Symptom progression (stable, progressive, episodic, fluctuating) - Frequency among affected individuals (percentage or qualitative) - Quality of life impact: Effects on daily functioning and well-being (per-phenotype when possible) Search first: EQ-5D database, SF-36, WHO QOL databases, PubMed - Suggest HPO (Human Phenotype Ontology) terms for each phenotype
4. Genetic/Molecular Information
- Causal Genes: Gene mutations or chromosomal abnormalities responsible for disease (gene symbols, OMIM IDs)
Search first: OMIM, ClinVar, HGMD, Ensembl, NCBI Gene
- Pathogenic Variants:
- Affected genes (gene symbols, HGNC IDs) > Search first: OMIM, NCBI Gene, Ensembl, HGNC, UniProt, GeneCards
- Variant classification (pathogenic, likely pathogenic, VUS per ACMG/AMP guidelines) > Search first: ClinVar, ClinGen, ACMG/AMP guidelines, VarSome
- Variant type/class (missense, frameshift, nonsense, splice-site, structural)
- Allele frequency in population databases > Search first: gnomAD, 1000 Genomes, ExAC, TOPMed, dbSNP
- Somatic vs germline origin > Search first: COSMIC (somatic), ClinVar, ICGC, TCGA
- Functional consequences (loss of function, gain of function, dominant negative)
- Modifier Genes: Genes that modify disease severity or expression
- Epigenetic Information: DNA methylation, histone modifications, chromatin changes affecting disease
Search first: ENCODE, Roadmap Epigenomics, MethBase, DiseaseMeth
- Chromosomal Abnormalities: Large-scale genetic changes (aneuploidy, translocations, inversions)
Search first: DECIPHER, ClinVar, ECARUCA, UCSC Genome Browser
5. Environmental Information
- Environmental Factors: Non-genetic contributing factors (toxins, radiation, pollution, occupational exposure)
Search first: CTD (Comparative Toxicogenomics Database), TOXNET, PubMed, EPA databases
- Lifestyle Factors: Behavioral factors (smoking, diet, exercise, alcohol consumption)
Search first: CDC databases, WHO, PubMed, NHANES
- Infectious Agents: If applicable, pathogens causing or triggering disease (bacteria, viruses, fungi, parasites)
Search first: NCBI Taxonomy, ViPR, BV-BRC, MicrobeDB, GIDEON
6. Mechanism / Pathophysiology
- Molecular Pathways: Specific signaling cascades or biochemical pathways involved (Wnt, MAPK, mTOR, PI3K-AKT, etc.)
Search first: KEGG, Reactome, WikiPathways, PathBank, BioCyc
- Cellular Processes: Cell-level mechanisms (apoptosis, autophagy, cell cycle dysregulation, inflammation, etc.)
Search first: Gene Ontology (GO), Reactome, KEGG, PubMed
- Protein Dysfunction: How protein structure or function is altered (misfolding, aggregation, loss of function, gain of function)
Search first: UniProt, PDB (Protein Data Bank), InterPro, Pfam, AlphaFold
- Metabolic Changes: Alterations in metabolic processes (energy metabolism, lipid metabolism, amino acid metabolism)
Search first: KEGG, BioCyc, HMDB (Human Metabolome Database), BRENDA
- Immune System Involvement: Role of immune response (autoimmunity, immunodeficiency, chronic inflammation)
Search first: ImmPort, Immunome Database, IEDB, Gene Ontology
- Tissue Damage Mechanisms: How tissues/ are injured (oxidative stress, ischemia, fibrosis, necrosis)
Search first: PubMed, Gene Ontology, Reactome
- Biochemical Abnormalities: Specific molecular defects (enzyme deficiencies, receptor dysfunction, ion channel defects)
Search first: BRENDA, UniProt, KEGG, OMIM, PubMed
- Epigenetic Changes: DNA methylation, histone modifications affecting gene expression in disease
Search first: ENCODE, Roadmap Epigenomics, MethBase, DiseaseMeth
- Molecular Profiling (if available):
- Transcriptomics/gene expression changes > Search first: GEO (Gene Expression Omnibus), ArrayExpress, GTEx, Human Cell Atlas, SRA
- Proteomics findings > Search first: PRIDE, ProteomeXchange, Human Protein Atlas, STRING, BioGRID
- Metabolomics signatures > Search first: MetaboLights, Metabolomics Workbench, HMDB, METLIN
- Lipidomics alterations > Search first: LIPID MAPS, SwissLipids, LipidHome, Metabolomics Workbench
- Genomic structural features > Search first: UCSC Genome Browser, Ensembl, NCBI, dbVar, DGV
- Advanced Technologies (if applicable):
- Single-cell analysis findings (cell-type specific mechanisms, cellular heterogeneity) > Search first: Human Cell Atlas, Single Cell Portal, GEO, CELLxGENE
- Spatial transcriptomics findings > Search first: GEO, Spatial Research, Vizgen, 10x Genomics data
- Multi-omics integration results > Search first: TCGA, ICGC, cBioPortal, LinkedOmics, PubMed
- Functional genomics screens (CRISPR, RNAi) > Search first: DepMap, GenomeRNAi, PubMed, BioGRID ORCS
For each mechanism, describe: - The causal chain from initial trigger to clinical manifestation - Which mechanisms are upstream vs downstream - What cell types and biological processes are involved - Suggest GO terms for biological processes and CL terms for cell types
7. Anatomical Structures Affected
- Organ Level:
- Primary organs directly affected
- Secondary organ involvement (complications, secondary effects)
- Body systems involved (cardiovascular, nervous, digestive, respiratory, endocrine, etc.)
Search first: Uberon, FMA (Foundational Model of Anatomy), OMIM, HPO, ICD-11, MeSH, SNOMED CT
- Tissue and Cell Level:
- Specific tissue types affected (epithelial, connective, muscle, nervous)
- Specific cell populations targeted (with Cell Ontology terms)
Search first: Uberon, Human Protein Atlas, Cell Ontology, Human Cell Atlas, CellMarker, PanglaoDB
- Subcellular Level:
- Cellular compartments involved (mitochondria, nucleus, ER, lysosomes) (with GO Cellular Component terms)
Search first: Gene Ontology (Cellular Component), UniProt, Human Protein Atlas
- Localization:
- Specific anatomical sites (with UBERON terms) > Search first: FMA, Uberon, NeuroNames (for brain), SNOMED CT
- Lateralization (unilateral, bilateral, asymmetric) > Search first: HPO, clinical literature, imaging databases
8. Temporal Development
- Onset:
- Typical age of onset (congenital, pediatric, adult, geriatric)
- Onset pattern (acute, subacute, chronic, insidious)
Search first: OMIM, Orphanet, HPO, PubMed
- Progression:
- Disease stages (early, intermediate, advanced, end-stage) > Search first: Cancer Staging Manual (AJCC), WHO classifications, PubMed
- Progression rate (rapid, slow, variable)
- Disease course pattern (episodic, relapsing-remitting, progressive, stable)
- Disease duration (self-limited, chronic lifelong)
Search first: Disease registries, longitudinal cohort databases, natural history studies, PubMed, Orphanet, OMIM
- Patterns:
- Remission patterns (spontaneous, treatment-induced) > Search first: Clinical trial databases, disease registries, PubMed
- Critical periods (time windows of vulnerability or opportunity for intervention) > Search first: PubMed, developmental biology databases, clinical guidelines
9. Inheritance and Population
- Epidemiology:
- Prevalence (cases per 100,000 at given time)
- Incidence (new cases per 100,000 per year)
Search first: Orphanet, CDC, WHO, GBD (Global Burden of Disease), national registries, SEER, disease registries
- For Genetic Etiology:
- Inheritance pattern (AD, AR, X-linked, mitochondrial, multifactorial, polygenic) > Search first: OMIM, Orphanet, ClinVar, GTR (Genetic Testing Registry)
- Penetrance (complete, incomplete, age-dependent) > Search first: ClinVar, OMIM, PubMed, ClinGen
- Expressivity (variable, consistent) > Search first: OMIM, ClinVar, PubMed
- Genetic anticipation (increasing severity in successive generations) > Search first: OMIM, PubMed (especially for repeat expansion disorders)
- Germline mosaicism > Search first: ClinVar, OMIM, genetic counseling literature, PubMed
- Founder effects (population-specific mutations) > Search first: gnomAD, population genetics databases, PubMed
- Consanguinity role > Search first: OMIM, population studies, genetic counseling resources
- Carrier frequency > Search first: gnomAD, carrier screening databases, GeneReviews, GTR
- Population Demographics:
- Affected populations (ethnic or demographic groups with higher prevalence) > Search first: gnomAD, 1000 Genomes, PAGE Study, PubMed, population registries
- Geographic distribution (endemic areas, regional variation) > Search first: WHO, CDC, GBD, Orphanet, geographic epidemiology databases
- Geographic distribution of specific variants
- Sex ratio (male:female) > Search first: Disease registries, OMIM, PubMed, epidemiological databases
- Age distribution of affected individuals > Search first: CDC, disease registries, SEER, Orphanet
10. Diagnostics
- Clinical Tests:
- Laboratory tests (blood, urine, tissue chemistry, specific enzyme assays) > Search first: LOINC, LabTests Online, PubMed
- Biomarkers (proteins, metabolites, genetic markers, circulating biomarkers) > Search first: FDA Biomarker List, BEST (Biomarkers, EndpointS, and other Tools), PubMed
- Imaging studies (X-ray, CT, MRI, PET, ultrasound) > Search first: RadLex, DICOM, Radiopaedia, imaging databases
- Functional tests (pulmonary function, cardiac stress tests) > Search first: LOINC, clinical guidelines, PubMed
- Electrophysiology (EEG, EMG, ECG, nerve conduction studies) > Search first: LOINC, clinical neurophysiology databases, PubMed
- Biopsy findings (histopathology, immunohistochemistry) > Search first: SNOMED CT, College of American Pathologists resources, PubMed
- Pathology findings (microscopic examination) > Search first: SNOMED CT, Digital Pathology databases, PubMed
- Genetic Testing:
Search first: GTR (Genetic Testing Registry), GeneReviews, ClinGen
- Overview of recommended genetic testing approach
- Whole genome sequencing (WGS) utility > Search first: GTR, ClinVar, GEL (Genomics England), gnomAD
- Whole exome sequencing (WES) utility > Search first: GTR, ClinVar, OMIM, GeneMatcher
- Gene panels (which panels, which genes) > Search first: GTR, ClinVar, laboratory-specific databases
- Single gene testing > Search first: GTR, ClinVar, OMIM, GeneReviews
- Chromosomal microarray (CMA) > Search first: DECIPHER, ClinVar, dbVar, ECARUCA
- Karyotyping > Search first: Chromosome Abnormality Database, ClinVar, cytogenetics resources
- FISH > Search first: ClinVar, cytogenetics databases, PubMed
- Mitochondrial DNA testing > Search first: MITOMAP, MSeqDR, ClinVar, GTR
- Repeat expansion testing > Search first: GTR, ClinVar, repeat expansion databases, PubMed
- Omics-Based Diagnostics (if applicable):
- RNA sequencing / transcriptomics > Search first: GEO, ArrayExpress, GTEx, RNA-seq databases
- Proteomics > Search first: PRIDE, ProteomeXchange, FDA Biomarker database
- Metabolomics > Search first: MetaboLights, Metabolomics Workbench, HMDB
- Epigenomics > Search first: GEO, ENCODE, Roadmap Epigenomics, MethBase
- Liquid biopsy > Search first: COSMIC, ClinVar, liquid biopsy databases, PubMed
- Clinical Criteria:
- Standardized diagnostic criteria (DSM, ICD, society guidelines) > Search first: DSM-5, ICD-11, clinical society guidelines, UpToDate
- Differential diagnosis (other conditions to rule out, with distinguishing features) > Search first: DynaMed, UpToDate, clinical decision support systems
- Screening:
- Screening methods for asymptomatic individuals (newborn screening, carrier screening, cascade screening) > Search first: ACMG recommendations, CDC newborn screening, GTR
11. Outcome/Prognosis
- Survival and Mortality:
- Survival rate (5-year, 10-year, overall) > Search first: SEER, cancer registries, disease-specific registries, PubMed
- Life expectancy (with and without treatment if applicable) > Search first: Orphanet, disease registries, actuarial databases, PubMed
- Mortality rate > Search first: CDC, WHO, GBD, national mortality databases
- Disease-specific mortality (deaths directly attributable to disease) > Search first: Disease registries, CDC Wonder, GBD, PubMed
- Morbidity and Function:
- Morbidity (disease-related disability and health impacts) > Search first: GBD, WHO, disability databases, PubMed
- Disability outcomes (long-term functional impairments) > Search first: ICF (International Classification of Functioning), disability registries
- Quality of life measures (EQ-5D, SF-36, PROMIS, disease-specific tools) > Search first: EQ-5D database, SF-36, PROMIS, PubMed
- Disease Course:
- Complications (secondary problems: infections, organ failure, etc.) > Search first: ICD codes, disease registries, clinical databases, PubMed
- Recovery potential (likelihood and extent of recovery, with vs without treatment) > Search first: Natural history studies, rehabilitation databases, PubMed
- Prediction:
- Prognostic factors (age, disease severity, biomarkers, treatment response) > Search first: Prognostic models databases, clinical calculators, PubMed
- Prognostic biomarkers (molecular markers predicting disease course) > Search first: FDA Biomarker database, PubMed, cancer prognostic databases
12. Treatment
- Pharmacotherapy:
- Pharmacological treatments (drug names, drug classes, mechanisms of action) > Search first: DrugBank, RxNorm, ATC classification, DailyMed, FDA databases
- Pharmacogenomics (how genetic variants affect drug metabolism, efficacy, toxicity) > Search first: PharmGKB, CPIC (Clinical Pharmacogenetics), FDA Table of PGx Biomarkers
- Advanced Therapeutics:
- Gene therapy (viral vectors, CRISPR, gene replacement, gene editing) > Search first: ClinicalTrials.gov, FDA gene therapy database, ASGCT resources
- Cell therapy (stem cell transplant, CAR-T, cellular therapeutics) > Search first: ClinicalTrials.gov, FDA cell therapy database, FACT standards
- RNA-based therapies (ASOs, siRNA, mRNA therapies) > Search first: ClinicalTrials.gov, FDA approvals, PubMed
- Targeted therapies (treatments directed at specific molecular targets) > Search first: My Cancer Genome, OncoKB, ClinicalTrials.gov, FDA approvals
- Immunotherapies (checkpoint inhibitors, monoclonal antibodies) > Search first: Cancer Immunotherapy Database, FDA approvals, ClinicalTrials.gov
- Surgical and Interventional:
- Surgical interventions (types of surgery, timing, outcomes) > Search first: CPT codes, surgical registries, clinical guidelines, PubMed
- Supportive and Rehabilitative:
- Supportive care (symptom management, pain control, nutrition) > Search first: Clinical guidelines, Cochrane Library, PubMed
- Rehabilitation (physical therapy, occupational therapy, speech therapy) > Search first: Rehabilitation medicine databases, clinical guidelines, PubMed
- Experimental:
- Experimental treatments in clinical trials (with NCT identifiers if available) > Search first: ClinicalTrials.gov, EU Clinical Trials Register, WHO ICTRP
- Treatment Outcomes:
- Treatment response rates > Search first: Clinical trial databases, FDA reviews, systematic reviews, PubMed
- Side effects and adverse events > Search first: FDA Adverse Event Reporting System (FAERS), MedWatch, PubMed
- Treatment Strategy:
- Treatment algorithms (clinical pathways, decision trees) > Search first: Clinical practice guidelines, NCCN Guidelines, UpToDate
- Combination therapies > Search first: ClinicalTrials.gov, treatment guidelines, PubMed
- Personalized medicine approaches (genotype-guided treatment) > Search first: My Cancer Genome, CIViC, PharmGKB, precision medicine databases
For each treatment, suggest MAXO (Medical Action Ontology) terms where applicable.
13. Prevention
- Prevention Levels:
- Primary prevention (preventing disease occurrence: vaccination, risk factor modification) > Search first: CDC, WHO, USPSTF recommendations, Cochrane Library
- Secondary prevention (early detection and treatment: screening programs, early intervention) > Search first: USPSTF, CDC screening guidelines, WHO
- Tertiary prevention (preventing complications in those with disease) > Search first: Clinical guidelines, disease management protocols, PubMed
- Immunization: Vaccine strategies (if applicable)
Search first: CDC vaccine schedules, WHO immunization, FDA vaccine database
- Screening and Early Detection:
- Screening programs (population-based: newborn screening, cancer screening) > Search first: CDC screening programs, USPSTF, cancer screening databases
- Genetic screening (carrier screening, preimplantation genetic diagnosis, prenatal testing) > Search first: ACMG recommendations, ACOG guidelines, GTR
- Risk stratification (identifying high-risk individuals for targeted prevention) > Search first: Risk prediction models, clinical calculators, PubMed
- Behavioral Interventions: Lifestyle modifications to reduce risk
Search first: CDC, WHO, behavioral intervention databases, Cochrane Library
- Counseling: Genetic counseling (risk assessment, family planning guidance)
Search first: NSGC resources, ACMG guidelines, GeneReviews
- Public Health:
- Public health interventions (sanitation, vector control, health education) > Search first: CDC, WHO, public health databases, PubMed
- Environmental interventions (reducing environmental risk factors) > Search first: EPA databases, WHO environmental health, PubMed
- Prophylaxis: Preventive medications or procedures
Search first: Clinical guidelines, FDA approvals, PubMed
14. Other Species / Natural Disease
- Taxonomy: Species affected (with NCBI Taxon identifiers)
Search first: NCBI Taxonomy
- Breed: Specific breeds affected (with VBO identifiers if applicable)
Search first: VBO (Vertebrate Breed Ontology)
- Gene: Orthologous genes in other species (with NCBI Gene IDs)
Search first: NCBI Gene
- Natural Disease:
- Naturally occurring disease in other species (companion animals, wildlife) > Search first: OMIA (Online Mendelian Inheritance in Animals), VetCompass, PubMed
- Veterinary relevance and importance in animal health > Search first: OMIA, veterinary databases, PubMed
- Comparative Biology:
- Comparative pathology (similarities and differences across species) > Search first: OMIA, comparative pathology databases, PubMed
- Evolutionary conservation of disease mechanisms > Search first: HomoloGene, OrthoMCL, Alliance of Genome Resources
- Transmission (if applicable):
- Zoonotic potential > Search first: CDC zoonotic diseases, WHO zoonoses, GIDEON
- Cross-species susceptibility > Search first: NCBI Taxonomy, veterinary databases, PubMed
15. Model Organisms
- Model Types:
- Model organism type (mammalian, invertebrate, cellular, in vitro) > Search first: Alliance of Genome Resources, model organism databases
- Specific model systems (mouse, rat, zebrafish, Drosophila, C. elegans, yeast, cell lines, organoids, iPSCs) > Search first: MGI, RGD, ZFIN, FlyBase, WormBase, SGD, ATCC, Cellosaurus
- Induced models (drug treatment, surgical intervention, environmental manipulation) > Search first: MGI, model organism databases, PubMed
- Genetic Models:
- Types available (knockout, knock-in, transgenic, conditional, humanized) > Search first: MGI, IMPC, KOMP, EuMMCR, IMSR
- Model Characteristics:
- Phenotype recapitulation (how well model reproduces human disease features) > Search first: Model organism databases, comparative studies, PubMed
- Model limitations (aspects of human disease not captured) > Search first: Model organism databases, PubMed, review articles
- Applications:
- Research applications (what aspects of disease can be studied) > Search first: Model organism databases, PubMed
- Resources:
- Model databases > Search first: MGI, RGD, ZFIN, FlyBase, WormBase, IMSR, EMMA, MMRRC
Citation Requirements
- Cite primary literature (PMID preferred) for all mechanistic and clinical claims
- Prioritize recent reviews and landmark papers
- Include direct quotes from abstracts where possible to support key statements
- Distinguish evidence source types: human clinical, model organism, in vitro, computational
Output Format
Structure your response as a comprehensive narrative organized by the sections above. For each section, provide: - Factual content with specific details (numbers, percentages, gene names, variant nomenclature) - Ontology term suggestions (HPO, GO, CL, UBERON, CHEBI, MAXO, MONDO) where applicable - Evidence citations with PMIDs - Direct quotes from abstracts to support key claims - Clear indication when information is not available or not applicable for this disease
This report will be used to populate a disease knowledge base entry with: - Pathophysiology descriptions with causal chains - Gene/protein annotations (HGNC, GO terms) - Phenotype associations (HP terms) with frequencies - Cell type involvement (CL terms) - Anatomical locations (UBERON terms) - Chemical entities (CHEBI terms) - Treatment annotations (MAXO terms) - Evidence items with PMIDs and exact abstract quotes - Epidemiology, prognosis, diagnostic, and prevention information - Animal model descriptions with phenotype recapitulation details
GNPTG‑Mucolipidosis (Mucolipidosis type III gamma; MLIIIγ): Comprehensive disease characteristics report
Executive summary
GNPTG‑mucolipidosis corresponds to mucolipidosis type III gamma (MLIIIγ), an autosomal recessive lysosomal storage disorder caused by biallelic pathogenic variants in GNPTG, encoding the γ subunit of N‑acetylglucosamine‑1‑phosphotransferase (GlcNAc‑1‑PT)—the initiating enzyme for mannose‑6‑phosphate (M6P) tagging of lysosomal hydrolases. Loss of proper M6P tagging causes missorting and hypersecretion of lysosomal enzymes into extracellular fluids (plasma/serum), with relative deficiency in lysosomes, leading to progressive connective‑tissue/bone/joint disease that is typically milder than GNPTAB‑related mucolipidosis. (velho2019thelysosomalstorage pages 1-3, dogterom2021mucolipidosistypeii pages 1-2)
Recent (2023–2024) advances relevant to GNPTG‑mucolipidosis include: (i) multiplex tandem mass spectrometry assays in dried blood spots to support screening/diagnosis of MLII/III; and (ii) whole‑genome sequencing (WGS) enabling diagnosis in atypical presentations and expanding the phenotype to include late‑onset retinal dystrophy/retinitis pigmentosa in at least one GNPTG‑confirmed case. (hong2023multiplextandemmass pages 1-2, geer2023unravelingmucolipidosistype pages 2-5)
1. Disease information
1.1 Definition and overview
Mucolipidosis type III gamma (MLIIIγ) is an attenuated mucolipidosis within the MLII/III spectrum, caused by defects in the lysosomal enzyme targeting pathway (M6P tagging). It is described as a variable but generally milder phenotype than MLII and MLIIIα/β, with major manifestations in the musculoskeletal system (progressive joint restriction, skeletal dysplasia/degeneration). (velho2019thelysosomalstorage pages 1-3, dogterom2021mucolipidosistypeii pages 1-2)
1.2 Key identifiers (OMIM/Orphanet/ICD/MeSH/MONDO)
The retrieved full‑text evidence did not include OMIM, Orphanet, ICD‑10/ICD‑11, MeSH, or MONDO identifiers; therefore these cannot be reported with tool‑verified citations in this run.
1.3 Synonyms and alternative names
- Mucolipidosis type III gamma (MLIIIγ). (velho2019thelysosomalstorage pages 1-3, dogterom2021mucolipidosistypeii pages 1-2)
- Historical MLIII literature uses pseudo‑Hurler polydystrophy (used in GNPTG mutation papers). (persichetti2009identificationandmolecular pages 1-2)
1.4 Evidence source types
- Aggregated disease‑level resources: systematic review of 843 cases and a GNPTAB/GNPTG mutation update compiling hundreds of individuals. (dogterom2021mucolipidosistypeii pages 1-2, velho2019thelysosomalstorage pages 1-3)
- Individual patient reports: e.g., WGS‑diagnosed late‑onset retinitis pigmentosa case. (geer2023unravelingmucolipidosistype pages 1-2)
- Model organism and cellular: Gnptg knockout mouse studies investigating cartilage/ECM and joint function. (westermann2020imbalancedcellularmetabolism pages 42-44)
2. Etiology
2.1 Disease causal factors
Primary cause: biallelic pathogenic variants in GNPTG (autosomal recessive). (velho2019thelysosomalstorage pages 1-3, dogterom2021mucolipidosistypeii pages 1-2)
Molecular cause: GNPTG encodes the γ subunit of the α2β2γ2 GlcNAc‑1‑phosphotransferase complex that initiates formation of M6P on lysosomal enzymes. Reduced function causes defective M6P tagging, missorting and extracellular secretion of lysosomal hydrolases, and downstream lysosomal substrate accumulation. (velho2019thelysosomalstorage pages 1-3, dogterom2021mucolipidosistypeii pages 1-2, khan2020mucolipidosesoverviewpast pages 3-6)
2.2 Risk factors
- Genetic: biallelic GNPTG pathogenic variants are causal. (dogterom2021mucolipidosistypeii pages 1-2, velho2019thelysosomalstorage pages 1-3)
Environmental, lifestyle, or infectious risk factors were not identified in the retrieved evidence and are not generally expected for this Mendelian disorder.
2.3 Protective factors / gene–environment interactions
No protective genetic variants or gene–environment interactions were identified in the retrieved evidence.
3. Phenotypes
3.1 Core clinical phenotype spectrum
Across primary reports and systematic synthesis, MLIIIγ is characterized by predominant musculoskeletal and connective‑tissue disease: * Joint stiffness / restricted joint mobility (often progressive). (meel2016mucolipidosisiiignptg pages 1-3, dogterom2021mucolipidosistypeii pages 1-2) * Claw‑hand deformity and hand stiffness. (meel2016mucolipidosisiiignptg pages 1-3) * Short stature and scoliosis/kyphosis. (meel2016mucolipidosisiiignptg pages 1-3, persichetti2009identificationandmolecular pages 1-2) * Progressive bone/joint disease, including hip destruction; orthopedic sequelae can be severe (e.g., hip replacement by age 30 in one GNPTG‑confirmed adult). (geer2023unravelingmucolipidosistype pages 1-2) * Carpal/tarsal tunnel syndrome is reported as common among MLIII patients in systematic synthesis. (dogterom2021mucolipidosistypeii pages 1-2) * Cardiac valve disease (valvular involvement) can occur. (meel2016mucolipidosisiiignptg pages 1-3) * Neurocognition: generally normal intellectual capacity or only mild impairment in MLIII (including MLIIIγ) in systematic synthesis. (dogterom2021mucolipidosistypeii pages 1-2)
3.2 Phenotype expansion (2023)
A 2023 WGS‑diagnosed case reported late‑onset retinal dystrophy/retinitis pigmentosa in a man with compound heterozygous GNPTG pathogenic variants, and explicitly noted that retinal dystrophy was not previously a common feature; the authors stated it was (to their knowledge) the first description of retinitis pigmentosa due to GNPTG compound heterozygosity. (geer2023unravelingmucolipidosistype pages 1-2)
3.3 Age of onset, severity, progression
In MLII/III overall, MLIII has later onset and milder course than MLII; in a systematic review, median age at diagnosis for MLIII was 9.0 years (contrasting with 0.7 years for MLII). (dogterom2021mucolipidosistypeii pages 1-2)
3.4 Quality of life impact (evidence‑based inference)
Quality of life is likely driven by progressive joint restriction, pain, and orthopedic morbidity (e.g., joint replacements). A direct quantitative QoL instrument (e.g., SF‑36/EQ‑5D) was not identified in the retrieved evidence; QoL assertions therefore cannot be made quantitatively.
3.5 Suggested HPO terms (non‑exhaustive)
Based on the evidence‑supported phenotype spectrum: * Joint stiffness / limitation of joint mobility (HP:0001376 / HP:0001385) (meel2016mucolipidosisiiignptg pages 1-3, dogterom2021mucolipidosistypeii pages 1-2) * Short stature (HP:0004322) (meel2016mucolipidosisiiignptg pages 1-3) * Scoliosis (HP:0002650) (meel2016mucolipidosisiiignptg pages 1-3) * Osteoarthritis / degenerative joint disease (HP:0002758) (westermann2020imbalancedcellularmetabolism pages 42-44) * Carpal tunnel syndrome (HP:0001240? note: HPO exists as HP:0001240 is “Seizures”; therefore do not treat this as a definitive code; term suggested conceptually from the review) (dogterom2021mucolipidosistypeii pages 1-2) * Valvular heart disease (HP:0001654 conceptually) (meel2016mucolipidosisiiignptg pages 1-3) * Retinitis pigmentosa (HP:0000510) (geer2023unravelingmucolipidosistype pages 1-2)
(Where HPO IDs are marked “conceptually,” exact IDs should be verified against the HPO database; the provided evidence supports the clinical concept but does not provide ontology mappings.)
4. Genetic / molecular information
4.1 Causal gene
- GNPTG encodes the soluble γ subunit of GlcNAc‑1‑phosphotransferase. (velho2019thelysosomalstorage pages 1-3, dogterom2021mucolipidosistypeii pages 1-2)
4.2 Pathogenic variant classes and functional consequences
Variant classes reported include missense, nonsense, splice‑site, and frameshift/microdeletion alleles. (persichetti2009identificationandmolecular pages 1-2)
Functional consequences (examples): * Multiple GNPTG missense variants (p.G106S, p.G126S, p.C142Y) cause misfolding, ER retention, and aggregation of the γ subunit, preventing functional rescue of lysosomal targeting. (meel2016mucolipidosisiiignptg pages 1-3) * Other pathogenic alleles can cause aberrant splicing or nonsense‑mediated decay, leading to loss of GNPTG protein synthesis. (persichetti2009identificationandmolecular pages 1-2)
4.3 Example variants from recent clinical reporting
In the 2023 late‑onset retinal dystrophy case, WGS identified compound heterozygous GNPTG pathogenic variants c.347_349del and c.607dup (protein consequences reported as p.Asn116del and p.Gln203fs in the evidence summary), motivating the MLIIIγ diagnosis. (geer2023unravelingmucolipidosistype pages 1-2)
4.4 Allele frequencies / population databases
Allele frequencies (gnomAD, 1000 Genomes, etc.) and carrier frequency estimates were not provided in the retrieved evidence.
4.5 Modifier genes / epigenetics / chromosomal abnormalities
No modifier genes, epigenetic mechanisms, or chromosomal abnormalities were identified in the retrieved evidence.
5. Environmental information
No environmental, lifestyle, or infectious contributors were identified in the retrieved evidence; MLIIIγ is primarily a Mendelian disorder driven by GNPTG genotype.
6. Mechanism / pathophysiology
6.1 Current mechanistic understanding
GlcNAc‑1‑phosphotransferase is the key enzyme initiating addition of the M6P targeting signal on lysosomal hydrolases. GNPTAB encodes the α/β precursor and GNPTG encodes the γ subunit in an α2β2γ2 complex; defective GlcNAc‑1‑PT leads to missorting of newly synthesized lysosomal enzymes into the extracellular space, causing a shortage in lysosomes and accumulation of non‑degradable macromolecules, impairing cellular function. (velho2019thelysosomalstorage pages 1-3, dogterom2021mucolipidosistypeii pages 1-2, khan2020mucolipidosesoverviewpast pages 3-6)
A systematic review emphasized that M6P is essential for targeting >70 soluble lysosomal enzymes, and without M6P, enzymes are secreted extracellularly. (dogterom2021mucolipidosistypeii pages 1-2)
6.2 Causal chain (trigger → molecular defect → cellular defect → organ disease)
1) Biallelic GNPTG pathogenic variants → defective γ subunit of GlcNAc‑1‑PT (often misfolding/ER retention or loss of transcript/protein). (meel2016mucolipidosisiiignptg pages 1-3, persichetti2009identificationandmolecular pages 1-2) 2) Reduced M6P tagging on subsets of lysosomal enzymes → missorting/hypersecretion of hydrolases into plasma/serum. (meel2016mucolipidosisiiignptg pages 1-3, dogterom2021mucolipidosistypeii pages 1-2) 3) Intralysosomal enzyme deficiency → storage of substrates in lysosomes (connective‑tissue relevant substrates include glycosaminoglycans/oligosaccharides, etc.). (dogterom2021mucolipidosistypeii pages 1-2, khan2020mucolipidosesoverviewpast pages 3-6) 4) Tissue dysfunction in cartilage/tendon/bone/connective tissue → progressive joint restriction, skeletal deformity/degeneration and pain. (westermann2020imbalancedcellularmetabolism pages 42-44)
6.3 Cellular processes and tissues/cell types implicated (supported by model data)
In a Gnptg knockout mouse model, joint pathology is linked to chondroitin sulfate accumulation in chondrocytes, impaired differentiation, and altered cartilage ECM microstructure; mechanosensitive connective tissues such as cartilage and tendon are highlighted. (westermann2020imbalancedcellularmetabolism pages 42-44)
Suggested GO biological process terms (conceptual; verify IDs): lysosomal enzyme targeting, lysosome organization, glycosaminoglycan catabolic process, extracellular matrix organization.
Suggested Cell Ontology (CL) cell types: chondrocyte; fibroblast.
7. Anatomical structures affected
7.1 Organ/tissue systems
Dominant involvement of musculoskeletal/connective tissue systems (joints, cartilage, tendon, bone) is supported by human phenotypes and mouse data. (dogterom2021mucolipidosistypeii pages 1-2, westermann2020imbalancedcellularmetabolism pages 42-44)
7.2 Subcellular localization
Primary dysfunction centers on the Golgi→lysosome trafficking axis (M6P tagging in Golgi) and downstream lysosomal substrate storage. (dogterom2021mucolipidosistypeii pages 1-2)
7.3 Suggested UBERON terms (conceptual)
Cartilage, tendon, bone, joint, retina.
8. Temporal development
8.1 Onset patterns
MLIII (including MLIIIγ) is typically recognized in childhood; MLIII average age‑of‑onset is cited as ~5 years in a 2023 diagnostic assay paper, but MLIIIγ may present variably and can be diagnosed later in atypical presentations. (hong2023multiplextandemmass pages 1-2, geer2023unravelingmucolipidosistype pages 1-2)
8.2 Disease course
MLIII is typically chronic and progressive. Median age at diagnosis in the systematic review was 9 years for MLIII, with long survival into adulthood for many patients. (dogterom2021mucolipidosistypeii pages 1-2)
9. Inheritance and population
9.1 Inheritance
Autosomal recessive. (velho2019thelysosomalstorage pages 1-3, dogterom2021mucolipidosistypeii pages 1-2)
9.2 Epidemiology (available data)
A 2023 MS/MS diagnostic assay paper reports combined MLII/III prevalence of 0.22–2.70 per 100,000 live births (not GNPTG‑specific). (hong2023multiplextandemmass pages 1-2)
9.3 Population genetics / founder effects
Carrier frequencies, penetrance estimates, and founder variants were not provided in the retrieved evidence.
10. Diagnostics
10.1 Biochemical testing (hallmark pattern)
A defining diagnostic hallmark across mucolipidosis II/III is elevated lysosomal enzyme activities in plasma/serum due to hypersecretion (with relative reduction/low‑normal activities in leukocytes/fibroblasts). (sohn2016moleculargeneticsand pages 1-2, meel2016mucolipidosisiiignptg pages 1-3)
Quantitative example (GNPTG case, 2023): plasma β‑glucuronidase 1370 µkat/L vs 42 control; Hexosaminidase B 5190 vs 135; β‑galactosidase 4.2 vs 0.9; leukocyte activities were slightly decreased or low‑normal. (geer2023unravelingmucolipidosistype pages 2-5)
10.2 Genetic testing
Molecular confirmation requires GNPTG sequencing (and GNPTAB to subtype within MLII/III), because biochemical enzyme elevations alone do not reliably distinguish MLII vs MLIII subtypes. (khan2020mucolipidosesoverviewpast pages 3-6, hong2023multiplextandemmass pages 2-4)
WGS as implementation example (2023): a 325‑gene retinal disease panel filter applied to WGS data unexpectedly identified GNPTG pathogenic variants, leading to MLIIIγ diagnosis in a retinitis pigmentosa presentation. (geer2023unravelingmucolipidosistype pages 1-2)
10.3 Screening / new diagnostic developments (2023)
A 2023 study developed a multiplex UPLC‑MS/MS enzyme activity assay using two 6‑plex panels on dried blood spots (12 enzymes total). In MLII/III DBS samples, ASM, IDS, and NAGLU were markedly elevated (about 20‑, 11‑, and 17‑fold vs newborn controls) and non‑overlapping with controls; this supports potential screening workflows (including newborn screening concepts), while emphasizing that genetic testing is still required for subtype assignment and that MLIIIγ stratification was limited by small sample size (n=1 MLIIIγ). (hong2023multiplextandemmass pages 2-4)
10.4 Differential diagnosis
MLII/III shares clinical overlap with mucopolysaccharidoses (MPS) (skeletal dysplasia/dysostosis multiplex), and biochemical enzyme panels alone cannot subtype; confirmation requires GNPTAB/GNPTG sequencing. (khan2020mucolipidosesoverviewpast pages 3-6)
11. Outcome / prognosis
A systematic review of 843 published MLII/III cases reported for MLIII (all subtypes): median survival 62.0 years and median age of death 33.0 years (not GNPTG‑specific, but MLIIIγ is described as the least severe ML phenotype). (dogterom2021mucolipidosistypeii pages 1-2)
12. Treatment
12.1 Current standard of care and real‑world implementations
No definitive disease‑modifying therapy was identified in the retrieved evidence; sources emphasize that treatment is largely supportive/symptomatic, with orthopedic management addressing joint disease (e.g., joint replacement in severe hip disease). (khan2020mucolipidosesoverviewpast pages 1-3, geer2023unravelingmucolipidosistype pages 1-2)
12.2 Clinical trials / advanced therapeutics
No GNPTG/MLIIIγ‑specific clinical trials or NCT identifiers were retrieved in this run.
12.3 Suggested MAXO terms (conceptual)
Supportive care; orthopedic surgery; physical therapy/rehabilitation; genetic counseling.
13. Prevention
Primary prevention is not applicable in the usual public‑health sense for a Mendelian disorder; prevention focuses on genetic counseling, carrier testing, and prenatal/preimplantation options, but these were not detailed in the retrieved evidence.
14. Other species / natural disease
No naturally occurring non‑human GNPTG‑mucolipidosis cases were identified in the retrieved evidence.
15. Model organisms
A Gnptg knockout mouse has been used as an MLIIIγ model. The study linked lysosomal enzyme missorting to cartilage homeostasis failure and joint dysfunction, including chondroitin sulfate accumulation in chondrocytes and altered ECM structure, with functional joint abnormalities and impaired motor performance under fatigue/pain conditions. (westermann2020imbalancedcellularmetabolism pages 42-44)
Visual evidence (phenotype)
A joint‑mobility figure illustrating range‑of‑motion abnormalities across MLII/MLIII subtypes (including MLIIIγ) was retrieved from the Gnptg mouse/human comparative study. (westermann2020imbalancedcellularmetabolism media df302704)
Evidence table (structured summary)
| Category | Key facts | Key sources (with PMID if known; otherwise DOI/URL) | Publication year |
|---|---|---|---|
| Disease entity / synonyms | GNPTG-associated disease is mucolipidosis type III gamma (MLIIIγ), an attenuated lysosomal storage disorder within the MLII/III spectrum; historical synonym pseudo-Hurler polydystrophy is used in MLIII literature. All pathogenic GNPTG variants reported in the systematic review resulted in the MLIII gamma phenotype. (velho2019thelysosomalstorage pages 1-3, persichetti2009identificationandmolecular pages 1-2, dogterom2021mucolipidosistypeii pages 1-2) | Velho et al., Hum Mutat 2019, DOI: https://doi.org/10.1002/humu.23748; Persichetti et al., Hum Mutat 2009, DOI: https://doi.org/10.1002/humu.20959; Dogterom et al., Genet Med 2021, DOI: https://doi.org/10.1038/s41436-021-01244-4 | 2009, 2019, 2021 |
| Causal gene | Causal gene: GNPTG, encoding the soluble γ-subunit of N-acetylglucosamine-1-phosphotransferase (GlcNAc-1-phosphotransferase), part of the α2β2γ2 hexameric complex. (velho2019thelysosomalstorage pages 1-3, sohn2016moleculargeneticsand pages 1-2, dogterom2021mucolipidosistypeii pages 1-2, khan2020mucolipidosesoverviewpast pages 3-6) | Velho et al., DOI: https://doi.org/10.1002/humu.23748; Sohn, DOI: https://doi.org/10.19125/jmrd.2016.2.1.13; Dogterom et al., DOI: https://doi.org/10.1038/s41436-021-01244-4 | 2016, 2019, 2021 |
| Inheritance | Autosomal recessive inheritance. MLIIIγ is consistently described as the mildest/least severe GlcNAc-1-phosphotransferase–related mucolipidosis phenotype. (velho2019thelysosomalstorage pages 1-3, sohn2016moleculargeneticsand pages 1-2, meel2016mucolipidosisiiignptg pages 1-3, hong2023multiplextandemmass pages 1-2) | Velho et al., DOI: https://doi.org/10.1002/humu.23748; Sohn, DOI: https://doi.org/10.19125/jmrd.2016.2.1.13; van Meel & Kornfeld, DOI: https://doi.org/10.1002/humu.22993; Hong et al., DOI: https://doi.org/10.1016/j.ymgmr.2023.100978 | 2016, 2019, 2023 |
| Key mechanism / pathophysiology | GlcNAc-1-phosphotransferase catalyzes the first step of mannose-6-phosphate (M6P) tagging of lysosomal hydrolases. GNPTG deficiency reduces phosphorylation of a subset of lysosomal enzymes, causing missorting into the extracellular space, lysosomal enzyme shortage within lysosomes, and accumulation of undegraded substrates in enlarged lysosomes. (velho2019thelysosomalstorage pages 1-3, meel2016mucolipidosisiiignptg pages 1-3, khan2020mucolipidosesoverviewpast pages 1-3, dogterom2021mucolipidosistypeii pages 1-2, khan2020mucolipidosesoverviewpast pages 3-6) | Velho et al., DOI: https://doi.org/10.1002/humu.23748; van Meel & Kornfeld, DOI: https://doi.org/10.1002/humu.22993; Khan & Tomatsu, DOI: https://doi.org/10.3390/ijms21186812; Dogterom et al., DOI: https://doi.org/10.1038/s41436-021-01244-4 | 2016, 2019, 2020, 2021 |
| Variant classes / molecular effects | Reported pathogenic GNPTG variants include missense, nonsense, splice-site, and frameshift/microdeletion alleles. Missense variants such as p.G106S, p.G126S, p.C142Y can cause misfolding, ER retention, and aggregation; other alleles produce aberrant splicing or nonsense-mediated decay. (persichetti2009identificationandmolecular pages 1-2, meel2016mucolipidosisiiignptg pages 1-3) | Persichetti et al., DOI: https://doi.org/10.1002/humu.20959; van Meel & Kornfeld, DOI: https://doi.org/10.1002/humu.22993 | 2009, 2016 |
| Hallmark biochemical findings | Diagnostic hallmark: elevated lysosomal enzyme activities in plasma/serum due to hypersecretion, with relatively decreased or low-normal leukocyte/fibroblast activities; urinary GAGs are often normal in MLII/III while oligosaccharides may be increased. (sohn2016moleculargeneticsand pages 1-2, meel2016mucolipidosisiiignptg pages 1-3, khan2020mucolipidosesoverviewpast pages 3-6, geer2023unravelingmucolipidosistype pages 2-5, geer2023unravelingmucolipidosistype pages 1-2) | Sohn, DOI: https://doi.org/10.19125/jmrd.2016.2.1.13; van Meel & Kornfeld, DOI: https://doi.org/10.1002/humu.22993; Khan & Tomatsu, DOI: https://doi.org/10.3390/ijms21186812; De Geer et al., DOI: https://doi.org/10.1186/s12886-023-03136-4 | 2016, 2020, 2023 |
| Quantitative biochemical examples | In the 2023 GNPTG case with retinitis pigmentosa, plasma enzymes were markedly elevated, e.g. β-glucuronidase 1370 µkat/L vs 42 control, Hexosaminidase B 5190 vs 135, β-galactosidase 4.2 vs 0.9; leukocyte activities were slightly decreased or low-normal. (geer2023unravelingmucolipidosistype pages 2-5) | De Geer et al., BMC Ophthalmol 2023, DOI: https://doi.org/10.1186/s12886-023-03136-4 | 2023 |
| Core clinical features | Typical MLIIIγ features include joint stiffness/restricted mobility, claw-hand deformity, short stature, scoliosis, progressive bone/joint disease, hip destruction/replacement, carpal/tarsal tunnel syndrome, and valvular heart disease; intellect is often normal or only mildly affected. (persichetti2009identificationandmolecular pages 1-2, meel2016mucolipidosisiiignptg pages 1-3, dogterom2021mucolipidosistypeii pages 1-2, geer2023unravelingmucolipidosistype pages 1-2, westermann2020imbalancedcellularmetabolism media df302704) | Persichetti et al., DOI: https://doi.org/10.1002/humu.20959; van Meel & Kornfeld, DOI: https://doi.org/10.1002/humu.22993; Dogterom et al., DOI: https://doi.org/10.1038/s41436-021-01244-4; De Geer et al., DOI: https://doi.org/10.1186/s12886-023-03136-4 | 2009, 2016, 2021, 2023 |
| Tissue/anatomic emphasis | Human and mouse data support major involvement of connective tissue, cartilage, tendon, bone, and joints; Gnptg knockout studies link enzyme missorting to chondroitin sulfate accumulation in chondrocytes, altered ECM structure, and functional joint abnormalities. (westermann2020imbalancedcellularmetabolism pages 42-44, westermann2020imbalancedcellularmetabolism media df302704) | Westermann et al., Dis Model Mech 2020, DOI: https://doi.org/10.1242/dmm.046425 | 2020 |
| Natural history / prognosis | MLIII is later-onset and longer-surviving than MLII. Systematic review statistics: median age at diagnosis 9.0 years for MLIII; median survival 62.0 years; median age at death 33.0 years in reported cases. (dogterom2021mucolipidosistypeii pages 1-2) | Dogterom et al., DOI: https://doi.org/10.1038/s41436-021-01244-4 | 2021 |
| Epidemiology | Reported combined prevalence for MLII/III is approximately 0.22–2.70 per 100,000 live births; GNPTG-specific prevalence was not separately quantified in the provided evidence. (hong2023multiplextandemmass pages 1-2) | Hong et al., DOI: https://doi.org/10.1016/j.ymgmr.2023.100978 | 2023 |
| Recent development: multiplex DBS MS/MS assay | 2023: multiplex UPLC-MS/MS dried blood spot assay measured 12 lysosomal enzymes in MLII/III and showed a characteristic pattern, supporting screening/diagnosis and potential newborn screening workflows. In DBS, ASM, IDS, and NAGLU were significantly elevated; in the cohort these were about 20-fold, 11-fold, and 17-fold above newborn controls, respectively. Molecular testing is still needed to distinguish MLIIIγ from other MLII/III subtypes. (hong2023multiplextandemmass pages 1-2, hong2023multiplextandemmass pages 2-4) | Hong et al., Mol Genet Metab Rep 2023, DOI: https://doi.org/10.1016/j.ymgmr.2023.100978 | 2023 |
| Recent development: WGS phenotype expansion | 2023: whole genome sequencing in a 47-year-old patient with late-onset retinal disease identified compound heterozygous GNPTG variants c.347_349del (p.Asn116del) and c.607dup (p.Gln203fs), leading to diagnosis of MLIIIγ. This was reported as the first description of retinitis pigmentosa/late-onset retinal dystrophy in GNPTG-related MLIIIγ, expanding the phenotype. (geer2023unravelingmucolipidosistype pages 2-5, geer2023unravelingmucolipidosistype pages 1-2) | De Geer et al., BMC Ophthalmol 2023, DOI: https://doi.org/10.1186/s12886-023-03136-4 | 2023 |
| Current diagnostic approach | Best-supported approach combines clinical suspicion, biochemical evidence of hypersecreted lysosomal enzymes (plasma/serum ± DBS), and molecular confirmation by GNPTG sequencing; biochemical testing alone cannot reliably subtype MLII vs MLIIIα/β vs MLIIIγ. (sohn2016moleculargeneticsand pages 1-2, khan2020mucolipidosesoverviewpast pages 3-6, hong2023multiplextandemmass pages 2-4) | Sohn, DOI: https://doi.org/10.19125/jmrd.2016.2.1.13; Khan & Tomatsu, DOI: https://doi.org/10.3390/ijms21186812; Hong et al., DOI: https://doi.org/10.1016/j.ymgmr.2023.100978 | 2016, 2020, 2023 |
| Treatment status | No curative or disease-modifying therapy was identified in the provided GNPTG-focused evidence; management remains largely supportive/symptomatic, and curative treatment was noted as unavailable in reviews/systematic summaries. (sohn2016moleculargeneticsand pages 1-2, khan2020mucolipidosesoverviewpast pages 1-3, dogterom2021mucolipidosistypeii pages 1-2) | Sohn, DOI: https://doi.org/10.19125/jmrd.2016.2.1.13; Khan & Tomatsu, DOI: https://doi.org/10.3390/ijms21186812; Dogterom et al., DOI: https://doi.org/10.1038/s41436-021-01244-4 | 2016, 2020, 2021 |
Table: This table condenses the core disease-defining facts for GNPTG-associated mucolipidosis type III gamma, including mechanism, clinical and biochemical hallmarks, natural history, and the most relevant 2023 diagnostic/phenotypic updates.
Expert synthesis / analysis (grounded in cited sources)
1) The convergent biochemical hallmark—high plasma lysosomal enzymes—is mechanistically coherent with M6P tagging failure and provides a high‑yield diagnostic clue, but molecular confirmation is required for subtype assignment and genetic counseling. (dogterom2021mucolipidosistypeii pages 1-2, hong2023multiplextandemmass pages 2-4) 2) 2023 data suggest practical pathways for earlier detection: DBS‑based MS/MS assays can provide a scalable biochemical screen, whereas WGS can resolve atypical phenotypes and broaden disease recognition (e.g., retinal dystrophy). Together, these support a “biochemical screen → genomic confirmation” diagnostic paradigm for MLII/III. (hong2023multiplextandemmass pages 2-4, geer2023unravelingmucolipidosistype pages 1-2)
Notes on evidence limits and gaps
- Identifiers (OMIM/Orphanet/ICD/MeSH/MONDO), carrier frequencies/gnomAD allele frequencies, and formal clinical guidelines were not present in the retrieved full‑text evidence, so they are intentionally not populated here.
- A 2024 mucolipidosis review was retrieved but primarily provides a broad overview; detailed GNPTG‑specific treatment innovations in 2024 were not available in the accessible evidence set. (westermann2020imbalancedcellularmetabolism media df302704)
References
-
(velho2019thelysosomalstorage pages 1-3): Renata Voltolini Velho, Frederike L. Harms, Tatyana Danyukova, Nataniel F. Ludwig, Michael J. Friez, Sara S. Cathey, Mirella Filocamo, Barbara Tappino, Nilay Güneş, Beyhan Tüysüz, Karen L. Tylee, Kathryn L. Brammeier, Lesley Heptinstall, Esmee Oussoren, Ans T. Ploeg, Christine Petersen, Sandra Alves, Gloria Durán Saavedra, Ida V. Schwartz, Nicole Muschol, Kerstin Kutsche, and Sandra Pohl. The lysosomal storage disorders mucolipidosis type ii, type iii alpha/beta, and type iii gamma: update on gnptab and gnptg mutations. Human Mutation, 40:842-864, Jul 2019. URL: https://doi.org/10.1002/humu.23748, doi:10.1002/humu.23748. This article has 81 citations and is from a domain leading peer-reviewed journal.
-
(dogterom2021mucolipidosistypeii pages 1-2): Emma J. Dogterom, Margreet A.E.M. Wagenmakers, Martina Wilke, Serwet Demirdas, Nicole M. Muschol, Sandra Pohl, Jan C. van der Meijden, Dimitris Rizopoulos, Ans T. van der Ploeg, and Esmée Oussoren. Mucolipidosis type ii and type iii: a systematic review of 843 published cases. Genetics in Medicine, 23:2047-2056, Nov 2021. URL: https://doi.org/10.1038/s41436-021-01244-4, doi:10.1038/s41436-021-01244-4. This article has 44 citations and is from a highest quality peer-reviewed journal.
-
(hong2023multiplextandemmass pages 1-2): Xinying Hong, Laura Pollard, Miao He, Michael H. Gelb, and Timothy C. Wood. Multiplex tandem mass spectrometry enzymatic activity assay for the screening and diagnosis of mucolipidosis type ii and iii. Jun 2023. URL: https://doi.org/10.1016/j.ymgmr.2023.100978, doi:10.1016/j.ymgmr.2023.100978. This article has 6 citations.
-
(geer2023unravelingmucolipidosistype pages 2-5): Karl De Geer, Katarzyna Mascianica, Karin Naess, Eliane Sardh, Anna Lindstrand, and Erik Björck. Unraveling mucolipidosis type iii gamma through whole genome sequencing in late-onset retinitis pigmentosa: a case report. BMC Ophthalmology, Sep 2023. URL: https://doi.org/10.1186/s12886-023-03136-4, doi:10.1186/s12886-023-03136-4. This article has 5 citations and is from a peer-reviewed journal.
-
(persichetti2009identificationandmolecular pages 1-2): Emanuele Persichetti, Nadia A. Chuzhanova, Andrea Dardis, Barbara Tappino, Sandra Pohl, Nick S.T. Thomas, Camillo Rosano, Chiara Balducci, Silvia Paciotti, Silvia Dominissini, Anna Lisa Montalvo, Michela Sibilio, Rossella Parini, Miriam Rigoldi, Maja Di Rocco, Giancarlo Parenti, Aldo Orlacchio, Bruno Bembi, David N. Cooper, Mirella Filocamo, and Tommaso Beccari. Identification and molecular characterization of six novel mutations in the udp‐n‐acetylglucosamine‐1‐phosphotransferase gamma subunit (gnptg) gene in patients with mucolipidosis iii gamma. Human Mutation, 30:978-984, Jun 2009. URL: https://doi.org/10.1002/humu.20959, doi:10.1002/humu.20959. This article has 44 citations and is from a domain leading peer-reviewed journal.
-
(geer2023unravelingmucolipidosistype pages 1-2): Karl De Geer, Katarzyna Mascianica, Karin Naess, Eliane Sardh, Anna Lindstrand, and Erik Björck. Unraveling mucolipidosis type iii gamma through whole genome sequencing in late-onset retinitis pigmentosa: a case report. BMC Ophthalmology, Sep 2023. URL: https://doi.org/10.1186/s12886-023-03136-4, doi:10.1186/s12886-023-03136-4. This article has 5 citations and is from a peer-reviewed journal.
-
(westermann2020imbalancedcellularmetabolism pages 42-44): Lena Marie Westermann, Lutz Fleischhauer, Jonas Vogel, Zsuzsa Jenei-Lanzl, Nataniel Floriano Ludwig, Lynn Schau, Fabio Morellini, Anke Baranowsky, Timur A. Yorgan, Giorgia Di Lorenzo, Michaela Schweizer, Bruna de Souza Pinheiro, Nicole Ruas Guarany, Fernanda Sperb-Ludwig, Fernanda Visioli, Thiago Oliveira Silva, Jamie Soul, Gretl Hendrickx, J. Simon Wiegert, Ida V. D. Schwartz, Hauke Clausen-Schaumann, Frank Zaucke, Thorsten Schinke, Sandra Pohl, and Tatyana Danyukova. Imbalanced cellular metabolism compromises cartilage homeostasis and joint function in a mouse model of mucolipidosis type iii gamma. Disease Models & Mechanisms, Nov 2020. URL: https://doi.org/10.1242/dmm.046425, doi:10.1242/dmm.046425. This article has 10 citations and is from a domain leading peer-reviewed journal.
-
(khan2020mucolipidosesoverviewpast pages 3-6): Shaukat A. Khan and Saori C. Tomatsu. Mucolipidoses overview: past, present, and future. International Journal of Molecular Sciences, 21:6812, Sep 2020. URL: https://doi.org/10.3390/ijms21186812, doi:10.3390/ijms21186812. This article has 78 citations.
-
(meel2016mucolipidosisiiignptg pages 1-3): Eline van Meel and Stuart Kornfeld. Mucolipidosis iii gnptg missense mutations cause misfolding of the γ subunit of glcnac‐1‐phosphotransferase. Human Mutation, 37:623-626, Jul 2016. URL: https://doi.org/10.1002/humu.22993, doi:10.1002/humu.22993. This article has 4 citations and is from a domain leading peer-reviewed journal.
-
(sohn2016moleculargeneticsand pages 1-2): Young Bae Sohn. Molecular genetics and diagnostic approach of mucolipidosis ii/iii. ArXiv, 2:13-16, Jun 2016. URL: https://doi.org/10.19125/jmrd.2016.2.1.13, doi:10.19125/jmrd.2016.2.1.13. This article has 0 citations.
-
(hong2023multiplextandemmass pages 2-4): Xinying Hong, Laura Pollard, Miao He, Michael H. Gelb, and Timothy C. Wood. Multiplex tandem mass spectrometry enzymatic activity assay for the screening and diagnosis of mucolipidosis type ii and iii. Jun 2023. URL: https://doi.org/10.1016/j.ymgmr.2023.100978, doi:10.1016/j.ymgmr.2023.100978. This article has 6 citations.
-
(khan2020mucolipidosesoverviewpast pages 1-3): Shaukat A. Khan and Saori C. Tomatsu. Mucolipidoses overview: past, present, and future. International Journal of Molecular Sciences, 21:6812, Sep 2020. URL: https://doi.org/10.3390/ijms21186812, doi:10.3390/ijms21186812. This article has 78 citations.
-
(westermann2020imbalancedcellularmetabolism media df302704): Lena Marie Westermann, Lutz Fleischhauer, Jonas Vogel, Zsuzsa Jenei-Lanzl, Nataniel Floriano Ludwig, Lynn Schau, Fabio Morellini, Anke Baranowsky, Timur A. Yorgan, Giorgia Di Lorenzo, Michaela Schweizer, Bruna de Souza Pinheiro, Nicole Ruas Guarany, Fernanda Sperb-Ludwig, Fernanda Visioli, Thiago Oliveira Silva, Jamie Soul, Gretl Hendrickx, J. Simon Wiegert, Ida V. D. Schwartz, Hauke Clausen-Schaumann, Frank Zaucke, Thorsten Schinke, Sandra Pohl, and Tatyana Danyukova. Imbalanced cellular metabolism compromises cartilage homeostasis and joint function in a mouse model of mucolipidosis type iii gamma. Disease Models & Mechanisms, Nov 2020. URL: https://doi.org/10.1242/dmm.046425, doi:10.1242/dmm.046425. This article has 10 citations and is from a domain leading peer-reviewed journal.