Autosomal Recessive Nonsyndromic Hearing Loss 28
DFNB28 is autosomal recessive nonsyndromic sensorineural hearing loss caused by biallelic pathogenic TRIOBP variants, commonly nonsense or frameshift alleles. Initial Palestinian and Indian/Pakistani families had prelingual severe-to-profound hearing loss. Dutch and Polish reports expanded the spectrum to congenital moderate and peri- or postlingual moderate-to-severe loss. The available families do not establish a general genotype-based prognosis or a uniform rate of progression. TRIOBP-4 and TRIOBP-5 organize densely bundled actin rootlets at the bases of cochlear hair-cell stereocilia. Combined loss of these isoforms in mice prevents rootlet formation, weakens bundle mechanics and precedes stereocilia degeneration. Selective TRIOBP-5 deficiency produces malformed rootlets and also reduces supporting-cell stiffness. These experimental lesions support the human disease mechanism, but their relative contributions and time courses have not been measured in patient cochleae. The widely expressed TRIOBP-1 isoform is spared by the established truncating alleles discussed here; isoform-specific expression alone does not prove why the clinical phenotype is predominantly auditory. Normal vestibular function has been reported in examined patients, without establishing its universal preservation. Cochlear implantation improved aided thresholds in three siblings from one Afghan family, with differing speech-perception outcomes during short follow-up. These results support consideration of implantation according to individual candidacy, rather than a guaranteed outcome for every TRIOBP genotype.
Ask OpenScientist
Ask a research question about Autosomal Recessive Nonsyndromic Hearing Loss 28. OpenScientist will conduct autonomous deep research using the Disorder Mechanisms Knowledge Base and PubMed literature (typically 10-30 minutes).
Do not include personal health information in your question. Questions and results are cached in your browser's local storage.
Classifications
Mappings
Inheritance
1Show evidence (3 references)
Discussions and Knowledge Gaps
2- Thresholds plateau at a moderate level with rootlets present but dysmorphic, matching the stable moderate human course.
- Hearing declines to profound on the same timescale as the TRIOBP-5-null mouse, which would point to a background or allele-specific explanation for the milder human cases.
Show evidence (3 references)
Show evidence (1 reference)
Pathophysiology
9Show evidence (6 references)
Show evidence (3 references)
Show evidence (5 references)
Show evidence (4 references)
Show evidence (3 references)
Show evidence (3 references)
Show evidence (1 reference)
Show evidence (1 reference)
Show evidence (1 reference)
Pathograph
Phenotypes
2Show evidence (5 references)
Show evidence (8 references)
Genetic Associations
1Show evidence (6 references)
Medical Actions
5Show evidence (3 references)
Show evidence (1 reference)
Show evidence (2 references)
Show evidence (1 reference)
Show evidence (1 reference)
Diagnosis
2Show evidence (4 references)
Show evidence (2 references)
Animal Models
3Show evidence (1 reference)
Show evidence (1 reference)
Show evidence (1 reference)
Show evidence (1 reference)
Show evidence (2 references)
Source YAML
click to showname: Autosomal Recessive Nonsyndromic Hearing Loss 28
category: Mendelian
creation_date: "2026-10-01T20:20:26Z"
synonyms:
- DFNB28
- deafness, autosomal recessive type 28
- deafness, autosomal recessive 28
- autosomal recessive deafness 28
- autosomal recessive nonsyndromic deafness 28
- autosomal recessive nonsyndromic deafness type 28
- autosomal recessive nonsyndromic deafness caused by mutation in TRIOBP
- TRIOBP autosomal recessive nonsyndromic deafness
description: >-
DFNB28 is autosomal recessive nonsyndromic sensorineural hearing loss caused by biallelic pathogenic TRIOBP variants, commonly nonsense or frameshift alleles. Initial Palestinian and Indian/Pakistani families had prelingual severe-to-profound hearing loss. Dutch and Polish reports expanded the spectrum to congenital moderate and peri- or postlingual moderate-to-severe loss. The available families do not establish a general genotype-based prognosis or a uniform rate of progression.
TRIOBP-4 and TRIOBP-5 organize densely bundled actin rootlets at the bases of cochlear hair-cell stereocilia. Combined loss of these isoforms in mice prevents rootlet formation, weakens bundle mechanics and precedes stereocilia degeneration. Selective TRIOBP-5 deficiency produces malformed rootlets and also reduces supporting-cell stiffness. These experimental lesions support the human disease mechanism, but their relative contributions and time courses have not been measured in patient cochleae. The widely expressed TRIOBP-1 isoform is spared by the established truncating alleles discussed here; isoform-specific expression alone does not prove why the clinical phenotype is predominantly auditory.
Normal vestibular function has been reported in examined patients, without establishing its universal preservation. Cochlear implantation improved aided thresholds in three siblings from one Afghan family, with differing speech-perception outcomes during short follow-up. These results support consideration of implantation according to individual candidacy, rather than a guaranteed outcome for every TRIOBP genotype.
disease_term:
preferred_term: autosomal recessive nonsyndromic hearing loss 28
term:
id: MONDO:0012355
label: autosomal recessive nonsyndromic hearing loss 28
parents:
- Autosomal Recessive Nonsyndromic Hearing Loss
classifications:
harrisons_chapter:
- classification_value: DISORDER_OF_EAR
- classification_value: GENETICS_ENVIRONMENT_DISEASE
mappings:
mondo_mappings:
- term:
id: MONDO:0012355
label: autosomal recessive nonsyndromic hearing loss 28
mapping_predicate: skos:exactMatch
mapping_source: MONDO
references:
- reference: PMID:16385458
title: "Mutations in a novel isoform of TRIOBP that encodes a filamentous-actin binding protein are responsible for DFNB28 recessive nonsyndromic hearing loss."
- reference: PMID:16385457
title: "Mutations in TRIOBP, which encodes a putative cytoskeletal-organizing protein, are associated with nonsyndromic recessive deafness."
- reference: PMID:20510926
title: "Actin-bundling protein TRIOBP forms resilient rootlets of hair cell stereocilia essential for hearing."
- reference: PMID:31217345
title: "TRIOBP-5 sculpts stereocilia rootlets and stiffens supporting cells enabling hearing."
- reference: PMID:35737845
title: "Unbalanced bidirectional radial stiffness gradients within the organ of Corti promoted by TRIOBP."
- reference: PMID:35175278
title: "ANKRD24 organizes TRIOBP to reinforce stereocilia insertion points."
- reference: PMID:23789641
title: "R1 motif is the major actin-binding domain of TRIOBP-4."
- reference: PMID:40471101
title: "Taperin bundles F-actin at stereocilia pivot points enabling optimal lifelong mechanosensitivity."
- reference: PMID:28089734
title: "Broadening the phenotype of DFNB28: Mutations in TRIOBP are associated with moderate, stable hereditary hearing impairment."
- reference: PMID:29197352
title: "Whole exome sequencing identifies TRIOBP pathogenic variants as a cause of post-lingual bilateral moderate-to-severe sensorineural hearing loss."
- reference: PMID:36029164
title: "Elucidation of repeat motifs R1- and R2-related TRIOBP variants in autosomal recessive nonsyndromic hearing loss DFNB28 among indigenous South African individuals."
- reference: PMID:32877897
title: "A New Pathogenic Variant in the TRIOBP Associated with Profound Deafness Is Remediable with Cochlear Implantation."
- reference: url:https://www.ncbi.nlm.nih.gov/sites/books/NBK1434/?report=reader
title: Genetic Hearing Loss Overview - GeneReviews® - NCBI Bookshelf
tags:
- GeneReviews
- reference: url:https://www.ncbi.nlm.nih.gov/sites/books/NBK1434/pdf/Bookshelf_NBK1434.pdf
title: https://www.ncbi.nlm.nih.gov/sites/books/NBK1434/pdf/Bookshelf_NBK1434.pdf
- reference: DOI:10.1186/s12881-020-01055-5
title: A novel mutation in TRIOBP gene leading to congenital deafness in a Chinese family
- reference: DOI:10.3389/fgene.2021.766973
title: 'Case Report: Novel Compound Heterozygous Variants in TRIOBP Associated With Congenital Deafness in a Chinese Family'
inheritance:
- name: Autosomal recessive
description: >-
Homozygous or compound heterozygous pathogenic TRIOBP variants segregate with hearing loss. The founding independent series included Palestinian families and families from India and Pakistan. Nonsense and frameshift variants have the strongest recurrent evidence; individual missense findings require separate assessment.
inheritance_term:
preferred_term: Autosomal recessive inheritance
term:
id: HP:0000007
label: Autosomal recessive inheritance
evidence:
- reference: PMID:16385458
reference_title: "Mutations in a novel isoform of TRIOBP that encodes a filamentous-actin binding protein are responsible for DFNB28 recessive nonsyndromic hearing loss."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "In seven families, 27 deaf individuals are homozygous for one of the nonsense mutations; in two other families, 3 deaf individuals are compound heterozygous for the two nonsense mutations or for Q581X and G1019R."
explanation: >-
The historical Palestinian series supports recessive segregation. Its abstract calls the second allele paired with G1019R Q581X, whereas the full-text case description and Figure 3C identify R347X. This inconsistency and the uncertain functional significance of G1019R preclude treating that child as a confirmed example of two truncating alleles.
- reference: PMID:16385457
reference_title: "Mutations in TRIOBP, which encodes a putative cytoskeletal-organizing protein, are associated with nonsyndromic recessive deafness."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "In seven families, six different mutant alleles of TRIOBP on chromosome 22q13 cosegregate with autosomal recessive nonsyndromic deafness."
explanation: The independent series identified six truncating alleles segregating in seven Indian/Pakistani families.
- reference: url:https://www.ncbi.nlm.nih.gov/sites/books/NBK1434/?report=reader
reference_title: Genetic Hearing Loss Overview - GeneReviews® - NCBI Bookshelf
supports: SUPPORT
evidence_source: OTHER
quote_role: REVIEW_SYNTHESIS
directness: DIRECT
snippet: If both parents are known to be heterozygous for an autosomal recessive hearing loss-related pathogenic variant, each sib of the proband has at conception a 25% chance of having hearing loss, a 50% chance of having no hearing loss and being a carrier, and a 25% chance of having no hearing loss and not being a carrier.
explanation: The Mendelian recurrence probabilities apply to confirmed heterozygous carrier parents; they are not measured DFNB28 penetrance estimates.
pathophysiology:
- name: TRIOBP-4/5 Loss of Function
description: >-
Biallelic loss-of-function TRIOBP variants disrupt the hearing-relevant TRIOBP-4 and/or TRIOBP-5 isoforms. Established alleles include nonsense and frameshift variants in sequence shared by TRIOBP-4/5 and later truncations affecting TRIOBP-5 alone. Exon numbers depend on the transcript and publication. Complete protein loss, nonsense-mediated decay and residual activity have not been measured for every human allele. The reported mouse embryonic lethality followed combined truncation of TRIOBP-1 and TRIOBP-5, rather than an isolated TRIOBP-1 knockout; its application to human viability or restriction of the clinical phenotype remains a hypothesis. Reported missense findings require separate variant-level assessment.
biological_scale: MOLECULAR
genes:
- preferred_term: TRIOBP
term:
id: hgnc:17009
label: TRIOBP
downstream:
- target: Loss of Dense F-Actin Bundling by TRIOBP-4/5
causal_link_type: DIRECT
- target: Impaired TRIOBP-5 Self-Association
description: Loss of TRIOBP-5 or truncation removing its C-terminal coiled-coil region may impair oligomerization; individual patient alleles were not tested in the cited assay.
causal_link_type: INDIRECT_UNKNOWN_INTERMEDIATES
- target: Disrupted Cuticular Plate Actin Organization
description: TRIOBP-5-deficient mouse hair cells have altered cuticular-plate actin; whether this is direct or secondary to rootlet disruption remains unresolved.
causal_link_type: INDIRECT_UNKNOWN_INTERMEDIATES
- target: Disrupted Supporting-Cell Apical Cytoskeleton
description: TRIOBP-5 deficiency is associated with disorganized supporting-cell apical filament structures in mouse cochlea.
causal_link_type: INDIRECT_UNKNOWN_INTERMEDIATES
evidence:
- reference: PMID:16385457
reference_title: "Mutations in TRIOBP, which encodes a putative cytoskeletal-organizing protein, are associated with nonsyndromic recessive deafness."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "These alleles include four nonsense (Q297X, R788X, R1068X, and R1117X) and two frameshift (D1069fsX1082 and R1078fsX1083) mutations, all located in exon 6 of TRIOBP."
explanation: The founding truncating alleles and their location in the TRIOBP-4/5 exon.
- reference: PMID:20510926
reference_title: "Actin-bundling protein TRIOBP forms resilient rootlets of hair cell stereocilia essential for hearing."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: BACKGROUND
snippet: "To date, all of the mutations of TRIOBP causing human deafness DFNB28 are located in exon 6 (Figure 1B), and only affect TRIOBP-4 and TRIOBP-5 (TRIOBP-4/5)."
explanation: >-
The isoform restriction of the human alleles as of 2010, stated as background in a mouse study. Graded by what the sentence describes (human genotypes) with quote_role BACKGROUND because the paper did not ascertain these patients.
- reference: PMID:20510926
reference_title: Actin-bundling protein TRIOBP forms resilient rootlets of hair cell stereocilia essential for hearing.
supports: SUPPORT
evidence_source: MODEL_ORGANISM
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: To determine the phenotype of mice deficient for TRIOBP-1, two different knockout alleles were generated that simultaneously truncated TRIOBP-1 and TRIOBP-5. Both alleles resulted in embryonic lethality (data not shown).
explanation: The reported lethal genotypes disrupted both TRIOBP-1 and TRIOBP-5. This is a mouse developmental observation; it does not demonstrate human lethality of isolated TRIOBP-1 loss.
- reference: PMID:28089734
reference_title: "Broadening the phenotype of DFNB28: Mutations in TRIOBP are associated with moderate, stable hereditary hearing impairment."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "To our knowledge, p.Gly1672* is the first pathogenic variant identified in DFNB28 that does not affect isoform class TRIOBP-4."
explanation: >-
Extends the allelic spectrum to a truncation affecting TRIOBP-5 only, which is the basis of the isoform-specific genotype-phenotype question recorded under discussions.
- reference: PMID:36029164
reference_title: "Elucidation of repeat motifs R1- and R2-related TRIOBP variants in autosomal recessive nonsyndromic hearing loss DFNB28 among indigenous South African individuals."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "Family TS005 carrying variants c.572delC, p.Pro191Argfs*50, and c.3510_3513dupTGCA, p.Pro1172Cysfs*13, demonstrated perfect cosegregation with the deafness phenotype."
explanation: Compound heterozygous frameshift alleles in a sub-Saharan African family.
- reference: PMID:29197352
reference_title: Whole exome sequencing identifies TRIOBP pathogenic variants as a cause of post-lingual bilateral moderate-to-severe sensorineural hearing loss.
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: The conclusive verification of this prediction needs further study; however, due to a restricted expression of TRIOBP-5 and TRIOBP-4 only to the inner ear and the retina we could not test the processing of TRIOBP transcripts in our patients.
explanation: The Polish study explicitly could not test predicted transcript processing in patients. Neither nonsense-mediated decay nor complete absence of the mutant proteins was demonstrated.
- name: Loss of Dense F-Actin Bundling by TRIOBP-4/5
description: >-
TRIOBP-4 and TRIOBP-5 share an N-terminal region carrying two repeat motifs, R1 and R2, of which R1 is the principal actin-binding domain. Purified TRIOBP-4 packs actin filaments into bundles that are denser than those made by espin, the cross-linker of the stereocilium shaft, and that resemble the rootlet. In the hair cell, the upper rootlet inside the stereocilium is bundled mainly by TRIOBP-4, while the lower rootlet inside the cuticular plate is bundled by both TRIOBP-4 and TRIOBP-5. TRIOBP-5 thickens the rootlet to its mature dimensions. Loss of the relevant isoforms is expected to impair this activity in hair cells and supporting cells; residual bundling by individual truncated human proteins has not been established for every allele.
biological_scale: MOLECULAR
genes:
- preferred_term: TRIOBP
term:
id: hgnc:17009
label: TRIOBP
cell_types:
- preferred_term: cochlear inner hair cell
term:
id: CL:0000589
label: cochlear inner hair cell
- preferred_term: cochlear outer hair cell
term:
id: CL:0000601
label: cochlear outer hair cell
molecular_functions:
- preferred_term: actin filament binding
term:
id: GO:0051015
label: actin filament binding
modifier: DECREASED
biological_processes:
- preferred_term: actin filament bundle assembly
term:
id: GO:0051017
label: actin filament bundle assembly
modifier: DECREASED
downstream:
- target: Abnormal Stereocilia Rootlet Assembly
causal_link_type: DIRECT
evidence:
- reference: PMID:20510926
reference_title: "Actin-bundling protein TRIOBP forms resilient rootlets of hair cell stereocilia essential for hearing."
supports: SUPPORT
evidence_source: IN_VITRO
snippet: "In vitro, purified TRIOBP isoform 4 protein organizes actin filaments into uniquely dense bundles reminiscent of rootlets but distinct from bundles formed by espin, an actin crosslinker in stereocilia."
explanation: >-
The biochemical activity this node names, measured on purified protein. PMID:20510926 is a mixed-source paper and its mouse results are carried by separate items.
- reference: PMID:23789641
reference_title: "R1 motif is the major actin-binding domain of TRIOBP-4."
supports: SUPPORT
evidence_source: IN_VITRO
snippet: "These results indicate that R1 motif is the major actin-binding domain of TRIOBP-4, and the binding of R2 motif with actin filaments is nonspecific."
explanation: Maps the bundling activity to the R1 repeat in the TRIOBP-4/5 N-terminal region.
- reference: PMID:31217345
reference_title: "TRIOBP-5 sculpts stereocilia rootlets and stiffens supporting cells enabling hearing."
supports: SUPPORT
evidence_source: MODEL_ORGANISM
snippet: "Using 3 new Triobp mouse models, we report that TRIOBP-5 is essential for thickening bundles of F-actin in rootlets, establishing their mature dimensions and for stiffening supporting cells of the auditory sensory epithelium."
explanation: >-
The two cellular sites where loss of TRIOBP-5 bundling acts, which is why this node has two downstream edges.
- name: Abnormal Stereocilia Rootlet Assembly
description: >-
Combined loss of TRIOBP-4/5 in mice prevents mature stereocilia rootlet formation, whereas selective TRIOBP-5 deficiency permits malformed rootlets that are thin or fragmented in the cuticular plate and expanded within stereocilia cores. These are distinct assembly defects, not a universal absence of rootlets across all human genotypes. ANKRD24 and TRIOBP-5 have reciprocal localization dependencies in mouse hair cells; re-expression of TRIOBP-5 in mutant cochlear explants restores ANKRD24 localization. TPRN-deficient mice also lose TRIOBP-5 and ANKRD24 from shorter-row rootlets during early development, illustrating a related molecular compartment rather than a second established cause or human modifier of DFNB28.
biological_scale: CELLULAR
cell_types:
- preferred_term: cochlear inner hair cell
term:
id: CL:0000589
label: cochlear inner hair cell
- preferred_term: cochlear outer hair cell
term:
id: CL:0000601
label: cochlear outer hair cell
biological_processes:
- preferred_term: auditory receptor cell stereocilium organization
term:
id: GO:0060088
label: auditory receptor cell stereocilium organization
modifier: DECREASED
downstream:
- target: Reduced Stereocilia Pivot Stiffness
causal_link_type: DIRECT
evidence:
- reference: PMID:20510926
reference_title: "Actin-bundling protein TRIOBP forms resilient rootlets of hair cell stereocilia essential for hearing."
supports: SUPPORT
evidence_source: MODEL_ORGANISM
snippet: "Stereocilia of Triobp(Deltaex8/Deltaex8) mice develop normally but fail to form rootlets and are easier to deflect and damage."
explanation: >-
Rootlets are absent in mice lacking TRIOBP-4/5 even though the stereocilia themselves develop, which places the lesion at the rootlet rather than at stereocilium growth.
- reference: PMID:31217345
reference_title: "TRIOBP-5 sculpts stereocilia rootlets and stiffens supporting cells enabling hearing."
supports: SUPPORT
evidence_source: MODEL_ORGANISM
snippet: "A loss of TRIOBP-5 in mouse results in dysmorphic rootlets that are abnormally thin in the cuticular plate but have increased widths and lengths within stereocilia cores, and causes progressive deafness recapitulating the human phenotype."
explanation: The rootlet lesion when only TRIOBP-5 is lost, the model for the TRIOBP-5-only human alleles.
- reference: PMID:35175278
reference_title: ANKRD24 organizes TRIOBP to reinforce stereocilia insertion points.
supports: SUPPORT
evidence_source: MODEL_ORGANISM
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: In the TriobpΔex9-ex10 line, where exons specific for TRIOBP-5 are deleted without consequence for TRIOBP-1 or TRIOBP-4 expression, ANKRD24 labeling was absent from rootlets of homozygous-null IHCs and OHCs (Fig. 8, P–S) and VHCs (Fig. S5, AA and AB).
explanation: The TRIOBP-5-specific knockout shows loss of ANKRD24 localization in mutant mouse hair cells.
- reference: PMID:40471101
reference_title: "Taperin bundles F-actin at stereocilia pivot points enabling optimal lifelong mechanosensitivity."
supports: SUPPORT
evidence_source: MODEL_ORGANISM
directness: INDIRECT
snippet: "Stereocilia that lack TPRN develop warped rootlets with gradual loss of TRIOBP-5 and ANKRD24 from mechanosensory rows starting postnatally."
explanation: >-
TPRN-deficient mouse hair cells lose TRIOBP-5 and ANKRD24 during development. This contextual comparison does not establish TPRN variants as a cause or modifier of human DFNB28.
- reference: PMID:35175278
reference_title: ANKRD24 organizes TRIOBP to reinforce stereocilia insertion points.
supports: SUPPORT
evidence_source: IN_VITRO
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: Thus, mislocalization of ANKRD24 in the TriobpΔex9–ex10 mouse was reversed by introduction of DsRed-tagged TRIOBP-5, which restored ANKRD24 to its normal location in rootlets.
explanation: Gene-gun transfection of P2-P3 cochlear explants rescued protein localization. This is not an in-vivo hearing rescue or a human treatment.
- name: Reduced Stereocilia Pivot Stiffness
description: >-
In early postnatal mouse cochlear explants lacking TRIOBP-4/5, inner-hair-cell bundles are more compliant to calibrated fluid-jet stimulation and can lose further stiffness after a large deflection. The magnitude depends on stimulus order and the presence of extracellular links. Mechanotransduction responses were preserved in P4-P9 hair-cell recordings before hearing onset, so the initial experimental defect concerns bundle mechanics rather than demonstrated failure of MET-channel assembly. These early ex vivo results do not show preserved transduction after bundle degeneration or directly measure patient hair cells.
biological_scale: CELLULAR
cell_types:
- preferred_term: cochlear inner hair cell
term:
id: CL:0000589
label: cochlear inner hair cell
downstream:
- target: Stereocilia Fusion and Hair Bundle Degeneration
causal_link_type: DIRECT
evidence:
- reference: PMID:20510926
reference_title: "Actin-bundling protein TRIOBP forms resilient rootlets of hair cell stereocilia essential for hearing."
supports: SUPPORT
evidence_source: IN_VITRO
snippet: "We found that TriobpΔex8/Δex8 stereocilia are about twice more flexible as compared to TriobpΔex8/+ stereocilia, when deflected by fluid-jet stimuli of progressively increasing intensity."
explanation: The measured loss of bundle stiffness without rootlets.
- reference: PMID:20510926
reference_title: "Actin-bundling protein TRIOBP forms resilient rootlets of hair cell stereocilia essential for hearing."
supports: SUPPORT
evidence_source: IN_VITRO
snippet: "Taken together, our data suggests that even though the MET machinery is operational in TriobpΔex8/Δex8 hair cells, these cells are unlikely to have normal mechanosensitivity in vivo due to both the decreased pivotal stiffness and increased fragility of the stereocilia bundle."
explanation: >-
Transduction is preserved, so the mechanical defect rather than the channel is the proposed cause of deafness.
- reference: PMID:20510926
reference_title: Actin-bundling protein TRIOBP forms resilient rootlets of hair cell stereocilia essential for hearing.
supports: SUPPORT
evidence_source: IN_VITRO
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: Thus, the absence of rootlets and TRIOBP-4/5 do not interfere with delivery, assembly or function of the MET machinery prior to the onset of hearing.
explanation: The experimental conclusion is limited to pre-hearing mouse hair cells, studied in cochlear explants.
- reference: PMID:31217345
reference_title: TRIOBP-5 sculpts stereocilia rootlets and stiffens supporting cells enabling hearing.
supports: SUPPORT
evidence_source: IN_VITRO
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: Deflections of the TriobpΔEx9-10/ΔEx9-10 IHC bundles did not differ from WT at low stimuli intensities. However, at higher stimuli intensities, the extent of the deflection of TriobpΔEx9-10/ΔEx9-10 IHC bundles was significantly larger compared with WT (Figure 8D and Supplemental Video 3).
explanation: P8 cochlear explants showed increased flexibility under stronger stimulation in the TRIOBP-5-specific knockout; the result depends on assay conditions.
- name: Stereocilia Fusion and Hair Bundle Degeneration
description: >-
Bundles that develop without rootlets then fuse and degenerate. In mice lacking TRIOBP-4/5 this is widespread by postnatal day 16, around the onset of hearing. When only TRIOBP-5 is lost the course is slower: stereocilia, mainly of the tallest outer hair cell row, are lost by about postnatal day 40, plausibly by detachment at fragile pivot points. These mouse timelines cannot predict the human trajectory: the milder reported human genotypes are not equivalent to homozygous TRIOBP-5 deletion, and the reviewed studies do not directly measure patient rootlet or bundle morphology.
biological_scale: CELLULAR
cell_types:
- preferred_term: cochlear inner hair cell
term:
id: CL:0000589
label: cochlear inner hair cell
- preferred_term: cochlear outer hair cell
term:
id: CL:0000601
label: cochlear outer hair cell
biological_processes:
- preferred_term: stereocilium maintenance
term:
id: GO:0120045
label: stereocilium maintenance
modifier: DECREASED
downstream:
- target: Prelingual Profound Sensorineural Hearing Impairment
causal_link_type: INDIRECT_UNKNOWN_INTERMEDIATES
description: Mouse bundle degeneration supports an indirect mechanism for human hearing impairment; patient rootlet morphology and the intervening human trajectory have not been directly measured.
- target: Moderate to Severe Sensorineural Hearing Impairment
causal_link_type: INDIRECT_UNKNOWN_INTERMEDIATES
description: Mouse bundle degeneration supports an indirect mechanism for human hearing impairment; patient rootlet morphology and the intervening human trajectory have not been directly measured.
evidence:
- reference: PMID:20510926
reference_title: "Actin-bundling protein TRIOBP forms resilient rootlets of hair cell stereocilia essential for hearing."
supports: SUPPORT
evidence_source: MODEL_ORGANISM
snippet: "By P16 however, widespread fusion and degeneration of stereocilia was evident throughout TriobpΔex8/Δex8 cochleae (Figures 6C and 6D)."
explanation: Widespread degeneration follows the early developmental period in mice lacking TRIOBP-4/5; some early bundle abnormalities were already detectable.
- reference: PMID:31217345
reference_title: "TRIOBP-5 sculpts stereocilia rootlets and stiffens supporting cells enabling hearing."
supports: SUPPORT
evidence_source: MODEL_ORGANISM
snippet: "By P40, we observed a loss of some stereocilia from OHC hair bundles, predominantly from the longest stereocilia row, perhaps due to their detachment at fragile pivot points near the apical surface of a hair cell (Figure 3D and Supplemental Figure 4)."
explanation: The slower, TRIOBP-5-only course and its proposed mechanism.
- reference: PMID:20510926
reference_title: Actin-bundling protein TRIOBP forms resilient rootlets of hair cell stereocilia essential for hearing.
supports: SUPPORT
evidence_source: MODEL_ORGANISM
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: However, TriobpΔex8 /Δex8 mice did not respond to either a 100 dB sound pressure level (SPL) click or tone bursts at 8 kHz to 32 kHz, indicating that they are profoundly deaf (Figure S4C), recapitulating human DFNB28 deafness.
explanation: Auditory brainstem testing demonstrated profound hearing loss in the combined-isoform-null mouse; this does not quantify hearing in human alleles.
- name: Reduced Apical Stiffness of Organ of Corti Supporting Cells
description: >-
In freshly dissected early postnatal mouse cochlear explants, TRIOBP deficiency reduces the apical stiffness of pillar and Deiters supporting cells. TRIOBP-5-only loss reduces opposing radial stiffness gradients across supporting-cell and hair-cell rows; combined TRIOBP-4/5 loss additionally reverses the hair-cell gradient. Some softening is also present in heterozygous mice with preserved hearing, so reduced stiffness alone is not sufficient evidence of deafness. Its contribution to human DFNB28 is unmeasured, and supporting-cell-specific perturbations are needed to separate this branch from the hair-bundle defect.
biological_scale: TISSUE
cell_types:
- preferred_term: Deiters' cell
term:
id: CL:0000635
label: Deiter's cell
- preferred_term: external pillar cell of cochlea
term:
id: CL:0002164
label: external pillar cell of cochlea
downstream:
- target: Prelingual Profound Sensorineural Hearing Impairment
causal_link_type: INDIRECT_UNKNOWN_INTERMEDIATES
evidence:
- reference: PMID:31217345
reference_title: "TRIOBP-5 sculpts stereocilia rootlets and stiffens supporting cells enabling hearing."
supports: SUPPORT
evidence_source: IN_VITRO
snippet: "Using PFT-AFM, we observed that the absence of TRIOBP-5 in the organ of Corti resulted in a significant decrease in the local axial stiffness of the supporting cells within the reticular lamina."
explanation: PFT-AFM measured reduced local supporting-cell stiffness in live P5 organ-of-Corti explants from TRIOBP-5-deficient mice.
- reference: PMID:35737845
reference_title: "Unbalanced bidirectional radial stiffness gradients within the organ of Corti promoted by TRIOBP."
supports: SUPPORT
evidence_source: IN_VITRO
snippet: "Deafness-associated TRIOBP deficiencies significantly disrupted the magnitude and orientation of these bidirectional radial stiffness gradients."
explanation: >-
PFT-AFM in live P5-P6 cochlear explants demonstrated altered radial stiffness gradients; an independent human causal contribution was not tested.
- reference: PMID:35737845
reference_title: Unbalanced bidirectional radial stiffness gradients within the organ of Corti promoted by TRIOBP.
supports: SUPPORT
evidence_source: MODEL_ORGANISM
quote_role: BACKGROUND
directness: DIRECT
snippet: However, the reduction in stiffness of the reticular lamina in young heterozygote mice does not significantly harm the hearing ability later on, as indicated by auditory brainstem response analyses at P14 (10).
explanation: The authors cite preserved hearing in heterozygous mice despite the softening measured in their explants. This background in-vivo observation limits causal interpretation of the stiffness result alone.
- name: Impaired TRIOBP-5 Self-Association
description: In transfected HeLa cells, TRIOBP-5 self-association depends on its C-terminal coiled-coil region. A construct lacking that region failed to oligomerize in a NanoSPD assay. Oligomerization is proposed to reinforce rootlet actin bundles; the corresponding defect has not been measured for each human truncating allele. This distinguishes a TRIOBP-5-specific function from the R1-mediated actin bundling retained by TRIOBP-4.
biological_scale: MOLECULAR
mechanism_confidence: PROVISIONAL
evidence:
- reference: PMID:31217345
reference_title: TRIOBP-5 sculpts stereocilia rootlets and stiffens supporting cells enabling hearing.
supports: SUPPORT
evidence_source: IN_VITRO
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: This internally deleted TRIOBP-5 variant failed to homo-oligomerize (Figure 7, B and D), consistent with the computational structural model showing that the coiled-coil domains permit homo-oligomerization (Figure 7 and Supplemental Figure 8).
explanation: The engineered coiled-coil deletion was tested by NanoSPD in HeLa cells, not by direct assays of patient proteins.
downstream:
- target: Abnormal Stereocilia Rootlet Assembly
description: The authors propose that coiled-coil-dependent oligomerization helps reinforce and maintain rootlet architecture; the precise causal contribution in patients remains unmeasured.
causal_link_type: INDIRECT_UNKNOWN_INTERMEDIATES
- name: Disrupted Cuticular Plate Actin Organization
description: In TRIOBP-5-deficient mouse outer hair cells, regular cortical actin tangles at the cuticular-plate surface are disrupted. This could be a direct consequence of TRIOBP-5 loss or a secondary response to rootlet abnormalities. Explant measurements show reduced cuticular-plate stiffness, without establishing the independent contribution of this branch to human hearing loss.
biological_scale: CELLULAR
mechanism_confidence: PROVISIONAL
evidence:
- reference: PMID:35737845
reference_title: Unbalanced bidirectional radial stiffness gradients within the organ of Corti promoted by TRIOBP.
supports: SUPPORT
evidence_source: MODEL_ORGANISM
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: The disruption of cuticular plate F-actin in TriobpΔEx9-10/ΔEx9-10 OHCs represents either a direct effect of TRIOBP-5 deficiency or F-actin rearrangement associated with the abnormalities of stereocilia rootlets in these mice (10).
explanation: Mouse ultrastructure identifies an additional actin compartment and explicitly leaves its causal position unresolved.
cell_types:
- preferred_term: cochlear outer hair cell
term:
id: CL:0000601
label: cochlear outer hair cell
downstream:
- target: Prelingual Profound Sensorineural Hearing Impairment
description: Altered cuticular-plate architecture may affect cochlear mechanics; this contribution has not been isolated from the rootlet defect in patients.
causal_link_type: INDIRECT_UNKNOWN_INTERMEDIATES
- name: Disrupted Supporting-Cell Apical Cytoskeleton
description: TRIOBP-5-deficient mouse pillar and Deiters cells have abnormalities in the density and organization of apical filament and microtubule structures. These ultrastructural findings accompany supporting-cell softening, but the experiments do not isolate an actin-only mechanism or prove that this branch independently causes human hearing loss.
biological_scale: CELLULAR
mechanism_confidence: PROVISIONAL
evidence:
- reference: PMID:35737845
reference_title: Unbalanced bidirectional radial stiffness gradients within the organ of Corti promoted by TRIOBP.
supports: SUPPORT
evidence_source: MODEL_ORGANISM
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: Thus, TRIOBP-5 deficiency results in ultrastructural defects in OHCs, PCs, and DCs, which are the three major types of cells that form reticular lamina of the organ of Corti.
explanation: FIB-SEM of mutant mouse cochlea documented cytoskeletal abnormalities in both sensory and supporting cells.
cell_types:
- preferred_term: Deiters' cell
term:
id: CL:0000635
label: Deiter's cell
- preferred_term: external pillar cell of cochlea
term:
id: CL:0002164
label: external pillar cell of cochlea
downstream:
- target: Reduced Apical Stiffness of Organ of Corti Supporting Cells
description: Disordered apical cytoskeletal structures are associated with softer supporting-cell surfaces; the relative mechanical roles of individual filaments remain unresolved.
causal_link_type: INDIRECT_UNKNOWN_INTERMEDIATES
phenotypes:
- name: Prelingual Profound Sensorineural Hearing Impairment
category: Auditory
description: >-
Bilateral prelingual severe-to-profound sensorineural hearing loss characterized the founding families; three Afghan siblings with homozygous p.Arg448* also had profound sensorineural loss. Symmetry, normal vision and absence of a conductive component were specifically documented in Palestinian family K. Those negative findings should not be generalized to every patient or used to exclude coincident conductive disease.
phenotype_term:
preferred_term: Prelingual profound sensorineural hearing impairment
term:
id: HP:0000399
label: Prelingual sensorineural hearing impairment
severity: SEVERE
evidence:
- reference: PMID:16385458
reference_title: "Mutations in a novel isoform of TRIOBP that encodes a filamentous-actin binding protein are responsible for DFNB28 recessive nonsyndromic hearing loss."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "we previously mapped DFNB28--a locus associated with recessively inherited, prelingual, profound sensorineural hearing impairment--to chromosome 22q13.1"
explanation: The phenotype that defined the locus.
- reference: PMID:16385458
reference_title: "Mutations in a novel isoform of TRIOBP that encodes a filamentous-actin binding protein are responsible for DFNB28 recessive nonsyndromic hearing loss."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "Hearing loss in family K was sensorineural, bilateral, symmetrical, and profound; the mode of inheritance appeared to be recessive."
explanation: The audiometric shape in the founding kindred.
- reference: PMID:16385458
reference_title: "Mutations in a novel isoform of TRIOBP that encodes a filamentous-actin binding protein are responsible for DFNB28 recessive nonsyndromic hearing loss."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "No signs of mixed hearing loss were noted, and vision was normal."
explanation: Normal vision and no mixed hearing loss were documented in Palestinian family K; this is a family-level observation, not a universal exclusion.
- reference: PMID:32877897
reference_title: "A New Pathogenic Variant in the TRIOBP Associated with Profound Deafness Is Remediable with Cochlear Implantation."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "Clinical audiometry demonstrated profound sensorineural hearing loss in all 3 affected siblings (2 males and 1 female), and they were implanted unilaterally."
explanation: The same profound phenotype in an Afghan family homozygous for an exon 7 nonsense allele.
- reference: DOI:10.1186/s12881-020-01055-5
reference_title: A novel mutation in TRIOBP gene leading to congenital deafness in a Chinese family
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: Tests of pure tone hearing thresholds showed a severe to profound symmetric hearing loss for the proband and her younger brother.
explanation: Two Chinese siblings with congenital hearing loss had severe-to-profound symmetric impairment, broadening the family evidence without supplying a population frequency.
- name: Moderate to Severe Sensorineural Hearing Impairment
category: Auditory
description: >-
Two unrelated Dutch patients had congenital moderate hearing loss; one had stable audiometry over 15 years. Three Polish siblings had moderate-to-severe hearing loss recognized at ages 3, 4.5 and 12 years. Two had no measured deterioration over two years; follow-up audiometry was unavailable for the youngest, who had passed newborn screening and later also had otitis-related conductive loss. One Dutch patient and the Polish siblings carried a TRIOBP-5-only truncation in trans with a TRIOBP-4/5 truncation. This does not explain every milder case or establish an isoform-based prognostic rule. Normal vestibular function was reported in the Dutch series.
phenotype_term:
preferred_term: Moderate to severe sensorineural hearing impairment
term:
id: HP:0000407
label: Sensorineural hearing impairment
evidence:
- reference: PMID:28089734
reference_title: "Broadening the phenotype of DFNB28: Mutations in TRIOBP are associated with moderate, stable hereditary hearing impairment."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "Here, we describe two isolated cases of Dutch origin with congenital, moderate HI and compound heterozygous mutations in TRIOBP."
explanation: Moderate congenital loss in two compound heterozygotes.
- reference: PMID:28089734
reference_title: "Broadening the phenotype of DFNB28: Mutations in TRIOBP are associated with moderate, stable hereditary hearing impairment."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "Longitudinal audiometric analyses in one of the subjects revealed that HI was stable over a period of 15 years."
explanation: Audiometric stability was documented in one Dutch patient during 15 years of observation; it does not establish lifelong stability for all DFNB28.
- reference: PMID:29197352
reference_title: "Whole exome sequencing identifies TRIOBP pathogenic variants as a cause of post-lingual bilateral moderate-to-severe sensorineural hearing loss."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "causative of nonsyndromic, peri- to postlingual, moderate-to-severe hearing loss in three siblings from a Polish family"
explanation: Later onset and moderate-to-severe degree in a second family carrying the same TRIOBP-5-only allele in trans.
- reference: PMID:29197352
reference_title: Whole exome sequencing identifies TRIOBP pathogenic variants as a cause of post-lingual bilateral moderate-to-severe sensorineural hearing loss.
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: The age of HI onset was 12 y for the proband and 4,5 y and 3 y for the sibling III.1 and III.3, respectively.
explanation: The three Polish siblings had distinct reported ages of onset; these were based on clinical histories and recognition of hearing difficulty.
- reference: PMID:29197352
reference_title: Whole exome sequencing identifies TRIOBP pathogenic variants as a cause of post-lingual bilateral moderate-to-severe sensorineural hearing loss.
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: The 2-year follow-up does not show hearing deterioration in sibling III.1 and III.2, for the patient III.3 the additional audiometric data were not available.
explanation: Stability was observed in two siblings over two years, with no longitudinal audiogram for the third.
- reference: PMID:29197352
reference_title: Whole exome sequencing identifies TRIOBP pathogenic variants as a cause of post-lingual bilateral moderate-to-severe sensorineural hearing loss.
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: Notably, the youngest brother (III.3) passed the newborn hearing screening test, the remaining sibs did not have this test.
explanation: A passed newborn screen in one sibling does not exclude later hearing impairment; it does not establish the onset of a cellular lesion.
- reference: PMID:29197352
reference_title: Whole exome sequencing identifies TRIOBP pathogenic variants as a cause of post-lingual bilateral moderate-to-severe sensorineural hearing loss.
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: Now his HI is classified as mixed comprising of a conductive and sensorineural component.
explanation: The youngest Polish sibling had frequent otitis media and an additional conductive component, demonstrating that coincident conductive disease can occur.
- reference: PMID:28089734
reference_title: 'Broadening the phenotype of DFNB28: Mutations in TRIOBP are associated with moderate, stable hereditary hearing impairment.'
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: Longitudinal audiometric analyses in one of the subjects revealed that HI was stable over a period of 15 years. Vestibular function was normal.
explanation: Normal vestibular function was reported in the small Dutch clinical series; this does not support disease-wide exclusion of vestibular abnormalities.
genetic:
- name: TRIOBP
relationship_type: CAUSATIVE
notes: >-
TRIOBP isoform classes have distinct promoters and coding sequences. TRIOBP-4 and TRIOBP-1 share no coding sequence; the long TRIOBP-5 isoform includes regions represented in both. The established truncating alleles discussed here affect TRIOBP-4/5 or TRIOBP-5 while sparing TRIOBP-1. The original expression study detected the long human transcript in cochlea, retina and fetal brain, so expression should not be described as exclusively inner ear and retina. Nonsense-mediated decay and protein-level consequences of individual patient alleles remain partly predicted. Transcript accession and version are needed when comparing variant coordinates across older studies. On NM_001039141.2, homozygous c.1342C>T (p.Arg448*) was reported in two Chinese siblings, separately from the Afghan implant family. A second Chinese report identified c.1170delC (p.Ser391Profs*488) and c.3764C>G (p.Ser1255*) in a woman with familial reports of bilateral prelingual deafness; detailed audiometry was unavailable, so that report cannot establish a quantified severity or course. The South African TS005 family supports two segregating frameshift alleles, although the deceased father was not genotyped. Lack of
segregation for particular missense variants in TS005/TS067 does not establish that every TRIOBP missense variant is benign.
gene_term:
preferred_term: TRIOBP
term:
id: hgnc:17009
label: TRIOBP
evidence:
- reference: PMID:16385458
reference_title: "Mutations in a novel isoform of TRIOBP that encodes a filamentous-actin binding protein are responsible for DFNB28 recessive nonsyndromic hearing loss."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "The novel long isoform of TRIOBP has a restricted expression profile, including cochlea, retina, and fetal brain, whereas the original short isoform is widely expressed."
explanation: The restricted expression of the long isoform hit by the deafness alleles.
- reference: PMID:29197352
reference_title: "Whole exome sequencing identifies TRIOBP pathogenic variants as a cause of post-lingual bilateral moderate-to-severe sensorineural hearing loss."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "The pathogenic variant p.Gln268Leufs*610 disrupts the TRIOBP-4 and TRIOBP-5 isoforms (both expressed exclusively in the inner ear and retina) whereas the second pathogenic variant c.514G>T, p.Gly1672* affects only TRIOBP-5."
explanation: >-
Isoform assignment of the two alleles in the Polish family. The source writes c.514G>T where the variant is c.5014G>T elsewhere in the same abstract; quoted as published.
- reference: DOI:10.1186/s12881-020-01055-5
reference_title: A novel mutation in TRIOBP gene leading to congenital deafness in a Chinese family
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: It was found that the two siblings carried a homozygous TRIOBP c.1342C > T (p.Arg448*) variant, and each of their parents carried a heterozygous TRIOBP c.1342C > T (p.Arg448*) variant (Fig. 2).
explanation: Segregation of p.Arg448* in a Chinese family provides human genetic evidence independent of the Afghan cochlear-implant family.
- reference: DOI:10.3389/fgene.2021.766973
reference_title: 'Case Report: Novel Compound Heterozygous Variants in TRIOBP Associated With Congenital Deafness in a Chinese Family'
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: Using next generation sequencing and bioinformatics analysis, we identified two novel TRIOBP c.1170delC (p.S391Pfs*488) and c.3764C > G (p.S1255*) variants.
explanation: The report adds a frameshift/nonsense genotype on NM_001039141.2; predicted protein truncation was not experimentally measured in the patient.
- reference: DOI:10.3389/fgene.2021.766973
reference_title: 'Case Report: Novel Compound Heterozygous Variants in TRIOBP Associated With Congenital Deafness in a Chinese Family'
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: Unfortunately, she had not received auditory tests at that time.
explanation: The reported bilateral prelingual phenotype relied on family history; detailed audiometric severity and trajectory cannot be inferred from this case.
- reference: PMID:36029164
reference_title: Elucidation of repeat motifs R1- and R2-related TRIOBP variants in autosomal recessive nonsyndromic hearing loss DFNB28 among indigenous South African individuals.
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: The mother is the carrier of the c.3510_3513dupTGCA variant while it is presumed that the deceased father was the carrier of the c.572delC deletion.
explanation: The South African family supports segregation, but the paternal allele assignment was inferred because the father was deceased.
diagnosis:
- name: TRIOBP sequencing on recessive hearing loss panels or exome
description: >-
Molecular confirmation requires biallelic pathogenic or likely pathogenic TRIOBP variants consistent with the phenotype and recessive inheritance, with parental testing to establish phase where needed. A hearing-loss panel or exome can identify such variants. Later-onset and less severe hearing loss can warrant TRIOBP analysis as well as congenital profound loss; two unclassified variants alone do not establish DFNB28.
evidence:
- reference: PMID:29197352
reference_title: "Whole exome sequencing identifies TRIOBP pathogenic variants as a cause of post-lingual bilateral moderate-to-severe sensorineural hearing loss."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "Although TRIOBP alterations are not a frequent cause of hearing impairment, this gene should be thoroughly analyzed especially in patients with a postlingual hearing loss."
explanation: The testing recommendation and its clinical trigger.
- reference: PMID:29197352
reference_title: Whole exome sequencing identifies TRIOBP pathogenic variants as a cause of post-lingual bilateral moderate-to-severe sensorineural hearing loss.
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: Analysis of parents’ DNA showed that they were both heterozygous carriers, thus establishing an in trans configuration of the variants in the affected children (Fig. 1a).
explanation: Parental segregation confirmed that the two variants were in trans in the Polish family.
- reference: url:https://www.ncbi.nlm.nih.gov/sites/books/NBK1434/?report=reader
reference_title: Genetic Hearing Loss Overview - GeneReviews® - NCBI Bookshelf
supports: SUPPORT
evidence_source: OTHER
quote_role: REVIEW_SYNTHESIS
directness: DIRECT
snippet: The identification of variant(s) of uncertain significance cannot be used to confirm or rule out the diagnosis.
explanation: General genetic hearing-loss guidance supplies the variant-interpretation limit; it is not a DFNB28-specific diagnostic-yield estimate.
- reference: url:https://www.ncbi.nlm.nih.gov/sites/books/NBK1434/pdf/Bookshelf_NBK1434.pdf
reference_title: https://www.ncbi.nlm.nih.gov/sites/books/NBK1434/pdf/Bookshelf_NBK1434.pdf
supports: SUPPORT
evidence_source: OTHER
quote_role: REVIEW_SYNTHESIS
directness: DIRECT
snippet: Recommended molecular genetic testing approaches include use of a multigene hearing loss panel and/or genomic testing
explanation: General GeneReviews guidance supports panel or genomic testing in a genetically heterogeneous hearing-loss presentation.
- name: Audiologic follow-up and evaluation of relatives
description: Serial audiologic assessment documents the individual hearing trajectory and identifies superimposed acquired loss, including otitis media. Relatives with familial risk warrant early diagnostic evaluation even after a passed newborn screen. The Polish family documents this limitation of newborn screening directly; follow-up frequency should be individualized rather than inferred from the mouse degeneration timeline.
evidence:
- reference: url:https://www.ncbi.nlm.nih.gov/sites/books/NBK1434/?report=reader
reference_title: Genetic Hearing Loss Overview - GeneReviews® - NCBI Bookshelf
supports: SUPPORT
evidence_source: OTHER
quote_role: REVIEW_SYNTHESIS
directness: DIRECT
snippet: Permanent childhood hearing loss in a parent or sib increases the probability of hearing loss in other family members and warrants early complete diagnostic evaluation of hearing regardless of the family member's newborn hearing screening result.
explanation: General genetic hearing-loss guidance supports early evaluation of relatives despite a passed newborn screen.
- reference: url:https://www.ncbi.nlm.nih.gov/sites/books/NBK1434/pdf/Bookshelf_NBK1434.pdf
reference_title: https://www.ncbi.nlm.nih.gov/sites/books/NBK1434/pdf/Bookshelf_NBK1434.pdf
supports: SUPPORT
evidence_source: OTHER
quote_role: REVIEW_SYNTHESIS
directness: DIRECT
snippet: 'Perform sequential audiologic examinations that: Document the stability or progression of the hearing loss; Are tailored to the genetic diagnosis and expected prognosis for hearing loss'
explanation: General surveillance guidance supports serial audiometry to establish the individual trajectory.
treatments:
- name: Cochlear Implantation
therapeutic_modality: DEVICE
treatment_term:
preferred_term: cochlear device implantation
term:
id: NCIT:C15329
label: Surgical Procedure
qualifiers:
- predicate:
preferred_term: medical device
term:
id: NCIT:C16830
label: Medical Device
value:
preferred_term: cochlear implant
term:
id: NCIT:C157820
label: Cochlear Implant
description: >-
Cochlear implantation can be considered for severe-to-profound hearing loss according to individual audiologic and anatomic candidacy. It electrically stimulates the auditory nerve downstream of hair-cell transduction. Three Afghan siblings homozygous for p.Arg448* received unilateral implants and reached aided pure-tone averages of 23 to 30 dB HL one month after activation. At ten months the two younger recipients achieved up to 77% correct phonemes, while the oldest could not yet perform open-set speech audiometry. This single-family, short-term series does not establish uniform benefit or localize every human lesion exclusively to the hair bundle.
target_mechanisms:
- target: Stereocilia Fusion and Hair Bundle Degeneration
treatment_effect: BYPASSES
description: >-
Electrical stimulation of the auditory nerve substitutes for transduction by degenerated hair bundles.
evidence:
- reference: PMID:32877897
reference_title: "A New Pathogenic Variant in the TRIOBP Associated with Profound Deafness Is Remediable with Cochlear Implantation."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "One month after activation, the pure-tone averages with the CI processor were between 30 and 23 dBHL."
explanation: Aided thresholds after implantation in DFNB28 patients.
- reference: PMID:32877897
reference_title: "A New Pathogenic Variant in the TRIOBP Associated with Profound Deafness Is Remediable with Cochlear Implantation."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "Ten months after the first activation of the implant, open-set speech audiometry test could be performed for the first time in the 2 younger CI recipients (S5 and S9), and they could identify up to a maximum 77% phonemes correctly."
explanation: Speech perception outcome in the same family.
- reference: PMID:32877897
reference_title: A New Pathogenic Variant in the TRIOBP Associated with Profound Deafness Is Remediable with Cochlear Implantation.
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: The oldest brother (S12) could not yet perform open-set speech audiometry at that moment.
explanation: At ten months, the oldest sibling could not perform the same speech test as the younger recipients; improved aided thresholds did not imply equivalent speech perception.
- name: Hearing Amplification
therapeutic_modality: DEVICE
treatment_term:
preferred_term: hearing aid amplification
term:
id: NCIT:C15315
label: Rehabilitation
qualifiers:
- predicate:
preferred_term: medical device
term:
id: NCIT:C16830
label: Medical Device
value:
preferred_term: hearing aid
term:
id: NCIT:C183182
label: Hearing Aid
description: >-
Hearing aids can be fitted according to residual hearing and audiometric configuration, including moderate-to-severe DFNB28 presentations. Amplification is supportive and does not repair the rootlet defect. The recommendation follows general genetic hearing-loss care; disease-specific hearing-aid outcome estimates are not established by the clinical reports reviewed here.
target_phenotypes:
- preferred_term: Moderate to Severe Sensorineural Hearing Impairment
term:
id: HP:0000407
label: Sensorineural hearing impairment
evidence:
- reference: url:https://www.ncbi.nlm.nih.gov/sites/books/NBK1434/pdf/Bookshelf_NBK1434.pdf
reference_title: https://www.ncbi.nlm.nih.gov/sites/books/NBK1434/pdf/Bookshelf_NBK1434.pdf
supports: SUPPORT
evidence_source: OTHER
quote_role: REVIEW_SYNTHESIS
directness: DIRECT
snippet: Hearing aids (sound amplification), customized by an audiologist to the degree and frequency of hearing loss, can be used in individuals with mild-to-severe hearing loss.
explanation: General genetic hearing-loss guidance supports individualized amplification for residual hearing, without claiming a TRIOBP-specific response rate.
- name: Genetic Counselling
therapeutic_modality: OTHER
treatment_term:
preferred_term: genetic counseling
term:
id: NCIT:C15240
label: Genetic Counseling
description: >-
Offer nondirective genetic counseling and confirm parental genotypes. If both parents are heterozygous carriers of the familial pathogenic TRIOBP variants, each pregnancy has a 25% chance of biallelic hearing loss, a 50% chance of carrier status and a 25% chance of inheriting neither familial variant. Other parental genotype combinations require a separate recurrence assessment. Onset, severity and progression cannot be reliably predicted from the small reported isoform-specific series. Discuss reproductive testing options according to individual preferences.
evidence:
- reference: url:https://www.ncbi.nlm.nih.gov/sites/books/NBK1434/?report=reader
reference_title: Genetic Hearing Loss Overview - GeneReviews® - NCBI Bookshelf
supports: SUPPORT
evidence_source: OTHER
quote_role: REVIEW_SYNTHESIS
directness: DIRECT
snippet: If both parents are known to be heterozygous for an autosomal recessive hearing loss-related pathogenic variant, each sib of the proband has at conception a 25% chance of having hearing loss, a 50% chance of having no hearing loss and being a carrier, and a 25% chance of having no hearing loss and not being a carrier.
explanation: The Mendelian recurrence probabilities apply to confirmed heterozygous carrier parents; they are not measured DFNB28 penetrance estimates.
- reference: url:https://www.ncbi.nlm.nih.gov/sites/books/NBK1434/?report=reader
reference_title: Genetic Hearing Loss Overview - GeneReviews® - NCBI Bookshelf
supports: SUPPORT
evidence_source: OTHER
quote_role: REVIEW_SYNTHESIS
directness: DIRECT
snippet: The optimal time for determination of genetic status and discussion of the availability of prenatal/preimplantation genetic testing is before pregnancy.
explanation: General counseling guidance supports discussing reproductive testing before pregnancy, according to family preferences.
- name: Communication and language support
therapeutic_modality: OTHER
treatment_term:
preferred_term: Communication and language rehabilitation
term:
id: NCIT:C15315
label: Rehabilitation
description: Establish communication goals with the person and family and arrange access to spoken-language, sign-language, educational and audiology services as appropriate. Early access to language is a care priority regardless of the hearing device selected. These recommendations come from general genetic hearing-loss guidance, not a TRIOBP-specific intervention trial.
target_phenotypes:
- preferred_term: Prelingual Profound Sensorineural Hearing Impairment
term:
id: HP:0000399
label: Prelingual sensorineural hearing impairment
- preferred_term: Moderate to Severe Sensorineural Hearing Impairment
term:
id: HP:0000407
label: Sensorineural hearing impairment
evidence:
- reference: url:https://www.ncbi.nlm.nih.gov/sites/books/NBK1434/?report=reader
reference_title: Genetic Hearing Loss Overview - GeneReviews® - NCBI Bookshelf
supports: SUPPORT
evidence_source: OTHER
quote_role: REVIEW_SYNTHESIS
directness: DIRECT
snippet: On initial evaluation of individuals with hearing loss, the goals for communication must be established with a focus on equipping individuals with language and appropriate access to language.
explanation: General hearing-loss guidance prioritizes access to language and individualized communication goals.
- name: Counseling to avoid additional noise injury
therapeutic_modality: OTHER
treatment_term:
preferred_term: Counseling
term:
id: NCIT:C61547
label: Counseling
description: Counsel people with hearing loss to avoid repeated excessive noise exposure. This aims to limit additional acquired injury; neither a DFNB28-specific human noise-susceptibility estimate nor prevention of the inherited rootlet defect is established.
evidence:
- reference: url:https://www.ncbi.nlm.nih.gov/sites/books/NBK1434/?report=reader
reference_title: Genetic Hearing Loss Overview - GeneReviews® - NCBI Bookshelf
supports: SUPPORT
evidence_source: OTHER
quote_role: REVIEW_SYNTHESIS
directness: DIRECT
snippet: Since this risk can be minimized by avoidance, persons with documented hearing loss should be counseled appropriately and repeated overexposure to loud noises should be avoided.
explanation: GeneReviews recommends avoiding additional noise injury in people with hearing loss generally; it does not establish a TRIOBP-specific gene-environment interaction.
animal_models:
- name: Triobp delta-exon8 mouse
species: Mouse
genotype: Triobp Δex8/Δex8 (exon 8, orthologous to human exon 6, replaced by lacZ)
publication: PMID:20510926
description: >-
Ablates TRIOBP-4 and TRIOBP-5 while keeping TRIOBP-1, the same isoform pattern as the founding human alleles. Homozygotes are profoundly deaf; stereocilia develop without rootlets, are more compliant and fragile, and fuse and degenerate by postnatal day 16.
genes:
- preferred_term: TRIOBP
term:
id: hgnc:17009
label: TRIOBP
modeled_mechanisms:
- target: Abnormal Stereocilia Rootlet Assembly
relationship: RECAPITULATES
fidelity: HIGH
model_scale: CELLULAR
description: >-
Same isoforms lost as in the founding human alleles, and the rootlet defect is directly observed.
limitations: >-
No human inner ear tissue has been examined, so that rootlets are absent in patients is inferred from the mouse. The mouse course is complete within weeks.
readouts:
- name: Inner hair cell bundle stiffness to fluid-jet deflection
target: Abnormal Stereocilia Rootlet Assembly
direction: DECREASED
interpretation: Bundles without rootlets are about twice as flexible.
evidence:
- reference: PMID:20510926
reference_title: "Actin-bundling protein TRIOBP forms resilient rootlets of hair cell stereocilia essential for hearing."
supports: SUPPORT
evidence_source: IN_VITRO
snippet: "We found that TriobpΔex8/Δex8 stereocilia are about twice more flexible as compared to TriobpΔex8/+ stereocilia, when deflected by fluid-jet stimuli of progressively increasing intensity."
explanation: The stiffness measurement behind this readout.
evidence:
- reference: PMID:20510926
reference_title: "Actin-bundling protein TRIOBP forms resilient rootlets of hair cell stereocilia essential for hearing."
supports: SUPPORT
evidence_source: MODEL_ORGANISM
snippet: "Stereocilia of Triobp(Deltaex8/Deltaex8) mice develop normally but fail to form rootlets and are easier to deflect and damage."
explanation: The model reproduces the rootlet lesion.
- name: Triobp delta-exon9-10 TRIOBP-5-deficient mouse
species: Mouse
genotype: Triobp ΔEx9-10/ΔEx9-10 (TRIOBP-5 ablated; TRIOBP-4 and TRIOBP-1 retained)
publication: PMID:31217345
description: >-
Removes TRIOBP-5 alone. Rootlets form but are dysmorphic, supporting cells are softer, and hearing declines from residual at four weeks to severe-to-profound by eight weeks. It is the model for the human alleles that truncate TRIOBP-5 only.
genes:
- preferred_term: TRIOBP
term:
id: hgnc:17009
label: TRIOBP
modeled_mechanisms:
- target: Stereocilia Fusion and Hair Bundle Degeneration
relationship: PARTIALLY_RECAPITULATES
fidelity: MODERATE
model_scale: CELLULAR
description: >-
Progressive bundle degeneration and hearing loss after rootlets form abnormally.
limitations: >-
The mouse loss becomes severe to profound within weeks, whereas the human patients with TRIOBP-5-only alleles had moderate or moderate-to-severe loss, stable for 15 years in one. Those patients were compound heterozygous with a second allele, not TRIOBP-5-null on both chromosomes as this mouse is, so the genotypes are not equivalent.
readouts:
- name: Auditory brainstem response threshold
target: Stereocilia Fusion and Hair Bundle Degeneration
direction: INCREASED
interpretation: Residual hearing at four weeks progresses to severe-to-profound loss by eight weeks.
evidence:
- reference: PMID:31217345
reference_title: "TRIOBP-5 sculpts stereocilia rootlets and stiffens supporting cells enabling hearing."
supports: SUPPORT
evidence_source: MODEL_ORGANISM
snippet: "Four-week-old TriobpΔEx9-10/ΔEx9-10 mice also had residual hearing at 8 and 16 kHz that progressed to severe to profound hearing loss by 8 weeks of age (Figure 3B)"
explanation: The functional time course in the TRIOBP-5-null mouse.
evidence:
- reference: PMID:31217345
reference_title: "TRIOBP-5 sculpts stereocilia rootlets and stiffens supporting cells enabling hearing."
supports: SUPPORT
evidence_source: MODEL_ORGANISM
snippet: "A loss of TRIOBP-5 in mouse results in dysmorphic rootlets that are abnormally thin in the cuticular plate but have increased widths and lengths within stereocilia cores, and causes progressive deafness recapitulating the human phenotype."
explanation: The authors' claim that the model is progressive; the fidelity grade reflects the severity mismatch above.
- name: Triobp delta-exon8/YHB226 compound-heterozygous mouse
species: Mouse
genotype: 'Triobp ΔEx8/YHB226: one exon-8 deletion and one exon-17 gene-trap allele'
publication: PMID:31217345
description: >-
This published compound-heterozygous model expresses wild-type TRIOBP-4 from one allele and TRIOBP-1 from the other. Full-length TRIOBP-5 is disrupted, but exon-skipping transcripts and rootlet immunoreactivity suggest residual shortened TRIOBP-5. Rootlets and auditory function deteriorate more slowly than in the exon9-10-null model; this is not a clean dosage-only comparison.
genes:
- preferred_term: TRIOBP
term:
id: hgnc:17009
label: TRIOBP
modeled_mechanisms:
- target: Stereocilia Fusion and Hair Bundle Degeneration
description: The existing compound heterozygote has progressive bundle degeneration and severe-to-profound hearing loss by 12 weeks.
evidence:
- reference: PMID:31217345
reference_title: TRIOBP-5 sculpts stereocilia rootlets and stiffens supporting cells enabling hearing.
supports: SUPPORT
evidence_source: MODEL_ORGANISM
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: At 4 and 8 weeks of age, TriobpΔEx8/YHB226 mice have residual hearing at 8 and 16 kHz. By 12 weeks of age hearing loss was severe to profound at all frequencies tested (Figure 3A).
explanation: The in-vivo ABR trajectory is slower than the exon9-10-null model.
- reference: PMID:31217345
reference_title: TRIOBP-5 sculpts stereocilia rootlets and stiffens supporting cells enabling hearing.
supports: SUPPORT
evidence_source: MODEL_ORGANISM
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: A similar but slower progressive degeneration of the stereocilia bundles was observed in the TriobpΔEx8/YHB226 TRIOBP-5–deficient mouse (Figure 3C and Supplemental Figure 5), consistent with a milder progressive loss of hearing.
explanation: The source directly reports the progressive bundle degeneration modeled by this link.
relationship: PARTIALLY_RECAPITULATES
model_scale: CELLULAR
limitations: The YHB226 allele may generate an internally truncated TRIOBP-5 protein. Residual protein and different TRIOBP-1 dosage confound comparison with human compound heterozygotes; human progression is not inferred from the mouse timeline.
fidelity: MODERATE
discussions:
- discussion_id: triobp5_only_alleles_and_milder_phenotype
kind: KNOWLEDGE_GAP
prompt: >-
Do alleles that truncate TRIOBP-5 but leave TRIOBP-4 intact cause a milder, later-onset or stable hearing loss, and if so, why does the TRIOBP-5-null mouse become profoundly deaf?
attaches_to:
- phenotypes#Moderate to Severe Sensorineural Hearing Impairment
- genetic#TRIOBP
- animal_models#Triobp delta-exon9-10 TRIOBP-5-deficient mouse
rationale: >-
A TRIOBP-5-only p.Gly1672* allele occurred in one Dutch patient and the Polish siblings, in trans with an allele affecting TRIOBP-4/5. The Dutch study also described another patient with moderate loss, and its authors stated that variant location might contribute in only one of their two cases. Preserving one TRIOBP-4-encoding copy therefore cannot explain the entire milder spectrum. The small number of families cannot separate isoform, allele-specific and background effects. The mouse lacking TRIOBP-5 on both alleles progresses to severe-to-profound loss within two months, which is a different genotype from those compound heterozygous patients. A published TriobpΔEx8/YHB226 compound heterozygote already expresses one wild-type TRIOBP-4 copy and has slower degeneration than the exon9-10-null mouse, but exon-skipping transcripts and TRIOBP-5 rootlet immunoreactivity suggest a residual shortened protein. A cleaner comparison must distinguish TRIOBP-4 dosage from residual TRIOBP-5 function and preserve comparable TRIOBP-1 expression.
evidence:
- reference: PMID:28089734
reference_title: "Broadening the phenotype of DFNB28: Mutations in TRIOBP are associated with moderate, stable hereditary hearing impairment."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "This suggests that a single TRIOBP copy to encode wildtype TRIOBP-4 is insufficient for normal hearing, and that at least one TRIOBP copy to encode TRIOBP-5 is indispensable for normal inner ear function."
explanation: The authors' isoform hypothesis, which this discussion records as open.
- reference: PMID:28089734
reference_title: "Broadening the phenotype of DFNB28: Mutations in TRIOBP are associated with moderate, stable hereditary hearing impairment."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "Predicted effects of the mutations do not explain the relatively mild phenotype in the presented subjects, whereas location of the mutation might well contribute to the milder HI in one of the subjects."
explanation: The same authors' caution that the genotype does not fully account for the phenotype.
- reference: PMID:31217345
reference_title: TRIOBP-5 sculpts stereocilia rootlets and stiffens supporting cells enabling hearing.
supports: SUPPORT
evidence_source: MODEL_ORGANISM
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: RT-PCR analysis of TriobpΔEx8/YHB226 mouse inner ear mRNA detected Triobp-5 transcripts skipping exons 17 to 23 or 17 to 24 (Supplemental Figure 8).
explanation: The published compound-heterozygous mouse is not a verified complete TRIOBP-5 null; immunoreactivity further suggests a residual protein.
proposed_experiments:
- experiment_id: exp_dfnb28_triobp5_only_compound_heterozygous_mouse
name: Separate TRIOBP-4 dosage from residual TRIOBP-5 activity
description: >-
Compare a verified TRIOBP-5-specific null/TRIOBP-4/5-null compound heterozygote with the published TriobpΔEx8/YHB226 and exon9-10-null models. Measure transcripts and proteins to exclude residual TRIOBP-5 and control TRIOBP-1 expression before comparing longitudinal ABR, DPOAE and rootlet ultrastructure. This tests a dosage hypothesis without claiming the existing gene-trap model is a clean null.
would_support:
- phenotypes#Moderate to Severe Sensorineural Hearing Impairment
supporting_outcome:
- >-
Thresholds plateau at a moderate level with rootlets present but dysmorphic, matching the stable moderate human course.
would_refute:
- phenotypes#Moderate to Severe Sensorineural Hearing Impairment
refuting_outcome:
- >-
Hearing declines to profound on the same timescale as the TRIOBP-5-null mouse, which would point to a background or allele-specific explanation for the milder human cases.
- discussion_id: triobp_supporting_cell_contribution
kind: HUMAN_MODEL_MISMATCH
prompt: >-
How much of DFNB28 hearing loss comes from softened supporting cells rather than from the hair bundle lesion, and does the supporting-cell defect exist in human cochleae?
attaches_to:
- pathophysiology#Reduced Apical Stiffness of Organ of Corti Supporting Cells
rationale: >-
The supporting-cell stiffness change was measured with atomic force microscopy in freshly dissected early postnatal mouse cochleae. Both TRIOBP-deficient mouse lines also lose their hair bundles, so the two lesions are confounded in every animal studied, and no method used in patients reads out reticular lamina mechanics. Improved aided thresholds after cochlear implantation in three siblings from one family are compatible with enough auditory neural function to benefit from electrical stimulation. The short follow-up and differing speech-perception outcomes do not establish universal neural preservation, and implantation does not measure supporting-cell mechanics. The edge from this node to hearing loss is therefore recorded with unknown intermediates.
evidence:
- reference: PMID:31217345
reference_title: "TRIOBP-5 sculpts stereocilia rootlets and stiffens supporting cells enabling hearing."
supports: SUPPORT
evidence_source: MODEL_ORGANISM
snippet: "Thus, in addition to hair cell stereocilia rootlets, TRIOBP-5 provides mechanical resilience to Deiters’ and pillar supporting cells of the organ of Corti that are also crucial for normal sound transduction."
explanation: The proposal that the supporting-cell defect contributes, stated in the mouse study.
References & Deep Research
References
16Deep Research
1Deep research results are used as seeds for research; they do not undergo the same validation as the main records and may contain errors. How we use deep research.
Evaluations and curation notes (1)
Create: Autosomal Recessive Nonsyndromic Hearing Loss 28 (DFNB28, TRIOBP) · 2026-10-01T20:51:02Z · View source
De novo curation of DFNB28 (MONDO:0012355), biallelic TRIOBP. Stub stubs/Autosomal_Recessive_Nonsyndromic_Hearing_Loss_28.yaml deleted. entry_type decision: DISEASE. One gene, MONDO records no descendants, and no existing entry covers TRIOBP (git grep of origin/main found TRIOBP only as a rootlet protein mentioned in the DFNB79/TPRN entry). Sources: own PubMed sweep ("TRIOBP AND (deafness OR hearing)", 48 records; "DFNB28", 12 records) plus an Edison/falcon deep-research report (research/Autosomal_Recessive_Nonsyndromic_Hearing_Loss_28-deep-research-falcon.md; 14/14 references verified, no confabulated terms). just preflight-dr could not run because it needs the local MONDO build, which was not fetched for disk reasons; a manual entity check found TRIOBP mentioned 59 times in the report and the report's phenotype OMIM 609823 matches MONDO's OMIM xref. The report added no claim that is not already in the entry from primary sources. Pathograph: TRIOBP-4/5 loss of function -> loss of dense F-actin bundling (GO:0051015 and GO:0051017 DECREASED) -> failure of stereocilia rootlet formation -> reduced pivot stiffness and fragility -> stereocilia fusion and bundle degeneration -> both hearing phenotypes. A second branch from the bundling node runs to reduced supporting-cell apical stiffness, linked to hearing loss with INDIRECT_UNKNOWN_INTERMEDIATES because its human contribution is unmeasured (recorded as a HUMAN_MODEL_MISMATCH discussion). Consistency with DFNB79: the rootlet node quotes the same PMID:40471101 sentence as DFNB79 about TPRN loss removing TRIOBP-5 and ANKRD24 from rootlets. The milder TRIOBP-5-only presentation (Dutch, Polish families) is curated as a separate phenotype and its isoform explanation recorded as a KNOWLEDGE_GAP, since both families share one allele (p.Gly1672*) in compound heterozygous form and the TRIOBP-5-null mouse becomes severe-to-profound. No GeneReviews or StatPearls chapter exists (just check-genereviews: NO_CHAPTER). Hearing aids and genetic counselling carry no evidence items, for the reasons stated in their notes. Uncited DOI cache files written by the deep-research reference validation were not committed. Validation: just validate, count-verified-snippets (42/42), validate-terms, check-duplicate-keys, check-entity-refs, check-causal-targets, check-qualifier-terms, check-enum-values, check-snippet-length, check-title-snippets, check-snippet-grading, check-coarse-phenotypes, check-reference-titles, check-folded-hyphens, check-environmental-evidence, check_gene_activity_grounding and validate-disorders.
Question: You are an expert researcher providing comprehensive, well-cited information.
Provide detailed information focusing on: 1. Key concepts and definitions with current understanding 2. Recent developments and latest research (prioritize 2023-2024 sources) 3. Current applications and real-world implementations 4. Expert opinions and analysis from authoritative sources 5. Relevant statistics and data from recent studies
Format as a comprehensive research report with proper citations. Include URLs and publication dates where available. Always prioritize recent, authoritative sources and provide specific citations for all major claims.
Disease Characteristics Research Template
Target Disease
- Disease Name: Autosomal Recessive Nonsyndromic Hearing Loss 28
- MONDO ID: MONDO:0012355 (if available)
- Category: Mendelian
Research Objectives
Please provide a comprehensive research report on Autosomal Recessive Nonsyndromic Hearing Loss 28 covering all of the disease characteristics listed below. This report will be used to populate a disease knowledge base entry. Be thorough and cite primary literature (PMID preferred) for all claims.
For each section, suggested databases/resources are listed. These are the first places you should search for information on each topic.
1. Disease Information
Search first: OMIM, Orphanet, ICD-10/ICD-11, MeSH, PubMed
- What is the disease? Provide a concise overview.
- What are the key identifiers? (OMIM, Orphanet, ICD-10/ICD-11, MeSH, Mondo)
- What are the common synonyms and alternative names?
- Is the information derived from individual patients (e.g., EHR) or aggregated disease-level resources?
2. Etiology
- Disease Causal Factors: What are the primary causes? (genetic, environmental, infectious, mechanistic)
- Risk Factors:
Search first: PubMed, Cochrane Library, UpToDate, clinical guidelines, ClinVar, ClinGen, GWAS Catalog, PheGenI, CTD, CDC, WHO, epidemiological databases
- Genetic risk factors (causal variants, susceptibility loci, modifier genes)
- Environmental risk factors (toxins, lifestyle, occupational exposures, age, sex, family history)
- Protective Factors:
Search first: PubMed, Cochrane Library, clinical trial databases, GWAS Catalog, gnomAD, WHO, CDC, nutrition databases
- Genetic protective factors (protective variants, modifier alleles)
- Environmental protective factors (diet, lifestyle, exposures that reduce risk)
- Gene-Environment Interactions: How do genetic and environmental factors interact to influence disease?
Search first: CTD, PubMed, PheGenI, GxE databases
3. Phenotypes
Search first: HPO (Human Phenotype Ontology), OMIM, Orphanet, PubMed, clinicaltrials.gov, MedDRA, SNOMED CT, DECIPHER, LOINC
For each phenotype, provide: - Phenotype type: symptoms, clinical signs, physical manifestations, behavioral changes, or laboratory abnormalities
For symptoms/signs: HPO, OMIM, Orphanet, PubMed For behavioral changes: HPO, DSM, RDoC (Research Domain Criteria), PubMed For laboratory abnormalities: LOINC, SNOMED CT, LabTests Online, PubMed - Phenotype characteristics: Search first: OMIM, Orphanet, HPO, PubMed - Age of symptom onset (neonatal, childhood, adult-onset, late-onset) - Symptom severity (mild, moderate, severe, variable) - Symptom progression (stable, progressive, episodic, fluctuating) - Frequency among affected individuals (percentage or qualitative) - Quality of life impact: Effects on daily functioning and well-being (per-phenotype when possible) Search first: EQ-5D database, SF-36, WHO QOL databases, PubMed - Suggest HPO (Human Phenotype Ontology) terms for each phenotype
4. Genetic/Molecular Information
- Causal Genes: Gene mutations or chromosomal abnormalities responsible for disease (gene symbols, OMIM IDs)
Search first: OMIM, ClinVar, HGMD, Ensembl, NCBI Gene
- Pathogenic Variants:
- Affected genes (gene symbols, HGNC IDs) > Search first: OMIM, NCBI Gene, Ensembl, HGNC, UniProt, GeneCards
- Variant classification (pathogenic, likely pathogenic, VUS per ACMG/AMP guidelines) > Search first: ClinVar, ClinGen, ACMG/AMP guidelines, VarSome
- Variant type/class (missense, frameshift, nonsense, splice-site, structural)
- Allele frequency in population databases > Search first: gnomAD, 1000 Genomes, ExAC, TOPMed, dbSNP
- Somatic vs germline origin > Search first: COSMIC (somatic), ClinVar, ICGC, TCGA
- Functional consequences (loss of function, gain of function, dominant negative)
- Modifier Genes: Genes that modify disease severity or expression
- Epigenetic Information: DNA methylation, histone modifications, chromatin changes affecting disease
Search first: ENCODE, Roadmap Epigenomics, MethBase, DiseaseMeth
- Chromosomal Abnormalities: Large-scale genetic changes (aneuploidy, translocations, inversions)
Search first: DECIPHER, ClinVar, ECARUCA, UCSC Genome Browser
5. Environmental Information
- Environmental Factors: Non-genetic contributing factors (toxins, radiation, pollution, occupational exposure)
Search first: CTD (Comparative Toxicogenomics Database), TOXNET, PubMed, EPA databases
- Lifestyle Factors: Behavioral factors (smoking, diet, exercise, alcohol consumption)
Search first: CDC databases, WHO, PubMed, NHANES
- Infectious Agents: If applicable, pathogens causing or triggering disease (bacteria, viruses, fungi, parasites)
Search first: NCBI Taxonomy, ViPR, BV-BRC, MicrobeDB, GIDEON
6. Mechanism / Pathophysiology
Present this section as an ordered causal chain first, then the detail below. Open with a numbered sequence of mechanistic steps running from the initiating lesion (mutation, exposure, infection) to the clinical manifestation, one step per line, each naming what it causes next. State the causal verb explicitly ("leads to", "results in") and say where a step is inferred rather than demonstrated. Where the mechanism branches, show the branch. The categories below are a checklist of what to cover within those steps, not the organizing structure — a step may draw on several of them, and a category may contribute to several steps.
- Molecular Pathways: Specific signaling cascades or biochemical pathways involved (Wnt, MAPK, mTOR, PI3K-AKT, etc.)
Search first: KEGG, Reactome, WikiPathways, PathBank, BioCyc
- Cellular Processes: Cell-level mechanisms (apoptosis, autophagy, cell cycle dysregulation, inflammation, etc.)
Search first: Gene Ontology (GO), Reactome, KEGG, PubMed
- Protein Dysfunction: How protein structure or function is altered (misfolding, aggregation, loss of function, gain of function)
Search first: UniProt, PDB (Protein Data Bank), InterPro, Pfam, AlphaFold
- Metabolic Changes: Alterations in metabolic processes (energy metabolism, lipid metabolism, amino acid metabolism)
Search first: KEGG, BioCyc, HMDB (Human Metabolome Database), BRENDA
- Immune System Involvement: Role of immune response (autoimmunity, immunodeficiency, chronic inflammation)
Search first: ImmPort, Immunome Database, IEDB, Gene Ontology
- Tissue Damage Mechanisms: How tissues/ are injured (oxidative stress, ischemia, fibrosis, necrosis)
Search first: PubMed, Gene Ontology, Reactome
- Biochemical Abnormalities: Specific molecular defects (enzyme deficiencies, receptor dysfunction, ion channel defects)
Search first: BRENDA, UniProt, KEGG, OMIM, PubMed
- Epigenetic Changes: DNA methylation, histone modifications affecting gene expression in disease
Search first: ENCODE, Roadmap Epigenomics, MethBase, DiseaseMeth
- Molecular Profiling (if available):
- Transcriptomics/gene expression changes > Search first: GEO (Gene Expression Omnibus), ArrayExpress, GTEx, Human Cell Atlas, SRA
- Proteomics findings > Search first: PRIDE, ProteomeXchange, Human Protein Atlas, STRING, BioGRID
- Metabolomics signatures > Search first: MetaboLights, Metabolomics Workbench, HMDB, METLIN
- Lipidomics alterations > Search first: LIPID MAPS, SwissLipids, LipidHome, Metabolomics Workbench
- Genomic structural features > Search first: UCSC Genome Browser, Ensembl, NCBI, dbVar, DGV
- Advanced Technologies (if applicable):
- Single-cell analysis findings (cell-type specific mechanisms, cellular heterogeneity) > Search first: Human Cell Atlas, Single Cell Portal, GEO, CELLxGENE
- Spatial transcriptomics findings > Search first: GEO, Spatial Research, Vizgen, 10x Genomics data
- Multi-omics integration results > Search first: TCGA, ICGC, cBioPortal, LinkedOmics, PubMed
- Functional genomics screens (CRISPR, RNAi) > Search first: DepMap, GenomeRNAi, PubMed, BioGRID ORCS
For each mechanism, describe: - The causal chain from initial trigger to clinical manifestation - Which mechanisms are upstream vs downstream - What cell types and biological processes are involved - Suggest GO terms for biological processes and CL terms for cell types
7. Anatomical Structures Affected
- Organ Level:
- Primary organs directly affected
- Secondary organ involvement (complications, secondary effects)
- Body systems involved (cardiovascular, nervous, digestive, respiratory, endocrine, etc.)
Search first: Uberon, FMA (Foundational Model of Anatomy), OMIM, HPO, ICD-11, MeSH, SNOMED CT
- Tissue and Cell Level:
- Specific tissue types affected (epithelial, connective, muscle, nervous)
- Specific cell populations targeted (with Cell Ontology terms)
Search first: Uberon, Human Protein Atlas, Cell Ontology, Human Cell Atlas, CellMarker, PanglaoDB
- Subcellular Level:
- Cellular compartments involved (mitochondria, nucleus, ER, lysosomes) (with GO Cellular Component terms)
Search first: Gene Ontology (Cellular Component), UniProt, Human Protein Atlas
- Localization:
- Specific anatomical sites (with UBERON terms) > Search first: FMA, Uberon, NeuroNames (for brain), SNOMED CT
- Lateralization (unilateral, bilateral, asymmetric) > Search first: HPO, clinical literature, imaging databases
8. Temporal Development
- Onset:
- Typical age of onset (congenital, pediatric, adult, geriatric)
- Onset pattern (acute, subacute, chronic, insidious)
Search first: OMIM, Orphanet, HPO, PubMed
- Progression:
- Disease stages (early, intermediate, advanced, end-stage) > Search first: Cancer Staging Manual (AJCC), WHO classifications, PubMed
- Progression rate (rapid, slow, variable)
- Disease course pattern (episodic, relapsing-remitting, progressive, stable)
- Disease duration (self-limited, chronic lifelong)
Search first: Disease registries, longitudinal cohort databases, natural history studies, PubMed, Orphanet, OMIM
- Patterns:
- Remission patterns (spontaneous, treatment-induced) > Search first: Clinical trial databases, disease registries, PubMed
- Critical periods (time windows of vulnerability or opportunity for intervention) > Search first: PubMed, developmental biology databases, clinical guidelines
9. Inheritance and Population
- Epidemiology:
- Prevalence (cases per 100,000 at given time)
- Incidence (new cases per 100,000 per year)
Search first: Orphanet, CDC, WHO, GBD (Global Burden of Disease), national registries, SEER, disease registries
- For Genetic Etiology:
- Inheritance pattern (AD, AR, X-linked, mitochondrial, multifactorial, polygenic) > Search first: OMIM, Orphanet, ClinVar, GTR (Genetic Testing Registry)
- Penetrance (complete, incomplete, age-dependent) > Search first: ClinVar, OMIM, PubMed, ClinGen
- Expressivity (variable, consistent) > Search first: OMIM, ClinVar, PubMed
- Genetic anticipation (increasing severity in successive generations) > Search first: OMIM, PubMed (especially for repeat expansion disorders)
- Germline mosaicism > Search first: ClinVar, OMIM, genetic counseling literature, PubMed
- Founder effects (population-specific mutations) > Search first: gnomAD, population genetics databases, PubMed
- Consanguinity role > Search first: OMIM, population studies, genetic counseling resources
- Carrier frequency > Search first: gnomAD, carrier screening databases, GeneReviews, GTR
- Population Demographics:
- Affected populations (ethnic or demographic groups with higher prevalence) > Search first: gnomAD, 1000 Genomes, PAGE Study, PubMed, population registries
- Geographic distribution (endemic areas, regional variation) > Search first: WHO, CDC, GBD, Orphanet, geographic epidemiology databases
- Geographic distribution of specific variants
- Sex ratio (male:female) > Search first: Disease registries, OMIM, PubMed, epidemiological databases
- Age distribution of affected individuals > Search first: CDC, disease registries, SEER, Orphanet
10. Diagnostics
- Clinical Tests:
- Laboratory tests (blood, urine, tissue chemistry, specific enzyme assays) > Search first: LOINC, LabTests Online, PubMed
- Biomarkers (proteins, metabolites, genetic markers, circulating biomarkers) > Search first: FDA Biomarker List, BEST (Biomarkers, EndpointS, and other Tools), PubMed
- Imaging studies (X-ray, CT, MRI, PET, ultrasound) > Search first: RadLex, DICOM, Radiopaedia, imaging databases
- Functional tests (pulmonary function, cardiac stress tests) > Search first: LOINC, clinical guidelines, PubMed
- Electrophysiology (EEG, EMG, ECG, nerve conduction studies) > Search first: LOINC, clinical neurophysiology databases, PubMed
- Biopsy findings (histopathology, immunohistochemistry) > Search first: SNOMED CT, College of American Pathologists resources, PubMed
- Pathology findings (microscopic examination) > Search first: SNOMED CT, Digital Pathology databases, PubMed
- Genetic Testing:
Search first: GTR (Genetic Testing Registry), GeneReviews, ClinGen
- Overview of recommended genetic testing approach
- Whole genome sequencing (WGS) utility > Search first: GTR, ClinVar, GEL (Genomics England), gnomAD
- Whole exome sequencing (WES) utility > Search first: GTR, ClinVar, OMIM, GeneMatcher
- Gene panels (which panels, which genes) > Search first: GTR, ClinVar, laboratory-specific databases
- Single gene testing > Search first: GTR, ClinVar, OMIM, GeneReviews
- Chromosomal microarray (CMA) > Search first: DECIPHER, ClinVar, dbVar, ECARUCA
- Karyotyping > Search first: Chromosome Abnormality Database, ClinVar, cytogenetics resources
- FISH > Search first: ClinVar, cytogenetics databases, PubMed
- Mitochondrial DNA testing > Search first: MITOMAP, MSeqDR, ClinVar, GTR
- Repeat expansion testing > Search first: GTR, ClinVar, repeat expansion databases, PubMed
- Omics-Based Diagnostics (if applicable):
- RNA sequencing / transcriptomics > Search first: GEO, ArrayExpress, GTEx, RNA-seq databases
- Proteomics > Search first: PRIDE, ProteomeXchange, FDA Biomarker database
- Metabolomics > Search first: MetaboLights, Metabolomics Workbench, HMDB
- Epigenomics > Search first: GEO, ENCODE, Roadmap Epigenomics, MethBase
- Liquid biopsy > Search first: COSMIC, ClinVar, liquid biopsy databases, PubMed
- Clinical Criteria:
- Standardized diagnostic criteria (DSM, ICD, society guidelines) > Search first: DSM-5, ICD-11, clinical society guidelines, UpToDate
- Differential diagnosis (other conditions to rule out, with distinguishing features) > Search first: DynaMed, UpToDate, clinical decision support systems
- Screening:
- Screening methods for asymptomatic individuals (newborn screening, carrier screening, cascade screening) > Search first: ACMG recommendations, CDC newborn screening, GTR
11. Outcome/Prognosis
- Survival and Mortality:
- Survival rate (5-year, 10-year, overall) > Search first: SEER, cancer registries, disease-specific registries, PubMed
- Life expectancy (with and without treatment if applicable) > Search first: Orphanet, disease registries, actuarial databases, PubMed
- Mortality rate > Search first: CDC, WHO, GBD, national mortality databases
- Disease-specific mortality (deaths directly attributable to disease) > Search first: Disease registries, CDC Wonder, GBD, PubMed
- Morbidity and Function:
- Morbidity (disease-related disability and health impacts) > Search first: GBD, WHO, disability databases, PubMed
- Disability outcomes (long-term functional impairments) > Search first: ICF (International Classification of Functioning), disability registries
- Quality of life measures (EQ-5D, SF-36, PROMIS, disease-specific tools) > Search first: EQ-5D database, SF-36, PROMIS, PubMed
- Disease Course:
- Complications (secondary problems: infections, organ failure, etc.) > Search first: ICD codes, disease registries, clinical databases, PubMed
- Recovery potential (likelihood and extent of recovery, with vs without treatment) > Search first: Natural history studies, rehabilitation databases, PubMed
- Prediction:
- Prognostic factors (age, disease severity, biomarkers, treatment response) > Search first: Prognostic models databases, clinical calculators, PubMed
- Prognostic biomarkers (molecular markers predicting disease course) > Search first: FDA Biomarker database, PubMed, cancer prognostic databases
12. Treatment
- Pharmacotherapy:
- Pharmacological treatments (drug names, drug classes, mechanisms of action) > Search first: DrugBank, RxNorm, ATC classification, DailyMed, FDA databases
- Pharmacogenomics (how genetic variants affect drug metabolism, efficacy, toxicity) > Search first: PharmGKB, CPIC (Clinical Pharmacogenetics), FDA Table of PGx Biomarkers
- Advanced Therapeutics:
- Gene therapy (viral vectors, CRISPR, gene replacement, gene editing) > Search first: ClinicalTrials.gov, FDA gene therapy database, ASGCT resources
- Cell therapy (stem cell transplant, CAR-T, cellular therapeutics) > Search first: ClinicalTrials.gov, FDA cell therapy database, FACT standards
- RNA-based therapies (ASOs, siRNA, mRNA therapies) > Search first: ClinicalTrials.gov, FDA approvals, PubMed
- Targeted therapies (treatments directed at specific molecular targets) > Search first: My Cancer Genome, OncoKB, ClinicalTrials.gov, FDA approvals
- Immunotherapies (checkpoint inhibitors, monoclonal antibodies) > Search first: Cancer Immunotherapy Database, FDA approvals, ClinicalTrials.gov
- Surgical and Interventional:
- Surgical interventions (types of surgery, timing, outcomes) > Search first: CPT codes, surgical registries, clinical guidelines, PubMed
- Supportive and Rehabilitative:
- Supportive care (symptom management, pain control, nutrition) > Search first: Clinical guidelines, Cochrane Library, PubMed
- Rehabilitation (physical therapy, occupational therapy, speech therapy) > Search first: Rehabilitation medicine databases, clinical guidelines, PubMed
- Experimental:
- Experimental treatments in clinical trials (with NCT identifiers if available) > Search first: ClinicalTrials.gov, EU Clinical Trials Register, WHO ICTRP
- Treatment Outcomes:
- Treatment response rates > Search first: Clinical trial databases, FDA reviews, systematic reviews, PubMed
- Side effects and adverse events > Search first: FDA Adverse Event Reporting System (FAERS), MedWatch, PubMed
- Treatment Strategy:
- Treatment algorithms (clinical pathways, decision trees) > Search first: Clinical practice guidelines, NCCN Guidelines, UpToDate
- Combination therapies > Search first: ClinicalTrials.gov, treatment guidelines, PubMed
- Personalized medicine approaches (genotype-guided treatment) > Search first: My Cancer Genome, CIViC, PharmGKB, precision medicine databases
For each treatment, suggest NCIT (NCI Thesaurus) clinical-intervention terms where applicable.
13. Prevention
- Prevention Levels:
- Primary prevention (preventing disease occurrence: vaccination, risk factor modification) > Search first: CDC, WHO, USPSTF recommendations, Cochrane Library
- Secondary prevention (early detection and treatment: screening programs, early intervention) > Search first: USPSTF, CDC screening guidelines, WHO
- Tertiary prevention (preventing complications in those with disease) > Search first: Clinical guidelines, disease management protocols, PubMed
- Immunization: Vaccine strategies (if applicable)
Search first: CDC vaccine schedules, WHO immunization, FDA vaccine database
- Screening and Early Detection:
- Screening programs (population-based: newborn screening, cancer screening) > Search first: CDC screening programs, USPSTF, cancer screening databases
- Genetic screening (carrier screening, preimplantation genetic diagnosis, prenatal testing) > Search first: ACMG recommendations, ACOG guidelines, GTR
- Risk stratification (identifying high-risk individuals for targeted prevention) > Search first: Risk prediction models, clinical calculators, PubMed
- Behavioral Interventions: Lifestyle modifications to reduce risk
Search first: CDC, WHO, behavioral intervention databases, Cochrane Library
- Counseling: Genetic counseling (risk assessment, family planning guidance)
Search first: NSGC resources, ACMG guidelines, GeneReviews
- Public Health:
- Public health interventions (sanitation, vector control, health education) > Search first: CDC, WHO, public health databases, PubMed
- Environmental interventions (reducing environmental risk factors) > Search first: EPA databases, WHO environmental health, PubMed
- Prophylaxis: Preventive medications or procedures
Search first: Clinical guidelines, FDA approvals, PubMed
14. Other Species / Natural Disease
- Taxonomy: Species affected (with NCBI Taxon identifiers)
Search first: NCBI Taxonomy
- Breed: Specific breeds affected (with VBO identifiers if applicable)
Search first: VBO (Vertebrate Breed Ontology)
- Gene: Orthologous genes in other species (with NCBI Gene IDs)
Search first: NCBI Gene
- Natural Disease:
- Naturally occurring disease in other species (companion animals, wildlife) > Search first: OMIA (Online Mendelian Inheritance in Animals), VetCompass, PubMed
- Veterinary relevance and importance in animal health > Search first: OMIA, veterinary databases, PubMed
- Comparative Biology:
- Comparative pathology (similarities and differences across species) > Search first: OMIA, comparative pathology databases, PubMed
- Evolutionary conservation of disease mechanisms > Search first: HomoloGene, OrthoMCL, Alliance of Genome Resources
- Transmission (if applicable):
- Zoonotic potential > Search first: CDC zoonotic diseases, WHO zoonoses, GIDEON
- Cross-species susceptibility > Search first: NCBI Taxonomy, veterinary databases, PubMed
15. Model Organisms
- Model Types:
- Model organism type (mammalian, invertebrate, cellular, in vitro) > Search first: Alliance of Genome Resources, model organism databases
- Specific model systems (mouse, rat, zebrafish, Drosophila, C. elegans, yeast, cell lines, organoids, iPSCs) > Search first: MGI, RGD, ZFIN, FlyBase, WormBase, SGD, ATCC, Cellosaurus
- Induced models (drug treatment, surgical intervention, environmental manipulation) > Search first: MGI, model organism databases, PubMed
- Genetic Models:
- Types available (knockout, knock-in, transgenic, conditional, humanized) > Search first: MGI, IMPC, KOMP, EuMMCR, IMSR
- Model Characteristics:
- Phenotype recapitulation (how well model reproduces human disease features) > Search first: Model organism databases, comparative studies, PubMed
- Model limitations (aspects of human disease not captured) > Search first: Model organism databases, PubMed, review articles
- Applications:
- Research applications (what aspects of disease can be studied) > Search first: Model organism databases, PubMed
- Resources:
- Model databases > Search first: MGI, RGD, ZFIN, FlyBase, WormBase, IMSR, EMMA, MMRRC
Citation Requirements
- Cite primary literature (PMID preferred) for all mechanistic and clinical claims
- Prioritize recent reviews and landmark papers
- Include direct quotes from abstracts where possible to support key statements
- Distinguish evidence source types: human clinical, model organism, in vitro, computational
Output Format
Structure your response as a comprehensive narrative organized by the sections above. For each section, provide: - Factual content with specific details (numbers, percentages, gene names, variant nomenclature) - Ontology term suggestions (HPO, GO, CL, UBERON, CHEBI, NCIT, MONDO) where applicable - Evidence citations with PMIDs - Direct quotes from abstracts to support key claims - Clear indication when information is not available or not applicable for this disease
This report will be used to populate a disease knowledge base entry with: - Pathophysiology descriptions with causal chains - Gene/protein annotations (HGNC, GO terms) - Phenotype associations (HP terms) with frequencies - Cell type involvement (CL terms) - Anatomical locations (UBERON terms) - Chemical entities (CHEBI terms) - Treatment annotations (NCIT terms) - Evidence items with PMIDs and exact abstract quotes - Epidemiology, prognosis, diagnostic, and prevention information - Animal model descriptions with phenotype recapitulation details
Autosomal recessive nonsyndromic hearing loss 28 (DFNB28): disease-characteristics report
Scope and evidence. DFNB28 is a rare, inherited form of predominantly bilateral sensorineural hearing loss caused by biallelic pathogenic variants in TRIOBP. The characteristic presentation is congenital or prelingual severe-to-profound loss, but independently documented families establish a milder, childhood-onset spectrum. The strongest disease-specific evidence comprises human segregation studies, biochemical experiments, genetically engineered mice, and a small cochlear-implant case series. Findings from general hearing-loss cohorts are identified below as such and are not DFNB28 prevalence or outcome estimates. (riazuddin2006mutationsintriobp pages 1-3, shahin2006mutationsina pages 1-2, pollak2017wholeexomesequencing pages 1-2, tekin2021anewpathogenic pages 1-2)
1. Disease information and identifiers
- Preferred name and synonyms: deafness, autosomal recessive 28; DFNB28; TRIOBP-related nonsyndromic hearing loss. “Nonsyndromic” denotes hearing impairment without a consistently associated extra-auditory syndrome; it does not exclude an unrelated additional diagnosis in an individual patient. DFNB28 is not autosomal dominant deafness 28 (DFNA28). (riazuddin2006mutationsintriobp pages 1-3, shahin2006mutationsina pages 1-2, margret2024unravelingthegenetic pages 1-2)
- Identifiers: phenotype OMIM 609823; causal gene TRIOBP, OMIM 609761; locus 22q13.1. The requested MONDO:0012355 is supplied by the question but could not be independently cross-checked in the retrieved material; retain it as a provisional mapping pending MONDO verification. No DFNB28-specific Orphanet, ICD-10/ICD-11 or MeSH identifier was verified; general hearing-loss codes should not be misrepresented as unique disease identifiers. Links: https://omim.org/entry/609823 ; https://omim.org/entry/609761 ; https://mondo.monarchinitiative.org/ . (kitajiri2010actinbundlingproteintriobp pages 1-3, zhou2021casereportnovel pages 1-2, pollak2017wholeexomesequencing pages 1-2)
- Data provenance: the evidence is published, aggregated disease-level research based on consenting pedigrees and experimental models, not a patient-specific electronic health record. The original studies reported seven TRIOBP-segregating Indian/Pakistani families and, separately, nine Palestinian families comprising 30 affected individuals. These ascertainment-based counts are not population frequencies. (riazuddin2006mutationsintriobp pages 1-3, shahin2006mutationsina pages 1-2)
2. Etiology and risk or protective factors
Established cause: two disease-causing germline TRIOBP alleles, either homozygous or compound heterozygous, usually disrupt inner-ear-relevant TRIOBP-4 and/or TRIOBP-5 isoforms. Reported classes include nonsense, frameshift and some missense variants; molecular interpretation is transcript-dependent. The original Pakistani/Indian families had four nonsense and two frameshift alleles, whereas the Palestinian study identified nonsense p.Arg347 and p.Gln581 and a candidate missense p.Gly1019Arg in a compound-heterozygous affected child. The older studies’ exon numbers and protein coordinates should not be silently converted to a modern reference transcript. (riazuddin2006mutationsintriobp pages 3-5, shahin2006mutationsina pages 1-2, shahin2006mutationsina pages 4-8)
Family history and ancestry: parental carrier status creates the Mendelian risk; consanguinity increases the chance that both parents share a rare allele but is not required, as compound-heterozygous cases show. Distinct Palestinian endogamous communities carried different recurrent alleles; shared haplotypes support ancestral enrichment in the sampled communities, not a measured population-wide founder frequency. Neither sex-specific risk nor anticipation is established. (shahin2006mutationsina pages 2-4, zhou2021casereportnovel pages 2-5, shahin2006mutationsina pages 4-8)
Environmental factors and modifiers: loud noise and ototoxic exposures can independently injure hearing, so minimizing them is prudent; a DFNB28-specific human gene–environment interaction, protective diet, protective variant, or validated severity-modifier gene has not been established. In an Ankrd24-null mouse, auditory recovery after noise was diminished; ANKRD24 physically organizes TRIOBP-5, but this does not demonstrate that human ANKRD24 alleles modify DFNB28. A 2025 noise-exposure/TRIOBP report concerns Ménière disease, not proven recessive DFNB28, and should not be used to assign DFNB28 environmental penetrance. (cruzgranados2025ararevariant pages 13-17, krey2022ankrd24organizestriobp pages 2-2)
3. Phenotypes
| Phenotype and type | Onset, severity, course and frequency | Functional impact and suggested HPO concept |
|---|---|---|
| Bilateral sensorineural hearing loss — cardinal clinical sign/audiometric abnormality | Usually congenital/prelingual and severe-to-profound in the originally ascertained families. The Palestinian index family had bilateral, symmetrical, profound loss. Bilaterality is well supported but a rigorous disease-wide percentage is unavailable. | Impaired access to sound and spoken communication; suggest HPO concepts sensorineural hearing impairment, bilateral hearing impairment. (riazuddin2006mutationsintriobp pages 1-3, shahin2006mutationsina pages 1-2) |
| Moderate-to-severe childhood-onset hearing loss — variant expression of the cardinal sign | Three Polish siblings had onset at 3, 4.5 and 12 years; two had no measured worsening over two years, while one passed newborn hearing screening. Thus congenital onset and inevitable progression should not be encoded as universal. Frequency is unknown. | Potential delayed recognition and educational/communication effects; suggest HPO concepts childhood-onset hearing impairment and moderate hearing impairment, where clinically appropriate. (pollak2017wholeexomesequencing pages 1-2) |
| Speech/language difficulty secondary to reduced hearing — functional/behavioral consequence, not a separate primary TRIOBP phenotype | Depends strongly on detection and access to communication and rehabilitation. Three affected siblings referred for implantation relied on gestures before intervention; this does not establish universal absence of speech. | Individualized spoken-language, sign-language and educational support; consider an HPO delayed speech and language development term only when actually observed. (tekin2021anewpathogenic pages 3-4) |
No consistent vestibular, retinal, cardiac, renal, metabolic or intellectual phenotype is established for DFNB28. In the original Indian/Pakistani report, retinitis pigmentosa in one family segregated independently from deafness; the Palestinian index family had normal vision. An affected child can nevertheless have unrelated conductive disease: one Polish sibling additionally had otitis media and an air–bone gap. Absence of reported abnormalities is not proof that every affected individual has normal vestibular function. Disease-specific EQ-5D, SF-36 and phenotype-percentage data were not found. HPO identifiers above are concept suggestions, not database-verified HP accession numbers. (riazuddin2006mutationsintriobp pages 1-3, shahin2006mutationsina pages 1-2, pollak2017wholeexomesequencing pages 1-2)
4. Genetic and molecular information
Causal gene annotation: TRIOBP, encoding TRIO- and F-actin-binding protein, is a chromosome-22 gene with alternatively initiated and spliced transcripts. TRIOBP-4 binds/bundles actin; TRIOBP-5 encompasses its actin-binding region plus additional domains needed for mature rootlet structure. TRIOBP-1 has distinct sequence/expression and is retained in a deaf mouse lacking TRIOBP-4/5. The HGNC accession number was not verified and should be resolved against https://www.genenames.org/ before knowledge-base import. No disease-causing constitutional aneuploidy, translocation or inversion, somatic driver, validated epigenetic lesion or clinically established TRIOBP dosage syndrome was identified for DFNB28. (kitajiri2010actinbundlingproteintriobp pages 1-3, kitajiri2010actinbundlingproteintriobp pages 6-9, katsuno2019triobp5sculptsstereocilia pages 1-2)
The following compact evidence register distinguishes established alleles from unvalidated candidates. Allele frequencies are historic study/database snapshots, not current pan-ancestry carrier frequencies. (riazuddin2006mutationsintriobp pages 3-5, pollak2017wholeexomesequencing pages 2-4, zhou2020anovelmutation pages 2-5, tlili2024geneticanalysisof pages 2-4)
| Study/year | Variant(s) and transcript nomenclature | Zygosity and patient count | Hearing phenotype | Evidence caveat |
|---|---|---|---|---|
| Shahin et al., 2006 (Palestinian families) | Legacy: R347X, Q581X, G1019R; reported as c.1039C>T, c.1741C>T, and c.3055G>A in the newly described long isoform; do not translate directly to current NM_001039141.2/.3 HGVS without transcript normalization | 27 affected people in 7 families homozygous for R347X or Q581X; 3 affected people in 2 families compound heterozygous (R347X/Q581X or Q581X/G1019R) | Prelingual, bilateral, symmetric, profound sensorineural hearing loss; normal vision reported | Strong segregation evidence. R347X and Q581X occurred in different endogamous Palestinian communities; ancestral relationships were suggested, but population-wide founder status and prevalence were not established. G1019R was found heterozygously in 1/300 hearing controls (shahin2006mutationsina pages 1-2, shahin2006mutationsina pages 4-8, shahin2006mutationsina pages 2-4) |
| Riazuddin et al., 2006 (Pakistani and Indian families) | Legacy exon-6 alleles: Q297X, R788X, R1068X, R1117X, D1069fsX1082, R1078fsX1083; original numbering used the first coding ATG of TRIOBP-6 and should not be presented as current NM_001039141.2/.3 HGVS without remapping | Six truncating alleles cosegregated in 7 consanguineous families; affected individuals were homozygous and obligate carriers heterozygous | Congenital/prelingual severe-to-profound nonsyndromic hearing loss in 11/12 studied linked families; heterozygous carriers had normal hearing | Strong familial segregation and absence from approximately 300 control DNA samples. Five additional linked families lacked detected TRIOBP mutations, indicating possible missed variants or locus heterogeneity (riazuddin2006mutationsintriobp pages 1-3, riazuddin2006mutationsintriobp pages 5-7, riazuddin2006mutationsintriobp pages 3-5) |
| Pollak et al., 2017 (Polish family) | NM_001039141.2:c.802_805delCAGG, p.(Gln268Leufs610), affecting TRIOBP-4/5; and c.5014G>T, p.(Gly1672), affecting TRIOBP-5 | Compound heterozygous in trans in 3 affected siblings; each parent was a heterozygous carrier | Bilateral moderate-to-severe sensorineural hearing loss; onset at 3, 4.5, and 12 years; two siblings showed no deterioration over 2 years; youngest passed newborn screening | Strong segregation but single family. Historical ExAC frequencies were 0.000008 and 0.0006, respectively; proposed residual-isoform explanation for milder disease remains a genotype–phenotype hypothesis (pollak2017wholeexomesequencing pages 1-2, pollak2017wholeexomesequencing pages 2-4) |
| Zhou et al., 2020 (Chinese family) | NM_001039141.2:c.1342C>T, p.(Arg448*) | Homozygous in 2 affected siblings; both consanguineous parents heterozygous | Congenital severe-to-profound, symmetric hearing loss; no additional symptoms reported | Classified likely pathogenic by the authors; historical ExAC allele frequency 0.00002 and South Asian gnomAD frequency reported as 0.0000. HOMER2 and TMC2 findings were VUS and should not be treated as causal (zhou2020anovelmutation pages 1-2, zhou2020anovelmutation pages 2-5) |
| Tekin et al., 2021 (Afghan family) | NM_001039141.2:c.1342C>T, p.(Arg448*) | Homozygous in 3 affected siblings | Profound sensorineural hearing loss; after unilateral cochlear implantation, aided pure-tone averages were 23–30 dBHL at 1 month; two younger recipients reached up to 77% phoneme identification at 10 months | Independent Afghan family—not the Zhou 2020 Chinese pedigree despite the same allele. Useful treatment evidence is limited to three siblings; earlier implantation appeared more favorable (tekin2021anewpathogenic pages 1-2, tekin2021anewpathogenic pages 6-7) |
| Zhou et al., 2021 (Chinese family) | NM_001039141.2:c.1170delC, p.(Ser391Profs488), and c.3764C>G, p.(Ser1255) | Compound heterozygous in 1 affected 33-year-old woman; each healthy parent carried one allele | Isolated bilateral prelingual/congenital profound hearing loss | Both variants were novel and absent from the population databases queried at publication; truncating effects and segregation support causality, but evidence derives from one family and detailed audiometry was unavailable (zhou2021casereportnovel pages 1-2, zhou2021casereportnovel pages 2-5) |
| Margret et al., 2024 (South Indian cohort) | NM_001039141.3:c.2320C>T, p.(Arg774Ter) | Homozygous in 1 male proband with hearing loss and infertility | Hearing loss attributed to TRIOBP; severity details were limited in the extracted report | Ultra-rare in gnomAD (MAF <0.0004%) and predicted deleterious. A separate homozygous LRGUK variant was assigned to male infertility; the authors concluded hearing loss and infertility were independent events, not a TRIOBP syndrome (margret2024unravelingthegenetic pages 1-2, margret2024unravelingthegenetic pages 2-3, margret2024unravelingthegenetic pages 5-6) |
| Tlili et al., 2024 (UAE cohort) | c.3133C>T, p.(Arg1045Cys); transcript not sufficiently specified in the extracted table for safe NM_001039141.2/.3 normalization | Homozygous in 1 sporadic case | Moderate-to-severe hearing loss | Candidate missense finding only: previously unreported, gnomAD frequency 0.0004364, computationally “possibly damaging/deleterious”; no family segregation or functional validation was reported, so it should not be labeled pathogenic or definitive DFNB28 (tlili2024geneticanalysisof pages 4-5, tlili2024geneticanalysisof pages 2-4) |
Table: Human TRIOBP findings underlying or proposed for DFNB28, with transcript-nomenclature cautions and evidence strength. The table separates well-segregated pathogenic truncating alleles from recent candidate missense findings lacking validation.
Additional ascertainment details: the original six Indian/Pakistani truncating alleles were each absent from approximately 608–662 control chromosomes, depending on allele; this cannot establish zero frequency. In 2024, a South Indian proband had homozygous NM_001039141.3:c.2320C>T, p.(Arg774Ter), reported at gnomAD MAF <0.0004%; a separate homozygous LRGUK variant was assigned to his infertility. In a 2024 UAE hearing-loss cohort, homozygous TRIOBP c.3133C>T, p.(Arg1045Cys) occurred in one person with moderate-to-severe hearing loss, with reported gnomAD frequency 0.0004364; computational prediction and a single case are insufficient to classify it as a proven pathogenic DFNB28 allele. Consult current ClinVar/ClinGen assessments and transcript-specific ACMG/AMP criteria before assigning an operational variant class. (riazuddin2006mutationsintriobp pages 3-5, margret2024unravelingthegenetic pages 2-3, margret2024unravelingthegenetic pages 5-6, tlili2024geneticanalysisof pages 2-4)
5. Environmental information
There is no pathogen, infection, dietary exposure, smoking pattern, occupational chemical or radiation exposure known to initiate this Mendelian condition. Congenital cytomegalovirus, meningitis, noise and drugs remain possible alternative or additional causes of hearing impairment and belong in an individual diagnostic history, not the DFNB28 causal definition. For exposure annotation, relevant general concepts include noise and ototoxic drugs, but no DFNB28-specific ChEBI chemical entity or exposure-response estimate is established. The presence of unrelated male infertility in one 2024 study must not be entered as a TRIOBP syndrome. (margret2024unravelingthegenetic pages 1-2, margret2024unravelingthegenetic pages 5-6, riazuddin2006mutationsintriobp pages 1-3)
6. Mechanism and pathophysiology
Ordered causal chain — direct observations distinguished from inference: (kitajiri2010actinbundlingproteintriobp pages 1-3, kitajiri2010actinbundlingproteintriobp pages 6-9, kitajiri2010actinbundlingproteintriobp pages 9-11, katsuno2019triobp5sculptsstereocilia pages 1-2)
- Biallelic pathogenic TRIOBP variants lead to loss or alteration of the cochlear TRIOBP-4/5 proteins; this is genetically supported in human families, although allele-specific protein loss is not measured for every patient variant. (riazuddin2006mutationsintriobp pages 1-3, shahin2006mutationsina pages 1-2, zhou2021casereportnovel pages 2-5)
- Insufficient TRIOBP-4 F-actin bundling and/or TRIOBP-5-dependent sculpting leads to absent rootlets when both isoforms are removed, or misshapen rootlets when TRIOBP-5 alone is lost; demonstrated in purified-protein experiments and isoform-specific mice. (kitajiri2010actinbundlingproteintriobp pages 1-3, kitajiri2010actinbundlingproteintriobp pages 6-9, katsuno2019triobp5sculptsstereocilia pages 1-2)
- Defective rootlets at stereocilia insertion points lead to reduced hair-bundle pivot stiffness and mechanical durability; directly measured in mutant mouse hair cells. Branch A: initially intact stereocilia retain near-normal mechanically evoked transduction currents in vitro, so primary loss of the MET channel is not the demonstrated initial defect. Branch B: repeated deflection and maturation lead to fusion and degeneration of stereocilia in the combined-isoform mouse; extrapolation of its exact time course to human patients is inferred. (kitajiri2010actinbundlingproteintriobp pages 9-11, kitajiri2010actinbundlingproteintriobp pages 6-9)
- Altered hair-bundle mechanics and subsequent structural damage lead to impaired cochlear sound detection/amplification and profound mouse auditory-brainstem-response deficits; applying the complete mouse cellular sequence to each human DFNB28 genotype is inferred from convergent human segregation and mouse findings. (kitajiri2010actinbundlingproteintriobp pages 6-9, shahin2006mutationsina pages 1-2)
- Reduced auditory input leads to bilateral sensorineural hearing loss and, without timely access to appropriate communication and intervention, potential secondary language/participation difficulties. The degree of language impact is not a fixed genetic phenotype. (shahin2006mutationsina pages 1-2, tekin2021anewpathogenic pages 1-2)
Quantitative mechanistic results: purified TRIOBP-4 packed actin at 8.2 ± 1.4 nm interfilament distance versus 11.9 ± 2.1 nm with espin 3A in the assay. In Triobp exon-8-deletion mice, no mature rootlets formed, stereocilia fusion/degeneration was widespread by postnatal day 16, and adults showed no auditory response to 100-dB SPL clicks or 8–32-kHz tones. Early maximum MET currents were approximately preserved; mutant bundles were approximately twofold to fourfold more compliant under different fluid-jet stimulus sequences, with about 64% reduced stiffness after removal of extracellular links. These values are mouse/in-vitro, not patient hearing thresholds. Cropped primary-paper microscopy/measurement panels support the rootlet and bundle observations. (kitajiri2010actinbundlingproteintriobp pages 5-6, kitajiri2010actinbundlingproteintriobp pages 6-9, kitajiri2010actinbundlingproteintriobp pages 9-11, kitajiri2010actinbundlingproteintriobp media 11bbc05c, kitajiri2010actinbundlingproteintriobp media 5b8af6d1)
Anatomical and molecular branches: TRIOBP-5 also stiffens cochlear supporting cells; its isoform-specific mouse deficiency causes dysmorphic, rather than entirely absent, rootlets and progressive hearing loss. ANKRD24 binds and spatially organizes TRIOBP-5 at insertion points, shown by microscopy and knockout/rescue work; this is a mechanistic partner not an established human DFNB28 modifier. A September 2024 bioRxiv preprint, integrating single-cell transcriptomics, ChIP-seq and ATAC-seq, proposed Rfx3 control of an intronic Triobp enhancer during mouse outer-hair-cell development. It is regulatory research, not evidence that RFX3 mutations or altered DNA methylation cause DFNB28. Specific pathogenic Wnt, MAPK, mTOR, PI3K–AKT, immune, mitochondrial, metabolic, lipidomic or proteomic signatures have not been demonstrated for DFNB28; do not import unrelated TRIOBP signaling from other diseases. (katsuno2019triobp5sculptsstereocilia pages 1-2, krey2022ankrd24organizestriobp pages 2-2, zhang2024rfx3controlsouter pages 1-6)
Suggested ontology annotations, subject to accession verification: GO biological processes actin filament bundle assembly/organization, stereocilium organization, sensory perception of sound; GO components stereocilium, stereocilium rootlet, actin cytoskeleton; CL cell types cochlear inner hair cell, cochlear outer hair cell, and, for TRIOBP-5 mouse findings, cochlear supporting cell. These are biologically motivated term labels, not verified GO/CL identifiers. An actin chemical annotation should distinguish F-actin polymer from a small-molecule ChEBI exposure. (kitajiri2010actinbundlingproteintriobp pages 1-3, katsuno2019triobp5sculptsstereocilia pages 1-2, krey2022ankrd24organizestriobp pages 2-2)
7. Anatomical structures affected
The primary organ is the inner ear, particularly the cochlea/organ of Corti. At tissue level, inner and outer sensory hair cells and their apical F-actin-filled stereocilia are implicated; rootlets extend into each hair cell’s cuticular plate. Supporting-cell mechanical involvement is demonstrated in an isoform-specific mouse model. Cochlear hair cells are the directly demonstrated disease-relevant cell population; mouse expression in vestibular sensory tissue or spiral ganglion does not, by itself, establish clinical vestibular or neural degeneration in DFNB28. The auditory nervous system is functionally downstream of reduced cochlear input. Suggested UBERON labels: inner ear, cochlea, organ of Corti, cuticular plate where an appropriate ontology class exists; lateralization: typically bilateral. Exact UBERON and GO-cellular-component accessions require ontology lookup. (kitajiri2010actinbundlingproteintriobp pages 1-3, kitajiri2010actinbundlingproteintriobp pages 6-9, shahin2006mutationsina pages 1-2, katsuno2019triobp5sculptsstereocilia pages 1-2)
8. Temporal development
The classical phenotype is congenital or prelingual and lifelong; it is neither episodic nor known to remit spontaneously. Nonetheless, the Polish family demonstrates later recognition/onset at 3–12 years and stability in two siblings during a two-year observation period. In the combined-isoform mouse, stereocilia initially develop, then degenerate around hearing onset; in the TRIOBP-5-specific mouse, hearing worsens progressively. These distinct experimental courses should not be collapsed into a single human progression rate or a formal staged classification. Newborn-to-early-childhood auditory and language development is the principal intervention window; passing one newborn screen does not exclude later DFNB28 hearing loss. (riazuddin2006mutationsintriobp pages 1-3, pollak2017wholeexomesequencing pages 1-2, kitajiri2010actinbundlingproteintriobp pages 6-9, katsuno2019triobp5sculptsstereocilia pages 1-2)
9. Inheritance and population characteristics
Inheritance: autosomal recessive; an affected person generally carries two pathogenic TRIOBP alleles in trans. If both parents are confirmed heterozygous carriers for pathogenic variants in the same gene, the theoretical risk per pregnancy is 25% affected, 50% carrier and 25% inheriting neither familial variant; this is Mendelian counseling, not a measured DFNB28 penetrance estimate. Healthy heterozygous parents and affected siblings support this pattern; complete, age-specific penetrance, germline-mosaicism rate and anticipation have not been established. Clinical expressivity ranges from congenital profound to later-onset moderate impairment. (shahin2006mutationsina pages 1-2, zhou2021casereportnovel pages 2-5, pollak2017wholeexomesequencing pages 1-2)
Epidemiology: no reliable DFNB28-specific prevalence per 100,000, annual incidence, population carrier frequency, sex ratio or geographic distribution estimate was identified. Reports document families from Palestine, Pakistan, India, Poland, China and Afghanistan, among others; ascertainment and consanguinity mean these cannot be compared as population rates. In the original Palestinian study, one of 300 unrelated hearing controls carried p.Gly1019Arg, while the sampled endogamous hearing-loss pedigrees showed recurrent p.Arg347 or p.Gln581; neither denominator estimates a global DFNB28 carrier rate. Broad congenital hearing-loss incidence figures must not be substituted for the subtype’s incidence. (shahin2006mutationsina pages 1-2, shahin2006mutationsina pages 4-8, shahin2006mutationsina pages 2-4, pollak2017wholeexomesequencing pages 1-2, zhou2020anovelmutation pages 1-2)
10. Diagnostics
Clinical work-up: universal newborn physiologic hearing screening followed, when indicated, by diagnostic auditory brainstem response, age-appropriate air- and bone-conduction pure-tone audiometry, tympanometry/otoscopy for conductive disease, and assessment of speech perception and communication needs. Otoacoustic emissions can characterize cochlear outer-hair-cell function but do not determine genotype. Examine for retinal, vestibular, renal or other findings if the history suggests a syndromic alternative; investigate infectious or acquired causes as appropriate. MRI/CT is selective—particularly for cochlear-implant planning or suspected malformation—not a diagnostic biomarker for TRIOBP, and biopsy is not indicated routinely. (tekin2021anewpathogenic pages 3-4, zhou2020anovelmutation pages 2-5, riazuddin2006mutationsintriobp pages 1-3)
Molecular confirmation: use a validated comprehensive hearing-loss multigene panel including TRIOBP, with coverage of its hearing-relevant exons/isoforms and suitable variant/copy-number analysis; consider exome or genome sequencing when panel testing is unrevealing or the phenotype is atypical. Interpret variants against a specified transcript, phenotype, population frequency, current ClinVar/ClinGen evidence and ACMG/AMP criteria; verify suspected biallelic variants in trans by parental/family testing. A VUS or unvalidated computational missense finding alone is not a molecular DFNB28 diagnosis. Single-gene TRIOBP testing is most useful when familial pathogenic alleles are already known. Chromosomal microarray, karyotype, FISH, mitochondrial sequencing or repeat-expansion testing are not routine DFNB28 confirmation tests; order for other differential diagnoses when indicated. No validated DFNB28-specific blood chemistry, RNA-seq, proteomic, metabolomic, epigenomic or liquid-biopsy assay was found. (zhou2021casereportnovel pages 2-5, pollak2017wholeexomesequencing pages 1-2, tekin2021anewpathogenic pages 3-4, tlili2024geneticanalysisof pages 2-4)
Contextual—not subtype-specific—diagnostic yields: a 2023 Spanish 155-person hearing-loss panel study reported 52/155 (34%) molecular diagnoses using 171 nuclear and eight mitochondrial genes. A 2023 Taiwanese bilateral-hearing-impairment cohort reported 52% diagnostic yield in 350 tested patients. These percentages describe heterogeneous hearing loss and must not be entered as TRIOBP test sensitivity or DFNB28 prevalence. Differential diagnoses include GJB2, STRC, OTOF, SLC26A4, Usher-spectrum genes, congenital infection and acquired conductive loss; their distinguishing features require audiology, examination and genotype. (lee2023revisitinggeneticepidemiology pages 9-10, pollak2017wholeexomesequencing pages 1-2, tekin2021anewpathogenic pages 3-4)
11. Outcome and prognosis
DFNB28 is predominantly a chronic sensory disability, with risks to communication, language acquisition, schooling and social participation when hearing and communication needs are unmet. There is no established disease-specific excess mortality or reduced life expectancy, and no legitimate DFNB28 five-year survival or fatality statistic. Severity and trajectory vary by allele/isoform and individual circumstance; an exact prognostic algorithm or validated blood biomarker does not exist. The Polish two-year stability observation cannot guarantee lifelong stability. Implant performance is individualized and affected by implantation age, residual hearing, auditory anatomy and rehabilitation. DFNB28-specific EQ-5D/SF-36 scores were not located. (pollak2017wholeexomesequencing pages 1-2, katsuno2019triobp5sculptsstereocilia pages 1-2, tekin2021anewpathogenic pages 1-2)
12. Treatment and current applications
Current strategy: prompt audiological characterization and family-centered communication support; trial hearing aids when residual hearing makes amplification useful; consider cochlear implantation when severe-to-profound loss and limited amplification benefit meet local criteria; provide sustained audiology, speech/language and educational services and access to sign language according to family preference. Implants bypass damaged cochlear hair-cell transduction rather than repairing TRIOBP or stereocilia. Suggested NCIT intervention labels for mapping—hearing aid, cochlear implantation, speech and language therapy, genetic counseling—require verification of exact NCIT codes before import. (tekin2021anewpathogenic pages 1-2, tekin2021anewpathogenic pages 3-4, katsuno2019triobp5sculptsstereocilia pages 1-2)
Disease-specific human outcome: three Afghan siblings homozygous for NM_001039141.2:c.1342C>T, p.(Arg448*) received unilateral cochlear implants. Their aided pure-tone averages were 23–30 dB HL at one month. At ten months, the two younger recipients attained up to 77% correct phonemes; the older brother was not yet able to perform open-set speech testing. These are outcomes for three people in one family, not a general DFNB28 response rate. The same named allele also occurred in an independently reported Chinese family. Implant surgery entails ordinary device/anesthesia/surgical risks, but variant-specific adverse-event rates were not reported. (tekin2021anewpathogenic pages 1-2, tekin2021anewpathogenic pages 6-7, zhou2020anovelmutation pages 1-2)
Experimental interventions: no TRIOBP/DFNB28-specific approved drug, pharmacogenomic regimen, RNA therapy, cellular therapy, gene therapy or confirmed interventional trial/NCT identifier was found in the searches. The important 2024 bilateral AAV-OTOF trial treated five children with DFNB9, all reporting bilateral hearing restoration in its interim analysis; it is a different genetic disease and not an efficacy result for TRIOBP. No supported claim can be made that DFNB28 is currently reversible by AAV, CRISPR or anti-inflammatory drugs. (katsuno2019triobp5sculptsstereocilia pages 1-2, tekin2021anewpathogenic pages 1-2)
13. Prevention
Primary genetic prevention: a person’s inherited TRIOBP genotype is not changed by diet, vaccination or ordinary exposure avoidance. Offer nondirective carrier/cascade testing and reproductive counseling when familial pathogenic alleles are known; prenatal or preimplantation genetic testing may be discussed according to preferences and local practice. Consanguinity raises shared-allele probability, but counseling should avoid stigmatization. Hearing protection and careful use of potentially ototoxic medication address additional avoidable auditory injury, not proven prevention of DFNB28 itself. There is no DFNB28-specific vaccine or drug prophylaxis. (shahin2006mutationsina pages 2-4, zhou2020anovelmutation pages 1-2, zhou2021casereportnovel pages 2-5)
Secondary/tertiary prevention: screen hearing in newborns and monitor children with familial risk even after a passed screen; arrange early audiological diagnosis and timely access to amplification, implants when appropriate, and communication/language services. General early-hearing-detection benchmarks target screening by one month, diagnostic assessment by three months, and early intervention by six months; these are general program targets, not DFNB28-specific trial outcomes. A 2024 expert consensus identified screening, audiologic management, amplification, medical care, early intervention, family support, Deaf/hard-of-hearing leadership and data management as components of integrated care. (pollak2017wholeexomesequencing pages 1-2, tekin2021anewpathogenic pages 1-2)
14. Other species and natural disease
The established natural human disease is in Homo sapiens (NCBI Taxon 9606). Mouse Triobp is the experimentally validated ortholog in Mus musculus (NCBI Taxon 10090); animal gene-accession numbers and breed-ontology identifiers were not verified. A naturally occurring TRIOBP-defined counterpart in a companion-animal breed or wildlife population, veterinary incidence and cross-species transmission were not established. DFNB28 is noninfectious and nonzoonotic. Conservation of TRIOBP-related actin/rootlet function is supported by the human–mouse comparison, while exon/transcript structure and expression differ between species. (riazuddin2006mutationsintriobp pages 3-5, kitajiri2010actinbundlingproteintriobp pages 6-9, katsuno2019triobp5sculptsstereocilia pages 1-2)
15. Model organisms and research resources
- Combined-isoform mouse knockout, Triobp exon-8 deletion: removes TRIOBP-4/5 while preserving TRIOBP-1; models rootlet absence, reduced hair-bundle stiffness, stereocilia degeneration and profound deafness, closely reproducing the classical severe auditory phenotype. Its postnatal developmental timing should not be assumed to equal human timing. Applications include ultrastructural electron microscopy, ABR testing, MET electrophysiology and hair-bundle mechanics. (kitajiri2010actinbundlingproteintriobp pages 6-9, kitajiri2010actinbundlingproteintriobp pages 9-11, kitajiri2010actinbundlingproteintriobp media 11bbc05c)
- TRIOBP-5-specific genetically engineered mice: show dysmorphic rootlets, changed supporting-cell stiffness and progressive hearing loss; especially informative for isoform-specific and milder human presentations, not proof that a particular human allele will progress at the same rate. Combined TRIOBP-1/-5 loss produced embryonic lethality in earlier mouse work, restricting its use for hearing studies. (katsuno2019triobp5sculptsstereocilia pages 1-2, kitajiri2010actinbundlingproteintriobp pages 6-9)
- Complementary systems: purified TRIOBP-4 plus F-actin establishes bundling in vitro; Ankrd24 knockout and exogenous TRIOBP-5 rescue probe rootlet organization. Mouse single-cell transcriptomics/epigenomic regulatory profiling in the 2024 unreviewed preprint are discovery tools, not diagnostic signatures. Model-resource search points include MGI, IMPC and IMSR; precise strain accessions should be checked there. No validated DFNB28-specific zebrafish, fly, patient-iPSC, organoid, spatial-transcriptomic or multi-omics clinical model was established in the consulted evidence. (kitajiri2010actinbundlingproteintriobp pages 5-6, krey2022ankrd24organizestriobp pages 2-2, zhang2024rfx3controlsouter pages 1-6)
Selected primary-source links and verbatim abstract evidence
- Shahin et al., January 2006, American Journal of Human Genetics, https://doi.org/10.1086/499495 — “In seven families, 27 deaf individuals are homozygous for one of the nonsense mutations”; this is familial human evidence, not a prevalence estimate. (shahin2006mutationsina pages 1-2)
- Riazuddin et al., January 2006, American Journal of Human Genetics, https://doi.org/10.1086/499164 — “In seven families, six different mutant alleles of TRIOBP on chromosome 22q13 cosegregate with autosomal recessive nonsyndromic deafness.” (riazuddin2006mutationsintriobp pages 1-3)
- Kitajiri et al., May 2010, Cell, https://doi.org/10.1016/j.cell.2010.03.049 — “Stereocilia of TriobpDex8/Dex8 mice develop normally but fail to form rootlets and are easier to deflect and damage.” This is mouse/in-vitro mechanism, not human histopathology. (kitajiri2010actinbundlingproteintriobp pages 1-3)
- Pollak et al., December 2017, BMC Medical Genetics, https://doi.org/10.1186/s12881-017-0499-z — “nonsyndromic, peri- to postlingual, moderate-to-severe hearing loss in three siblings”; this establishes the expanded human spectrum. (pollak2017wholeexomesequencing pages 1-2)
- Katsuno et al., June 2019, JCI Insight, https://doi.org/10.1172/jci.insight.128561 — “TRIOBP-5 is essential for thickening bundles of F-actin in rootlets”; isoform-specific mouse evidence. (katsuno2019triobp5sculptsstereocilia pages 1-2)
- Zhou et al., June 2020, BMC Medical Genetics, https://doi.org/10.1186/s12881-020-01055-5 — “severe to profound symmetric hearing loss” in two siblings carrying p.Arg448*. (zhou2020anovelmutation pages 1-2)
- Tekin et al., 2021, Audiology and Neurotology, https://doi.org/10.1159/000508434 — “One month after activation, the pure-tone averages with the CI processor were between 30 and 23 dBHL.” Human, three-sibling treatment report. (tekin2021anewpathogenic pages 1-2)
- Krey et al., February 2022, Journal of Cell Biology, https://doi.org/10.1083/jcb.202109134 — “TRIOBP-5 is mislocalized in Ankrd24KO/KO hair cells”; mouse protein-network evidence, not a human modifier association. (krey2022ankrd24organizestriobp pages 2-2)
- Margret et al., May 2024, Advanced Genetics, https://doi.org/10.1002/ggn2.202300206 — “deafness and infertility are independent events”; human sequencing evidence against conflating these phenotypes. (margret2024unravelingthegenetic pages 1-2, margret2024unravelingthegenetic pages 2-3)
- Tlili et al., June 2024, Human Genomics, https://doi.org/10.1186/s40246-024-00630-8 — a candidate homozygous p.Arg1045Cys in one UAE hearing-loss case; segregation and functional confirmation are lacking. (tlili2024geneticanalysisof pages 2-4)
- Zhang et al., posted September 2024, bioRxiv preprint, https://doi.org/10.1101/2024.09.24.614849 — “Rfx3 regulates the spatiotemporal expression of hair bundle gene Triobp”; proposed mouse regulatory biology, not peer-reviewed evidence of a new DFNB28 cause. (zhang2024rfx3controlsouter pages 1-6)
Knowledge-base limitation: precise PMID numbers, HGNC accession, most requested HP/GO/CL/UBERON/ChEBI/NCIT accession numbers, Orphanet cross-reference and the question-supplied MONDO mapping were not independently verified from the retrieved sources. DOI URLs above identify the underlying publications; resolve accession-level ontology and PMID fields against their issuing databases rather than fabricating values. (kitajiri2010actinbundlingproteintriobp pages 1-3, zhou2021casereportnovel pages 1-2, OpenTargets Search: autosomal recessive nonsyndromic hearing loss 28-TRIOBP)
References
-
(riazuddin2006mutationsintriobp pages 1-3): Saima Riazuddin, Shaheen N. Khan, Zubair M. Ahmed, Manju Ghosh, Kyle Caution, Sabiha Nazli, Madhulika Kabra, Ahmad U. Zafar, Kevin Chen, Sadaf Naz, Anthony Antonellis, William J. Pavan, Eric D. Green, Edward R. Wilcox, Penelope L. Friedman, Robert J. Morell, Sheikh Riazuddin, and Thomas B. Friedman. Mutations in triobp, which encodes a putative cytoskeletal-organizing protein, are associated with nonsyndromic recessive deafness. American journal of human genetics, 78 1:137-43, Jan 2006. URL: https://doi.org/10.1086/499164, doi:10.1086/499164. This article has 141 citations and is from a highest quality peer-reviewed journal.
-
(shahin2006mutationsina pages 1-2): Hashem Shahin, Tom Walsh, Tama Sobe, Judeh Abu Sa’ed, Amal Abu Rayan, Eric D. Lynch, Ming K. Lee, Karen B. Avraham, Mary-Claire King, and Moein Kanaan. Mutations in a novel isoform of triobp that encodes a filamentous-actin binding protein are responsible for dfnb28 recessive nonsyndromic hearing loss. American journal of human genetics, 78 1:144-52, Jan 2006. URL: https://doi.org/10.1086/499495, doi:10.1086/499495. This article has 147 citations and is from a highest quality peer-reviewed journal.
-
(pollak2017wholeexomesequencing pages 1-2): Agnieszka Pollak, Urszula Lechowicz, Victor Abel Murcia Pieńkowski, Piotr Stawiński, Joanna Kosińska, Henryk Skarżyński, Monika Ołdak, and Rafał Płoski. Whole exome sequencing identifies triobp pathogenic variants as a cause of post-lingual bilateral moderate-to-severe sensorineural hearing loss. BMC Medical Genetics, Dec 2017. URL: https://doi.org/10.1186/s12881-017-0499-z, doi:10.1186/s12881-017-0499-z. This article has 26 citations and is from a peer-reviewed journal.
-
(tekin2021anewpathogenic pages 1-2): Ahmet M. Tekin, Geert de Ceulaer, Paul Govaerts, Yıldırım Bayazit, Wim Wuyts, Paul Van de Heyning, and Vedat Topsakal. A new pathogenic variant in the triobp associated with profound deafness is remediable with cochlear implantation. Sep 2021. URL: https://doi.org/10.1159/000508434, doi:10.1159/000508434. This article has 5 citations.
-
(margret2024unravelingthegenetic pages 1-2): Jeffrey Justin Margret, Chandru Jayasankaran, Pavithra Amritkumar, Hela Azaiez, and C. R. Srikumari Srisailapathy. Unraveling the genetic basis of combined deafness and male infertility phenotypes through high‐throughput sequencing in a unique cohort from south india. Advanced Genetics, May 2024. URL: https://doi.org/10.1002/ggn2.202300206, doi:10.1002/ggn2.202300206. This article has 5 citations.
-
(kitajiri2010actinbundlingproteintriobp pages 1-3): Shin-ichiro Kitajiri, Takeshi Sakamoto, Inna A. Belyantseva, Richard J. Goodyear, Ruben Stepanyan, Ikuko Fujiwara, Jonathan E. Bird, Saima Riazuddin, Sheikh Riazuddin, Zubair M. Ahmed, Jenny E. Hinshaw, James Sellers, James R. Bartles, John A. Hammer, Guy P. Richardson, Andrew J. Griffith, Gregory I. Frolenkov, and Thomas B. Friedman. Actin-bundling protein triobp forms resilient rootlets of hair cell stereocilia essential for hearing. Cell, 141:786-798, May 2010. URL: https://doi.org/10.1016/j.cell.2010.03.049, doi:10.1016/j.cell.2010.03.049. This article has 234 citations and is from a highest quality peer-reviewed journal.
-
(zhou2021casereportnovel pages 1-2): Cong Zhou, Yuanyuan Xiao, Hanbing Xie, Jing Wang, and Shanling Liu. Case report: novel compound heterozygous variants in triobp associated with congenital deafness in a chinese family. Frontiers in Genetics, Nov 2021. URL: https://doi.org/10.3389/fgene.2021.766973, doi:10.3389/fgene.2021.766973. This article has 2 citations and is from a peer-reviewed journal.
-
(riazuddin2006mutationsintriobp pages 3-5): Saima Riazuddin, Shaheen N. Khan, Zubair M. Ahmed, Manju Ghosh, Kyle Caution, Sabiha Nazli, Madhulika Kabra, Ahmad U. Zafar, Kevin Chen, Sadaf Naz, Anthony Antonellis, William J. Pavan, Eric D. Green, Edward R. Wilcox, Penelope L. Friedman, Robert J. Morell, Sheikh Riazuddin, and Thomas B. Friedman. Mutations in triobp, which encodes a putative cytoskeletal-organizing protein, are associated with nonsyndromic recessive deafness. American journal of human genetics, 78 1:137-43, Jan 2006. URL: https://doi.org/10.1086/499164, doi:10.1086/499164. This article has 141 citations and is from a highest quality peer-reviewed journal.
-
(shahin2006mutationsina pages 4-8): Hashem Shahin, Tom Walsh, Tama Sobe, Judeh Abu Sa’ed, Amal Abu Rayan, Eric D. Lynch, Ming K. Lee, Karen B. Avraham, Mary-Claire King, and Moein Kanaan. Mutations in a novel isoform of triobp that encodes a filamentous-actin binding protein are responsible for dfnb28 recessive nonsyndromic hearing loss. American journal of human genetics, 78 1:144-52, Jan 2006. URL: https://doi.org/10.1086/499495, doi:10.1086/499495. This article has 147 citations and is from a highest quality peer-reviewed journal.
-
(shahin2006mutationsina pages 2-4): Hashem Shahin, Tom Walsh, Tama Sobe, Judeh Abu Sa’ed, Amal Abu Rayan, Eric D. Lynch, Ming K. Lee, Karen B. Avraham, Mary-Claire King, and Moein Kanaan. Mutations in a novel isoform of triobp that encodes a filamentous-actin binding protein are responsible for dfnb28 recessive nonsyndromic hearing loss. American journal of human genetics, 78 1:144-52, Jan 2006. URL: https://doi.org/10.1086/499495, doi:10.1086/499495. This article has 147 citations and is from a highest quality peer-reviewed journal.
-
(zhou2021casereportnovel pages 2-5): Cong Zhou, Yuanyuan Xiao, Hanbing Xie, Jing Wang, and Shanling Liu. Case report: novel compound heterozygous variants in triobp associated with congenital deafness in a chinese family. Frontiers in Genetics, Nov 2021. URL: https://doi.org/10.3389/fgene.2021.766973, doi:10.3389/fgene.2021.766973. This article has 2 citations and is from a peer-reviewed journal.
-
(cruzgranados2025ararevariant pages 13-17): Pablo Cruz-Granados, Giselle Bianco-Bortoletto, Yuzhong Zhang, Prathamesh T Nadar-Ponniah, Kiana Bagheri-Loftabad, Edi Lúcia Sartorato, Inés Sánchez-Sellero, Andrés Soto-Varela, J. A. Lopez-Escamez, A. Prof.Jose, and Lopez-Escamez. A rare variant in triobp linked to occupational noise exposure in meniere disease. MedRxiv, Jul 2025. URL: https://doi.org/10.1101/2025.07.23.25332035, doi:10.1101/2025.07.23.25332035. This article has 2 citations.
-
(krey2022ankrd24organizestriobp pages 2-2): Jocelyn F. Krey, Chang Liu, Inna A. Belyantseva, Michael Bateschell, Rachel A. Dumont, Jennifer Goldsmith, Paroma Chatterjee, Rachel S. Morrill, Lev M. Fedorov, Sarah Foster, Jinkyung Kim, Alfred L. Nuttall, Sherri M. Jones, Dongseok Choi, Thomas B. Friedman, Anthony J. Ricci, Bo Zhao, and Peter G. Barr-Gillespie. Ankrd24 organizes triobp to reinforce stereocilia insertion points. The Journal of Cell Biology, Feb 2022. URL: https://doi.org/10.1083/jcb.202109134, doi:10.1083/jcb.202109134. This article has 19 citations.
-
(tekin2021anewpathogenic pages 3-4): Ahmet M. Tekin, Geert de Ceulaer, Paul Govaerts, Yıldırım Bayazit, Wim Wuyts, Paul Van de Heyning, and Vedat Topsakal. A new pathogenic variant in the triobp associated with profound deafness is remediable with cochlear implantation. Sep 2021. URL: https://doi.org/10.1159/000508434, doi:10.1159/000508434. This article has 5 citations.
-
(kitajiri2010actinbundlingproteintriobp pages 6-9): Shin-ichiro Kitajiri, Takeshi Sakamoto, Inna A. Belyantseva, Richard J. Goodyear, Ruben Stepanyan, Ikuko Fujiwara, Jonathan E. Bird, Saima Riazuddin, Sheikh Riazuddin, Zubair M. Ahmed, Jenny E. Hinshaw, James Sellers, James R. Bartles, John A. Hammer, Guy P. Richardson, Andrew J. Griffith, Gregory I. Frolenkov, and Thomas B. Friedman. Actin-bundling protein triobp forms resilient rootlets of hair cell stereocilia essential for hearing. Cell, 141:786-798, May 2010. URL: https://doi.org/10.1016/j.cell.2010.03.049, doi:10.1016/j.cell.2010.03.049. This article has 234 citations and is from a highest quality peer-reviewed journal.
-
(katsuno2019triobp5sculptsstereocilia pages 1-2): Tatsuya Katsuno, I. A. Belyantseva, Alexander X. Cartagena-Rivera, Keisuke Ohta, S. Crump, R. Petralia, Kazuya Ono, R. Tona, A. Imtiaz, A. Rehman, H. Kiyonari, Mari Kaneko, Ya-Xian Wang, T. Abe, M. Ikeya, C. Fenollar-Ferrer, G. Riordan, Elizabeth Wilson, Tracy S. Fitzgerald, Kohei Segawa, K. Omori, J. Ito, G. Frolenkov, T. Friedman, and S. Kitajiri. Triobp-5 sculpts stereocilia rootlets and stiffens supporting cells enabling hearing. JCI insight, Jun 2019. URL: https://doi.org/10.1172/jci.insight.128561, doi:10.1172/jci.insight.128561. This article has 40 citations and is from a domain leading peer-reviewed journal.
-
(pollak2017wholeexomesequencing pages 2-4): Agnieszka Pollak, Urszula Lechowicz, Victor Abel Murcia Pieńkowski, Piotr Stawiński, Joanna Kosińska, Henryk Skarżyński, Monika Ołdak, and Rafał Płoski. Whole exome sequencing identifies triobp pathogenic variants as a cause of post-lingual bilateral moderate-to-severe sensorineural hearing loss. BMC Medical Genetics, Dec 2017. URL: https://doi.org/10.1186/s12881-017-0499-z, doi:10.1186/s12881-017-0499-z. This article has 26 citations and is from a peer-reviewed journal.
-
(zhou2020anovelmutation pages 2-5): Bingxin Zhou, Lili Yu, Yan Wang, Wenjing Shang, Yi Xie, Xiong Wang, and Fengchan Han. A novel mutation in triobp gene leading to congenital deafness in a chinese family. BMC Medical Genetics, Jun 2020. URL: https://doi.org/10.1186/s12881-020-01055-5, doi:10.1186/s12881-020-01055-5. This article has 8 citations and is from a peer-reviewed journal.
-
(tlili2024geneticanalysisof pages 2-4): Abdelaziz Tlili, Mona Mahfood, Abdullah Al Mutery, and Jihen Chouchen. Genetic analysis of 106 sporadic cases with hearing loss in the uae population. Human Genomics, Jun 2024. URL: https://doi.org/10.1186/s40246-024-00630-8, doi:10.1186/s40246-024-00630-8. This article has 11 citations and is from a peer-reviewed journal.
-
(riazuddin2006mutationsintriobp pages 5-7): Saima Riazuddin, Shaheen N. Khan, Zubair M. Ahmed, Manju Ghosh, Kyle Caution, Sabiha Nazli, Madhulika Kabra, Ahmad U. Zafar, Kevin Chen, Sadaf Naz, Anthony Antonellis, William J. Pavan, Eric D. Green, Edward R. Wilcox, Penelope L. Friedman, Robert J. Morell, Sheikh Riazuddin, and Thomas B. Friedman. Mutations in triobp, which encodes a putative cytoskeletal-organizing protein, are associated with nonsyndromic recessive deafness. American journal of human genetics, 78 1:137-43, Jan 2006. URL: https://doi.org/10.1086/499164, doi:10.1086/499164. This article has 141 citations and is from a highest quality peer-reviewed journal.
-
(zhou2020anovelmutation pages 1-2): Bingxin Zhou, Lili Yu, Yan Wang, Wenjing Shang, Yi Xie, Xiong Wang, and Fengchan Han. A novel mutation in triobp gene leading to congenital deafness in a chinese family. BMC Medical Genetics, Jun 2020. URL: https://doi.org/10.1186/s12881-020-01055-5, doi:10.1186/s12881-020-01055-5. This article has 8 citations and is from a peer-reviewed journal.
-
(tekin2021anewpathogenic pages 6-7): Ahmet M. Tekin, Geert de Ceulaer, Paul Govaerts, Yıldırım Bayazit, Wim Wuyts, Paul Van de Heyning, and Vedat Topsakal. A new pathogenic variant in the triobp associated with profound deafness is remediable with cochlear implantation. Sep 2021. URL: https://doi.org/10.1159/000508434, doi:10.1159/000508434. This article has 5 citations.
-
(margret2024unravelingthegenetic pages 2-3): Jeffrey Justin Margret, Chandru Jayasankaran, Pavithra Amritkumar, Hela Azaiez, and C. R. Srikumari Srisailapathy. Unraveling the genetic basis of combined deafness and male infertility phenotypes through high‐throughput sequencing in a unique cohort from south india. Advanced Genetics, May 2024. URL: https://doi.org/10.1002/ggn2.202300206, doi:10.1002/ggn2.202300206. This article has 5 citations.
-
(margret2024unravelingthegenetic pages 5-6): Jeffrey Justin Margret, Chandru Jayasankaran, Pavithra Amritkumar, Hela Azaiez, and C. R. Srikumari Srisailapathy. Unraveling the genetic basis of combined deafness and male infertility phenotypes through high‐throughput sequencing in a unique cohort from south india. Advanced Genetics, May 2024. URL: https://doi.org/10.1002/ggn2.202300206, doi:10.1002/ggn2.202300206. This article has 5 citations.
-
(tlili2024geneticanalysisof pages 4-5): Abdelaziz Tlili, Mona Mahfood, Abdullah Al Mutery, and Jihen Chouchen. Genetic analysis of 106 sporadic cases with hearing loss in the uae population. Human Genomics, Jun 2024. URL: https://doi.org/10.1186/s40246-024-00630-8, doi:10.1186/s40246-024-00630-8. This article has 11 citations and is from a peer-reviewed journal.
-
(kitajiri2010actinbundlingproteintriobp pages 9-11): Shin-ichiro Kitajiri, Takeshi Sakamoto, Inna A. Belyantseva, Richard J. Goodyear, Ruben Stepanyan, Ikuko Fujiwara, Jonathan E. Bird, Saima Riazuddin, Sheikh Riazuddin, Zubair M. Ahmed, Jenny E. Hinshaw, James Sellers, James R. Bartles, John A. Hammer, Guy P. Richardson, Andrew J. Griffith, Gregory I. Frolenkov, and Thomas B. Friedman. Actin-bundling protein triobp forms resilient rootlets of hair cell stereocilia essential for hearing. Cell, 141:786-798, May 2010. URL: https://doi.org/10.1016/j.cell.2010.03.049, doi:10.1016/j.cell.2010.03.049. This article has 234 citations and is from a highest quality peer-reviewed journal.
-
(kitajiri2010actinbundlingproteintriobp pages 5-6): Shin-ichiro Kitajiri, Takeshi Sakamoto, Inna A. Belyantseva, Richard J. Goodyear, Ruben Stepanyan, Ikuko Fujiwara, Jonathan E. Bird, Saima Riazuddin, Sheikh Riazuddin, Zubair M. Ahmed, Jenny E. Hinshaw, James Sellers, James R. Bartles, John A. Hammer, Guy P. Richardson, Andrew J. Griffith, Gregory I. Frolenkov, and Thomas B. Friedman. Actin-bundling protein triobp forms resilient rootlets of hair cell stereocilia essential for hearing. Cell, 141:786-798, May 2010. URL: https://doi.org/10.1016/j.cell.2010.03.049, doi:10.1016/j.cell.2010.03.049. This article has 234 citations and is from a highest quality peer-reviewed journal.
-
(kitajiri2010actinbundlingproteintriobp media 11bbc05c): Shin-ichiro Kitajiri, Takeshi Sakamoto, Inna A. Belyantseva, Richard J. Goodyear, Ruben Stepanyan, Ikuko Fujiwara, Jonathan E. Bird, Saima Riazuddin, Sheikh Riazuddin, Zubair M. Ahmed, Jenny E. Hinshaw, James Sellers, James R. Bartles, John A. Hammer, Guy P. Richardson, Andrew J. Griffith, Gregory I. Frolenkov, and Thomas B. Friedman. Actin-bundling protein triobp forms resilient rootlets of hair cell stereocilia essential for hearing. Cell, 141:786-798, May 2010. URL: https://doi.org/10.1016/j.cell.2010.03.049, doi:10.1016/j.cell.2010.03.049. This article has 234 citations and is from a highest quality peer-reviewed journal.
-
(kitajiri2010actinbundlingproteintriobp media 5b8af6d1): Shin-ichiro Kitajiri, Takeshi Sakamoto, Inna A. Belyantseva, Richard J. Goodyear, Ruben Stepanyan, Ikuko Fujiwara, Jonathan E. Bird, Saima Riazuddin, Sheikh Riazuddin, Zubair M. Ahmed, Jenny E. Hinshaw, James Sellers, James R. Bartles, John A. Hammer, Guy P. Richardson, Andrew J. Griffith, Gregory I. Frolenkov, and Thomas B. Friedman. Actin-bundling protein triobp forms resilient rootlets of hair cell stereocilia essential for hearing. Cell, 141:786-798, May 2010. URL: https://doi.org/10.1016/j.cell.2010.03.049, doi:10.1016/j.cell.2010.03.049. This article has 234 citations and is from a highest quality peer-reviewed journal.
-
(zhang2024rfx3controlsouter pages 1-6): Penghui Zhang, Yafan Wang, Xiang Guo, Lu Ma, Xiangyao Zeng, Zhili Feng, Jinlei Liu, Mengzhen Yan, Yushan Gao, Jieran Dong, Junhong Li, Jie Ling, Hong Wu, Qianchen Jing, Yong Feng, and Jun Li. Rfx3 controls outer hair cell differentiation, maintenance, and hair bundle formation by regulating the expression of insm1, ikzf2, and triobp genes. bioRxiv, Sep 2024. URL: https://doi.org/10.1101/2024.09.24.614849, doi:10.1101/2024.09.24.614849. This article has 1 citations.
-
(lee2023revisitinggeneticepidemiology pages 9-10): Yen-Hui Lee, Cheng-Yu Tsai, Yue-Sheng Lu, Pei-Hsuan Lin, Yu-Ting Chiang, Ting-Hua Yang, Jacob Shu-Jui Hsu, Chuan-Jen Hsu, Pei-Lung Chen, Tien-Chen Liu, and Chen-Chi Wu. Revisiting genetic epidemiology with a refined targeted gene panel for hereditary hearing impairment in the taiwanese population. Genes, 14:880, Apr 2023. URL: https://doi.org/10.3390/genes14040880, doi:10.3390/genes14040880. This article has 10 citations.
-
(OpenTargets Search: autosomal recessive nonsyndromic hearing loss 28-TRIOBP): Open Targets Query (autosomal recessive nonsyndromic hearing loss 28-TRIOBP, 0 results). Buniello, A. et al. (2025). Open Targets Platform: facilitating therapeutic hypotheses building in drug discovery. Nucleic Acids Research.
Artifacts
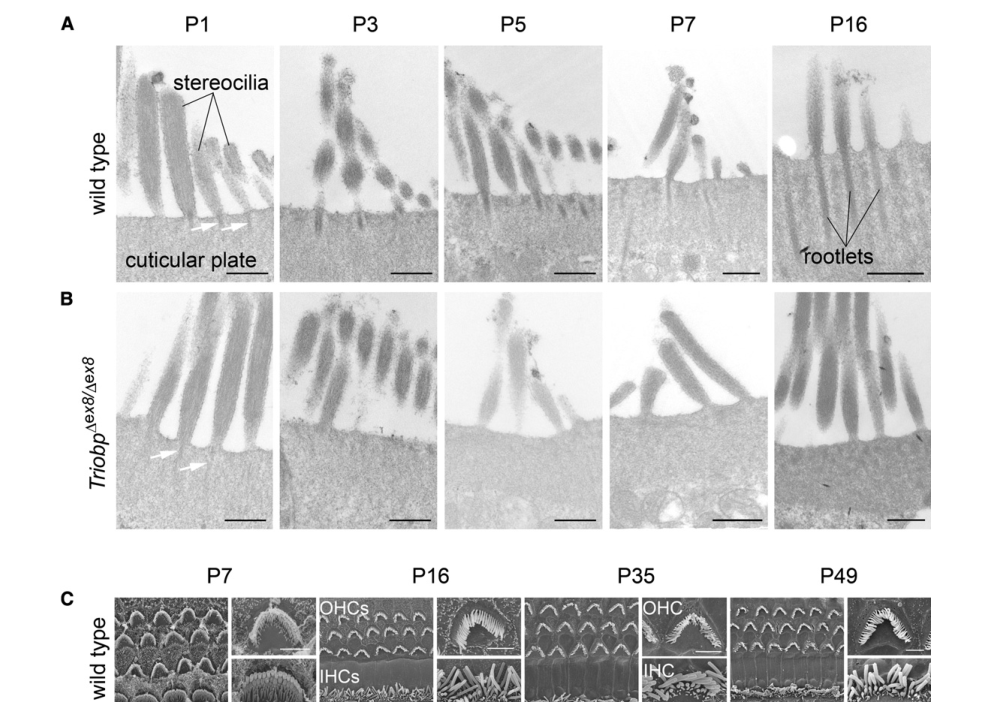
Reference Validation
Checked with linkml-reference-validator 0.3.0rc3.
| Outcome | Count |
|---|---|
| References checked | 14 |
| Resolved | 14 |
| Unresolved (possible confabulation) | 0 |
| Unverifiable | 0 |
| References weighed for topical relevance | 14 |
| On topic | 9 |
| Off topic | 0 |
All extracted references resolved successfully.
Term Validation
Checked with linkml-term-validator 0.4.5, through the ols: adapter.
| Outcome | Count |
|---|---|
| Terms checked | 1 |
| Resolved | 1 |
| Unresolved (possible confabulation) | 0 |
| Obsolete | 0 |
| Unverifiable | 0 |
| Terms whose name was checked | 1 |
| Terms named correctly | 0 |
| Terms named as a different term | 1 |
Terms the report names something else
These identifiers resolve, so nothing about them looks wrong, and the ontology calls them something unrelated to what the report calls them. That usually means the identifier is not the one the sentence needs:
MONDO:0012355(2 mentions) - the report calls it "if available"; MONDO calls it autosomal recessive nonsyndromic hearing loss 28