Autosomal Recessive Nonsyndromic Hearing Loss 124
DFNB124 is autosomal recessive nonsyndromic sensorineural hearing loss associated with biallelic PKHD1L1 variants. The founding human series described four unrelated probands with bilateral congenital or presumed-congenital hearing loss, ranging from mild-moderate to severe. Quantitative progression over nine years was documented in one proband; another was described as progressive. Attribution in the most severely affected proband remains uncertain because of a competing homozygous MYO7A variant. PKHD1L1 is a large, predominantly extracellular membrane protein associated with the developmental stereocilia surface coat. Knockout mice lose the upper stereociliary coat and subsequently develop stereocilia loss, bundle disorganization and progressive hearing impairment. Gross early bundle morphology and FM1-43 uptake were preserved under the tested conditions; these results do not establish normal mechanotransduction throughout life. Increased vulnerability to noise was demonstrated in mice, but has not been established in the reported human families. Human cochlear coat pathology and the relationship between developmental protein expression and later bundle maintenance remain unresolved.
Ask OpenScientist
Ask a research question about Autosomal Recessive Nonsyndromic Hearing Loss 124. OpenScientist will conduct autonomous deep research using the Disorder Mechanisms Knowledge Base and PubMed literature (typically 10-30 minutes).
Do not include personal health information in your question. Questions and results are cached in your browser's local storage.
Inheritance
1Show evidence (3 references)
Discussions and Knowledge Gaps
3Show evidence (4 references)
Show evidence (2 references)
Show evidence (2 references)
Pathophysiology
9Show evidence (4 references)
Show evidence (5 references)
Show evidence (8 references)
Show evidence (2 references)
Show evidence (1 reference)
Show evidence (2 references)
Show evidence (3 references)
Show evidence (1 reference)
Show evidence (2 references)
Pathograph
Phenotypes
5Show evidence (2 references)
Show evidence (2 references)
Show evidence (2 references)
Show evidence (2 references)
Show evidence (2 references)
Genetic Associations
1Show evidence (2 references)
External Assertions
1Show evidence (1 reference)
Medical Actions
5Show evidence (2 references)
Show evidence (2 references)
Show evidence (1 reference)
Show evidence (1 reference)
Show evidence (1 reference)
Environmental Factors
1Show evidence (1 reference)
Diagnosis
2Show evidence (4 references)
Show evidence (2 references)
Progression
2Show evidence (1 reference)
Show evidence (1 reference)
Prevalence
1Show evidence (4 references)
Animal Models
2Show evidence (1 reference)
Show evidence (1 reference)
Show evidence (1 reference)
Show evidence (1 reference)
Show evidence (1 reference)
Show evidence (1 reference)
Show evidence (1 reference)
Show evidence (1 reference)
Show evidence (1 reference)
Show evidence (2 references)
Show evidence (1 reference)
Source YAML
click to showname: Autosomal Recessive Nonsyndromic Hearing Loss 124
category: Mendelian
creation_date: "2026-09-03T00:00:00Z"
synonyms:
- DFNB124
- deafness, autosomal recessive 124
- autosomal recessive deafness 124
- PKHD1L1-related hearing loss
description: >-
DFNB124 is autosomal recessive nonsyndromic sensorineural hearing loss associated with biallelic PKHD1L1
variants. The founding human series described four unrelated probands with bilateral congenital or presumed-congenital
hearing loss, ranging from mild-moderate to severe. Quantitative progression over nine years was documented
in one proband; another was described as progressive. Attribution in the most severely affected proband remains
uncertain because of a competing homozygous MYO7A variant. PKHD1L1 is a large, predominantly extracellular
membrane protein associated with the developmental stereocilia surface coat. Knockout mice lose the upper
stereociliary coat and subsequently develop stereocilia loss, bundle disorganization and progressive hearing
impairment. Gross early bundle morphology and FM1-43 uptake were preserved under the tested conditions; these
results do not establish normal mechanotransduction throughout life. Increased vulnerability to noise was
demonstrated in mice, but has not been established in the reported human families. Human cochlear coat pathology
and the relationship between developmental protein expression and later bundle maintenance remain unresolved.
disease_term:
preferred_term: autosomal recessive nonsyndromic hearing loss 124
term:
id: MONDO:0968981
label: autosomal recessive nonsyndromic hearing loss 124
parents:
- Autosomal Recessive Nonsyndromic Hearing Loss
references:
- reference: PMID:38459354
title: "PKHD1L1, a gene involved in the stereocilia coat, causes autosomal recessive nonsyndromic hearing loss."
- reference: PMID:31444330
title: "PKHD1L1 is a coat protein of hair-cell stereocilia and is required for normal hearing."
- reference: PMID:39482437
title: "PKHD1L1 is required for stereocilia bundle maintenance, durable hearing function and resilience to noise exposure."
- reference: PMID:36960824
title: "A conserved function of Pkhd1l1, a mammalian hair cell stereociliary coat protein, in regulating hearing in zebrafish."
- reference: PMID:32555200
title: "Serial scanning electron microscopy of anti-PKHD1L1 immuno-gold labeled mouse hair cell stereocilia bundles."
- reference: url:https://www.ncbi.nlm.nih.gov/sites/books/NBK1434/?report=reader
title: Genetic Hearing Loss Overview - GeneReviews® - NCBI Bookshelf
tags:
- GeneReviews
inheritance:
- name: Autosomal recessive
description: >-
Two homozygous and two compound heterozygous PKHD1L1 genotypes were reported in four unrelated families.
Segregation was confirmed in Families 1, 2 and 4. The parents in Family 3 did not participate, and a competing
homozygous MYO7A variant limits causal attribution in that proband. The reported pedigrees support recessive
inheritance, without establishing population penetrance.
inheritance_term:
preferred_term: Autosomal recessive inheritance
term:
id: HP:0000007
label: Autosomal recessive inheritance
evidence:
- reference: PMID:38459354
reference_title: "PKHD1L1, a gene involved in the stereocilia coat, causes autosomal recessive nonsyndromic hearing loss."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "Exome sequencing was performed on DNA of four families segregating autosomal recessive nonsyndromic sensorineural hearing loss."
explanation: The founding study evaluated four unrelated families; segregation confirmation was available for three.
- reference: PMID:38459354
reference_title: "PKHD1L1, a gene involved in the stereocilia coat, causes autosomal recessive nonsyndromic hearing loss."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "Sanger sequencing at this locus confirmed the homozygous variant and revealed that the parents were both heterozygous carriers of the c.10141C>T, p.(Arg3381Ter) substitution."
explanation: >-
Carrier confirmation in the parents of the family with the clearest null allele, which is what makes
the recessive model explicit rather than inferred from homozygosity alone.
- reference: PMID:38459354
reference_title: "PKHD1L1, a gene involved in the stereocilia coat, causes autosomal recessive nonsyndromic hearing loss."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: PKHD1L1 variant segregation in Families 1, 2, and 4 was confirmed using Sanger sequencing, but not for the proband in Family 3.
explanation: Distinguishes confirmed segregation from the untested parents in Family 3.
pathophysiology:
- name: Biallelic PKHD1L1 Functional Impairment
biological_scale: MOLECULAR
description: >-
Reported biallelic PKHD1L1 variants are consistent with reduced protein function, with allele-dependent
evidence. p.Arg3381Ter and p.Leu2818TyrfsTer5 are predicted truncating alleles. p.Gly129Ser and p.Gly1314Val
destabilized recombinant mouse PKHD1L1 fragments carrying the corresponding substitutions; full-length
protein function was not measured. c.1813G>A, annotated p.Gly605Arg, caused exon 17 skipping in HEK293
and HeLa minigene assays. The contribution of p.His2479Gln remains uncertain because functional testing
and parental segregation were unavailable and the proband also carried a homozygous MYO7A variant.
genes:
- preferred_term: PKHD1L1
term:
id: hgnc:20313
label: PKHD1L1
genetic_context:
genes:
- preferred_term: PKHD1L1
term:
id: hgnc:20313
label: PKHD1L1
allele_type: nonsense, frameshift, splice-disrupting and destabilising missense variants
variant_origin: GERMLINE
functional_impact_category: LOSS_OF_FUNCTION
description: >-
Two reported genotypes were homozygous and two compound heterozygous; no single zygosity value represents
the disease-level node.
notes: >-
Reduced function is the proposed shared mechanism, not a measured property of every allele. Nonsense-mediated
decay of p.Arg3381Ter is predicted. NanoDSF measured decreased folding stability of recombinant fragments,
not full-length activity in a hair cell. Exon skipping was measured in minigenes, not patient cochlear
RNA. p.His2479Gln lacks direct functional evidence.
evidence:
- reference: PMID:38459354
reference_title: "PKHD1L1, a gene involved in the stereocilia coat, causes autosomal recessive nonsyndromic hearing loss."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "The proband was compound heterozygous for missense variants, c.385G>A, p.(Gly129Ser) and c.3941G>T, p.(Gly1314Val)"
explanation: >-
Compound heterozygous genotype of Family 1.
- reference: PMID:38459354
reference_title: "PKHD1L1, a gene involved in the stereocilia coat, causes autosomal recessive nonsyndromic hearing loss."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "Exome sequencing revealed that the proband was homozygous for the c.10141C>T, p.(Arg3381Ter) nonsense variant"
explanation: >-
Homozygous nonsense genotype of Family 2; its molecular consequence is predicted.
- reference: PMID:38459354
reference_title: "PKHD1L1, a gene involved in the stereocilia coat, causes autosomal recessive nonsyndromic hearing loss."
supports: SUPPORT
evidence_source: IN_VITRO
snippet: "In vitro functional assessment indicated that both engineered PKHD1L1 p.(Gly129Ser) and p.(Gly1314Val) mutant constructs significantly reduced the folding and structural stabilities of the expressed protein fragments, providing further evidence to support pathogenicity of these variants."
explanation: >-
The functional evidence for the two missense alleles, and its precise scope: folding and structural stability
of expressed fragments, measured by nanoDSF.
- reference: PMID:38459354
reference_title: "PKHD1L1, a gene involved in the stereocilia coat, causes autosomal recessive nonsyndromic hearing loss."
supports: SUPPORT
evidence_source: IN_VITRO
snippet: "Minigene assay of the c.1813G>A p.(Gly605Arg) variant, located at the boundary of exon 17, revealed exon skipping leading to an in-frame deletion of 48 amino acids."
explanation: >-
Exon skipping in HEK293 and HeLa minigene assays; patient cochlear splicing was not measured.
downstream:
- target: Loss of the Stereocilia Surface Coat at Stereocilia Tips
causal_link_type: INDIRECT_KNOWN_INTERMEDIATES
description: >-
Loss of Pkhd1l1 causes upper coat deficiency in knockout mice. Extrapolation to the effects of individual
human alleles is provisional; human cochlear coat morphology has not been examined.
- name: Loss of the Stereocilia Surface Coat at Stereocilia Tips
biological_scale: MOLECULAR
description: >-
PKHD1L1 immunogold labeling is enriched near stereocilia tips in developing mouse cochlear hair cells.
Conditional knockout markedly reduces tannic-acid-stained coat material at the upper stereocilia while
preserving the lower coat. The six reconstructed P4 outer hair-cell bundles came from one cochlear sample.
Immunogold density measures relative labeling, not molecule counts, and does not establish that PKHD1L1
alone comprises the upper coat.
cellular_components:
- preferred_term: upper stereocilium coat
modifier: DECREASED
term:
id: GO:0120234
label: stereocilium coat
- preferred_term: stereocilium tip
term:
id: GO:0032426
label: stereocilium tip
cell_types:
- preferred_term: cochlear outer hair cell
term:
id: CL:0000601
label: cochlear outer hair cell
- preferred_term: cochlear inner hair cell
term:
id: CL:0000589
label: cochlear inner hair cell
locations:
- preferred_term: organ of Corti
term:
id: UBERON:0002227
label: spiral organ of cochlea
evidence:
- reference: PMID:31444330
reference_title: "PKHD1L1 is a coat protein of hair-cell stereocilia and is required for normal hearing."
supports: SUPPORT
evidence_source: MODEL_ORGANISM
directness: INDIRECT
snippet: "PKHD1L1-deficient mice lack the surface coat at the upper but not lower regions of stereocilia, and they develop progressive hearing loss."
explanation: >-
The compartmental specificity of the lesion - upper stereocilia only - and the hearing consequence, in
one sentence.
- reference: PMID:31444330
reference_title: "PKHD1L1 is a coat protein of hair-cell stereocilia and is required for normal hearing."
supports: SUPPORT
evidence_source: MODEL_ORGANISM
directness: INDIRECT
snippet: "We conclude that PKHD1L1 is the major component of the upper surface coat of hair-cell stereocilia, and is essential for normal hearing especially at high frequencies."
explanation: The authors' summary of what the protein is and where it acts.
- reference: PMID:31444330
reference_title: "PKHD1L1 is a coat protein of hair-cell stereocilia and is required for normal hearing."
supports: SUPPORT
evidence_source: MODEL_ORGANISM
directness: INDIRECT
snippet: "Using serial immunogold scanning electron microscopy, we show that PKHD1L1 is expressed at the tips of stereocilia, especially in the high-frequency regions of the cochlea."
explanation: >-
The localisation, and the basal-to-apical gradient that matches where the hearing loss begins.
- reference: PMID:32555200
reference_title: "Serial scanning electron microscopy of anti-PKHD1L1 immuno-gold labeled mouse hair cell stereocilia bundles."
supports: SUPPORT
evidence_source: MODEL_ORGANISM
snippet: The serial EM datasets in this submission are of six closely located OHCs, collected from the middle cochlear turn of a P4 mouse cochlea.
explanation: The methods/data paper documents the sampling underlying the earlier localization result; it is not an independent biological replication.
- reference: PMID:31444330
reference_title: "PKHD1L1 is a coat protein of hair-cell stereocilia and is required for normal hearing."
supports: SUPPORT
evidence_source: MODEL_ORGANISM
snippet: Thus PKHD1L1 either might itself form the bulk of the surface coat at P4 at the tips, or facilitate the presence of other proteins that compose the coat.
explanation: The molecular composition and recruitment mechanism remain unresolved.
downstream:
- target: Impaired Stereocilia Bundle Maintenance
causal_link_type: INDIRECT_UNKNOWN_INTERMEDIATES
description: >-
What the coat does mechanically is not known, so the step from its absence to the loss of bundle integrity
is a demonstrated association without an identified intermediate.
- name: Impaired Stereocilia Bundle Maintenance
biological_scale: CELLULAR
description: >-
Mouse knockout findings support impaired long-term bundle maintenance despite preserved gross early morphology.
P3 planar polarity, P4 explants cultured for three days before FM1-43 uptake assessment, stereocilin localization
and tectorial-membrane imprints did not show major deficits in the tested assays. FM1-43 uptake is a proxy
for channel permeability and does not establish normal mechanotransduction currents at all ages. Immunolabeling
declines during early postnatal development and is undetectable by P21 with the antibody used. The authors
propose that early PKHD1L1 expression establishes durable bundles, but adult protein below detection or
lacking the antibody epitope cannot be excluded.
biological_processes:
- preferred_term: auditory receptor cell stereocilium organization
modifier: ABNORMAL
term:
id: GO:0060088
label: auditory receptor cell stereocilium organization
cellular_components:
- preferred_term: stereocilium bundle
term:
id: GO:0032421
label: stereocilium bundle
cell_types:
- preferred_term: cochlear outer hair cell
term:
id: CL:0000601
label: cochlear outer hair cell
evidence:
- reference: PMID:39482437
reference_title: "PKHD1L1 is required for stereocilia bundle maintenance, durable hearing function and resilience to noise exposure."
supports: SUPPORT
evidence_source: MODEL_ORGANISM
directness: INDIRECT
snippet: "Our findings reveal that PKHD1L1-deficient mice display no disruption to bundle cohesion or tectorial membrane attachment-crown formation during development."
explanation: >-
Preserved gross developmental morphology supports the maintenance hypothesis without excluding subtler
developmental abnormalities.
- reference: PMID:39482437
reference_title: "PKHD1L1 is required for stereocilia bundle maintenance, durable hearing function and resilience to noise exposure."
supports: SUPPORT
evidence_source: MODEL_ORGANISM
directness: INDIRECT
snippet: "However, starting from 6 weeks of age, PKHD1L1-deficient mice display missing stereocilia and disruptions to bundle coherence."
explanation: The onset of the lesion, in the same sentence structure as the negative above.
- reference: PMID:39482437
reference_title: "PKHD1L1 is required for stereocilia bundle maintenance, durable hearing function and resilience to noise exposure."
supports: SUPPORT
evidence_source: MODEL_ORGANISM
directness: INDIRECT
snippet: "We therefore conclude that PKHD1L1 is not required for the localization of STRC to attachment crowns or horizontal top connectors."
explanation: >-
Preserved stereocilin localization argues against defective initial attachment-crown assembly; later
attachment weakening remains possible.
- reference: PMID:31444330
reference_title: "PKHD1L1 is a coat protein of hair-cell stereocilia and is required for normal hearing."
supports: SUPPORT
evidence_source: IN_VITRO
directness: INDIRECT
snippet: "FM1-43 loading by PKHD1L1-deficient hair cells was similar to that in normal hearing littermates, suggesting no major deficiency of the hair-cell mechanotransduction complex"
explanation: >-
FM1-43 uptake in neonatal mouse cochlear explants supports retained channel permeability at this stage,
not normal lifetime mechanotransduction.
- reference: PMID:39482437
reference_title: "PKHD1L1 is required for stereocilia bundle maintenance, durable hearing function and resilience to noise exposure."
supports: SUPPORT
evidence_source: MODEL_ORGANISM
directness: INDIRECT
snippet: "Anti-PKHD1L1 labeling intensity gradually decreased with development, was largely gone by P10, and completely cleared in mature hair cells at P21."
explanation: >-
The measured endpoint is antibody labeling, not proof that all protein or fragments are absent.
- reference: PMID:39482437
reference_title: "PKHD1L1 is required for stereocilia bundle maintenance, durable hearing function and resilience to noise exposure."
supports: SUPPORT
evidence_source: MODEL_ORGANISM
directness: INDIRECT
snippet: "PKHD1L1 protein expression labeling is restricted to early postnatal development and localizes to the hair bundles of both IHCs and OHCs."
explanation: >-
The same restriction stated as a conclusion, and the note that both hair cell types carry the protein
even though the phenotype is read out mainly in outer hair cells.
- reference: PMID:39482437
reference_title: "PKHD1L1 is required for stereocilia bundle maintenance, durable hearing function and resilience to noise exposure."
supports: SUPPORT
evidence_source: MODEL_ORGANISM
snippet: TM imprints formed during development are likely permanent however47, therefore these data do not preclude the possibility of TM attachments weakening in PKHD1L1-deficient mice as they age and/or are exposed to noise insult.
explanation: The full-text discussion explicitly limits inference from preserved imprints.
- reference: PMID:39482437
reference_title: "PKHD1L1 is required for stereocilia bundle maintenance, durable hearing function and resilience to noise exposure."
supports: SUPPORT
evidence_source: MODEL_ORGANISM
snippet: PKHD1L1 splice variants lacking the NBP2-13765 epitope region in adult IHCs cannot be ruled out.
explanation: Alternative explanations for undetectable adult immunolabeling remain hypotheses, not demonstrated isoforms.
downstream:
- target: Progressive Stereocilia Loss
- target: Reduced Resilience to Acoustic Overexposure
mechanism_confidence: PROVISIONAL
- name: Progressive Stereocilia Loss
biological_scale: CELLULAR
description: >-
In conditional knockout mice, missing outer hair-cell stereocilia are detectable from six weeks. Loss is
most pronounced in the shortest row, progressing to taller rows with age, especially in the basal cochlea.
This distribution suggests a role beyond tall-row tectorial-membrane attachment. Human stereocilia loss
has not been directly examined.
cellular_components:
- preferred_term: stereocilium bundle
term:
id: GO:0032421
label: stereocilium bundle
cell_types:
- preferred_term: cochlear outer hair cell
term:
id: CL:0000601
label: cochlear outer hair cell
evidence:
- reference: PMID:39482437
reference_title: "PKHD1L1 is required for stereocilia bundle maintenance, durable hearing function and resilience to noise exposure."
supports: SUPPORT
evidence_source: MODEL_ORGANISM
directness: INDIRECT
snippet: "stereocilia loss was most significant in the shortest row, progressing to the taller rows with age; most prominently in, but not limited to, the high frequency basal region"
explanation: The spatial pattern of the loss, by row and by cochlear region.
- reference: PMID:39482437
reference_title: "PKHD1L1 is required for stereocilia bundle maintenance, durable hearing function and resilience to noise exposure."
supports: SUPPORT
evidence_source: MODEL_ORGANISM
directness: INDIRECT
snippet: "This phenotype is unlikely to be caused by disruption of tall row stereocilia attachment to the TM."
explanation: >-
The authors' own reading of why short-row-first loss argues against the tectorial-membrane explanation.
downstream:
- target: Stereocilia Bundle Disorganization
causal_link_type: INDIRECT_UNKNOWN_INTERMEDIATES
description: Loss and disorganization occur together in the knockout; the intervening process has not been experimentally isolated.
- name: Stereocilia Bundle Disorganization
biological_scale: CELLULAR
description: >-
Outer hair-cell bundle disorganization is observed at fourteen weeks and is more pronounced at nine months
in conditional knockout mice. Its relationship to individual stereocilia loss and declining function is
inferred from their co-occurrence; these morphological endpoints have not been measured in affected humans.
cellular_components:
- preferred_term: stereocilium bundle
term:
id: GO:0032421
label: stereocilium bundle
cell_types:
- preferred_term: cochlear outer hair cell
term:
id: CL:0000601
label: cochlear outer hair cell
evidence:
- reference: PMID:39482437
reference_title: "PKHD1L1 is required for stereocilia bundle maintenance, durable hearing function and resilience to noise exposure."
supports: SUPPORT
evidence_source: MODEL_ORGANISM
snippet: bundle disorganization is observed at 14 weeks increasing in severity by 9 months of age.
explanation: The structural endpoint and its time course in conditional knockout mice.
downstream:
- target: Outer Hair Cell Dysfunction
causal_link_type: INDIRECT_UNKNOWN_INTERMEDIATES
description: Bundle pathology accompanies elevated DPOAE thresholds in knockout mice; direct mediation has not been isolated.
- name: Reduced Resilience to Acoustic Overexposure
biological_scale: CELLULAR
description: >-
In six-week-old conditional knockout mice, 94 dB SPL octave-band noise at 8-16 kHz for two hours produces
ABR and DPOAE threshold shifts persisting to eight weeks. Cre-negative floxed littermates recover to baseline
by two weeks. Post-noise mutant bundles show missing or shortened tall-row stereocilia and disorganization.
This genotype-dependent vulnerability is demonstrated in mice; increased susceptibility in humans remains
untested.
cellular_components:
- preferred_term: stereocilium bundle
term:
id: GO:0032421
label: stereocilium bundle
cell_types:
- preferred_term: cochlear outer hair cell
term:
id: CL:0000601
label: cochlear outer hair cell
mechanism_confidence: PROVISIONAL
notes: >-
Human transfer is provisional. The exposure experiment does not establish a clinical noise threshold or
a DFNB124-specific prevention effect.
evidence:
- reference: PMID:39482437
reference_title: "PKHD1L1 is required for stereocilia bundle maintenance, durable hearing function and resilience to noise exposure."
supports: SUPPORT
evidence_source: MODEL_ORGANISM
directness: INDIRECT
snippet: "Furthermore, PKHD1L1-deficient mice are susceptible to permanent hearing loss following moderate acoustic overexposure, which induces only temporary hearing threshold shifts in wild-type mice."
explanation: >-
The result itself, stated as the contrast that defines it: the same dose, two different outcomes by genotype.
- reference: PMID:39482437
reference_title: "PKHD1L1 is required for stereocilia bundle maintenance, durable hearing function and resilience to noise exposure."
supports: SUPPORT
evidence_source: MODEL_ORGANISM
directness: INDIRECT
snippet: "These findings demonstrate that deficiency in PKHD1L1 increases susceptibility to bundle disruption during moderate acoustic overexposure, leading to permanent threshold shifts."
explanation: >-
The morphological correlate, which is what connects this node to the bundle nodes rather than leaving
it a threshold observation.
downstream:
- target: Progressive Stereocilia Loss
- target: Outer Hair Cell Dysfunction
- name: Outer Hair Cell Dysfunction
biological_scale: CELLULAR
description: >-
Elevated DPOAE thresholds in Pkhd1l1 knockout mice support outer hair-cell dysfunction. In the human series,
Family 4 had absent DPOAEs with normal tympanograms, while Family 2 had preserved bilateral otoacoustic
emissions despite moderate-to-severe hearing loss. Thus outer hair-cell dysfunction is supported in one
reported patient and in mice, but is not established as a uniform human lesion.
cell_types:
- preferred_term: cochlear outer hair cell
term:
id: CL:0000601
label: cochlear outer hair cell
locations:
- preferred_term: cochlea
term:
id: UBERON:0001844
label: cochlea
evidence:
- reference: PMID:38459354
reference_title: "PKHD1L1, a gene involved in the stereocilia coat, causes autosomal recessive nonsyndromic hearing loss."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "DPOAEs were absent in both ears, and the tympanograms were normal, suggesting dysfunction of the outer hair cells."
explanation: >-
The only human measurement in this entry that speaks to the cellular level, and the authors' interpretation
of it.
- reference: PMID:31444330
reference_title: "PKHD1L1 is a coat protein of hair-cell stereocilia and is required for normal hearing."
supports: SUPPORT
evidence_source: MODEL_ORGANISM
directness: INDIRECT
snippet: "PKHD1L1-deficient mice show elevated thresholds in response to high-frequency tones"
explanation: >-
The mouse counterpart. Both auditory brainstem response and distortion product thresholds rise together
in this model, which is what places the lesion in the outer hair cells rather than centrally.
- reference: PMID:38459354
reference_title: "PKHD1L1, a gene involved in the stereocilia coat, causes autosomal recessive nonsyndromic hearing loss."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "Speech audiometry understanding is 100% at a comfortable listening level, and the otoacoustic emissions were present bilaterally."
explanation: >-
The contrary observation in another proband, recorded so the phenotype is not read as uniform across
the series.
downstream:
- target: Reduced Cochlear Amplification
- target: Absent Otoacoustic Emissions
- name: Reduced Cochlear Amplification
biological_scale: TISSUE
locations:
- preferred_term: cochlea
term:
id: UBERON:0001844
label: cochlea
description: >-
Reduced cochlear amplification is inferred from elevated DPOAE thresholds accompanying ABR threshold elevations
in knockout mice. The measurements implicate an outer hair-cell contribution to hearing loss without establishing
an exclusive cellular lesion or directly measuring electromotility.
evidence:
- reference: PMID:31444330
reference_title: "PKHD1L1 is a coat protein of hair-cell stereocilia and is required for normal hearing."
supports: SUPPORT
evidence_source: MODEL_ORGANISM
snippet: Overall, elevated ABR thresholds of Pkhd1l1fl/fl, Atoh1-Cre+ animals closely followed the DPOAE thresholds, suggesting the hearing phenotype is likely to arise primarily from an OHC deficit.
explanation: Coupled threshold elevations support an outer hair-cell amplification deficit in mice.
downstream:
- target: Cochlear Hearing Loss
- name: Cochlear Hearing Loss
biological_scale: ORGANISM
conforms_to: "sensorineural_hair_cell_loss#Progressive Sensorineural Hearing Loss"
description: >-
The human endpoint is bilateral congenital or presumed-congenital sensorineural hearing loss with variable
severity. Slow progression was quantified in Family 1 and progression was also described in Family 2. The
adult course and general progression rate are unknown. Conditional and constitutive knockout mice show
high-frequency hearing impairment extending to lower frequencies with age.
notes: >-
Conformance is limited to the module endpoint, with progression documented in a subset of humans and in
mice. The module intermediate involving mechanotransduction failure and hair-cell death is not asserted:
neonatal FM1-43 uptake does not establish normal lifetime transduction, and the cited studies do not demonstrate
this combined intermediate in DFNB124. Human congenital onset is a timing descriptor, not a consequence
of progressive hearing loss.
evidence:
- reference: PMID:38459354
reference_title: "PKHD1L1, a gene involved in the stereocilia coat, causes autosomal recessive nonsyndromic hearing loss."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "Multiple lines of evidence collectively associate PKHD1L1 with nonsyndromic mild-moderate to severe sensorineural hearing loss."
explanation: The human endpoint and its severity range, as the defining paper states it.
- reference: PMID:39482437
reference_title: "PKHD1L1 is required for stereocilia bundle maintenance, durable hearing function and resilience to noise exposure."
supports: SUPPORT
evidence_source: MODEL_ORGANISM
directness: INDIRECT
snippet: "Both conditional and constitutive PKHD1L1 knockout mice develop high-frequency hearing loss progressing to lower frequencies with age."
explanation: >-
The mouse endpoint in two independent knockout designs, which rules out a Cre-transgene artefact as the
source of the phenotype.
downstream:
- target: Bilateral Sensorineural Hearing Impairment
- target: Slowly Progressive Hearing Impairment
- target: Severe Hearing Impairment
phenotypes:
- name: Bilateral Sensorineural Hearing Impairment
category: Ear
description: >-
The defining and, in all four reported families, the only feature. Loss is bilateral and sensorineural,
spanning mild-moderate to severe across the four probands.
phenotype_term:
preferred_term: Bilateral sensorineural hearing impairment
term:
id: HP:0008619
label: Bilateral sensorineural hearing impairment
notes: >-
All four probands had bilateral sensorineural hearing loss, but this ascertained series does not estimate
penetrance or population phenotype frequency.
evidence:
- reference: PMID:38459354
reference_title: "PKHD1L1, a gene involved in the stereocilia coat, causes autosomal recessive nonsyndromic hearing loss."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "bilateral congenital SNHL which is nonsyndromic and mild–moderate to severe"
explanation: The phenotype, its laterality, and its severity range in one clause.
- reference: PMID:38459354
reference_title: "PKHD1L1, a gene involved in the stereocilia coat, causes autosomal recessive nonsyndromic hearing loss."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "Additional syndromic involvement was excluded in all four probands."
explanation: >-
The statement that makes the entity nonsyndromic, and the reason the entry curates no eye, cardiac or
cognitive phenotype.
- name: Congenital Onset of Hearing Impairment
category: Ear
description: >-
Hearing loss is present from birth or infancy. The proband of Family 1 failed newborn hearing screening
bilaterally; the proband of Family 2 was clinically diagnosed at two and a half months. In Family 4 the
loss is believed congenital by parental report but newborn screening was not performed.
phenotype_term:
preferred_term: Congenital sensorineural hearing impairment
term:
id: HP:0008527
label: Congenital sensorineural hearing impairment
notes: >-
The paper describes congenital or presumed-congenital onset. Family 4 lacked newborn screening and onset
relies on parental report. These observations do not justify a quantitative frequency band.
evidence:
- reference: PMID:38459354
reference_title: "PKHD1L1, a gene involved in the stereocilia coat, causes autosomal recessive nonsyndromic hearing loss."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "Although presenting congenitally in the majority of patients, the degree of hearing impairment in the patients we present is fairly broad"
explanation: >-
The cautious form of the onset claim, and the same sentence's statement that severity varies widely.
- reference: PMID:38459354
reference_title: "PKHD1L1, a gene involved in the stereocilia coat, causes autosomal recessive nonsyndromic hearing loss."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "She did not pass a newborn hearing screen bilaterally."
explanation: >-
The one proband in whom congenital onset is documented by a screening test rather than inferred.
- name: Slowly Progressive Hearing Impairment
category: Ear
description: >-
Family 1 had serial audiometry approximately every six months. From ages 4.3 to 13.3 years, pure-tone average
increased by 5 dB in the right ear and 8 dB in the left. Family 2 was also described as progressing to
moderate-to-severe loss, without a comparable numerical longitudinal series.
phenotype_term:
preferred_term: Progressive sensorineural hearing impairment
term:
id: HP:0000408
label: Progressive sensorineural hearing impairment
notes: >-
The rate in Family 1 cannot be generalized to all genotypes or adulthood. Progression frequency cannot
be estimated from this small series.
evidence:
- reference: PMID:38459354
reference_title: "PKHD1L1, a gene involved in the stereocilia coat, causes autosomal recessive nonsyndromic hearing loss."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "consistently demonstrated a slowly progressive mild to moderate SNHL bilaterally"
explanation: >-
The progression claim, from the one proband with nine years of serial pure-tone audiometry.
- reference: PMID:38459354
reference_title: "PKHD1L1, a gene involved in the stereocilia coat, causes autosomal recessive nonsyndromic hearing loss."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "It has progressed to a bilateral moderate to severe degree. Pure-tone audiometry shows moderate to severe SNHL at all frequencies."
explanation: >-
The Family 2 proband, whose loss reached moderate-to-severe by age nine and who has the cleanest genotype
in the series, a homozygous nonsense allele.
- name: Severe Hearing Impairment
category: Ear
description: >-
The severe end of the range. The proband of Family 3 had bilateral severe loss, and the proband of Family
2 moderate-to-severe loss at all frequencies by age nine.
phenotype_term:
preferred_term: Severe sensorineural hearing impairment
term:
id: HP:0008625
label: Severe sensorineural hearing impairment
notes: >-
Family 3 had a pure-tone average of 85 dB HL, measured in ambient noise without a soundproof testing environment.
Attribution is uncertain because homozygous MYO7A p.Leu375Val accompanied PKHD1L1 p.His2479Gln. Family
2 independently had moderate-to-severe loss. Neither severity frequency nor genotype-severity correlations
can be inferred from these cases.
evidence:
- reference: PMID:38459354
reference_title: "PKHD1L1, a gene involved in the stereocilia coat, causes autosomal recessive nonsyndromic hearing loss."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "his audiometric testing demonstrated a bilateral severe SNHL"
explanation: The severe pole of the reported range, in the Family 3 proband.
- reference: PMID:38459354
reference_title: "PKHD1L1, a gene involved in the stereocilia coat, causes autosomal recessive nonsyndromic hearing loss."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "It has progressed to a bilateral moderate to severe degree. Pure-tone audiometry shows moderate to severe SNHL at all frequencies."
explanation: >-
The Family 2 proband, whose loss reached moderate-to-severe by age nine and who has the cleanest genotype
in the series, a homozygous nonsense allele.
- name: Absent Otoacoustic Emissions
category: Ear
description: >-
Absent distortion product otoacoustic emissions with normal tympanometry in the proband of Family 4, which
localises the failure to the outer hair cells rather than the middle ear. Emissions were by contrast present
bilaterally in the proband of Family 2 despite moderate-to-severe loss.
phenotype_term:
preferred_term: Absent otoacoustic emissions
term:
id: HP:6000182
label: Absent otoacoustic emissions
notes: >-
Curated because it is the only human measurement in the series that speaks to cell type, and because it
is not uniform: one proband has absent emissions and another has preserved ones. Both facts are recorded
rather than the convenient one alone.
evidence:
- reference: PMID:38459354
reference_title: "PKHD1L1, a gene involved in the stereocilia coat, causes autosomal recessive nonsyndromic hearing loss."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "DPOAEs were absent in both ears, and the tympanograms were normal, suggesting dysfunction of the outer hair cells."
explanation: The finding and the inference the clinicians drew from it.
- reference: PMID:38459354
reference_title: "PKHD1L1, a gene involved in the stereocilia coat, causes autosomal recessive nonsyndromic hearing loss."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "Speech audiometry understanding is 100% at a comfortable listening level, and the otoacoustic emissions were present bilaterally."
explanation: >-
The contrary observation in another proband, recorded so the phenotype is not read as uniform across
the series.
environmental:
- name: Moderate acoustic overexposure
description: >-
In six-week-old conditional knockout mice, 94 dB SPL octave-band noise at 8-16 kHz for two hours produces
ABR and DPOAE threshold shifts persisting to eight weeks. Cre-negative floxed littermates recover to baseline
by two weeks. Post-noise mutant bundles show missing or shortened tall-row stereocilia and disorganization.
This genotype-dependent vulnerability is demonstrated in mice; increased susceptibility in humans remains
untested.
exposure_term:
preferred_term: exposure to sound radiation
term:
id: ECTO:8000044
label: exposure to sound radiation
influences_mechanisms:
- target: Reduced Resilience to Acoustic Overexposure
environmental_effect: EXACERBATES
causal_link_type: DIRECT
description: >-
The exposure is what converts the latent bundle fragility into permanent damage. It does not cause the
disease - the mutant bundle deteriorates with age without any noise challenge - so the effect is EXACERBATES
rather than TRIGGERS.
evidence:
- reference: PMID:39482437
reference_title: "PKHD1L1 is required for stereocilia bundle maintenance, durable hearing function and resilience to noise exposure."
supports: SUPPORT
evidence_source: MODEL_ORGANISM
directness: INDIRECT
snippet: "These data therefore show the noise exposure, normally only sufficient to cause TTS in normal mice, leads to PTS in PKHD1L1-deficient mice."
explanation: >-
The exposure acting on the mechanism, stated as a genotype-dependent conversion of a temporary shift
into a permanent one.
evidence:
- reference: PMID:39482437
reference_title: "PKHD1L1 is required for stereocilia bundle maintenance, durable hearing function and resilience to noise exposure."
supports: SUPPORT
evidence_source: MODEL_ORGANISM
directness: INDIRECT
snippet: "Furthermore, PKHD1L1-deficient mice are susceptible to permanent hearing loss following moderate acoustic overexposure, which induces only temporary hearing threshold shifts in wild-type mice."
explanation: >-
Entry-level evidence that acoustic overexposure is a factor in this disease at all, separate from the
claim about which mechanism it acts on.
notes: >-
This is a mouse modifier experiment in a genetic hearing disorder. Human DFNB124 noise susceptibility and
prevention efficacy have not been established in the cited studies.
prevalence:
- population: Worldwide
measure_type: CASES_IN_LITERATURE
notes: >-
The 2024 founding paper described four unrelated probands from the United States, Iran, Pakistan and China.
Population prevalence, penetrance and founder effects cannot be inferred from this series. Family 4 came
from a subset of 449 exome-sequenced probands within a 1450-person hearing-loss cohort; testing was not
uniform. Neither one in 1450 nor one in 449 establishes a population rate or an upper bound on disease
frequency.
evidence:
- reference: PMID:38459354
reference_title: "PKHD1L1, a gene involved in the stereocilia coat, causes autosomal recessive nonsyndromic hearing loss."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "Exome sequencing was performed on DNA of four families segregating autosomal recessive nonsyndromic sensorineural hearing loss."
explanation: Size of the founding case series, not a population denominator.
- reference: PMID:38459354
reference_title: "PKHD1L1, a gene involved in the stereocilia coat, causes autosomal recessive nonsyndromic hearing loss."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "PKHD1L1 testing in individuals with mild-moderate hearing loss may identify further affected families."
explanation: >-
The authors' own ascertainment argument, which is why the case count should not be read as an upper bound.
- reference: PMID:38459354
reference_title: "PKHD1L1, a gene involved in the stereocilia coat, causes autosomal recessive nonsyndromic hearing loss."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "This set comprises a total of 1450 hearing loss probands mainly from Henan Province, China, and it includes syndromic and nonsyndromic hearing loss."
explanation: >-
Parent cohort size; only a subset underwent exome sequencing.
- reference: PMID:38459354
reference_title: "PKHD1L1, a gene involved in the stereocilia coat, causes autosomal recessive nonsyndromic hearing loss."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: The proband in Family 4 was identified from a subset of 449 probands for whom exome sequencing data were available.
explanation: Correct ascertainment denominator for the exome-sequenced subset.
progression:
- phase: Congenital and early childhood
notes: >-
The reported onset was congenital or presumed congenital, but age at recognition varied; Family 4 presented
at age eight without a newborn screen.
evidence:
- reference: PMID:38459354
reference_title: "PKHD1L1, a gene involved in the stereocilia coat, causes autosomal recessive nonsyndromic hearing loss."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "bilateral congenital SNHL which is nonsyndromic and mild–moderate to severe"
explanation: The state of hearing at presentation.
- phase: Slow progression through childhood
notes: >-
Slow progression was quantified over nine years in Family 1; progression was also reported in Family 2.
The oldest proband in this 2024 series was thirteen. The series does not establish adult natural history
or a general rate.
evidence:
- reference: PMID:38459354
reference_title: "PKHD1L1, a gene involved in the stereocilia coat, causes autosomal recessive nonsyndromic hearing loss."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "consistently demonstrated a slowly progressive mild to moderate SNHL bilaterally"
explanation: The pace of change over the longest documented follow-up in the series.
genetic:
- name: PKHD1L1
relationship_type: CAUSATIVE
variant_origin: GERMLINE
gene_term:
preferred_term: PKHD1L1
term:
id: hgnc:20313
label: PKHD1L1
notes: >-
Reported genotypes, using NM_177531.6 and NP_803875.2, were c.385G>A/p.Gly129Ser in trans with c.3941G>T/p.Gly1314Val;
homozygous c.10141C>T/p.Arg3381Ter; homozygous c.7437C>A/p.His2479Gln; and c.1813G>A/p.Gly605Arg in trans
with c.8452_8468del/p.Leu2818TyrfsTer5. Functional evidence differs among alleles: recombinant mouse-fragment
stability assays support the first pair, and minigene splicing supports c.1813G>A. Truncation/decay effects
are predicted; p.His2479Gln remains particularly uncertain because of the competing MYO7A variant and absent
parental segregation. These are reported disease-associated alleles, not a blanket clinical classification
of pathogenicity. The human paper describes a 4243-residue protein; the mouse papers use a 4249-residue
sequence. PKHD1L1 is distinct from PKHD1, the polycystic-kidney-disease gene.
evidence:
- reference: PMID:38459354
reference_title: "PKHD1L1, a gene involved in the stereocilia coat, causes autosomal recessive nonsyndromic hearing loss."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "Here we provide data to support that mutations in PKHD1L1 cause human nonsyndromic autosomal recessive congenital, mild–moderate to severe SNHL."
explanation: The gene-disease relationship as established by the defining paper.
- reference: PMID:31444330
reference_title: "PKHD1L1 is a coat protein of hair-cell stereocilia and is required for normal hearing."
supports: SUPPORT
evidence_source: MODEL_ORGANISM
directness: INDIRECT
snippet: "we identify Polycystic Kidney and Hepatic Disease 1-Like 1 (PKHD1L1), a large, mostly extracellular protein of 4249 amino acids with a single transmembrane domain"
explanation: >-
Architecture of the mouse protein; the human and mouse sequence lengths must not be conflated or attributed
to signal-peptide inclusion without evidence.
diagnosis:
- name: PKHD1L1 sequencing in unexplained bilateral congenital sensorineural hearing loss
description: >-
Exome sequencing identified the four reported biallelic genotypes, including Family 4 through reanalysis.
General genetic-hearing-loss guidance supports a multigene panel or genomic testing, with explicit confirmation
that PKHD1L1 is covered. Interpret variants with segregation, phase, current pathogenicity assessment and
the phenotype; a pair of uncertain variants does not by itself establish causation. Family 3 illustrates
the need to investigate competing genes.
evidence:
- reference: PMID:38459354
reference_title: "PKHD1L1, a gene involved in the stereocilia coat, causes autosomal recessive nonsyndromic hearing loss."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "This study serves as a call to clinical laboratories to include careful screening of PKHD1L1 biallelic variants in patients with a hearing loss ranging from mild–moderate to severe."
explanation: The diagnostic recommendation the paper makes, and the severity range it targets.
- reference: PMID:38459354
reference_title: "PKHD1L1, a gene involved in the stereocilia coat, causes autosomal recessive nonsyndromic hearing loss."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "The homozygous variant in MYO7A, with a coverage of 198 high quality reads, was deprioritized given uncertain and neutral in silico predictions with respect to impact on protein structure and function"
explanation: >-
How the second candidate was handled in the one family where two plausible homozygous variants were present.
It was deprioritised on prediction, not excluded by function.
- reference: PMID:38459354
reference_title: "PKHD1L1, a gene involved in the stereocilia coat, causes autosomal recessive nonsyndromic hearing loss."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "The initial exome analysis was negative; sequencing data were reanalyzed after this manuscript was deposited as a preprint in medRxiv"
explanation: >-
The fourth family was solved by reanalysing an exome that had already been reported as negative, once
the gene became known. That is the practical argument for periodic reinterpretation in undiagnosed hearing
loss, and it is the reason a negative exome predating 2024 does not exclude this diagnosis.
- reference: url:https://www.ncbi.nlm.nih.gov/sites/books/NBK1434/?report=reader
reference_title: Genetic Hearing Loss Overview - GeneReviews® - NCBI Bookshelf
supports: SUPPORT
evidence_source: OTHER
snippet: can often identify the cause of genetic hearing loss while limiting identification of pathogenic variants and variants of uncertain significance in genes that are irrelevant to the underlying phenotype.
explanation: General hearing-loss testing guidance; panel coverage and diagnostic interpretation remain laboratory dependent.
- name: Audiologic assessment and serial follow-up
description: Document hearing thresholds, type and severity with age-appropriate audiometry or diagnostic ABR, supplemented by tympanometry, otoacoustic emissions and speech testing. Repeat assessment to document progression and guide management. This is general hearing-loss care; no DFNB124-specific surveillance interval is established.
evidence:
- reference: url:https://www.ncbi.nlm.nih.gov/sites/books/NBK1434/?report=reader
reference_title: Genetic Hearing Loss Overview - GeneReviews® - NCBI Bookshelf
supports: SUPPORT
evidence_source: OTHER
snippet: 'Regular follow up is recommended for all individuals with genetic hearing loss in order to:'
explanation: General surveillance recommendation, not a disorder-specific interval.
- reference: PMID:38459354
reference_title: "PKHD1L1, a gene involved in the stereocilia coat, causes autosomal recessive nonsyndromic hearing loss."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: Routine pure-tone audiometry was performed according to current standards in all probands and measured hearing thresholds at 0.25, 0.5, 1, 2, 4, 6, and 8 kHz.
explanation: Audiologic characterization of the human series.
treatments:
- name: Hearing Amplification and Audiological Management
description: >-
Bilateral hearing aids were reported for Family 2, but aided outcome measurements were not provided. General
hearing-loss guidance supports individualized amplification for mild-to-severe hearing loss. This is symptomatic
habilitation; correction of PKHD1L1 dysfunction or slowed disease progression has not been demonstrated.
therapeutic_modality: DEVICE
treatment_term:
preferred_term: hearing amplification with a hearing aid
term:
id: NCIT:C15315
label: Rehabilitation
qualifiers:
- predicate:
preferred_term: medical device
term:
id: NCIT:C16830
label: Medical Device
value:
preferred_term: hearing aid
term:
id: NCIT:C183182
label: Hearing Aid
evidence:
- reference: PMID:38459354
reference_title: "PKHD1L1, a gene involved in the stereocilia coat, causes autosomal recessive nonsyndromic hearing loss."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "The proband currently uses hearing aids bilaterally."
explanation: >-
Documented hearing-aid use in Family 2; this sentence does not report an efficacy result.
- reference: url:https://www.ncbi.nlm.nih.gov/sites/books/NBK1434/?report=reader
reference_title: Genetic Hearing Loss Overview - GeneReviews® - NCBI Bookshelf
supports: SUPPORT
evidence_source: OTHER
snippet: customized by an audiologist to the degree and frequency of hearing loss, can be used in individuals with mild-to-severe hearing loss.
explanation: General phenotype-directed management, not a DFNB124 treatment trial.
target_phenotypes:
- preferred_term: Bilateral sensorineural hearing impairment
term:
id: HP:0008619
label: Bilateral sensorineural hearing impairment
- name: Genetic Counselling and Cascade Testing
description: >-
Provide nondirective counseling and family testing after establishing a molecular diagnosis. When both
parents carry confirmed pathogenic alleles for the same autosomal recessive hearing disorder, each pregnancy
has a 25% chance of an affected child. Verify the familial variants and their segregation before applying
this risk; uncertain causation in Family 3 requires particular caution. Counseling is relevant to consanguineous
and non-consanguineous families alike.
treatment_term:
preferred_term: genetic counseling
term:
id: NCIT:C15240
label: Genetic Counseling
evidence:
- reference: url:https://www.ncbi.nlm.nih.gov/sites/books/NBK1434/?report=reader
reference_title: Genetic Hearing Loss Overview - GeneReviews® - NCBI Bookshelf
supports: SUPPORT
evidence_source: OTHER
snippet: If both parents are known to be heterozygous for an autosomal recessive hearing loss-related pathogenic variant, each sib of the proband has at conception a 25% chance of having hearing loss, a 50% chance of having no hearing loss and being a carrier, and a 25% chance of having no hearing loss and not being a carrier.
explanation: General recessive recurrence risk, conditional on established parental pathogenic variants.
- reference: url:https://www.ncbi.nlm.nih.gov/sites/books/NBK1434/?report=reader
reference_title: Genetic Hearing Loss Overview - GeneReviews® - NCBI Bookshelf
supports: SUPPORT
evidence_source: OTHER
snippet: Carrier testing for relatives who may have a hearing loss-related pathogenic variant requires prior identification of the pathogenic variants in the family.
explanation: Condition for informative cascade testing.
- name: Cochlear implantation when clinically indicated
description: Consider specialist cochlear-implant candidacy assessment for severe-to-profound hearing loss according to hearing ability, communication goals and clinical evaluation. This follows general hearing-loss guidance; the founding DFNB124 series did not report implant outcomes and does not establish genotype-specific benefit.
therapeutic_modality: DEVICE
treatment_term:
preferred_term: cochlear device implantation
term:
id: NCIT:C15329
label: Surgical Procedure
qualifiers:
- predicate:
preferred_term: medical device
term:
id: NCIT:C16830
label: Medical Device
value:
preferred_term: cochlear implant
term:
id: NCIT:C157820
label: Cochlear Implant
target_phenotypes:
- preferred_term: Severe sensorineural hearing impairment
term:
id: HP:0008625
label: Severe sensorineural hearing impairment
evidence:
- reference: url:https://www.ncbi.nlm.nih.gov/sites/books/NBK1434/?report=reader
reference_title: Genetic Hearing Loss Overview - GeneReviews® - NCBI Bookshelf
supports: SUPPORT
evidence_source: OTHER
snippet: Cochlear implantation can be considered in children with severe-to-profound hearing loss who are older than age nine months.
explanation: General candidacy guidance, not evidence of implantation or outcomes in DFNB124.
- name: Communication and language support
description: Establish communication goals with the individual and family and provide early access to language, speech-language services and educational support as appropriate. These are general measures for childhood hearing loss, not evidence of an additional neurodevelopmental phenotype in DFNB124.
evidence:
- reference: url:https://www.ncbi.nlm.nih.gov/sites/books/NBK1434/?report=reader
reference_title: Genetic Hearing Loss Overview - GeneReviews® - NCBI Bookshelf
supports: SUPPORT
evidence_source: OTHER
snippet: On initial evaluation of individuals with hearing loss, the goals for communication must be established with a focus on equipping individuals with language and appropriate access to language.
explanation: General management guidance applicable to the auditory phenotype.
- name: Hearing-conservation counseling
description: Counsel about avoiding repeated hazardous noise exposure as general care for documented hearing loss. The knockout-mouse finding strengthens a research rationale but does not establish increased human DFNB124 susceptibility or a disease-specific protective effect.
evidence:
- reference: url:https://www.ncbi.nlm.nih.gov/sites/books/NBK1434/?report=reader
reference_title: Genetic Hearing Loss Overview - GeneReviews® - NCBI Bookshelf
supports: SUPPORT
evidence_source: OTHER
snippet: Since this risk can be minimized by avoidance, persons with documented hearing loss should be counseled appropriately and repeated overexposure to loud noises should be avoided.
explanation: General hearing conservation advice; no DFNB124-specific clinical efficacy claim.
animal_models:
- name: Pkhd1l1 conditional knockout mouse using Atoh1-Cre
species: Mouse
genotype: Pkhd1l1 fl/fl; Atoh1-Cre+ (conditional exon 10 deletion)
background: Mixed C57BL/6N/129S4/CBA; homozygous Cdh23ahl
publication: PMID:31444330
genes:
- preferred_term: PKHD1L1
term:
id: hgnc:20313
label: PKHD1L1
description: >-
Conditional exon 10 deletion causes loss of the upper stereociliary coat, progressive auditory impairment,
later bundle pathology and increased noise vulnerability. Early gross morphology, FM1-43 uptake and initial
attachment-crown formation are preserved in the tested assays. Atoh1-Cre activity is not strictly limited
to hair cells; the constitutive knockout corroborates hearing impairment independently of persistent Cre
expression.
modeled_mechanisms:
- target: Loss of the Stereocilia Surface Coat at Stereocilia Tips
relationship: RECAPITULATES
fidelity: MODERATE
description: >-
This model is how the coat was identified as a PKHD1L1 structure at all: the coat at the tips is absent
in the mutant and present in littermate controls, on tannic-acid stained transmission electron microscopy.
limitations: >-
The coat has never been examined in human hair cells, in a patient or otherwise, so the identity of the
human structure with the mouse one is assumed from protein orthology. The loss is also partial in a specific
sense that should not be lost: only the upper coat goes, and the paper suggests the lower coat is made
of different proteins.
readouts:
- name: Density of the electron-dense surface coat at stereocilia tips
target: Loss of the Stereocilia Surface Coat at Stereocilia Tips
direction: DECREASED
interpretation: >-
Tannic-acid staining intensity across the stereocilia membrane falls at the tips in mutants and is
unchanged at the bases.
evidence:
- reference: PMID:31444330
reference_title: "PKHD1L1 is a coat protein of hair-cell stereocilia and is required for normal hearing."
supports: SUPPORT
evidence_source: MODEL_ORGANISM
snippet: "PKHD1L1-deficient mice show a large decrease of the surface coat intensity only at the tips of stereocilia"
explanation: The measurement and its restriction to the upper stereocilia.
evidence:
- reference: PMID:31444330
reference_title: "PKHD1L1 is a coat protein of hair-cell stereocilia and is required for normal hearing."
supports: SUPPORT
evidence_source: MODEL_ORGANISM
snippet: "PKHD1L1-deficient mice lack the surface coat at the upper but not lower regions of stereocilia, and they develop progressive hearing loss."
explanation: >-
Establishes this model as the system in which the coat-PKHD1L1 relationship was demonstrated.
- target: Impaired Stereocilia Bundle Maintenance
relationship: RECAPITULATES
fidelity: MODERATE
description: >-
This line supports the maintenance hypothesis through preserved early gross morphology followed by stereocilia
loss and later bundle disorganization. Neonatal dye uptake and initial attachment findings do not rule
out all transduction or attachment abnormalities.
limitations: >-
Mice carry Cdh23ahl on a mixed background, and littermate comparisons do not eliminate interaction with
that sensitizing allele. Stereocilia counts come from small numbers of cochleae. Human bundle pathology
has not been measured. Atoh1-Cre also acts in some supporting cells and sensory neurons.
readouts:
- name: Missing stereocilia per outer hair cell bundle by row
target: Impaired Stereocilia Bundle Maintenance
direction: INCREASED
interpretation: >-
Stereocilia are lost from the shortest row first and from taller rows with age, most in the high-frequency
base.
evidence:
- reference: PMID:39482437
reference_title: "PKHD1L1 is required for stereocilia bundle maintenance, durable hearing function and resilience to noise exposure."
supports: SUPPORT
evidence_source: MODEL_ORGANISM
snippet: "stereocilia loss was most significant in the shortest row, progressing to the taller rows with age; most prominently in, but not limited to, the high frequency basal region"
explanation: The readout, its direction and its spatial pattern.
- name: Stereocilin localisation at tectorial-membrane attachment crowns
target: Impaired Stereocilia Bundle Maintenance
direction: UNCHANGED
interpretation: >-
Stereocilin localization is preserved at the tested ages, supporting initial attachment-crown formation
without proving lifelong normal mechanical attachment.
evidence:
- reference: PMID:39482437
reference_title: "PKHD1L1 is required for stereocilia bundle maintenance, durable hearing function and resilience to noise exposure."
supports: SUPPORT
evidence_source: MODEL_ORGANISM
snippet: "We therefore conclude that PKHD1L1 is not required for the localization of STRC to attachment crowns or horizontal top connectors."
explanation: The negative readout, as the authors state it.
evidence:
- reference: PMID:39482437
reference_title: "PKHD1L1 is required for stereocilia bundle maintenance, durable hearing function and resilience to noise exposure."
supports: SUPPORT
evidence_source: MODEL_ORGANISM
snippet: "These results suggest a role for PKHD1L1 in establishing robust sensory hair bundles during development, necessary for maintaining bundle cohesion and function in response to acoustic trauma and aging."
explanation: >-
The authors' summary of what this model shows, which is the claim the node makes.
- target: Reduced Resilience to Acoustic Overexposure
relationship: RECAPITULATES
fidelity: UNKNOWN
description: >-
The noise arm. Six-week-old mutants and littermate controls received the same two-hour 94 dB SPL bandpass
exposure; controls recovered to baseline by two weeks and mutants did not, at two weeks or at eight.
limitations: >-
Fidelity is UNKNOWN, not LOW, and the distinction is deliberate: the experiment is well controlled and
the result is clear, but there is no human observation of any kind to compare it against, so how faithfully
it represents a human patient cannot be assessed. Two further limits belong on the result itself. Threshold
shifts were measured against an 80 dB SPL ceiling, and the mutants' baseline high-frequency thresholds
already sit near that ceiling, so the measured shift understates the damage at exactly the frequencies
where the disease is worst. And the exposure paradigm is a single laboratory dose, not the chronic, variable
noise history a person accumulates.
readouts:
- name: Auditory brainstem response threshold shift at 8 weeks after noise
target: Reduced Resilience to Acoustic Overexposure
direction: INCREASED
interpretation: >-
Mutant thresholds remain elevated 8 weeks after an exposure from which controls fully recover - a permanent
rather than temporary threshold shift.
evidence:
- reference: PMID:39482437
reference_title: "PKHD1L1 is required for stereocilia bundle maintenance, durable hearing function and resilience to noise exposure."
supports: SUPPORT
evidence_source: MODEL_ORGANISM
snippet: "These data therefore show the noise exposure, normally only sufficient to cause TTS in normal mice, leads to PTS in PKHD1L1-deficient mice."
explanation: The readout stated as the genotype contrast.
evidence:
- reference: PMID:38459354
reference_title: "PKHD1L1, a gene involved in the stereocilia coat, causes autosomal recessive nonsyndromic hearing loss."
supports: NO_EVIDENCE
evidence_source: HUMAN_CLINICAL
snippet: "Further research will be needed to determine the effect of age or noise trauma on the potential progression of PKHD1L1-linked hearing loss."
explanation: >-
Recorded on the link rather than omitted, because it is what fixes the fidelity call: the human paper
names age and noise as untested, so the model result has no human comparator. Graded NO_EVIDENCE because
the sentence reports the absence of a study, not a result.
- target: Cochlear Hearing Loss
relationship: PARTIALLY_RECAPITULATES
fidelity: MODERATE
description: >-
Both models and humans have auditory impairment, but their measured onset and course differ. The 2019
conditional-mouse study detected high-frequency threshold elevations at three weeks; the 2024 study detected
them at six weeks with slower spread to lower frequencies. Human loss was congenital or presumed congenital,
with quantitatively slow progression documented in one proband.
limitations: >-
The onset divergence is the substantive limitation and is the subject of an open human-model mismatch
discussion. There is a second, quieter one: the two studies of this same line disagree in degree, the
later one finding threshold elevations less pronounced and slower to spread than the earlier, which the
authors attribute to the recordings being made in different institutions. Any quantitative claim taken
from this model should carry that.
readouts:
- name: Auditory brainstem response and distortion product thresholds across frequency and age
target: Cochlear Hearing Loss
direction: INCREASED
interpretation: >-
Thresholds rise - worse hearing - from three weeks at high frequencies and across all frequencies by
six months.
evidence:
- reference: PMID:31444330
reference_title: "PKHD1L1 is a coat protein of hair-cell stereocilia and is required for normal hearing."
supports: SUPPORT
evidence_source: MODEL_ORGANISM
snippet: "PKHD1L1-deficient mice show elevated thresholds in response to high-frequency tones"
explanation: The readout and where in the frequency range it starts.
evidence:
- reference: PMID:39482437
reference_title: "PKHD1L1 is required for stereocilia bundle maintenance, durable hearing function and resilience to noise exposure."
supports: SUPPORT
evidence_source: MODEL_ORGANISM
snippet: "Both conditional and constitutive PKHD1L1 knockout mice develop high-frequency hearing loss progressing to lower frequencies with age."
explanation: >-
Two independent knockout designs give the same endpoint, which is what supports treating this model
as informative for the human phenotype despite the timing difference.
- name: pkhd1l1a/pkhd1l1b double mutant zebrafish
species: Zebrafish
genotype: pkhd1l1a and pkhd1l1b CRISPR-Cas9 loss-of-function alleles, double homozygous
publication: PMID:36960824
genes:
- preferred_term: PKHD1L1
term:
id: hgnc:20313
label: PKHD1L1
description: >-
CRISPR loss of both zebrafish pkhd1l1 paralogs produced impaired auditory behavior from the larval stage.
This supports a conserved auditory role but does not establish the mammalian cochlear mechanism or resolve
the difference in onset among species.
modeled_mechanisms:
- target: Cochlear Hearing Loss
relationship: PARTIALLY_RECAPITULATES
fidelity: LOW
description: >-
Auditory function is impaired, establishing that the gene's role in hearing is conserved across vertebrates,
and the early onset is the closer match to the human congenital presentation.
limitations: >-
Zebrafish lack the mammalian cochlea and outer-hair-cell amplifier. Behavioral responses are indirect
hearing measures. The cached primary source is abstract-only; the human paper describes the startle assay
at six days post fertilization. Detailed dose-response and motor controls were not independently assessed
from the zebrafish full text.
readouts:
- name: Auditory-evoked startle response at the larval stage
target: Cochlear Hearing Loss
direction: DECREASED
interpretation: >-
Reduced larval auditory-evoked responses indicate early auditory dysfunction; they do not exclude deterioration
before the measured stage.
evidence:
- reference: PMID:36960824
reference_title: "A conserved function of Pkhd1l1, a mammalian hair cell stereociliary coat protein, in regulating hearing in zebrafish."
supports: SUPPORT
evidence_source: MODEL_ORGANISM
snippet: "the double mutant zebrafish exhibited statistically significant hearing loss even from the larval stage"
explanation: The readout and the age at which it is present.
- reference: PMID:38459354
reference_title: "PKHD1L1, a gene involved in the stereocilia coat, causes autosomal recessive nonsyndromic hearing loss."
supports: SUPPORT
evidence_source: OTHER
snippet: More recent data from zebrafish (Danio rerio, Dr) with a double knockout of pkhd1l1a and pkhd1l1b (orthologs of human (Hs) PKHD1L1) show significant deficits in auditory startle responses at the larval stage, consistent with an early-onset auditory phenotype in zebrafish (Makrogkikas et al. 2023).
explanation: The full-text human paper describes the zebrafish readout; this is background synthesis, not another experiment.
evidence:
- reference: PMID:36960824
reference_title: "A conserved function of Pkhd1l1, a mammalian hair cell stereociliary coat protein, in regulating hearing in zebrafish."
supports: SUPPORT
evidence_source: MODEL_ORGANISM
snippet: "in contrast to Pkhd1l1 mutant mice, which develop progressive hearing loss, the double mutant zebrafish exhibited statistically significant hearing loss even from the larval stage"
explanation: >-
The comparison that makes this model informative here: the two animal models disagree about onset,
and the human phenotype sits on the zebrafish side of that disagreement.
discussions:
- discussion_id: dfnb124_onset_species_mismatch
kind: HUMAN_MODEL_MISMATCH
status: OPEN
attaches_to:
- "pathophysiology#Impaired Stereocilia Bundle Maintenance"
- "pathophysiology#Cochlear Hearing Loss"
prompt: >-
How do developmental timing and allele-specific effects contribute to differences in observed auditory
onset among humans, mice and zebrafish?
rationale: >-
Human hearing loss was congenital or presumed congenital; the best-followed proband had slow childhood
progression. Mouse high-frequency deficits were detected at three or six weeks in different studies, and
later stereocilia pathology supports a maintenance defect after grossly preserved early development. Zebrafish
double mutants show larval auditory impairment. These observations use different developmental stages,
alleles and measurements. Developmental timing, residual allele function and species-specific biology are
plausible explanations, but none is established. Undetectable adult antibody labeling does not prove absence
of all PKHD1L1 protein, and the neonatal FM1-43 assay does not establish normal lifetime transduction.
evidence:
- reference: PMID:39482437
reference_title: "PKHD1L1 is required for stereocilia bundle maintenance, durable hearing function and resilience to noise exposure."
supports: SUPPORT
evidence_source: MODEL_ORGANISM
snippet: "Our findings reveal that PKHD1L1-deficient mice display no disruption to bundle cohesion or tectorial membrane attachment-crown formation during development."
explanation: "Preserved gross developmental morphology in the tested mouse assays."
- reference: PMID:39482437
reference_title: "PKHD1L1 is required for stereocilia bundle maintenance, durable hearing function and resilience to noise exposure."
supports: SUPPORT
evidence_source: MODEL_ORGANISM
snippet: "Anti-PKHD1L1 labeling intensity gradually decreased with development, was largely gone by P10, and completely cleared in mature hair cells at P21."
explanation: >-
Loss of immunolabeling supports temporally restricted detectability; it does not rank explanations for
human onset.
- reference: PMID:38459354
reference_title: "PKHD1L1, a gene involved in the stereocilia coat, causes autosomal recessive nonsyndromic hearing loss."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "bilateral congenital SNHL which is nonsyndromic and mild–moderate to severe"
explanation: "The human half: present at birth."
- reference: PMID:36960824
reference_title: "A conserved function of Pkhd1l1, a mammalian hair cell stereociliary coat protein, in regulating hearing in zebrafish."
supports: SUPPORT
evidence_source: MODEL_ORGANISM
snippet: "in contrast to Pkhd1l1 mutant mice, which develop progressive hearing loss, the double mutant zebrafish exhibited statistically significant hearing loss even from the larval stage"
explanation: >-
Larval impairment provides an additional species comparison using a different auditory assay.
proposed_experiments:
- experiment_id: exp_dfnb124_developmental_timing
name: Establish when the human lesion appears
description: >-
Two complementary approaches. First, examine the surface coat and bundle morphology in PKHD1L1-deficient
mouse cochlea at prenatal and early postnatal ages with the sensitive methods used later, to test whether
a subtle developmental deficit precedes the six-week onset. Second, differentiate hair-cell-like cells
from patient-derived induced pluripotent stem cells carrying the human alleles and ask whether the coat
forms at all, which would separate a construction defect from a maintenance defect without needing patient
cochlea.
- discussion_id: dfnb124_noise_untested_in_humans
kind: KNOWLEDGE_GAP
status: OPEN
attaches_to:
- pathophysiology#Reduced Resilience to Acoustic Overexposure
- treatments#Hearing-conservation counseling
prompt: >-
Does DFNB124 increase human noise susceptibility beyond the general risks associated with hazardous noise?
rationale: >-
Conditional knockout mice retain hearing deficits after a noise exposure from which control littermates
recover. The human series did not establish genotype-specific noise susceptibility. General hearing-conservation
counseling is appropriate under genetic-hearing-loss guidance, independently of this unresolved mouse-to-human
extrapolation. Longitudinal audiometry and exposure histories could test whether an additional DFNB124-specific
vulnerability exists.
evidence:
- reference: PMID:38459354
reference_title: "PKHD1L1, a gene involved in the stereocilia coat, causes autosomal recessive nonsyndromic hearing loss."
supports: NO_EVIDENCE
evidence_source: HUMAN_CLINICAL
snippet: "Further research will be needed to determine the effect of age or noise trauma on the potential progression of PKHD1L1-linked hearing loss."
explanation: >-
The gap stated by the authors of the human paper. Graded NO_EVIDENCE because the sentence reports that
no study exists, which neither supports nor refutes the claim.
- reference: PMID:39482437
reference_title: "PKHD1L1 is required for stereocilia bundle maintenance, durable hearing function and resilience to noise exposure."
supports: SUPPORT
evidence_source: MODEL_ORGANISM
snippet: "Furthermore, PKHD1L1-deficient mice are susceptible to permanent hearing loss following moderate acoustic overexposure, which induces only temporary hearing threshold shifts in wild-type mice."
explanation: The animal result that makes the untested human question worth asking.
- discussion_id: dfnb124_family3_myo7a_confound
kind: OPEN_QUESTION
status: OPEN
attaches_to:
- "phenotypes#Severe Hearing Impairment"
- "genetic#PKHD1L1"
prompt: >-
Is the severe end of the DFNB124 range real, or does it belong to a second gene?
rationale: >-
Family 3 carries homozygous PKHD1L1 p.His2479Gln and MYO7A p.Leu375Val. Both calls had high read coverage,
but parental segregation and functional testing were unavailable. MYO7A was deprioritized using prediction
and conservation, which does not exclude pathogenicity. The source explicitly allows PKHD1L1 causation,
incidental PKHD1L1 variation, or contributions from both genes. Audiometry was performed in ambient noise
without a soundproof environment. Severe loss is retained as a reported observation with attribution uncertainty
and without a frequency estimate.
evidence:
- reference: PMID:38459354
reference_title: "PKHD1L1, a gene involved in the stereocilia coat, causes autosomal recessive nonsyndromic hearing loss."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "Exome analysis revealed two homozygous missense variants of interest: one in PKHD1L1 c.7437C>A, p.(His2479Gln) and one in MYO7A (NM_000260.4:c.1123C>G, p.(Leu375Val))."
explanation: The two candidate variants in this proband, both homozygous.
- reference: PMID:38459354
reference_title: "PKHD1L1, a gene involved in the stereocilia coat, causes autosomal recessive nonsyndromic hearing loss."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "The homozygous variant in MYO7A, with a coverage of 198 high quality reads, was deprioritized given uncertain and neutral in silico predictions with respect to impact on protein structure and function"
explanation: >-
How the alternative was handled, and the basis on which it was set aside - prediction rather than experiment.
proposed_experiments:
- experiment_id: exp_dfnb124_his2479gln_function
name: Functional test of PKHD1L1 p.(His2479Gln) and MYO7A p.(Leu375Val)
description: >-
Assess both variants in appropriately validated functional systems, alongside segregation and additional
unrelated cases when available. Fragment stability and MYO7A motor/trafficking assays could add evidence,
but neither a positive nor a negative isolated assay would by itself settle clinical causation.
external_assertions:
- name: OMIM deafness, autosomal recessive 124 record
source: OMIM
assertion_type: disease_record
external_id: OMIM:620794
url: https://omim.org/entry/620794
description: >-
The OMIM phenotype record for DFNB124, and the identifier under which the entity was designated. Recorded
here rather than under mappings because the DiseaseMappings class carries only ICD-10-CM, ICD-11, MONDO
and NCIT slots.
evidence:
- reference: PMID:39482437
reference_title: "PKHD1L1 is required for stereocilia bundle maintenance, durable hearing function and resilience to noise exposure."
supports: SUPPORT
evidence_source: OTHER
snippet: "recently designated as autosomal recessive deafness-124 (DFNB124) in the OMIM database (entry #620794)"
explanation: >-
The OMIM number, cited from a paper that states it. Graded OTHER because the sentence is a nomenclature
statement in an introduction, not a study result.
notes: >-
The mechanism is supported chiefly by knockout mice and allele-level in-vitro assays; human cochlear pathology
has not been directly observed. The nonsyndromic scope reflects the founding human series, not an assertion
that all PKHD1L1 biology is confined to hearing. General genetic-hearing-loss guidance supplies testing,
habilitation, counseling and noise-avoidance principles; it does not supply DFNB124-specific frequencies
or treatment outcomes. OTOF-directed therapy and unrelated noncochlear PKHD1L1 associations do not establish
treatment or syndromic manifestations for DFNB124. PMID:32555200 describes reuse of six P4 mouse bundles
from the earlier localization study, rather than independent biological replication. The human paper reports
normal eye, cardiac and developmental assessments; positive ontology annotations for these features are therefore
not adopted as DFNB124 manifestations.
References & Deep Research
References
6Deep Research
1Deep research results are used as seeds for research; they do not undergo the same validation as the main records and may contain errors. How we use deep research.
Evaluations and curation notes (1)
Create: Autosomal Recessive Nonsyndromic Hearing Loss 124 (MONDO:0968981, PKHD1L1) · 2026-09-04T00:08:32Z · View source
Created kb/disorders/Autosomal_Recessive_Nonsyndromic_Hearing_Loss_124.yaml (MONDO:0968981, PKHD1L1, OMIM 620794). Scope decision: DISEASE - one gene, one proximal lesion (loss of the stereocilia surface coat on the upper stereocilia), one clinical picture. Stub deleted. Deep research: falcon (Edison), run through scripts/run_deep_research_client.sh directly. NEC preflight passes (PKHD1L1 mentioned 50 times; MYO7A correctly identified as the competing gene in one family rather than a second cause). It added nothing the primary literature does not carry. Sources: PMID:38459354 (Redfield 2024, full text) is the defining human paper - four families; PMID:31444330 (Wu 2019) identifies PKHD1L1 as the coat protein; PMID:39482437 (Strelkova 2024) supplies the bundle-maintenance time course, the negative controls and the noise result; PMID:36960824 is the zebrafish model. Curation decisions worth recording. Only the terminal pathophysiology node conforms to sensorineural_hair_cell_loss: conforming to that module's mechanotransduction node would assert a mechanism the data contradict, since FM1-43 loads normally in PKHD1L1-deficient hair cells, and its ionic/oxidative node has never been investigated here. The noise result is curated structurally as an environmental entry with an EXACERBATES influences_mechanisms link plus a PROVISIONAL node, and deliberately NOT as a treatment, because it is a mouse result and the human paper names it as an open question. The mouse-human onset divergence is a HUMAN_MODEL_MISMATCH discussion. The Family 3 MYO7A confound is recorded on the severe phenotype and in its own discussion rather than glossed. The four HPOA annotations on MONDO:0968981 that contradict "nonsyndromic" are not curated; the reasoning is in the entry notes and is worth reporting upstream. Validation: just validate exit 0; 62/62 snippets verified; check-duplicate-keys, check-entity-refs, check-causal-targets, check-snippet-length, check-title-snippets, check-snippet-grading, check-environmental-evidence, check-folded-hyphens all exit 0. One bracketed-phase snippet from the abstract was replaced with two case-description quotes because the reference validator strips bracketed spans before matching.
Question: You are an expert researcher providing comprehensive, well-cited information.
Provide detailed information focusing on: 1. Key concepts and definitions with current understanding 2. Recent developments and latest research (prioritize 2023-2024 sources) 3. Current applications and real-world implementations 4. Expert opinions and analysis from authoritative sources 5. Relevant statistics and data from recent studies
Format as a comprehensive research report with proper citations. Include URLs and publication dates where available. Always prioritize recent, authoritative sources and provide specific citations for all major claims.
Disease Characteristics Research Template
Target Disease
- Disease Name: Autosomal Recessive Nonsyndromic Hearing Loss 124 (DFNB124, PKHD1L1-related deafness)
- MONDO ID: MONDO:0968981 (if available)
- Category: Mendelian
Research Objectives
Please provide a comprehensive research report on Autosomal Recessive Nonsyndromic Hearing Loss 124 (DFNB124, PKHD1L1-related deafness) covering all of the disease characteristics listed below. This report will be used to populate a disease knowledge base entry. Be thorough and cite primary literature (PMID preferred) for all claims.
For each section, suggested databases/resources are listed. These are the first places you should search for information on each topic.
1. Disease Information
Search first: OMIM, Orphanet, ICD-10/ICD-11, MeSH, PubMed
- What is the disease? Provide a concise overview.
- What are the key identifiers? (OMIM, Orphanet, ICD-10/ICD-11, MeSH, Mondo)
- What are the common synonyms and alternative names?
- Is the information derived from individual patients (e.g., EHR) or aggregated disease-level resources?
2. Etiology
- Disease Causal Factors: What are the primary causes? (genetic, environmental, infectious, mechanistic)
- Risk Factors:
Search first: PubMed, Cochrane Library, UpToDate, clinical guidelines, ClinVar, ClinGen, GWAS Catalog, PheGenI, CTD, CDC, WHO, epidemiological databases
- Genetic risk factors (causal variants, susceptibility loci, modifier genes)
- Environmental risk factors (toxins, lifestyle, occupational exposures, age, sex, family history)
- Protective Factors:
Search first: PubMed, Cochrane Library, clinical trial databases, GWAS Catalog, gnomAD, WHO, CDC, nutrition databases
- Genetic protective factors (protective variants, modifier alleles)
- Environmental protective factors (diet, lifestyle, exposures that reduce risk)
- Gene-Environment Interactions: How do genetic and environmental factors interact to influence disease?
Search first: CTD, PubMed, PheGenI, GxE databases
3. Phenotypes
Search first: HPO (Human Phenotype Ontology), OMIM, Orphanet, PubMed, clinicaltrials.gov, MedDRA, SNOMED CT, DECIPHER, LOINC
For each phenotype, provide: - Phenotype type: symptoms, clinical signs, physical manifestations, behavioral changes, or laboratory abnormalities
For symptoms/signs: HPO, OMIM, Orphanet, PubMed For behavioral changes: HPO, DSM, RDoC (Research Domain Criteria), PubMed For laboratory abnormalities: LOINC, SNOMED CT, LabTests Online, PubMed - Phenotype characteristics: Search first: OMIM, Orphanet, HPO, PubMed - Age of symptom onset (neonatal, childhood, adult-onset, late-onset) - Symptom severity (mild, moderate, severe, variable) - Symptom progression (stable, progressive, episodic, fluctuating) - Frequency among affected individuals (percentage or qualitative) - Quality of life impact: Effects on daily functioning and well-being (per-phenotype when possible) Search first: EQ-5D database, SF-36, WHO QOL databases, PubMed - Suggest HPO (Human Phenotype Ontology) terms for each phenotype
4. Genetic/Molecular Information
- Causal Genes: Gene mutations or chromosomal abnormalities responsible for disease (gene symbols, OMIM IDs)
Search first: OMIM, ClinVar, HGMD, Ensembl, NCBI Gene
- Pathogenic Variants:
- Affected genes (gene symbols, HGNC IDs) > Search first: OMIM, NCBI Gene, Ensembl, HGNC, UniProt, GeneCards
- Variant classification (pathogenic, likely pathogenic, VUS per ACMG/AMP guidelines) > Search first: ClinVar, ClinGen, ACMG/AMP guidelines, VarSome
- Variant type/class (missense, frameshift, nonsense, splice-site, structural)
- Allele frequency in population databases > Search first: gnomAD, 1000 Genomes, ExAC, TOPMed, dbSNP
- Somatic vs germline origin > Search first: COSMIC (somatic), ClinVar, ICGC, TCGA
- Functional consequences (loss of function, gain of function, dominant negative)
- Modifier Genes: Genes that modify disease severity or expression
- Epigenetic Information: DNA methylation, histone modifications, chromatin changes affecting disease
Search first: ENCODE, Roadmap Epigenomics, MethBase, DiseaseMeth
- Chromosomal Abnormalities: Large-scale genetic changes (aneuploidy, translocations, inversions)
Search first: DECIPHER, ClinVar, ECARUCA, UCSC Genome Browser
5. Environmental Information
- Environmental Factors: Non-genetic contributing factors (toxins, radiation, pollution, occupational exposure)
Search first: CTD (Comparative Toxicogenomics Database), TOXNET, PubMed, EPA databases
- Lifestyle Factors: Behavioral factors (smoking, diet, exercise, alcohol consumption)
Search first: CDC databases, WHO, PubMed, NHANES
- Infectious Agents: If applicable, pathogens causing or triggering disease (bacteria, viruses, fungi, parasites)
Search first: NCBI Taxonomy, ViPR, BV-BRC, MicrobeDB, GIDEON
6. Mechanism / Pathophysiology
Present this section as an ordered causal chain first, then the detail below. Open with a numbered sequence of mechanistic steps running from the initiating lesion (mutation, exposure, infection) to the clinical manifestation, one step per line, each naming what it causes next. State the causal verb explicitly ("leads to", "results in") and say where a step is inferred rather than demonstrated. Where the mechanism branches, show the branch. The categories below are a checklist of what to cover within those steps, not the organizing structure — a step may draw on several of them, and a category may contribute to several steps.
- Molecular Pathways: Specific signaling cascades or biochemical pathways involved (Wnt, MAPK, mTOR, PI3K-AKT, etc.)
Search first: KEGG, Reactome, WikiPathways, PathBank, BioCyc
- Cellular Processes: Cell-level mechanisms (apoptosis, autophagy, cell cycle dysregulation, inflammation, etc.)
Search first: Gene Ontology (GO), Reactome, KEGG, PubMed
- Protein Dysfunction: How protein structure or function is altered (misfolding, aggregation, loss of function, gain of function)
Search first: UniProt, PDB (Protein Data Bank), InterPro, Pfam, AlphaFold
- Metabolic Changes: Alterations in metabolic processes (energy metabolism, lipid metabolism, amino acid metabolism)
Search first: KEGG, BioCyc, HMDB (Human Metabolome Database), BRENDA
- Immune System Involvement: Role of immune response (autoimmunity, immunodeficiency, chronic inflammation)
Search first: ImmPort, Immunome Database, IEDB, Gene Ontology
- Tissue Damage Mechanisms: How tissues/ are injured (oxidative stress, ischemia, fibrosis, necrosis)
Search first: PubMed, Gene Ontology, Reactome
- Biochemical Abnormalities: Specific molecular defects (enzyme deficiencies, receptor dysfunction, ion channel defects)
Search first: BRENDA, UniProt, KEGG, OMIM, PubMed
- Epigenetic Changes: DNA methylation, histone modifications affecting gene expression in disease
Search first: ENCODE, Roadmap Epigenomics, MethBase, DiseaseMeth
- Molecular Profiling (if available):
- Transcriptomics/gene expression changes > Search first: GEO (Gene Expression Omnibus), ArrayExpress, GTEx, Human Cell Atlas, SRA
- Proteomics findings > Search first: PRIDE, ProteomeXchange, Human Protein Atlas, STRING, BioGRID
- Metabolomics signatures > Search first: MetaboLights, Metabolomics Workbench, HMDB, METLIN
- Lipidomics alterations > Search first: LIPID MAPS, SwissLipids, LipidHome, Metabolomics Workbench
- Genomic structural features > Search first: UCSC Genome Browser, Ensembl, NCBI, dbVar, DGV
- Advanced Technologies (if applicable):
- Single-cell analysis findings (cell-type specific mechanisms, cellular heterogeneity) > Search first: Human Cell Atlas, Single Cell Portal, GEO, CELLxGENE
- Spatial transcriptomics findings > Search first: GEO, Spatial Research, Vizgen, 10x Genomics data
- Multi-omics integration results > Search first: TCGA, ICGC, cBioPortal, LinkedOmics, PubMed
- Functional genomics screens (CRISPR, RNAi) > Search first: DepMap, GenomeRNAi, PubMed, BioGRID ORCS
For each mechanism, describe: - The causal chain from initial trigger to clinical manifestation - Which mechanisms are upstream vs downstream - What cell types and biological processes are involved - Suggest GO terms for biological processes and CL terms for cell types
7. Anatomical Structures Affected
- Organ Level:
- Primary organs directly affected
- Secondary organ involvement (complications, secondary effects)
- Body systems involved (cardiovascular, nervous, digestive, respiratory, endocrine, etc.)
Search first: Uberon, FMA (Foundational Model of Anatomy), OMIM, HPO, ICD-11, MeSH, SNOMED CT
- Tissue and Cell Level:
- Specific tissue types affected (epithelial, connective, muscle, nervous)
- Specific cell populations targeted (with Cell Ontology terms)
Search first: Uberon, Human Protein Atlas, Cell Ontology, Human Cell Atlas, CellMarker, PanglaoDB
- Subcellular Level:
- Cellular compartments involved (mitochondria, nucleus, ER, lysosomes) (with GO Cellular Component terms)
Search first: Gene Ontology (Cellular Component), UniProt, Human Protein Atlas
- Localization:
- Specific anatomical sites (with UBERON terms) > Search first: FMA, Uberon, NeuroNames (for brain), SNOMED CT
- Lateralization (unilateral, bilateral, asymmetric) > Search first: HPO, clinical literature, imaging databases
8. Temporal Development
- Onset:
- Typical age of onset (congenital, pediatric, adult, geriatric)
- Onset pattern (acute, subacute, chronic, insidious)
Search first: OMIM, Orphanet, HPO, PubMed
- Progression:
- Disease stages (early, intermediate, advanced, end-stage) > Search first: Cancer Staging Manual (AJCC), WHO classifications, PubMed
- Progression rate (rapid, slow, variable)
- Disease course pattern (episodic, relapsing-remitting, progressive, stable)
- Disease duration (self-limited, chronic lifelong)
Search first: Disease registries, longitudinal cohort databases, natural history studies, PubMed, Orphanet, OMIM
- Patterns:
- Remission patterns (spontaneous, treatment-induced) > Search first: Clinical trial databases, disease registries, PubMed
- Critical periods (time windows of vulnerability or opportunity for intervention) > Search first: PubMed, developmental biology databases, clinical guidelines
9. Inheritance and Population
- Epidemiology:
- Prevalence (cases per 100,000 at given time)
- Incidence (new cases per 100,000 per year)
Search first: Orphanet, CDC, WHO, GBD (Global Burden of Disease), national registries, SEER, disease registries
- For Genetic Etiology:
- Inheritance pattern (AD, AR, X-linked, mitochondrial, multifactorial, polygenic) > Search first: OMIM, Orphanet, ClinVar, GTR (Genetic Testing Registry)
- Penetrance (complete, incomplete, age-dependent) > Search first: ClinVar, OMIM, PubMed, ClinGen
- Expressivity (variable, consistent) > Search first: OMIM, ClinVar, PubMed
- Genetic anticipation (increasing severity in successive generations) > Search first: OMIM, PubMed (especially for repeat expansion disorders)
- Germline mosaicism > Search first: ClinVar, OMIM, genetic counseling literature, PubMed
- Founder effects (population-specific mutations) > Search first: gnomAD, population genetics databases, PubMed
- Consanguinity role > Search first: OMIM, population studies, genetic counseling resources
- Carrier frequency > Search first: gnomAD, carrier screening databases, GeneReviews, GTR
- Population Demographics:
- Affected populations (ethnic or demographic groups with higher prevalence) > Search first: gnomAD, 1000 Genomes, PAGE Study, PubMed, population registries
- Geographic distribution (endemic areas, regional variation) > Search first: WHO, CDC, GBD, Orphanet, geographic epidemiology databases
- Geographic distribution of specific variants
- Sex ratio (male:female) > Search first: Disease registries, OMIM, PubMed, epidemiological databases
- Age distribution of affected individuals > Search first: CDC, disease registries, SEER, Orphanet
10. Diagnostics
- Clinical Tests:
- Laboratory tests (blood, urine, tissue chemistry, specific enzyme assays) > Search first: LOINC, LabTests Online, PubMed
- Biomarkers (proteins, metabolites, genetic markers, circulating biomarkers) > Search first: FDA Biomarker List, BEST (Biomarkers, EndpointS, and other Tools), PubMed
- Imaging studies (X-ray, CT, MRI, PET, ultrasound) > Search first: RadLex, DICOM, Radiopaedia, imaging databases
- Functional tests (pulmonary function, cardiac stress tests) > Search first: LOINC, clinical guidelines, PubMed
- Electrophysiology (EEG, EMG, ECG, nerve conduction studies) > Search first: LOINC, clinical neurophysiology databases, PubMed
- Biopsy findings (histopathology, immunohistochemistry) > Search first: SNOMED CT, College of American Pathologists resources, PubMed
- Pathology findings (microscopic examination) > Search first: SNOMED CT, Digital Pathology databases, PubMed
- Genetic Testing:
Search first: GTR (Genetic Testing Registry), GeneReviews, ClinGen
- Overview of recommended genetic testing approach
- Whole genome sequencing (WGS) utility > Search first: GTR, ClinVar, GEL (Genomics England), gnomAD
- Whole exome sequencing (WES) utility > Search first: GTR, ClinVar, OMIM, GeneMatcher
- Gene panels (which panels, which genes) > Search first: GTR, ClinVar, laboratory-specific databases
- Single gene testing > Search first: GTR, ClinVar, OMIM, GeneReviews
- Chromosomal microarray (CMA) > Search first: DECIPHER, ClinVar, dbVar, ECARUCA
- Karyotyping > Search first: Chromosome Abnormality Database, ClinVar, cytogenetics resources
- FISH > Search first: ClinVar, cytogenetics databases, PubMed
- Mitochondrial DNA testing > Search first: MITOMAP, MSeqDR, ClinVar, GTR
- Repeat expansion testing > Search first: GTR, ClinVar, repeat expansion databases, PubMed
- Omics-Based Diagnostics (if applicable):
- RNA sequencing / transcriptomics > Search first: GEO, ArrayExpress, GTEx, RNA-seq databases
- Proteomics > Search first: PRIDE, ProteomeXchange, FDA Biomarker database
- Metabolomics > Search first: MetaboLights, Metabolomics Workbench, HMDB
- Epigenomics > Search first: GEO, ENCODE, Roadmap Epigenomics, MethBase
- Liquid biopsy > Search first: COSMIC, ClinVar, liquid biopsy databases, PubMed
- Clinical Criteria:
- Standardized diagnostic criteria (DSM, ICD, society guidelines) > Search first: DSM-5, ICD-11, clinical society guidelines, UpToDate
- Differential diagnosis (other conditions to rule out, with distinguishing features) > Search first: DynaMed, UpToDate, clinical decision support systems
- Screening:
- Screening methods for asymptomatic individuals (newborn screening, carrier screening, cascade screening) > Search first: ACMG recommendations, CDC newborn screening, GTR
11. Outcome/Prognosis
- Survival and Mortality:
- Survival rate (5-year, 10-year, overall) > Search first: SEER, cancer registries, disease-specific registries, PubMed
- Life expectancy (with and without treatment if applicable) > Search first: Orphanet, disease registries, actuarial databases, PubMed
- Mortality rate > Search first: CDC, WHO, GBD, national mortality databases
- Disease-specific mortality (deaths directly attributable to disease) > Search first: Disease registries, CDC Wonder, GBD, PubMed
- Morbidity and Function:
- Morbidity (disease-related disability and health impacts) > Search first: GBD, WHO, disability databases, PubMed
- Disability outcomes (long-term functional impairments) > Search first: ICF (International Classification of Functioning), disability registries
- Quality of life measures (EQ-5D, SF-36, PROMIS, disease-specific tools) > Search first: EQ-5D database, SF-36, PROMIS, PubMed
- Disease Course:
- Complications (secondary problems: infections, organ failure, etc.) > Search first: ICD codes, disease registries, clinical databases, PubMed
- Recovery potential (likelihood and extent of recovery, with vs without treatment) > Search first: Natural history studies, rehabilitation databases, PubMed
- Prediction:
- Prognostic factors (age, disease severity, biomarkers, treatment response) > Search first: Prognostic models databases, clinical calculators, PubMed
- Prognostic biomarkers (molecular markers predicting disease course) > Search first: FDA Biomarker database, PubMed, cancer prognostic databases
12. Treatment
- Pharmacotherapy:
- Pharmacological treatments (drug names, drug classes, mechanisms of action) > Search first: DrugBank, RxNorm, ATC classification, DailyMed, FDA databases
- Pharmacogenomics (how genetic variants affect drug metabolism, efficacy, toxicity) > Search first: PharmGKB, CPIC (Clinical Pharmacogenetics), FDA Table of PGx Biomarkers
- Advanced Therapeutics:
- Gene therapy (viral vectors, CRISPR, gene replacement, gene editing) > Search first: ClinicalTrials.gov, FDA gene therapy database, ASGCT resources
- Cell therapy (stem cell transplant, CAR-T, cellular therapeutics) > Search first: ClinicalTrials.gov, FDA cell therapy database, FACT standards
- RNA-based therapies (ASOs, siRNA, mRNA therapies) > Search first: ClinicalTrials.gov, FDA approvals, PubMed
- Targeted therapies (treatments directed at specific molecular targets) > Search first: My Cancer Genome, OncoKB, ClinicalTrials.gov, FDA approvals
- Immunotherapies (checkpoint inhibitors, monoclonal antibodies) > Search first: Cancer Immunotherapy Database, FDA approvals, ClinicalTrials.gov
- Surgical and Interventional:
- Surgical interventions (types of surgery, timing, outcomes) > Search first: CPT codes, surgical registries, clinical guidelines, PubMed
- Supportive and Rehabilitative:
- Supportive care (symptom management, pain control, nutrition) > Search first: Clinical guidelines, Cochrane Library, PubMed
- Rehabilitation (physical therapy, occupational therapy, speech therapy) > Search first: Rehabilitation medicine databases, clinical guidelines, PubMed
- Experimental:
- Experimental treatments in clinical trials (with NCT identifiers if available) > Search first: ClinicalTrials.gov, EU Clinical Trials Register, WHO ICTRP
- Treatment Outcomes:
- Treatment response rates > Search first: Clinical trial databases, FDA reviews, systematic reviews, PubMed
- Side effects and adverse events > Search first: FDA Adverse Event Reporting System (FAERS), MedWatch, PubMed
- Treatment Strategy:
- Treatment algorithms (clinical pathways, decision trees) > Search first: Clinical practice guidelines, NCCN Guidelines, UpToDate
- Combination therapies > Search first: ClinicalTrials.gov, treatment guidelines, PubMed
- Personalized medicine approaches (genotype-guided treatment) > Search first: My Cancer Genome, CIViC, PharmGKB, precision medicine databases
For each treatment, suggest NCIT (NCI Thesaurus) clinical-intervention terms where applicable.
13. Prevention
- Prevention Levels:
- Primary prevention (preventing disease occurrence: vaccination, risk factor modification) > Search first: CDC, WHO, USPSTF recommendations, Cochrane Library
- Secondary prevention (early detection and treatment: screening programs, early intervention) > Search first: USPSTF, CDC screening guidelines, WHO
- Tertiary prevention (preventing complications in those with disease) > Search first: Clinical guidelines, disease management protocols, PubMed
- Immunization: Vaccine strategies (if applicable)
Search first: CDC vaccine schedules, WHO immunization, FDA vaccine database
- Screening and Early Detection:
- Screening programs (population-based: newborn screening, cancer screening) > Search first: CDC screening programs, USPSTF, cancer screening databases
- Genetic screening (carrier screening, preimplantation genetic diagnosis, prenatal testing) > Search first: ACMG recommendations, ACOG guidelines, GTR
- Risk stratification (identifying high-risk individuals for targeted prevention) > Search first: Risk prediction models, clinical calculators, PubMed
- Behavioral Interventions: Lifestyle modifications to reduce risk
Search first: CDC, WHO, behavioral intervention databases, Cochrane Library
- Counseling: Genetic counseling (risk assessment, family planning guidance)
Search first: NSGC resources, ACMG guidelines, GeneReviews
- Public Health:
- Public health interventions (sanitation, vector control, health education) > Search first: CDC, WHO, public health databases, PubMed
- Environmental interventions (reducing environmental risk factors) > Search first: EPA databases, WHO environmental health, PubMed
- Prophylaxis: Preventive medications or procedures
Search first: Clinical guidelines, FDA approvals, PubMed
14. Other Species / Natural Disease
- Taxonomy: Species affected (with NCBI Taxon identifiers)
Search first: NCBI Taxonomy
- Breed: Specific breeds affected (with VBO identifiers if applicable)
Search first: VBO (Vertebrate Breed Ontology)
- Gene: Orthologous genes in other species (with NCBI Gene IDs)
Search first: NCBI Gene
- Natural Disease:
- Naturally occurring disease in other species (companion animals, wildlife) > Search first: OMIA (Online Mendelian Inheritance in Animals), VetCompass, PubMed
- Veterinary relevance and importance in animal health > Search first: OMIA, veterinary databases, PubMed
- Comparative Biology:
- Comparative pathology (similarities and differences across species) > Search first: OMIA, comparative pathology databases, PubMed
- Evolutionary conservation of disease mechanisms > Search first: HomoloGene, OrthoMCL, Alliance of Genome Resources
- Transmission (if applicable):
- Zoonotic potential > Search first: CDC zoonotic diseases, WHO zoonoses, GIDEON
- Cross-species susceptibility > Search first: NCBI Taxonomy, veterinary databases, PubMed
15. Model Organisms
- Model Types:
- Model organism type (mammalian, invertebrate, cellular, in vitro) > Search first: Alliance of Genome Resources, model organism databases
- Specific model systems (mouse, rat, zebrafish, Drosophila, C. elegans, yeast, cell lines, organoids, iPSCs) > Search first: MGI, RGD, ZFIN, FlyBase, WormBase, SGD, ATCC, Cellosaurus
- Induced models (drug treatment, surgical intervention, environmental manipulation) > Search first: MGI, model organism databases, PubMed
- Genetic Models:
- Types available (knockout, knock-in, transgenic, conditional, humanized) > Search first: MGI, IMPC, KOMP, EuMMCR, IMSR
- Model Characteristics:
- Phenotype recapitulation (how well model reproduces human disease features) > Search first: Model organism databases, comparative studies, PubMed
- Model limitations (aspects of human disease not captured) > Search first: Model organism databases, PubMed, review articles
- Applications:
- Research applications (what aspects of disease can be studied) > Search first: Model organism databases, PubMed
- Resources:
- Model databases > Search first: MGI, RGD, ZFIN, FlyBase, WormBase, IMSR, EMMA, MMRRC
Citation Requirements
- Cite primary literature (PMID preferred) for all mechanistic and clinical claims
- Prioritize recent reviews and landmark papers
- Include direct quotes from abstracts where possible to support key statements
- Distinguish evidence source types: human clinical, model organism, in vitro, computational
Output Format
Structure your response as a comprehensive narrative organized by the sections above. For each section, provide: - Factual content with specific details (numbers, percentages, gene names, variant nomenclature) - Ontology term suggestions (HPO, GO, CL, UBERON, CHEBI, NCIT, MONDO) where applicable - Evidence citations with PMIDs - Direct quotes from abstracts to support key claims - Clear indication when information is not available or not applicable for this disease
This report will be used to populate a disease knowledge base entry with: - Pathophysiology descriptions with causal chains - Gene/protein annotations (HGNC, GO terms) - Phenotype associations (HP terms) with frequencies - Cell type involvement (CL terms) - Anatomical locations (UBERON terms) - Chemical entities (CHEBI terms) - Treatment annotations (NCIT terms) - Evidence items with PMIDs and exact abstract quotes - Epidemiology, prognosis, diagnostic, and prevention information - Animal model descriptions with phenotype recapitulation details
Autosomal Recessive Nonsyndromic Hearing Loss 124 (DFNB124)
PKHD1L1-related deafness: disease-characteristics research report
Evidence cutoff and scope. DFNB124 is a newly delineated, exceptionally rare Mendelian hearing disorder. The disease-specific human literature located for this report consists principally of a 2024 four-family report; consequently, percentages derived from those four probands must not be treated as population frequencies. Mechanistic support comes mainly from in-vitro studies and mouse and zebrafish models. General congenital sensorineural-hearing-loss (SNHL) evidence is explicitly labeled as such.
| Domain | Established finding | Evidence type/sample | Quantitative detail | Confidence/limitation |
|---|---|---|---|---|
| Disease identity | Autosomal recessive nonsyndromic hearing loss 124; PKHD1L1-related deafness; DFNB124 | Curated disease-resource and peer-reviewed evidence | MONDO:0968981; OMIM phenotype 620794 | Exact identifiers supported; no disease-specific Orphanet, ICD, or MeSH identifier was established in the retrieved sources. (strelkova2024pkhd1l1isrequired pages 1-2, OpenTargets Search: autosomal recessive nonsyndromic hearing loss 124-PKHD1L1) |
| Human phenotype | Bilateral congenital or early-onset sensorineural hearing loss ranging from mild–moderate to severe | Human series of four unrelated probands | Audiometry reported at ages 13, 9, 12, and 8 years | Foundational but very small cohort; population-level phenotype frequencies cannot be estimated. (redfield2024pkhd1l1agene pages 1-2, redfield2024pkhd1l1agene pages 5-8, redfield2024pkhd1l1agene pages 8-9) |
| Family 1 | Slowly progressive bilateral mild–moderate SNHL in a White American female who failed newborn screening | Human longitudinal audiology | PTA increased 5 dB right and 8 dB left from ages 4.3–13.3 years; latest PTA 45.00/48.75 dB; word recognition 90%; SRT 45 dB bilaterally | Best longitudinal human evidence; episodic BPPV resolved with Epley maneuver, but association with DFNB124 is uncertain. (redfield2024pkhd1l1agene pages 5-8) |
| Families 2–4 | Family 2: Iranian Lur boy with progressive moderate–severe SNHL; Family 3: Pakistani boy with severe SNHL; Family 4: Chinese boy with moderate SNHL | Human cases in the four-family series | Family 2: diagnosed at 2.5 months, SRT 60 dB, SDS 100% at 80 dB; Family 3: PTA 85 dB HL; Family 4: absent DPOAEs with normal tympanograms | Follow-up was limited; Family 3 also had a homozygous MYO7A variant, weakening attribution of severity solely to PKHD1L1. (redfield2024pkhd1l1agene pages 8-9, redfield2024pkhd1l1agene pages 11-13) |
| Reported genotypes | Six alleles occur in four biallelic genotypes: p.[Gly129Ser];[Gly1314Val], homozygous p.Arg3381Ter, homozygous p.His2479Gln, and p.[Gly605Arg];[Leu2818TyrfsTer5] | Human exome sequencing and segregation | Alleles: c.385G>A, c.3941G>T, c.10141C>T, c.7437C>A, c.1813G>A, and c.8452_8468del; protein reference NP_803875.2 | Only four disease-associated genotypes were reported; current ClinVar and laboratory ACMG classifications should be checked before clinical use. (redfield2024pkhd1l1agene pages 1-2, redfield2024pkhd1l1agene pages 5-8, redfield2024pkhd1l1agene pages 8-9) |
| Population frequency | Reported alleles are rare but not uniformly ultra-rare | gnomAD frequencies reported in the discovery study | Gly129Ser 0.001471%; Gly1314Val 0.07204%; Arg3381Ter 0.02067%; His2479Gln 0.3107% | Frequencies vary by database version and ancestry; the higher His2479Gln frequency and competing MYO7A finding require caution. (redfield2024pkhd1l1agene pages 5-8, redfield2024pkhd1l1agene pages 8-9) |
| Functional assays | Gly129Ser and Gly1314Val destabilize recombinant PKHD1L1 fragments; Gly605Arg alters splicing | In-vitro NanoDSF and HEK293/HeLa minigene assays | Gly129Ser reduced unfolding onset by about 6 °C and melting transitions by about 4 °C; Gly1314Val reduced onset by about 7 °C and melting temperature by 9.1 °C; Gly605Arg caused exon 17 skipping and p.Val557_Arg604del | Direct molecular effects were demonstrated in protein fragments or cultured cells, not full-length protein in human cochlear tissue. (redfield2024pkhd1l1agene pages 9-11, redfield2024pkhd1l1agene pages 11-13, redfield2024pkhd1l1agene pages 13-15) |
| Protein mechanism | PKHD1L1 is a large, predominantly extracellular, single-pass membrane component of the transient stereocilia surface coat | Mouse immunolocalization, immunogold SEM, and structural modeling | Approximately 4,249 amino acids; 14 predicted IPT repeats; enriched near stereocilia tips, especially in high-frequency cochlear regions | Coat localization is demonstrated in mice; binding partners and precise biochemical function remain unknown. (wu2019pkhd1l1isa pages 1-2, strelkova2024pkhd1l1isrequired pages 1-2, redfield2024pkhd1l1agene pages 9-11) |
| Mouse models | Hair-cell-specific and constitutive Pkhd1l1 loss causes progressive stereocilia loss, bundle disorganization, and hearing impairment | Conditional Pkhd1l1 floxed/Atoh1-Cre-positive and constitutive knockout mice | High-frequency ABR and DPOAE deficits were evident by about 6 weeks and extended toward lower frequencies with age; stereocilia loss was assessed at 6 weeks, 14 weeks, and 9 months | Strong causal animal evidence, although mouse onset is delayed relative to congenital hearing loss in humans. (strelkova2024pkhd1l1isrequired pages 10-11, strelkova2024pkhd1l1isrequired pages 1-2, strelkova2024pkhd1l1isrequired pages 12-13) |
| Zebrafish model | Combined loss of the pkhd1l1 paralogs impairs auditory behavior in larvae | pkhd1l1a/pkhd1l1b double-mutant zebrafish | Reduced auditory-evoked startle at 6 days post-fertilization | Supports conserved auditory function, but startle behavior is an indirect hearing measure. (strelkova2024pkhd1l1isrequired pages 1-2, redfield2024pkhd1l1agene pages 11-13, redfield2024pkhd1l1agene pages 1-2) |
| Gene–environment interaction | PKHD1L1 deficiency increases susceptibility to acoustic overexposure | Controlled mouse noise-exposure experiment | Moderate octave-band exposure caused permanent threshold shifts in deficient mice but temporary shifts in controls; follow-up extended from 1 day to 8 weeks | Compelling preclinical interaction; increased human noise susceptibility remains unproven. (strelkova2024pkhd1l1isrequired pages 1-2, strelkova2024pkhd1l1isrequired pages 12-13, strelkova2024pkhd1l1isrequireda pages 19-21, strelkova2024pkhd1l1isrequired pages 10-11) |
| Diagnostics | Diagnosis requires confirmation of bilateral SNHL and biallelic PKHD1L1 variants with segregation and careful variant interpretation | Human audiology and exome sequencing; general congenital-SNHL practice | Relevant tests include newborn screening, ABR or behavioral audiometry, tympanometry, OAEs, speech testing, and panel/WES/WGS analysis with CNV detection | No DFNB124-specific clinical criteria or biomarker exists; exome reanalysis identified Family 4, showing the value of periodic reinterpretation. (redfield2024pkhd1l1agene pages 5-8, redfield2024pkhd1l1agene pages 8-9, redfield2024pkhd1l1agene pages 1-2) |
| Management | Care is phenotype-directed: hearing aids, speech-language and communication intervention, educational accommodations, serial audiology, and cochlear-implant evaluation when indicated | General congenital-SNHL care; one DFNB124 proband used bilateral hearing aids | Family 2 used bilateral hearing aids; no disease-specific response rate is available | No evidence shows that standard hearing devices perform differently in DFNB124; no therapy currently corrects PKHD1L1 dysfunction. (redfield2024pkhd1l1agene pages 8-9, fan2026internationalexpertconsensus pages 1-3, redfield2024pkhd1l1agene pages 1-2) |
| Epidemiology | Disease-specific prevalence, incidence, penetrance, carrier frequency, sex ratio, founder effects, and geographic distribution are unknown | Four unrelated families from the United States, Iran, Pakistan, and China | Four published probands in the founding series; two came from consanguineous families | Geographic diversity suggests the disease is not confined to one population, but the sample cannot establish demographic risks or prevalence. (redfield2024pkhd1l1agene pages 1-2, redfield2024pkhd1l1agene pages 8-9) |
| Clinical trials | No PKHD1L1/DFNB124-specific interventional trial or targeted therapy was identified | ClinicalTrials.gov and literature searches | Zero relevant disease-specific trials among retrieved records | A search-negative finding is not proof that no unregistered or newly initiated study exists; current hereditary-hearing-loss gene-therapy trials chiefly target other genes such as OTOF. (fan2026internationalexpertconsensus pages 1-3, li2024advancedmanagementof pages 3-4, redfield2024pkhd1l1agene pages 1-2) |
Table: Compact evidence map for PKHD1L1-related deafness, separating human, in-vitro, animal, and inferred findings. It also highlights the major epidemiologic, diagnostic, therapeutic, and clinical-trial evidence gaps.
1. Disease information
Definition
DFNB124 is an autosomal-recessive, nonsyndromic, usually congenital or very-early-onset bilateral sensorineural hearing loss caused by biallelic variants in PKHD1L1. In the founding series, severity ranged from mild–moderate to severe. The disorder was established through segregation of biallelic variants in four unrelated families, functional testing of selected alleles, and concordant animal models (PMID 38459354; published online 9 March 2024; DOI/URL: https://doi.org/10.1007/s00439-024-02649-2). (redfield2024pkhd1l1agene pages 1-2)
A key abstract conclusion was: “Multiple lines of evidence collectively associate PKHD1L1 with nonsyndromic mild–moderate to severe sensorineural hearing loss.” (redfield2024pkhd1l1agene pages 2-4)
Identifiers and synonyms
- Preferred name: autosomal recessive nonsyndromic hearing loss 124.
- Synonyms: DFNB124; PKHD1L1-related deafness; PKHD1L1-related autosomal-recessive nonsyndromic hearing loss.
- MONDO: MONDO:0968981.
- OMIM phenotype: #620794, cited in the November 2024 mechanistic paper.
- Gene: PKHD1L1, approved name PKHD1 like 1; Ensembl ENSG00000205038. Protein aliases include fibrocystin-L/FPC-L. (strelkova2024pkhd1l1isrequired pages 1-2, OpenTargets Search: autosomal recessive nonsyndromic hearing loss 124-PKHD1L1)
- Orphanet: no disease-specific entry was established in the retrieved evidence.
- ICD-10/ICD-11 and MeSH: no DFNB124-specific code or descriptor was established. Broader congenital/bilateral SNHL codes should not be represented as uniquely identifying DFNB124.
The source data are aggregated disease-level resources plus published individual research participants, not routine EHR-derived evidence. Open Targets records one PKHD1L1–DFNB124 association, supported by PMID 38459354 and variant-condition records. (OpenTargets Search: autosomal recessive nonsyndromic hearing loss 124-PKHD1L1)
2. Etiology
Causal factor
The primary cause is germline biallelic PKHD1L1 variation, consistent with loss or impairment of PKHD1L1 function. Reported allele classes include missense, nonsense, frameshift, and a missense substitution that disrupts splicing. No infectious or purely environmental cause defines DFNB124. (redfield2024pkhd1l1agene pages 1-2, redfield2024pkhd1l1agene pages 8-9)
Genetic risk factors
Risk is determined chiefly by inheriting pathogenic or likely pathogenic alleles in trans. Consanguinity increases the probability that both parents carry the same rare allele: two of four founding probands had consanguineous parents. Family history can be absent because heterozygous parents have normal hearing. (redfield2024pkhd1l1agene pages 1-2, redfield2024pkhd1l1agene pages 8-9)
No validated susceptibility loci, modifier genes, protective variants, founder mutations, or epigenetic risk factors are known. Family 3 also carried homozygous MYO7A p.(Leu375Val); its possible contribution makes that proband’s severe phenotype less securely attributable solely to PKHD1L1. (redfield2024pkhd1l1agene pages 11-13)
Environmental and protective factors
No human environmental risk or protective factor has been demonstrated specifically for DFNB124. Mouse evidence shows a plausible gene–noise interaction: moderate acoustic overexposure caused permanent threshold loss in deficient mice but only temporary shifts in controls. Thus, avoiding hazardous noise is prudent, but a human PKHD1L1-specific benefit has not been measured. No diet, exercise, medication, or genetic protective allele is known. (strelkova2024pkhd1l1isrequired pages 1-2, strelkova2024pkhd1l1isrequireda pages 19-21)
3. Phenotypes
Core auditory phenotype
All four reported probands had bilateral congenital or presumed-congenital SNHL. On this ascertainment-limited sample, bilateral SNHL was 4/4, but this is a study inclusion characteristic rather than a reliable population frequency. Severity was mild–moderate in Family 1, moderate–severe in Family 2, severe in Family 3, and moderate in Family 4. Their pedigrees and audiograms demonstrate bilateral but variable configurations. (redfield2024pkhd1l1agene pages 1-2, redfield2024pkhd1l1agene pages 8-9, redfield2024pkhd1l1agene media e885a2a5)
Suggested HPO terms: - Sensorineural hearing impairment — HP:0000407. - Bilateral sensorineural hearing impairment — HP:0008619. - Congenital hearing impairment — HP:0008527. - Progressive hearing impairment — HP:0001730, where longitudinally demonstrated. - Moderate hearing impairment — HP:0012712; severe hearing impairment — HP:0012714. - Absent otoacoustic emissions — use the current HPO term after ontology validation.
Case-level characteristics
- Family 1: 13-year-old White American female; failed bilateral newborn screening. Mild–moderate bilateral SNHL progressed slowly: PTA rose 5 dB right and 8 dB left between ages 4.3 and 13.3 years. Latest PTAs were 45.00/48.75 dB; word recognition was 90%, and SRT 45 dB bilaterally. Episodic benign paroxysmal positional vertigo resolved with an Epley maneuver. Normal ECG and ophthalmology; no dysmorphism, neurologic, or developmental abnormality. The vertigo cannot presently be assigned to DFNB124. (redfield2024pkhd1l1agene pages 5-8)
- Family 2: 9-year-old Iranian Lur male; congenital loss clinically diagnosed at 2.5 months and progressing to bilateral moderate–severe SNHL across frequencies. SRT was 60 dB; speech discrimination 100% at 80 dB; OAEs were present bilaterally. No vestibular abnormality or motor delay; bilateral hearing-aid use was reported. (redfield2024pkhd1l1agene pages 8-9)
- Family 3: 12-year-old Pakistani male with congenital bilateral severe SNHL; PTA 85 dB HL. Follow-up was unavailable, and interpretation is complicated by the additional MYO7A variant. (redfield2024pkhd1l1agene pages 8-9, redfield2024pkhd1l1agene pages 11-13)
- Family 4: 8-year-old boy from Henan, China; presumed congenital bilateral moderate SNHL. DPOAEs were absent and tympanograms normal, supporting outer-hair-cell dysfunction; newborn screening had not been performed. (redfield2024pkhd1l1agene pages 8-9)
Non-auditory manifestations and quality of life
The four probands lacked consistent syndromic involvement; no human evidence currently links PKHD1L1 variants to seizures. No disease-specific behavioral, laboratory, renal, hepatic, ophthalmic, or neurodevelopmental phenotype is established despite the gene’s name. (redfield2024pkhd1l1agene pages 15-16, redfield2024pkhd1l1agene pages 13-15)
DFNB124-specific quality-of-life instruments have not been reported. By extrapolation from pediatric hearing loss, consequences can include impaired speech perception, communication, education, psychosocial well-being, and participation, depending on severity, access to language, and intervention. These are anticipated consequences of hearing impairment, not additional PKHD1L1 manifestations.
4. Genetic and molecular information
Gene and protein
PKHD1L1 encodes a very large, predominantly extracellular, single-pass membrane protein of approximately 4,243–4,249 amino acids, depending on reference annotation. The human protein contains a signal peptide, approximately 14 predicted extracellular IPT/plexin-like repeats, a TMEM2-like region, one transmembrane segment, and a very short cytoplasmic tail. Protein numbering in the 2024 human study used NP_803875.2. (redfield2024pkhd1l1agene pages 1-2, wu2019pkhd1l1isa pages 1-2, strelkova2024pkhd1l1isrequired pages 1-2, redfield2024pkhd1l1agene pages 9-11)
Reported biallelic genotypes
- c.385G>A, p.(Gly129Ser) in trans with c.3941G>T, p.(Gly1314Val).
- Homozygous c.10141C>T, p.(Arg3381Ter).
- Homozygous c.7437C>A, p.(His2479Gln).
- c.1813G>A, p.(Gly605Arg) in trans with c.8452_8468del, p.(Leu2818TyrfsTer5). (redfield2024pkhd1l1agene pages 1-2, redfield2024pkhd1l1agene pages 8-9)
Reported gnomAD maximum allele frequencies were 0.001471% for Gly129Ser, 0.07204% for Gly1314Val, 0.02067% for Arg3381Ter, and 0.3107% for His2479Gln. Frequencies are database-version and ancestry dependent and should be refreshed directly before clinical interpretation. The comparatively high His2479Gln frequency and the competing MYO7A finding warrant particular caution. (redfield2024pkhd1l1agene pages 5-8, redfield2024pkhd1l1agene pages 8-9)
These are germline variants. No somatic DFNB124 mechanism is recognized. The retrieved text did not provide definitive current ClinVar classifications for every allele; laboratories should apply current ACMG/AMP criteria rather than treating all six alleles as equivalently established.
Functional consequences
- Gly129Ser reduced recombinant-fragment unfolding onset by approximately 6°C and shifted melting transitions by about 4°C.
- Gly1314Val reduced unfolding onset by about 7°C and melting temperature by 9.1°C.
- Gly605Arg, at an exon boundary, caused exon 17 skipping in HEK293 and HeLa minigene assays, yielding r.1670_1813del; p.Val557_Arg604del, an in-frame 48-residue deletion.
- Arg3381Ter is predicted to trigger nonsense-mediated decay; any escaping truncated protein would lack about 882 residues, including the transmembrane domain, impairing membrane insertion or causing abnormal secretion.
- Leu2818TyrfsTer5 is a truncating frameshift and presumptive loss-of-function allele.
- His2479Gln affects a conserved residue in a modeled TMEM2-like, possible cation-binding region; this consequence remains computational rather than experimentally proven. (redfield2024pkhd1l1agene pages 13-15, redfield2024pkhd1l1agene pages 9-11, redfield2024pkhd1l1agene pages 11-13)
No validated modifier gene, disease-specific methylation signature, chromatin abnormality, recurrent CNV, inversion, translocation, or aneuploidy has been reported.
5. Environmental information
No toxin, infection, smoking pattern, diet, alcohol exposure, occupation, or lifestyle factor causes this Mendelian disorder. Standard acquired-hearing-loss exposures can independently worsen auditory function and confound phenotype assessment. Mouse data specifically support enhanced vulnerability to acoustic overexposure, whereas human noise susceptibility is untested. Ototoxic medications and infections should therefore be recorded as potential competing or additive causes, not labeled established DFNB124 modifiers. (strelkova2024pkhd1l1isrequired pages 1-2, strelkova2024pkhd1l1isrequireda pages 19-21)
6. Mechanism and pathophysiology
Ordered causal chain
- Biallelic PKHD1L1 variants lead to absent, truncated, misspliced, destabilized, or otherwise dysfunctional PKHD1L1 protein.
- PKHD1L1 dysfunction leads to deficient formation or altered material properties of the transient developmental stereocilia surface coat in cochlear hair cells; direct demonstration exists in knockout mice, while extrapolation to human cochlea is inferred.
- An abnormal developmental coat leads to hair bundles that initially form and transduce relatively normally but are less mechanically durable; this durability step is inferred from the temporal mouse phenotype.
- Reduced bundle durability leads to progressive loss or shortening of stereocilia and loss of bundle coherence, first prominent in basal/high-frequency outer hair cells in mice.
- Outer-hair-cell bundle damage leads to reduced cochlear amplification, reflected by elevated DPOAE thresholds.
- Reduced amplification and hair-bundle function lead to elevated auditory thresholds and bilateral SNHL.
- Branch—aging leads to extension of high-frequency deficits toward lower frequencies in mice.
- Branch—moderate noise exposure leads to disproportionately persistent stereocilia damage and permanent threshold shift in deficient mice; the corresponding human interaction remains unproven. (wu2019pkhd1l1isa pages 1-2, strelkova2024pkhd1l1isrequired pages 10-11, strelkova2024pkhd1l1isrequired pages 1-2)
Molecular and cellular detail
Mouse immunogold SEM localized PKHD1L1 over the stereocilia surface, especially near tips and in high-frequency cochlear regions. Knockout removed the upper stereociliary coat but did not abolish the lower coating. Early planar polarity, FM1-43 uptake, gross cochlear anatomy, STRC localization, and tectorial-membrane attachment-crown formation were largely preserved. Thus, PKHD1L1 is not established as a MET-channel component or canonical signaling-pathway protein; no Wnt, MAPK, mTOR, PI3K–AKT, metabolic, immune, inflammatory, apoptotic, or autophagic pathway has been causally implicated in DFNB124. (wu2019pkhd1l1isa pages 1-2, strelkova2024pkhd1l1isrequired pages 5-6, wu2019pkhd1l1isa pages 8-9)
In 2024 mice, Pkhd1l1 mRNA was detected in inner and outer hair cells. Protein was prominent on bundles during P4–P8, mostly gone by P10, and undetectable by P21, although later structural failure began around six weeks. This temporal separation supports a developmental “build quality” or resilience function rather than a requirement for continuous abundant adult protein. (strelkova2024pkhd1l1isrequired pages 3-4, strelkova2024pkhd1l1isrequireda pages 3-5)
Suggested GO biological processes: sensory perception of sound (GO:0007605); inner-ear development (GO:0048839); stereocilium organization (GO:0032429); actin-filament-based process (GO:0030029); mechanosensory behavior (GO:0007638). Use “maintenance of stereocilia bundle” only if available in the ontology release.
Suggested cellular components: stereocilium (GO:0032420); stereocilium tip (GO:0032426); plasma membrane (GO:0005886); extracellular region (GO:0005576); cell projection membrane (GO:0031253).
Suggested cell types: auditory hair cell (CL:0000202, verify current label); cochlear inner hair cell and cochlear outer hair cell using current CL terms. The strongest evidence points to outer hair cells, but inner-hair-cell expression is also documented.
No human DFNB124 transcriptomic, proteomic, metabolomic, lipidomic, epigenomic, spatial-transcriptomic, organoid, iPSC, or CRISPR-screen signature has been reported. Single-cell auditory datasets provide context for hair-cell regulation but do not constitute a disease profile.
7. Anatomical structures affected
- Organ: inner ear, specifically the cochlea; no consistent secondary-organ disease.
- Tissue: organ of Corti sensory epithelium.
- Cells: inner and outer cochlear hair cells, with especially strong protein localization and physiological evidence in outer hair cells.
- Subcellular site: apical actin-rich stereocilia bundles, their extracellular surface coat, and plasma membrane anchoring of PKHD1L1.
- Tonotopic localization: mouse damage and hearing loss begin preferentially in basal/high-frequency regions and spread toward lower-frequency regions with age.
- Lateralization: bilateral in all four reported humans. (wu2019pkhd1l1isa pages 1-2, strelkova2024pkhd1l1isrequired pages 10-11, redfield2024pkhd1l1agene pages 8-9)
Suggested UBERON terms: inner ear (UBERON:0001846); cochlea (UBERON:0001844); organ of Corti (UBERON:0002227, verify); cochlear duct (UBERON:0002292, verify); tectorial membrane using the current UBERON term. No structural imaging abnormality is established.
8. Temporal development
Human onset was congenital or presumed congenital. Family 1 failed newborn screening, Family 2 was diagnosed at 2.5 months, and Families 3–4 were described as congenital/presumed congenital. The course may be stable or progressive, but only Family 1 had quantitative longitudinal data; Family 2 was described as progressive. Disease stages have not been formally defined. (redfield2024pkhd1l1agene pages 5-8, redfield2024pkhd1l1agene pages 8-9)
The disorder is expected to be lifelong because mammalian auditory hair cells do not regenerate. There is no evidence for episodic remission. The critical clinical period is early childhood, when hearing access supports language development; this is a general congenital-SNHL principle rather than a PKHD1L1-specific trial result. The critical biological period in mice is early postnatal stereocilia-coat formation. (strelkova2024pkhd1l1isrequired pages 1-2, rajanbabu2024earlyhearingdetection pages 5-6)
9. Inheritance and population
Inheritance
Inheritance is autosomal recessive. For two carrier parents, each pregnancy has a theoretical 25% affected, 50% carrier, and 25% unaffected/non-carrier probability. Heterozygous parents in reported pedigrees had normal or subjectively normal hearing. (redfield2024pkhd1l1agene pages 8-9, redfield2024pkhd1l1agene media e885a2a5)
Penetrance cannot be estimated from four ascertained probands. Expressivity is variable, spanning mild–moderate to severe hearing loss. There is no evidence of anticipation, parent-of-origin effect, germline mosaicism, or sex linkage.
Epidemiology
No reliable disease-specific prevalence, incidence, carrier frequency, sex ratio, founder effect, or geographic distribution is available. Four probands were reported from the United States, Iran, Pakistan, and China; this diversity argues against confinement to one population but cannot establish relative risk. Two of four families were consanguineous. (redfield2024pkhd1l1agene pages 1-2, redfield2024pkhd1l1agene pages 8-9)
For context only, GBD 2021 estimated 97.83 million people under 20 had hearing loss in 2021, producing 3.91 million YLDs; prevalence was 3,711 per 100,000 and 62.1% of cases were mild. These figures cover hearing loss of all causes and must not be assigned to DFNB124. (guo2024globalregionaland pages 11-11)
10. Diagnostics
Clinical evaluation
There are no disease-specific clinical criteria. Evaluation should document congenital/early bilateral SNHL with: - universal newborn screening using OAE and/or automated ABR; - diagnostic ABR in infants or developmentally appropriate pure-tone audiometry; - air- and bone-conduction thresholds, tympanometry, OAEs, speech-reception and word-recognition testing; - serial audiograms to detect progression; - otologic, vestibular, developmental, ophthalmologic, and family-history assessment; - MRI/CT only when clinically indicated to assess anatomy or implantation planning, not to diagnose DFNB124.
European programs reviewed in 2024 generally exceeded 90% coverage and commonly used staged TEOAE/automated ABR followed by diagnostic ABR, illustrating real-world screening implementation rather than a DFNB124-specific protocol. (hatzopoulos2024theotoacousticemissions pages 4-5)
Genetic testing
A contemporary comprehensive hearing-loss panel that includes PKHD1L1, with sequence and exon-level CNV analysis, is a practical first-line test. Trio WES or WGS is appropriate when panel testing is negative, the phenotype is atypical, or novel/splice/structural variants are suspected. Family 4’s initially negative exome was solved after reanalysis, demonstrating the value of periodic reinterpretation as gene–disease knowledge changes. Sanger or equivalent orthogonal confirmation and segregation testing are advisable for reportable biallelic variants. (redfield2024pkhd1l1agene pages 8-9, redfield2024pkhd1l1agene pages 1-2)
Single-gene sequencing can be used for known familial variants but is inefficient for an unsolved proband because hereditary hearing loss is highly heterogeneous. CMA, karyotyping, FISH, mitochondrial testing, and repeat-expansion assays are not primary DFNB124 tests unless other findings suggest those etiologies. RNA studies may resolve suspected splice variants, but patient cochlear tissue is inaccessible; minigene assays remain research-level evidence.
Differential diagnosis
The differential includes other autosomal-recessive nonsyndromic deafness genes—particularly GJB2, STRC, OTOF, SLC26A4, MYO7A, TMC1, PCDH15, LOXHD1, and many others—plus congenital CMV, ototoxic exposure, inner-ear malformation, auditory neuropathy, and syndromic hearing loss. Distinguishing DFNB124 requires a convincing biallelic genotype and phenotype compatibility, not audiometry alone. Family 3 illustrates the need to evaluate competing variants. (redfield2024pkhd1l1agene pages 5-8, redfield2024pkhd1l1agene pages 11-13)
Screen relatives by cascade testing after a molecular diagnosis. Newborn screening detects hearing loss but not genotype and may miss mild or delayed/progressive cases.
11. Outcome and prognosis
No mortality or shortened-life-expectancy signal exists; DFNB124 is not known to affect survival. The major morbidity is lifelong auditory disability with possible communication, educational, psychosocial, and occupational effects. No DFNB124-specific EQ-5D, SF-36, PROMIS, language, academic, or cochlear-implant outcome data exist.
Residual hearing and speech discrimination can be substantial: Family 1 had 90% word recognition, and Family 2 had 100% speech discrimination at 80 dB. Prognosis is nevertheless uncertain because only one individual had detailed decade-long audiometric follow-up. Potential adverse prognostic factors—severe initial thresholds, truncating genotypes, aging, and noise—remain hypotheses rather than validated human predictors. (redfield2024pkhd1l1agene pages 5-8, redfield2024pkhd1l1agene pages 8-9, redfield2024pkhd1l1agene pages 11-13)
12. Treatment
Current management
There is no PKHD1L1-directed drug, approved gene therapy, pharmacogenomic recommendation, enzyme replacement, cell therapy, ASO, siRNA, immunotherapy, or disease-modifying surgery. Management follows general bilateral pediatric SNHL practice:
- Prompt audiologic confirmation and serial monitoring.
- Appropriately fitted hearing aids for aidable loss; Family 2 used bilateral aids.
- Speech-language, listening, sign-language/total-communication, family-centered, and educational support according to family goals.
- Remote-microphone systems and classroom accommodations.
- Cochlear-implant candidacy evaluation when optimized hearing aids provide inadequate access, based on severity, speech perception, anatomy, development, and local criteria.
- Vestibular therapy for independently documented vestibular disorders; Family 1’s BPPV responded to Epley repositioning. (redfield2024pkhd1l1agene pages 5-8, redfield2024pkhd1l1agene pages 8-9, fan2026internationalexpertconsensus pages 1-3)
Suggested NCIT intervention concepts, with identifiers verified against the current release before ingestion: hearing-aid device; cochlear implantation; audiologic rehabilitation; speech therapy; genetic counseling; preimplantation genetic testing; prenatal diagnosis.
Experimental treatments and recent developments
No PKHD1L1/DFNB124-specific interventional trial was found in ClinicalTrials.gov or the retrieved literature. Hereditary-hearing-loss gene-therapy trials showing early benefit chiefly target OTOF/DFNB9, not PKHD1L1, and cannot be generalized to this gene. PKHD1L1’s approximately 13-kb coding sequence and large extracellular protein would complicate conventional single-AAV replacement; no validated dual-vector, editing, or RNA strategy has been reported for DFNB124. (fan2026internationalexpertconsensus pages 1-3, li2024advancedmanagementof pages 3-4, redfield2024pkhd1l1agene pages 1-2)
13. Prevention
Primary prevention
Because the causal alleles are inherited, ordinary lifestyle changes cannot prevent genotype occurrence. Reproductive options after familial-variant confirmation include genetic counseling, partner testing when appropriate, prenatal diagnosis, and IVF with preimplantation genetic testing. These require nondirective counseling.
Avoidance of damaging noise and unnecessary ototoxic exposure is sensible tertiary risk reduction. The noise recommendation is biologically strengthened by mice but is not backed by human DFNB124 outcomes. Vaccination and infection prevention reduce some acquired hearing loss but do not prevent PKHD1L1-related disease. (strelkova2024pkhd1l1isrequired pages 1-2, strelkova2024pkhd1l1isrequireda pages 19-21)
Secondary and tertiary prevention
Universal newborn hearing screening, prompt diagnostic testing, early amplification/communication access, cascade testing, and regular audiometry can reduce delayed detection and functional complications. They do not reverse the genetic lesion. Hearing-conservation counseling, device optimization, educational accommodations, and speech-language support aim to preserve participation and development. A 2024 review of low- and middle-income programs found heterogeneous implementation and limited long-term intervention-outcome reporting, emphasizing access gaps. (hatzopoulos2024theotoacousticemissions pages 4-5, rajanbabu2024earlyhearingdetection pages 5-6)
14. Other species and natural disease
No naturally occurring veterinary PKHD1L1 deafness syndrome, breed predisposition, zoonotic transmission, or cross-species infectious risk was identified. DFNB124 is not transmissible.
Orthologous auditory function is conserved in Mus musculus (NCBI Taxonomy 10090) and duplicated paralogs occur in Danio rerio (7955). Human and mouse PKHD1L1 proteins share approximately 81.8% amino-acid identity excluding signal peptide, supporting comparative modeling. Exact NCBI Gene IDs and VBO terms should be imported from authoritative current database records rather than inferred here. (redfield2024pkhd1l1agene pages 8-9, redfield2024pkhd1l1agene pages 13-15)
15. Model organisms
Mouse
Two principal models are available: - hair-cell-enriched conditional knockout Pkhd1l1fl/fl;Atoh1-Cre+; - constitutive Pkhd1l1−/− knockout generated by germline Cre deletion.
The conditional allele deletes exon 10, producing a frameshift/premature stop. Both models develop progressive hearing loss. At early stages, planar polarity, gross anatomy, and FM1-43 uptake are relatively preserved. By about six weeks, basal outer-hair-cell bundles exhibit missing stereocilia and disorganization; ABR and DPOAE deficits begin at high frequencies and extend lower with age. Noise exposure produces persistent deficits not seen in controls. These models reproduce auditory dysfunction and progressive bundle pathology but differ from the mostly congenital human presentation and do not model individual human missense alleles. (strelkova2024pkhd1l1isrequired pages 12-13, wu2019pkhd1l1isa pages 8-9, strelkova2024pkhd1l1isrequired pages 10-11)
The 2019 abstract stated: “PKHD1L1-deficient mice lack the surface coat at the upper but not lower regions of stereocilia, and they develop progressive hearing loss.” DOI/URL: https://doi.org/10.1038/s41467-019-11712-w. (wu2019pkhd1l1isa pages 1-2)
The peer-reviewed November 2024 study concluded that deficient mice develop high-frequency loss progressing to lower frequencies and are unusually susceptible to permanent loss after moderate acoustic overexposure (Communications Biology 7:1423; DOI/URL: https://doi.org/10.1038/s42003-024-07121-5). (strelkova2024pkhd1l1isrequired pages 1-2)
Zebrafish
Combined disruption of pkhd1l1a and pkhd1l1b reduces auditory-evoked startle at six days post-fertilization, supporting evolutionarily conserved auditory function. Limitations include paralog redundancy and use of a behavioral surrogate rather than mammalian audiometry. The report was published in Journal of Neurogenetics in March 2023; DOI/URL: https://doi.org/10.1080/01677063.2023.2187792. (redfield2024pkhd1l1agene pages 11-13, redfield2024pkhd1l1agene pages 1-2)
In-vitro and computational systems
Recombinant mouse PKHD1L1 fragments enabled NanoDSF stability testing; HEK293 and HeLa minigene systems demonstrated Gly605Arg-associated exon skipping; AlphaFold2 supported structural hypotheses. These systems establish allele-level molecular effects but do not reproduce full-length protein trafficking, human hair-cell biomechanics, or clinical severity. (redfield2024pkhd1l1agene pages 9-11, redfield2024pkhd1l1agene pages 13-15)
Overall expert assessment
The disease–gene relationship is supported by moderate but still early evidence: four unrelated human families with recessive segregation, multiple variant classes, functional confirmation for selected variants, and concordant loss-of-function phenotypes in two vertebrate models. The strongest established phenotype is bilateral congenital/early-onset SNHL with variable severity. The strongest mechanistic model is failure of a developmentally transient stereocilia surface-coat protein to establish mechanically durable cochlear hair bundles. Major unresolved questions are penetrance, full allelic spectrum, genotype–phenotype correlation, human progression and noise susceptibility, binding partners and coat biochemistry, population prevalence, and suitability for molecular therapy. Extended case series, longitudinal audiology, variant-specific knock-in models, and full-length protein or human hair-cell systems are priorities. (redfield2024pkhd1l1agene pages 11-13, strelkova2024pkhd1l1isrequired pages 1-2, redfield2024pkhd1l1agene pages 1-2)
References
-
(strelkova2024pkhd1l1isrequired pages 1-2): Olga S. Strelkova, Richard T. Osgood, Chunjie J. Tian, Xinyuan Zhang, Evan Hale, Pedro De-la-Torre, Daniel M. Hathaway, and Artur A. Indzhykulian. Pkhd1l1 is required for stereocilia bundle maintenance, durable hearing function and resilience to noise exposure. Communications Biology, Nov 2024. URL: https://doi.org/10.1038/s42003-024-07121-5, doi:10.1038/s42003-024-07121-5. This article has 7 citations and is from a peer-reviewed journal.
-
(OpenTargets Search: autosomal recessive nonsyndromic hearing loss 124-PKHD1L1): Open Targets Query (autosomal recessive nonsyndromic hearing loss 124-PKHD1L1, 1 results). Buniello, A. et al. (2025). Open Targets Platform: facilitating therapeutic hypotheses building in drug discovery. Nucleic Acids Research.
-
(redfield2024pkhd1l1agene pages 1-2): Shelby E. Redfield, Pedro De-la-Torre, Mina Zamani, Hanjun Wang, Hina Khan, Tyler Morris, Gholamreza Shariati, Majid Karimi, Margaret A. Kenna, Go Hun Seo, Hongen Xu, Wei Lu, Sadaf Naz, Hamid Galehdari, Artur A. Indzhykulian, A. Eliot Shearer, and Barbara Vona. Pkhd1l1, a gene involved in the stereocilia coat, causes autosomal recessive nonsyndromic hearing loss. Human Genetics, 143:311-329, Mar 2024. URL: https://doi.org/10.1007/s00439-024-02649-2, doi:10.1007/s00439-024-02649-2. This article has 8 citations and is from a peer-reviewed journal.
-
(redfield2024pkhd1l1agene pages 5-8): Shelby E. Redfield, Pedro De-la-Torre, Mina Zamani, Hanjun Wang, Hina Khan, Tyler Morris, Gholamreza Shariati, Majid Karimi, Margaret A. Kenna, Go Hun Seo, Hongen Xu, Wei Lu, Sadaf Naz, Hamid Galehdari, Artur A. Indzhykulian, A. Eliot Shearer, and Barbara Vona. Pkhd1l1, a gene involved in the stereocilia coat, causes autosomal recessive nonsyndromic hearing loss. Human Genetics, 143:311-329, Mar 2024. URL: https://doi.org/10.1007/s00439-024-02649-2, doi:10.1007/s00439-024-02649-2. This article has 8 citations and is from a peer-reviewed journal.
-
(redfield2024pkhd1l1agene pages 8-9): Shelby E. Redfield, Pedro De-la-Torre, Mina Zamani, Hanjun Wang, Hina Khan, Tyler Morris, Gholamreza Shariati, Majid Karimi, Margaret A. Kenna, Go Hun Seo, Hongen Xu, Wei Lu, Sadaf Naz, Hamid Galehdari, Artur A. Indzhykulian, A. Eliot Shearer, and Barbara Vona. Pkhd1l1, a gene involved in the stereocilia coat, causes autosomal recessive nonsyndromic hearing loss. Human Genetics, 143:311-329, Mar 2024. URL: https://doi.org/10.1007/s00439-024-02649-2, doi:10.1007/s00439-024-02649-2. This article has 8 citations and is from a peer-reviewed journal.
-
(redfield2024pkhd1l1agene pages 11-13): Shelby E. Redfield, Pedro De-la-Torre, Mina Zamani, Hanjun Wang, Hina Khan, Tyler Morris, Gholamreza Shariati, Majid Karimi, Margaret A. Kenna, Go Hun Seo, Hongen Xu, Wei Lu, Sadaf Naz, Hamid Galehdari, Artur A. Indzhykulian, A. Eliot Shearer, and Barbara Vona. Pkhd1l1, a gene involved in the stereocilia coat, causes autosomal recessive nonsyndromic hearing loss. Human Genetics, 143:311-329, Mar 2024. URL: https://doi.org/10.1007/s00439-024-02649-2, doi:10.1007/s00439-024-02649-2. This article has 8 citations and is from a peer-reviewed journal.
-
(redfield2024pkhd1l1agene pages 9-11): Shelby E. Redfield, Pedro De-la-Torre, Mina Zamani, Hanjun Wang, Hina Khan, Tyler Morris, Gholamreza Shariati, Majid Karimi, Margaret A. Kenna, Go Hun Seo, Hongen Xu, Wei Lu, Sadaf Naz, Hamid Galehdari, Artur A. Indzhykulian, A. Eliot Shearer, and Barbara Vona. Pkhd1l1, a gene involved in the stereocilia coat, causes autosomal recessive nonsyndromic hearing loss. Human Genetics, 143:311-329, Mar 2024. URL: https://doi.org/10.1007/s00439-024-02649-2, doi:10.1007/s00439-024-02649-2. This article has 8 citations and is from a peer-reviewed journal.
-
(redfield2024pkhd1l1agene pages 13-15): Shelby E. Redfield, Pedro De-la-Torre, Mina Zamani, Hanjun Wang, Hina Khan, Tyler Morris, Gholamreza Shariati, Majid Karimi, Margaret A. Kenna, Go Hun Seo, Hongen Xu, Wei Lu, Sadaf Naz, Hamid Galehdari, Artur A. Indzhykulian, A. Eliot Shearer, and Barbara Vona. Pkhd1l1, a gene involved in the stereocilia coat, causes autosomal recessive nonsyndromic hearing loss. Human Genetics, 143:311-329, Mar 2024. URL: https://doi.org/10.1007/s00439-024-02649-2, doi:10.1007/s00439-024-02649-2. This article has 8 citations and is from a peer-reviewed journal.
-
(wu2019pkhd1l1isa pages 1-2): Xudong Wu, Maryna V. Ivanchenko, Hoor Al Jandal, Marcelo Cicconet, Artur A. Indzhykulian, and David P. Corey. Pkhd1l1 is a coat protein of hair-cell stereocilia and is required for normal hearing. Nature Communications, Aug 2019. URL: https://doi.org/10.1038/s41467-019-11712-w, doi:10.1038/s41467-019-11712-w. This article has 50 citations and is from a highest quality peer-reviewed journal.
-
(strelkova2024pkhd1l1isrequired pages 10-11): Olga S. Strelkova, Richard T. Osgood, Chunjie J. Tian, Xinyuan Zhang, Evan Hale, Pedro De-la-Torre, Daniel M. Hathaway, and Artur A. Indzhykulian. Pkhd1l1 is required for stereocilia bundle maintenance, durable hearing function and resilience to noise exposure. Communications Biology, Nov 2024. URL: https://doi.org/10.1038/s42003-024-07121-5, doi:10.1038/s42003-024-07121-5. This article has 7 citations and is from a peer-reviewed journal.
-
(strelkova2024pkhd1l1isrequired pages 12-13): Olga S. Strelkova, Richard T. Osgood, Chunjie J. Tian, Xinyuan Zhang, Evan Hale, Pedro De-la-Torre, Daniel M. Hathaway, and Artur A. Indzhykulian. Pkhd1l1 is required for stereocilia bundle maintenance, durable hearing function and resilience to noise exposure. Communications Biology, Nov 2024. URL: https://doi.org/10.1038/s42003-024-07121-5, doi:10.1038/s42003-024-07121-5. This article has 7 citations and is from a peer-reviewed journal.
-
(strelkova2024pkhd1l1isrequireda pages 19-21): Olga S. Strelkova, Richard T. Osgood, Chunjie J. Tian, Xinyuan Zhang, Evan Hale, Pedro De-la-Torre, Daniel M. Hathaway, and Artur A. Indzhykulian. Pkhd1l1 is required for stereocilia bundle maintenance, durable hearing function and resilience to noise exposure. bioRxiv, Mar 2024. URL: https://doi.org/10.1101/2024.02.29.582786, doi:10.1101/2024.02.29.582786. This article has 0 citations.
-
(fan2026internationalexpertconsensus pages 1-3): Xintai Fan, Ziwen Gao, Jiake Zhong, Yuxin Chen, Xiaoyun Chen, Lukas D. Landegger, Tobias Moser, Fan-Gang Zeng, Yu Sun, Xin Jin, Robert Nash, Wade W. Chien, Dan Jiang, John H. Greinwald, Manohar Bance, Manuel Manrique Rodríguez, Sang-Yeon Lee, Guodong Feng, Haidi Yang, Chen-Chi Wu, Lei Xu, Wei Yuan, Yong Feng, Yu Zhao, Barbara Vona, Nicola Strenzke, Dirk Beutner, Nikul Amin, James Arwyn-Jones, Deepak Chandrasekeharan, Dazhi Shi, Di Zhang, Jianming Yang, Jieyu Qi, Qin Wang, Yanbo Yin, Yen-Fu Cheng, Yong Tao, Yongfu Yu, Daqi Wang, Luoying Jiang, Luo Guo, Liheng Chen, Xiaoting Cheng, Chong Cui, Jun Lv, Shuang Han, Wuqing Wang, Yongxin Li, Xia Gao, Xue Zhong Liu, Dingjun Zha, Haibo Shi, Bing Chen, Qiuju Wang, Huijun Yuan, Shiming Yang, Shankai Yin, Hao Wu, Zhengmin Wang, Huawei Li, Jay T. Rubinstein, Lawrence R. Lustig, Renjie Chai, Zheng-Yi Chen, and Yilai Shu. International expert consensus on gene therapy for hereditary hearing loss: based on clinical trials. Med, 7(1):100886, Jan 2026. URL: https://doi.org/10.1016/j.medj.2025.100886, doi:10.1016/j.medj.2025.100886. This article has 13 citations and is from a domain leading peer-reviewed journal.
-
(li2024advancedmanagementof pages 3-4): Shuna Li, Ling Lu, Jun Yang, and Maoli Duan. Advanced management of hearing loss: a comprehensive review of the special issue. Dec 2024. URL: https://doi.org/10.3390/jcm13237409, doi:10.3390/jcm13237409. This article has 2 citations.
-
(redfield2024pkhd1l1agene pages 2-4): Shelby E. Redfield, Pedro De-la-Torre, Mina Zamani, Hanjun Wang, Hina Khan, Tyler Morris, Gholamreza Shariati, Majid Karimi, Margaret A. Kenna, Go Hun Seo, Hongen Xu, Wei Lu, Sadaf Naz, Hamid Galehdari, Artur A. Indzhykulian, A. Eliot Shearer, and Barbara Vona. Pkhd1l1, a gene involved in the stereocilia coat, causes autosomal recessive nonsyndromic hearing loss. Human Genetics, 143:311-329, Mar 2024. URL: https://doi.org/10.1007/s00439-024-02649-2, doi:10.1007/s00439-024-02649-2. This article has 8 citations and is from a peer-reviewed journal.
-
(redfield2024pkhd1l1agene media e885a2a5): Shelby E. Redfield, Pedro De-la-Torre, Mina Zamani, Hanjun Wang, Hina Khan, Tyler Morris, Gholamreza Shariati, Majid Karimi, Margaret A. Kenna, Go Hun Seo, Hongen Xu, Wei Lu, Sadaf Naz, Hamid Galehdari, Artur A. Indzhykulian, A. Eliot Shearer, and Barbara Vona. Pkhd1l1, a gene involved in the stereocilia coat, causes autosomal recessive nonsyndromic hearing loss. Human Genetics, 143:311-329, Mar 2024. URL: https://doi.org/10.1007/s00439-024-02649-2, doi:10.1007/s00439-024-02649-2. This article has 8 citations and is from a peer-reviewed journal.
-
(redfield2024pkhd1l1agene pages 15-16): Shelby E. Redfield, Pedro De-la-Torre, Mina Zamani, Hanjun Wang, Hina Khan, Tyler Morris, Gholamreza Shariati, Majid Karimi, Margaret A. Kenna, Go Hun Seo, Hongen Xu, Wei Lu, Sadaf Naz, Hamid Galehdari, Artur A. Indzhykulian, A. Eliot Shearer, and Barbara Vona. Pkhd1l1, a gene involved in the stereocilia coat, causes autosomal recessive nonsyndromic hearing loss. Human Genetics, 143:311-329, Mar 2024. URL: https://doi.org/10.1007/s00439-024-02649-2, doi:10.1007/s00439-024-02649-2. This article has 8 citations and is from a peer-reviewed journal.
-
(strelkova2024pkhd1l1isrequired pages 5-6): Olga S. Strelkova, Richard T. Osgood, Chunjie J. Tian, Xinyuan Zhang, Evan Hale, Pedro De-la-Torre, Daniel M. Hathaway, and Artur A. Indzhykulian. Pkhd1l1 is required for stereocilia bundle maintenance, durable hearing function and resilience to noise exposure. Communications Biology, Nov 2024. URL: https://doi.org/10.1038/s42003-024-07121-5, doi:10.1038/s42003-024-07121-5. This article has 7 citations and is from a peer-reviewed journal.
-
(wu2019pkhd1l1isa pages 8-9): Xudong Wu, Maryna V. Ivanchenko, Hoor Al Jandal, Marcelo Cicconet, Artur A. Indzhykulian, and David P. Corey. Pkhd1l1 is a coat protein of hair-cell stereocilia and is required for normal hearing. Nature Communications, Aug 2019. URL: https://doi.org/10.1038/s41467-019-11712-w, doi:10.1038/s41467-019-11712-w. This article has 50 citations and is from a highest quality peer-reviewed journal.
-
(strelkova2024pkhd1l1isrequired pages 3-4): Olga S. Strelkova, Richard T. Osgood, Chunjie J. Tian, Xinyuan Zhang, Evan Hale, Pedro De-la-Torre, Daniel M. Hathaway, and Artur A. Indzhykulian. Pkhd1l1 is required for stereocilia bundle maintenance, durable hearing function and resilience to noise exposure. Communications Biology, Nov 2024. URL: https://doi.org/10.1038/s42003-024-07121-5, doi:10.1038/s42003-024-07121-5. This article has 7 citations and is from a peer-reviewed journal.
-
(strelkova2024pkhd1l1isrequireda pages 3-5): Olga S. Strelkova, Richard T. Osgood, Chunjie J. Tian, Xinyuan Zhang, Evan Hale, Pedro De-la-Torre, Daniel M. Hathaway, and Artur A. Indzhykulian. Pkhd1l1 is required for stereocilia bundle maintenance, durable hearing function and resilience to noise exposure. bioRxiv, Mar 2024. URL: https://doi.org/10.1101/2024.02.29.582786, doi:10.1101/2024.02.29.582786. This article has 0 citations.
-
(rajanbabu2024earlyhearingdetection pages 5-6): Keerthana Rajanbabu, Deepashree Joshi B, Vidya Ramkumar, Hannah Kuper, and Ramya Vaidyanath. Early hearing detection and intervention programmes for neonates, infants and children in non-asian low-income and middle-income countries: a systematic review. BMJ Paediatrics Open, 8(1):e002794, Nov 2024. URL: https://doi.org/10.1136/bmjpo-2024-002794, doi:10.1136/bmjpo-2024-002794. This article has 14 citations and is from a peer-reviewed journal.
-
(guo2024globalregionaland pages 11-11): Zhifeng Guo, Wangquan Ji, Ping Song, Jingli Zhao, Mengqing Yan, Xianan Zou, Fanghui Bai, Yu Wu, Zhe Guo, and Linlin Song. Global, regional, and national burden of hearing loss in children and adolescents, 1990–2021: a systematic analysis from the global burden of disease study 2021. BMC Public Health, Sep 2024. URL: https://doi.org/10.1186/s12889-024-20010-0, doi:10.1186/s12889-024-20010-0. This article has 70 citations and is from a peer-reviewed journal.
-
(hatzopoulos2024theotoacousticemissions pages 4-5): Stavros Hatzopoulos, Ludovica Cardinali, Piotr Henryk Skarżyński, and Giovanna Zimatore. The otoacoustic emissions in the universal neonatal hearing screening: an update on the european data (2004 to 2024). Children, 11:1276, Oct 2024. URL: https://doi.org/10.3390/children11111276, doi:10.3390/children11111276. This article has 12 citations.
Artifacts
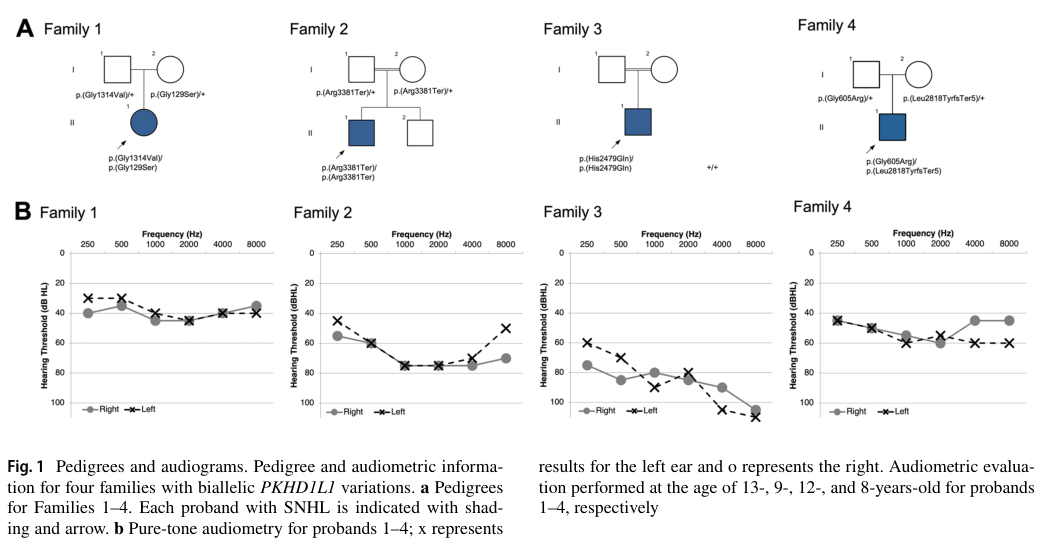
Reference Validation
Checked with linkml-reference-validator 0.2.1.
| Outcome | Count |
|---|---|
| References checked | 11 |
| Resolved | 11 |
| Unresolved (possible confabulation) | 0 |
| Unverifiable | 0 |
| References weighed for topical relevance | 11 |
| On topic | 4 |
| Off topic | 0 |
All extracted references resolved successfully.
Term Validation
Checked with linkml-term-validator 0.4.5, through the ols: adapter.
| Outcome | Count |
|---|---|
| Terms checked | 22 |
| Resolved | 21 |
| Unresolved (possible confabulation) | 0 |
| Obsolete | 1 |
| Unverifiable | 0 |
| Terms whose name was checked | 1 |
| Terms named correctly | 0 |
| Terms named as a different term | 1 |
Terms the report names something else
These identifiers resolve, so nothing about them looks wrong, and the ontology calls them something unrelated to what the report calls them. That usually means the identifier is not the one the sentence needs:
MONDO:0968981(3 mentions) - the report calls it "if available"; MONDO calls it autosomal recessive nonsyndromic hearing loss 124
Obsolete terms
These terms are real but deprecated. Citing one is not a fabrication; it does mean the report is naming something the ontology has retired:
GO:0032429(obsolete regulation of phospholipase A2 activity) (1 mention)