Autosomal Dominant Nonsyndromic Hearing Loss 4B
DFNA4B is CEACAM16-associated autosomal dominant nonsyndromic sensorineural hearing loss, usually postlingual and progressive. High-frequency onset is common in reported families, but a de novo case had a flat audiogram and no progression during one year of follow-up. Tinnitus varies between families and can precede measurable hearing loss. CEACAM16 is a secreted tectorial-membrane glycoprotein. Later mouse localization studies identify nonsensory supporting and limbal cells as sources, correcting the initial outer-hair-cell expression interpretation. Dominant missense variants have allele-specific biochemical effects; a universal dominant-negative mechanism has not been established. Null mice demonstrate matrix disorganization, altered material properties and auditory dysfunction, but the tissue consequences in human heterozygous missense carriers remain inferred. Biallelic loss-of-function variants cause the separate recessive disorder DFNB113, which can overlap clinically with DFNA4B.
Ask OpenScientist
Ask a research question about Autosomal Dominant Nonsyndromic Hearing Loss 4B. OpenScientist will conduct autonomous deep research using the Disorder Mechanisms Knowledge Base and PubMed literature (typically 10-30 minutes).
Do not include personal health information in your question. Questions and results are cached in your browser's local storage.
Classifications
Inheritance
1Show evidence (3 references)
Mechanistic Hypotheses
1Discussions and Knowledge Gaps
2Pathophysiology
5Show evidence (3 references)
Show evidence (3 references)
Show evidence (4 references)
Show evidence (1 reference)
Show evidence (2 references)
Pathograph
Phenotypes
5Ear 4
Show evidence (4 references)
Show evidence (2 references)
Show evidence (3 references)
Show evidence (2 references)
Nervous System 1
Show evidence (1 reference)
Genetic Associations
1Show evidence (6 references)
External Assertions
1Medical Actions
4Show evidence (3 references)
Show evidence (1 reference)
Show evidence (2 references)
Show evidence (2 references)
Diagnosis
1Show evidence (3 references)
Animal Models
2Show evidence (1 reference)
Show evidence (1 reference)
Show evidence (1 reference)
Show evidence (1 reference)
Show evidence (1 reference)
Show evidence (1 reference)
Show evidence (1 reference)
Show evidence (1 reference)
Source YAML
click to showname: Autosomal Dominant Nonsyndromic Hearing Loss 4B
category: Mendelian
creation_date: '2026-09-14T00:00:00Z'
synonyms:
- DFNA4B
- deafness, autosomal dominant 4B
- CEACAM16-related autosomal dominant hearing loss
description: DFNA4B is CEACAM16-associated autosomal dominant nonsyndromic sensorineural hearing loss, usually postlingual and progressive. High-frequency onset is common in reported families, but a de novo case had a flat audiogram and no progression during one year of follow-up. Tinnitus varies between families and can precede measurable hearing loss. CEACAM16 is a secreted tectorial-membrane glycoprotein. Later mouse localization studies identify nonsensory supporting and limbal cells as sources, correcting the initial outer-hair-cell expression interpretation. Dominant missense variants have allele-specific biochemical effects; a universal dominant-negative mechanism has not been established. Null mice demonstrate matrix disorganization, altered material properties and auditory dysfunction, but the tissue consequences in human heterozygous missense carriers remain inferred. Biallelic loss-of-function variants cause the separate recessive disorder DFNB113, which can overlap clinically with DFNA4B.
disease_term:
preferred_term: autosomal dominant nonsyndromic hearing loss 4B
term:
id: MONDO:0013823
label: autosomal dominant nonsyndromic hearing loss 4B
parents:
- Autosomal Dominant Nonsyndromic Hearing Loss
references:
- reference: PMID:25589040
title: Exome sequencing identifies a novel CEACAM16 mutation associated with autosomal dominant nonsyndromic hearing loss DFNA4B in a Chinese family.
- reference: PMID:26648831
title: A Novel de novo Mutation in CEACAM16 Associated with Postlingual Hearing Impairment.
- reference: url:https://www.ncbi.nlm.nih.gov/sites/books/NBK1434/?report=reader
title: Genetic Hearing Loss Overview - GeneReviews® - NCBI Bookshelf
tags:
- GeneReviews
- reference: PMID:21368133
title: Carcinoembryonic antigen-related cell adhesion molecule 16 interacts with alpha-tectorin and is mutated in autosomal dominant hearing loss (DFNA4).
- reference: PMID:30514912
title: Further evidence for loss-of-function mutations in the CEACAM16 gene causing nonsyndromic autosomal recessive hearing loss in humans.
- reference: PMID:33040498
title: '[Identification of a novel mutation of CEACAM16 gene in a Chinese family with autosomal dominant nonsyndromic hearing loss].'
- reference: PMID:29703829
title: 'Old gene, new phenotype: splice-altering variants in CEACAM16 cause recessive non-syndromic hearing impairment.'
- reference: url:https://search.clinicalgenome.org/kb/gene-validity/CGGV:assertion_e25cadf1-752a-495d-aaf2-30565519518a-2022-09-21T160000.000Z
title: curation results for Gene-Disease Validity
- reference: PMID:35292975
title: A novel missense variant in CEACAM16 gene causes autosomal dominant nonsyndromic hearing loss.
- reference: PMID:25080593
title: Loss of the tectorial membrane protein CEACAM16 enhances spontaneous, stimulus-frequency, and transiently evoked otoacoustic emissions.
- reference: url:https://pmc.ncbi.nlm.nih.gov/articles/PMC4115139/?report=xml
title: Loss of the Tectorial Membrane Protein CEACAM16 Enhances Spontaneous, Stimulus-Frequency, and Transiently Evoked Otoacoustic Emissions - PMC
- reference: PMID:34555361
title: Age-related degradation of tectorial membrane dynamics with loss of CEACAM16.
- reference: url:https://pmc.ncbi.nlm.nih.gov/articles/PMC4662267/?report=xml
title: A Novel de novo Mutation in CEACAM16 Associated with Postlingual Hearing Impairment - PMC
- reference: PMID:39157884
title: Auditory Phenotype of a Novel Missense Variant in the CEACAM16 Gene in a Large Russian Family With Autosomal Dominant Nonsyndromic Hearing Loss.
- reference: PMID:31249509
title: Accelerated Age-Related Degradation of the Tectorial Membrane in the Ceacam16(βgal/βgal) Null Mutant Mouse, a Model for Late-Onset Human Hereditary Deafness DFNB113.
- reference: url:https://pmc.ncbi.nlm.nih.gov/articles/PMC3381124/?pdf=1
title: Loss of Mammal-specific Tectorial Membrane Component Carcinoembryonic Antigen Cell Adhesion Molecule 16 (CEACAM16) Leads to Hearing Impairment at Low and High Frequencies - PMC
- reference: PMID:22544735
title: Loss of mammal-specific tectorial membrane component carcinoembryonic antigen cell adhesion molecule 16 (CEACAM16) leads to hearing impairment at low and high frequencies.
inheritance:
- name: Autosomal dominant
description: Heterozygous CEACAM16 missense variants segregating with hearing loss in multigenerational families, plus at least one de novo case. Biallelic loss-of-function variants in the same gene cause a separate recessive disease and are not part of this entry.
inheritance_term:
preferred_term: Autosomal dominant inheritance
term:
id: HP:0000006
label: Autosomal dominant inheritance
evidence:
- reference: PMID:25589040
reference_title: Exome sequencing identifies a novel CEACAM16 mutation associated with autosomal dominant nonsyndromic hearing loss DFNA4B in a Chinese family.
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: Sanger sequencing verified that the mutation co-segregated with hearing loss in the family and that it was not present in 200 unrelated control subjects with matched ancestry.
explanation: Co-segregation across a five-generation family with absence in matched controls, which is the genetic argument for dominant inheritance of a heterozygous missense allele.
quote_role: PRIMARY_RESULT
directness: DIRECT
- reference: PMID:26648831
reference_title: A Novel de novo Mutation in CEACAM16 Associated with Postlingual Hearing Impairment.
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: we discovered a de novo CEACAM16 mutation (c.1094T>G, p.Leu365Arg) as the sole likely pathogenic variant
explanation: A de novo dominant allele in a trio without family history, which is independent evidence for dominance that does not rest on segregation.
quote_role: PRIMARY_RESULT
directness: DIRECT
- reference: url:https://www.ncbi.nlm.nih.gov/sites/books/NBK1434/?report=reader
reference_title: Genetic Hearing Loss Overview - GeneReviews® - NCBI Bookshelf
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: REVIEW_SYNTHESIS
directness: DIRECT
snippet: Individuals with autosomal dominant hearing loss have a 50% chance of transmitting the pathogenic variant to each child.
explanation: General dominant-inheritance counseling; the risk refers to offspring, including offspring of a proband with a de novo variant.
genetic:
- name: CEACAM16
gene_term:
preferred_term: CEACAM16
term:
id: hgnc:31948
label: CEACAM16
relationship_type: CAUSATIVE
notes: Reported dominant missense alleles include c.418A>C (p.Thr140Pro), c.418A>G (p.Thr140Ala), c.419C>T (p.Thr140Ile), c.505G>A (p.Gly169Arg), c.763A>G (p.Arg255Gly), and de novo c.1094T>G (p.Leu365Arg). The Chinese p.Thr140Ala family adds onset in the twenties. These reports do not establish equal pathogenic certainty or a shared molecular mechanism for every allele. ClinGen retained Moderate gene-disease validity in September 2022. Its evidence table gave a heterozygous frameshift case with onset at 50 years zero points, so this entry does not treat all heterozygous truncating variants as causal. Hearing-normal human carriers in recessive splice-variant families argue against simple haploinsufficiency, without proving dominant-negative action. Penetrance cannot be quantified from the small, variably ascertained families; the Russian report is internally inconsistent about the genotype of hearing-normal child V-11. A meningitis/aminoglycoside-associated noncarrier in the p.Gly169Arg family and age-related hearing loss in the noncarrier father of the p.Thr140Ala family illustrate phenocopies.
evidence:
- reference: PMID:21368133
reference_title: Carcinoembryonic antigen-related cell adhesion molecule 16 interacts with alpha-tectorin and is mutated in autosomal dominant hearing loss (DFNA4).
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: we show that mutation of CEACAM16 leads to autosomal dominant nonsyndromic deafness (ADNSHL) at the autosomal dominant hearing loss (DFNA4) locus
explanation: The original assignment of dominant nonsyndromic deafness at the DFNA4 locus to CEACAM16.
quote_role: PRIMARY_RESULT
directness: DIRECT
- reference: PMID:30514912
reference_title: Further evidence for loss-of-function mutations in the CEACAM16 gene causing nonsyndromic autosomal recessive hearing loss in humans.
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: Our results give additional support that loss-of-function variants in CEACAM16 cause autosomal recessive hearing loss in humans.
explanation: Establishes the allelic recessive disease, which is what makes the dominant missense mechanism a distinct claim rather than the severe end of one spectrum.
quote_role: PRIMARY_RESULT
directness: DIRECT
- reference: PMID:33040498
reference_title: '[Identification of a novel mutation of CEACAM16 gene in a Chinese family with autosomal dominant nonsyndromic hearing loss].'
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: The affected family members appeared to have developed a high-frequency hearing loss with the onset of twenties.
explanation: Clinical segregation report for p.Thr140Ala; no functional assay established the effect of this substitution.
- reference: PMID:29703829
reference_title: 'Old gene, new phenotype: splice-altering variants in CEACAM16 cause recessive non-syndromic hearing impairment.'
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: Individuals homozygous for either of these variants have progressive hearing loss, whereas heterozygous carriers have normal hearing (Figure 1).
explanation: Human recessive splice-variant families distinguish inheritance; the hearing-loss phenotype can overlap with DFNA4B.
- reference: url:https://search.clinicalgenome.org/kb/gene-validity/CGGV:assertion_e25cadf1-752a-495d-aaf2-30565519518a-2022-09-21T160000.000Z
reference_title: curation results for Gene-Disease Validity
supports: SUPPORT
evidence_source: OTHER
quote_role: REVIEW_SYNTHESIS
directness: DIRECT
snippet: In summary, there is moderate evidence to support this gene-disease relationship.
explanation: ClinGen Hearing Loss GCEP assessment dated 2022-09-21.
- reference: url:https://search.clinicalgenome.org/kb/gene-validity/CGGV:assertion_e25cadf1-752a-495d-aaf2-30565519518a-2022-09-21T160000.000Z
reference_title: curation results for Gene-Disease Validity
supports: SUPPORT
evidence_source: OTHER
quote_role: REVIEW_SYNTHESIS
directness: DIRECT
snippet: Other variants associated with AD hearing loss are missense variants and this age of onset is not consistent with the other cases.
explanation: Reason the late-onset frameshift case was assigned zero points; not evidence for universal dominant loss of function.
pathophysiology:
- name: Heterozygous CEACAM16 Missense Alteration
description: Reported DFNA4B alleles alter conserved residues of the secreted CEACAM16 protein. Variant location and segregation establish the initiating genetic lesion; altered folding, stability and poisoning of wild-type protein are not uniformly demonstrated.
biological_scale: MOLECULAR
genetic_context:
variant_origin: GERMLINE
zygosity: HETEROZYGOUS
functional_impact: Allele-specific biochemical effects; dominant disease mechanism unresolved.
downstream:
- target: Altered CEACAM16 Secretion
causal_link_type: DIRECT
description: Secretion assays support this link for p.Gly169Arg and p.Arg255Gly; it is not established for every reported allele.
evidence:
- reference: PMID:25589040
reference_title: Exome sequencing identifies a novel CEACAM16 mutation associated with autosomal dominant nonsyndromic hearing loss DFNA4B in a Chinese family.
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: a heterozygous missense mutation, c.505G>A (p.G169R) in exon 3 of the CEACAM16 gene
explanation: A representative dominant allele, heterozygous and missense, which is what this node describes.
quote_role: PRIMARY_RESULT
directness: DIRECT
- reference: PMID:26648831
reference_title: A Novel de novo Mutation in CEACAM16 Associated with Postlingual Hearing Impairment.
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: affects a highly conserved amino acid in the most strongly conserved CEACAM16 N2 domain
explanation: The de novo p.Leu365Arg allele affects a conserved N2-domain residue; this does not itself demonstrate abnormal folding.
quote_role: PRIMARY_RESULT
directness: DIRECT
- reference: url:https://search.clinicalgenome.org/kb/gene-validity/CGGV:assertion_e25cadf1-752a-495d-aaf2-30565519518a-2022-09-21T160000.000Z
reference_title: curation results for Gene-Disease Validity
supports: SUPPORT
evidence_source: OTHER
quote_role: REVIEW_SYNTHESIS
directness: DIRECT
snippet: The mechanism of pathogenicity is unknown.
explanation: Expert synthesis limits any universal dominant-negative or gain-of-function classification.
- name: Altered CEACAM16 Secretion
description: HEK293T assays show reduced secretion and altered extracellular oligomer recovery for p.Gly169Arg, but increased intracellular and extracellular protein levels for p.Arg255Gly despite lower transcript abundance. These measurements do not establish a common trafficking defect or interference with wild-type matrix assembly. ClinGen reports that p.Thr140Pro had preserved localization, secretion and TECTA binding in the assays reviewed.
biological_scale: MOLECULAR
downstream:
- target: Destabilized Tectorial Membrane Matrix Assembly
causal_link_type: INDIRECT_UNKNOWN_INTERMEDIATES
description: Altered protein availability may disturb matrix organization; neither secretion study directly measured mutant/wild-type matrix assembly.
hypothesis_groups:
- dfna4b_missense_to_matrix
evidence:
- reference: PMID:25589040
reference_title: Exome sequencing identifies a novel CEACAM16 mutation associated with autosomal dominant nonsyndromic hearing loss DFNA4B in a Chinese family.
supports: SUPPORT
evidence_source: IN_VITRO
snippet: our studies in transfected HEK293T cells show that the secretion efficacy of the mutant CEACAM16 is much lower than that of the wild type, suggesting a deleterious effect of the sequence variant
explanation: 'The reduced-secretion arm, measured in transfected cells. Note the authors'' own hedge: they infer a deleterious effect rather than demonstrating a matrix consequence.'
quote_role: PRIMARY_RESULT
directness: DIRECT
- reference: PMID:35292975
reference_title: A novel missense variant in CEACAM16 gene causes autosomal dominant nonsyndromic hearing loss.
supports: SUPPORT
evidence_source: IN_VITRO
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: The amount of mutant CEACAM16 protein was significantly higher than that of WT protein (p < 0.01)
explanation: ELISA of cell lysate and medium supports increased protein levels for p.Arg255Gly, opposite to p.Gly169Arg secretion; this is not a demonstrated dominant-negative mechanism.
- reference: url:https://search.clinicalgenome.org/kb/gene-validity/CGGV:assertion_e25cadf1-752a-495d-aaf2-30565519518a-2022-09-21T160000.000Z
reference_title: curation results for Gene-Disease Validity
supports: SUPPORT
evidence_source: OTHER
quote_role: REVIEW_SYNTHESIS
directness: DIRECT
snippet: All protein functions examined were similar to WT. There was no alteration in localization, binding with TECTA or secretion.
explanation: ClinGen summary of p.Thr140Pro assays limits generalization of altered secretion to that allele.
mechanism_confidence: PROVISIONAL
- name: Destabilized Tectorial Membrane Matrix Assembly
description: CEACAM16 interacts with alpha- and beta-tectorin in transfected-cell assays. Homozygous null mice have reduced beta-tectorin, deficient striated-sheet organization and absent Hensen's stripe in regions where it normally occurs. Progressive matrix loss is biased toward the apical, low-frequency cochlea. These experiments support a matrix-maintenance role, but the same structural lesion has not been demonstrated in human dominant missense carriers.
biological_scale: TISSUE
biological_processes:
- preferred_term: extracellular matrix organization
term:
id: GO:0030198
label: extracellular matrix organization
modifier: ABNORMAL
cellular_components:
- preferred_term: extracellular matrix
term:
id: GO:0031012
label: extracellular matrix
locations:
- preferred_term: tectorial membrane of cochlea
term:
id: UBERON:0002233
label: tectorial membrane of cochlea
downstream:
- target: Altered Tectorial Membrane Mechanics
causal_link_type: DIRECT
description: Null-mouse structural and ex vivo mechanical measurements support this relation; translation to dominant missense disease is provisional.
hypothesis_groups:
- dfna4b_missense_to_matrix
evidence:
- reference: PMID:25080593
reference_title: Loss of the tectorial membrane protein CEACAM16 enhances spontaneous, stimulus-frequency, and transiently evoked otoacoustic emissions.
supports: SUPPORT
evidence_source: MODEL_ORGANISM
snippet: In the absence of CEACAM16, TECTB levels are reduced, a clearly defined striated-sheet matrix does not develop, and Hensen's stripe, a prominent feature in the basal two-thirds of the TM in WT mice, is absent.
explanation: The three structural consequences of losing CEACAM16, measured in the null mouse tectorial membrane.
quote_role: PRIMARY_RESULT
directness: DIRECT
- reference: PMID:25080593
reference_title: Loss of the tectorial membrane protein CEACAM16 enhances spontaneous, stimulus-frequency, and transiently evoked otoacoustic emissions.
supports: SUPPORT
evidence_source: IN_VITRO
snippet: CEACAM16 is also shown to interact with TECTB, indicating that it may stabilize interactions between TECTA and TECTB.
explanation: Interaction inferred from coimmunoprecipitation in transfected HEK293-EBNA cells, not directly measured tectorin cross-linking in human tissue.
quote_role: PRIMARY_RESULT
directness: DIRECT
- reference: url:https://pmc.ncbi.nlm.nih.gov/articles/PMC4115139/?report=xml
reference_title: Loss of the Tectorial Membrane Protein CEACAM16 Enhances Spontaneous, Stimulus-Frequency, and Transiently Evoked Otoacoustic Emissions - PMC
supports: SUPPORT
evidence_source: MODEL_ORGANISM
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: Strong X-Gal staining is seen in epithelial cells of the spiral limbus and inner sulcus, border cells, inner and outer pillar cells, and Deiters' cells. Staining is not detected within the IHCs or OHCs
explanation: Reporter localization identifies nonsensory source cells; Deiters cells are one of several sources.
- reference: url:https://pmc.ncbi.nlm.nih.gov/articles/PMC3381124/?pdf=1
reference_title: Loss of Mammal-specific Tectorial Membrane Component Carcinoembryonic Antigen Cell Adhesion Molecule 16 (CEACAM16) Leads to Hearing Impairment at Low and High Frequencies - PMC
supports: SUPPORT
evidence_source: MODEL_ORGANISM
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: CEACAM16 was found in interdental and Deiters cells and was deposited in the tectorial membrane of the cochlea between postnatal days 12 and 15, when hearing starts in mice.
explanation: The independent localization study supports interdental and Deiters cells as sources of CEACAM16.
cell_types:
- preferred_term: Deiter's cell
term:
id: CL:0000635
label: Deiter's cell
- preferred_term: internal pillar cell of cochlea
term:
id: CL:0002163
label: internal pillar cell of cochlea
- preferred_term: external pillar cell of cochlea
term:
id: CL:0002164
label: external pillar cell of cochlea
- preferred_term: border cell of cochlea
term:
id: CL:0002168
label: border cell of cochlea
- preferred_term: interdental cell of cochlea
term:
id: CL:0002172
label: interdental cell of cochlea
mechanism_confidence: PROVISIONAL
- name: Altered Tectorial Membrane Mechanics
description: 'Isolated tectorial membranes from adult Ceacam16-null mice have slower, shorter-range traveling waves and reduced stiffness and viscosity relative to adult wild-type tissue. This directly supports altered material properties. It does not establish progressive detachment from hair bundles: the earlier BALB/c model retained attachment during tissue preparation. Human missense-carrier mechanics remain unmeasured.'
biological_scale: TISSUE
locations:
- preferred_term: tectorial membrane of cochlea
term:
id: UBERON:0002233
label: tectorial membrane of cochlea
downstream:
- target: Destabilized Cochlear Amplification
causal_link_type: INDIRECT_UNKNOWN_INTERMEDIATES
description: Changes in membrane mechanics can alter cochlear feedback, but wave measurements alone do not account for every age-dependent emission or threshold effect.
hypothesis_groups:
- dfna4b_missense_to_matrix
evidence:
- reference: PMID:34555361
reference_title: Age-related degradation of tectorial membrane dynamics with loss of CEACAM16.
supports: SUPPORT
evidence_source: IN_VITRO
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: Here, we show that TM stiffness and viscosity are significantly reduced in adult mice that lack functional CEACAM16 relative to age-matched wild-type controls.
explanation: Measurements on isolated mouse tissue establish material-property changes. The experiment used adult null and wild-type tissue plus two juvenile null preparations, without a juvenile wild-type group.
notes: The 2012 study describes preparation-associated contraction/detachment in wild-type tissue while null membranes often remained attached. That observation is not evidence of faster in vivo detachment in CEACAM16 deficiency.
mechanism_confidence: PROVISIONAL
- name: Destabilized Cochlear Amplification
description: Young C57BL/6J beta-galactosidase-null mice show enlarged stimulus-frequency and transient emissions and frequent spontaneous emissions despite largely preserved thresholds. With aging, emissions decline and thresholds increase; genotype-dependent matrix loss and sensitivity deficits are especially evident in low-frequency regions. Similar hair-cell complements between genotypes indicate an additional matrix contribution, not complete absence of age-related hair-cell loss. The separate BALB/c null model has a different early threshold phenotype.
biological_scale: TISSUE
biological_processes:
- preferred_term: sensory perception of sound
term:
id: GO:0007605
label: sensory perception of sound
modifier: ABNORMAL
cell_types:
- preferred_term: cochlear outer hair cell
term:
id: CL:0000601
label: cochlear outer hair cell
downstream:
- target: Bilateral sensorineural hearing impairment
causal_link_type: INDIRECT_UNKNOWN_INTERMEDIATES
description: Proposed translation from matrix-dependent auditory dysfunction in null mice to the dominant human hearing-loss phenotype; the causal sequence in missense carriers is unresolved.
hypothesis_groups:
- dfna4b_missense_to_matrix
- target: High-frequency sensorineural hearing impairment
causal_link_type: INDIRECT_UNKNOWN_INTERMEDIATES
description: Reported human families often begin with high-frequency loss, whereas the beta-galactosidase-null matrix phenotype is apically biased. Frequency-specific human susceptibility is not explained by the null data.
hypothesis_groups:
- dfna4b_missense_to_matrix
- target: Progressive hearing impairment
causal_link_type: INDIRECT_UNKNOWN_INTERMEDIATES
description: Age-related null-mouse matrix degeneration supports a maintenance hypothesis; it does not establish the decade-scale human trajectory for every allele.
hypothesis_groups:
- dfna4b_missense_to_matrix
evidence:
- reference: PMID:25080593
reference_title: Loss of the tectorial membrane protein CEACAM16 enhances spontaneous, stimulus-frequency, and transiently evoked otoacoustic emissions.
supports: SUPPORT
evidence_source: MODEL_ORGANISM
snippet: We also observed spontaneous otoacoustic emissions (SOAEs) in 70% of the homozygous mice. This incidence is remarkable considering that <3% of WT controls have SOAEs.
explanation: The quantified instability signature, with its wild-type comparison.
quote_role: PRIMARY_RESULT
directness: DIRECT
- reference: PMID:25080593
reference_title: Loss of the tectorial membrane protein CEACAM16 enhances spontaneous, stimulus-frequency, and transiently evoked otoacoustic emissions.
supports: SUPPORT
evidence_source: MODEL_ORGANISM
snippet: Changes in TM structure appear to influence the balance between energy generation and dissipation such that the system becomes unstable.
explanation: The authors' interpretation of why a matrix defect destabilises rather than merely weakens the amplifier, which is the claim this node makes.
quote_role: PRIMARY_RESULT
directness: DIRECT
mechanism_confidence: PROVISIONAL
phenotypes:
- category: Auditory
name: Bilateral sensorineural hearing impairment
description: Bilateral sensorineural hearing loss is the defining phenotype. Available families have no consistent syndromic pattern. Vestibular symptoms were absent in several reports, but testing was limited and the de novo child did not undergo formal vestibular assessment.
frequency: OBLIGATE
phenotype_term:
preferred_term: Bilateral sensorineural hearing impairment
term:
id: HP:0008619
label: Bilateral sensorineural hearing impairment
onset:
notes: Usually postlingual; Russian family onset was 5–20 years, the Chinese p.Gly169Arg table spans 10–28 years, and p.Thr140Ala family onset was in the twenties.
evidence:
- reference: url:https://pmc.ncbi.nlm.nih.gov/articles/PMC4662267/?report=xml
reference_title: A Novel de novo Mutation in CEACAM16 Associated with Postlingual Hearing Impairment - PMC
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: The child has a bilateral hearing loss that moderately affects all frequencies except 0.125 kHz.
explanation: Figure legend explicitly documents bilateral involvement in the de novo child.
- reference: url:https://pmc.ncbi.nlm.nih.gov/articles/PMC4662267/?report=xml
reference_title: A Novel de novo Mutation in CEACAM16 Associated with Postlingual Hearing Impairment - PMC
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: His audiogram exhibits a stable flat threshold throughout all frequencies except 0.125 kHz in the 40-60 dB range
explanation: The de novo child had bilateral flat rather than high-frequency-first loss; the clinical report includes bilateral audiometry.
- reference: url:https://www.ncbi.nlm.nih.gov/sites/books/NBK1434/?report=reader
reference_title: Genetic Hearing Loss Overview - GeneReviews® - NCBI Bookshelf
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: REVIEW_SYNTHESIS
directness: DIRECT
snippet: Sensorineural hearing loss due to malfunction of the inner ear structures (i.e., cochlea or auditory nerve, with potential for vestibular dysfunction)
explanation: General clinical definition, not a claim of DFNA4B-specific vestibular dysfunction.
- reference: PMID:39157884
reference_title: Auditory Phenotype of a Novel Missense Variant in the CEACAM16 Gene in a Large Russian Family With Autosomal Dominant Nonsyndromic Hearing Loss.
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: The hearing impairment manifests in the age of 5-20 years with tinnitus and thresholds increase at high frequencies. It should be differentiated with age-related hearing loss if identified late.
explanation: Gives the age-at-onset range that makes the loss postlingual, and the authors' own statement of the presbycusis differential.
quote_role: PRIMARY_RESULT
directness: DIRECT
notes: Onset is reported after language acquisition in the principal families; neither a fixed childhood range nor normal hearing throughout childhood applies to every reported allele.
- category: Auditory
name: High-frequency sensorineural hearing impairment
description: High-frequency loss with later involvement of lower frequencies is documented in several families, but the de novo p.Leu365Arg child had a flat audiogram.
phenotype_term:
preferred_term: High-frequency sensorineural hearing impairment
term:
id: HP:0001757
label: High-frequency sensorineural hearing impairment
evidence:
- reference: PMID:39157884
reference_title: Auditory Phenotype of a Novel Missense Variant in the CEACAM16 Gene in a Large Russian Family With Autosomal Dominant Nonsyndromic Hearing Loss.
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: Eleven family members suffer from hearing impairment, which starts with tinnitus and threshold increase at high frequencies, since the age of 5-20 years.
explanation: Documents the high-frequency onset in eleven affected members of a five-generation family, with the age range.
quote_role: PRIMARY_RESULT
directness: DIRECT
- reference: PMID:33040498
reference_title: '[Identification of a novel mutation of CEACAM16 gene in a Chinese family with autosomal dominant nonsyndromic hearing loss].'
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: The affected family members appeared to have developed a high-frequency hearing loss with the onset of twenties.
explanation: The p.Thr140Ala family provides independent clinical evidence of high-frequency hearing loss with onset in the twenties.
- category: Auditory
name: Progressive hearing impairment
description: Progressive loss over years or decades is documented in multiple families. The Russian longitudinal observations include a later stable interval in an older relative; the de novo child had no progression during one year of observation, too short to establish lifelong stability.
phenotype_term:
preferred_term: Progressive hearing impairment
term:
id: HP:0001730
label: Progressive hearing impairment
evidence:
- reference: PMID:39157884
reference_title: Auditory Phenotype of a Novel Missense Variant in the CEACAM16 Gene in a Large Russian Family With Autosomal Dominant Nonsyndromic Hearing Loss.
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: Hearing loss slowly progresses with age in each person and is similar to age-related hearing loss.
explanation: States both the progressive course and the resemblance to presbycusis that makes it a diagnostic trap.
quote_role: PRIMARY_RESULT
directness: DIRECT
- reference: PMID:25589040
reference_title: Exome sequencing identifies a novel CEACAM16 mutation associated with autosomal dominant nonsyndromic hearing loss DFNA4B in a Chinese family.
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: a five-generation nonconsanguineous Chinese family affected by late-onset and progressive ADNSHL
explanation: Independent confirmation of the late-onset progressive course in a second family.
quote_role: PRIMARY_RESULT
directness: DIRECT
- reference: url:https://pmc.ncbi.nlm.nih.gov/articles/PMC4662267/?report=xml
reference_title: A Novel de novo Mutation in CEACAM16 Associated with Postlingual Hearing Impairment - PMC
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: His hearing loss has not progressed in the short span of one year since his diagnosis.
explanation: Limits universal progression at every observed interval; long-term course remains unknown.
- category: Auditory
name: Tinnitus
description: Tinnitus varies by family and can precede measurable threshold elevation. It was reported by nine of twelve variant-positive affected people in the p.Gly169Arg family and by many members of the Russian p.Thr140Ile family, but was absent in the p.Arg255Gly family and the de novo child. No pooled frequency is established.
phenotype_term:
preferred_term: Tinnitus
term:
id: HP:0000360
label: Tinnitus
evidence:
- reference: PMID:39157884
reference_title: Auditory Phenotype of a Novel Missense Variant in the CEACAM16 Gene in a Large Russian Family With Autosomal Dominant Nonsyndromic Hearing Loss.
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: Most family members had high-pitched ringing in their ears prior to the development of hearing loss.
explanation: Tinnitus can precede the hearing-loss phenotype in this family.
- reference: PMID:35292975
reference_title: A novel missense variant in CEACAM16 gene causes autosomal dominant nonsyndromic hearing loss.
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: None of the affected individuals reported complains consistent with tinnitus or vestibular dysfunction.
explanation: Family-specific absence limits generalization of tinnitus prevalence.
- name: Hyperacusis
category: Auditory
description: Noise sensitivity preceded measurable hearing loss in the single de novo p.Leu365Arg case. This is a subjective symptom; it does not establish increased susceptibility to acoustic injury.
phenotype_term:
preferred_term: Hyperacusis
term:
id: HP:0010780
label: Hyperacusis
evidence:
- reference: url:https://pmc.ncbi.nlm.nih.gov/articles/PMC4662267/?report=xml
reference_title: A Novel de novo Mutation in CEACAM16 Associated with Postlingual Hearing Impairment - PMC
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: Prior to developing hearing impairment, he reported noise sensitivity and headaches that correlated with strained hearing.
explanation: Single-case noise sensitivity; headaches and allergic comorbidities are not generalized as CEACAM16 disease features.
animal_models:
- name: Ceacam16-null mutant mouse
species: Mouse
genotype: Ceacam16 null (beta-galactosidase knock-in allele, homozygous)
publication: PMID:25080593
description: Ceacam16 beta-galactosidase-null mice on a C57BL/6J background show matrix disorganization and early emission abnormalities, followed by age-related loss of sensitivity. The perturbation is biallelic absence of protein, not the heterozygous missense state in DFNA4B.
modeled_mechanisms:
- target: Destabilized Tectorial Membrane Matrix Assembly
relationship: PARTIALLY_RECAPITULATES
fidelity: MODERATE
model_scale: TISSUE
description: Reproduces the structural endpoint - reduced beta-tectorin, no striated-sheet matrix, absent Hensen's stripe - at the scale the node describes.
limitations: Direct mouse matrix pathology supports CEACAM16 function. It does not establish the corresponding histopathology or dominant molecular mechanism in human missense carriers.
divergences:
- divergence_type: PROXY_QUANTITY
materiality: QUALIFYING
description: Complete protein absence differs from an altered protein with allele-specific biochemical behavior in DFNA4B.
evidence:
- reference: PMID:25080593
reference_title: Loss of the tectorial membrane protein CEACAM16 enhances spontaneous, stimulus-frequency, and transiently evoked otoacoustic emissions.
supports: SUPPORT
evidence_source: MODEL_ORGANISM
snippet: To learn more about the structural and functional significance of CEACAM16, we created a Ceacam16-null mutant mouse.
explanation: Establishes the model and its purpose, which is what makes it informative for this node.
quote_role: PRIMARY_RESULT
directness: DIRECT
- target: Destabilized Cochlear Amplification
relationship: PARTIALLY_RECAPITULATES
fidelity: MODERATE
model_scale: TISSUE
description: 'Reproduces the functional signature directly: enlarged stimulus-frequency and transiently evoked emissions, and spontaneous emissions in most homozygotes.'
limitations: Young beta-galactosidase-null mice have nearly normal thresholds, and age-related matrix loss is biased toward the low-frequency apex. This differs from common high-frequency-first human onset. Aged heterozygous null mice also develop frequent spontaneous emissions despite thresholds resembling wild type, so they are not physiologically normal in every respect.
readouts:
- name: Spontaneous otoacoustic emission incidence
target: Destabilized Cochlear Amplification
direction: INCREASED
interpretation: Spontaneous emissions are the direct physiological readout of an under-damped cochlear amplifier, which is what this node asserts.
evidence:
- reference: PMID:25080593
reference_title: Loss of the tectorial membrane protein CEACAM16 enhances spontaneous, stimulus-frequency, and transiently evoked otoacoustic emissions.
supports: SUPPORT
evidence_source: MODEL_ORGANISM
snippet: We also observed spontaneous otoacoustic emissions (SOAEs) in 70% of the homozygous mice.
explanation: The measurement itself, with its incidence in homozygotes.
quote_role: PRIMARY_RESULT
directness: DIRECT
- name: Auditory brainstem response threshold
target: Destabilized Cochlear Amplification
direction: UNCHANGED
interpretation: 'A real negative result, and an informative one: thresholds are largely normal in young nulls while the emission measures are already abnormal, which is what identifies the early defect as instability rather than loss of sensitivity.'
evidence:
- reference: PMID:25080593
reference_title: Loss of the tectorial membrane protein CEACAM16 enhances spontaneous, stimulus-frequency, and transiently evoked otoacoustic emissions.
supports: SUPPORT
evidence_source: MODEL_ORGANISM
snippet: Although brain-stem evoked responses and distortion product otoacoustic emissions are, for most frequencies, normal in young mice lacking CEACAM16, stimulus-frequency and transiently evoked emissions are larger.
explanation: Reports both the normal thresholds and the abnormal emissions in the same sentence. This is the young-animal timepoint; see the neighbouring readout for the late one, where thresholds do rise.
quote_role: PRIMARY_RESULT
directness: DIRECT
- name: Auditory brainstem response threshold at 12 months
target: Destabilized Cochlear Amplification
direction: INCREASED
interpretation: The measured quantity is ABR threshold, which increases as sensitivity worsens. Hair-cell counts were similar between genotypes, although both had age-related hair-cell loss. The additional null-associated deficit was strongest at low frequencies; high-frequency genotype differences were limited by shared background degeneration.
evidence:
- reference: PMID:31249509
reference_title: Accelerated Age-Related Degradation of the Tectorial Membrane in the Ceacam16(βgal/βgal) Null Mutant Mouse, a Model for Late-Onset Human Hereditary Deafness DFNB113.
supports: SUPPORT
evidence_source: MODEL_ORGANISM
snippet: SOAEs and DPOAEs are not detected in Ceacam16βgal/βgal mice and ABR thresholds are increased by up to ∼40 dB across frequency, despite a complement of hair cells similar to that present in Ceacam16+/+ mice
explanation: Late threshold elevation despite similar hair-cell complements between genotypes; this does not mean every hair cell remained intact.
quote_role: PRIMARY_RESULT
directness: DIRECT
evidence:
- reference: PMID:25080593
reference_title: Loss of the tectorial membrane protein CEACAM16 enhances spontaneous, stimulus-frequency, and transiently evoked otoacoustic emissions.
supports: SUPPORT
evidence_source: MODEL_ORGANISM
snippet: Results from mice lacking CEACAM16 are consistent with the idea that the organ of Corti evolved to maximize the gain of the cochlear amplifier while preventing large oscillations.
explanation: The authors' statement of what the model shows about amplifier stability.
quote_role: PRIMARY_RESULT
directness: DIRECT
- target: Heterozygous CEACAM16 Missense Alteration
relationship: FAILS_TO_RECAPITULATE
fidelity: LOW
model_scale: MOLECULAR
description: A null allele removes CEACAM16 and cannot reproduce the initiating heterozygous missense perturbation.
limitations: The reviewed studies do not provide a dominant human-allele knock-in; secretion assays and family segregation do not substitute for that model.
divergences:
- divergence_type: BOUNDARY_OMISSION
materiality: INVALIDATING
description: The missense protein is absent from this model, preventing assessment of allele-specific secretion, glycosylation or mutant/wild-type interaction.
evidence:
- reference: PMID:25080593
reference_title: Loss of the tectorial membrane protein CEACAM16 enhances spontaneous, stimulus-frequency, and transiently evoked otoacoustic emissions.
supports: SUPPORT
evidence_source: MODEL_ORGANISM
snippet: we created a Ceacam16-null mutant mouse
explanation: 'States the allele type. The negative claim rests on the model''s construction, not on a negative experimental result: a null cannot express a missense monomer.'
quote_role: PRIMARY_RESULT
directness: DIRECT
- target: Altered Tectorial Membrane Mechanics
relationship: PARTIALLY_RECAPITULATES
fidelity: MODERATE
model_scale: TISSUE
description: Ex vivo wave measurements on isolated membranes from adult null mice demonstrate reduced stiffness and viscosity.
limitations: Five adult null, six adult wild-type and two juvenile null preparations were studied; repeated spatial/frequency measurements are not independent animals. The juvenile comparison lacks juvenile wild-type controls, and tissue was isolated from the middle cochlear turn.
evidence:
- reference: PMID:34555361
reference_title: Age-related degradation of tectorial membrane dynamics with loss of CEACAM16.
supports: SUPPORT
evidence_source: IN_VITRO
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: Here, we show that TM stiffness and viscosity are significantly reduced in adult mice that lack functional CEACAM16 relative to age-matched wild-type controls.
explanation: Measurements on isolated mouse tissue establish material-property changes. The experiment used adult null and wild-type tissue plus two juvenile null preparations, without a juvenile wild-type group.
- name: Ceacam16 deletion-null mouse on BALB/c background
species: Mouse
genotype: Homozygous deletion of exons 2–4 and part of exon 5
publication: PMID:22544735
description: A distinct null line showed low- and high-frequency threshold elevation and reduced suprathreshold emissions early in life. Subsequent age-related decline was broadly parallel to wild type. Noise exposure did not reveal increased vulnerability under the tested conditions.
modeled_mechanisms:
- target: Destabilized Cochlear Amplification
relationship: PARTIALLY_RECAPITULATES
fidelity: LOW
model_scale: TISSUE
description: Supports a role for CEACAM16 in auditory sensitivity, with a phenotype differing from the beta-galactosidase-null line.
limitations: Different genetic background, deletion construct and tests preclude treating the two null studies as one uniform longitudinal experiment. Neither models dominant missense disease.
evidence:
- reference: url:https://pmc.ncbi.nlm.nih.gov/articles/PMC3381124/?pdf=1
reference_title: Loss of Mammal-specific Tectorial Membrane Component Carcinoembryonic Antigen Cell Adhesion Molecule 16 (CEACAM16) Leads to Hearing Impairment at Low and High Frequencies - PMC
supports: SUPPORT
evidence_source: MODEL_ORGANISM
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: indicating that vulnerability to cochlear trauma is not increased by loss of the CEACAM16 protein in the cochlea.
explanation: Noise challenge in this null model did not increase injury; this is not a clinical study of dominant carriers.
treatments:
- name: Hearing Amplification
description: Hearing aids and individualized auditory/communication rehabilitation are used according to audiologic need. Hearing-aid fitting is documented in the Russian family and the de novo child; these reports do not quantify comparative treatment efficacy.
therapeutic_modality: DEVICE
treatment_term:
preferred_term: hearing aid fitting and auditory rehabilitation
term:
id: NCIT:C15315
label: Rehabilitation
qualifiers:
- predicate:
preferred_term: medical device
term:
id: NCIT:C16830
label: Medical Device
value:
preferred_term: hearing aid
term:
id: NCIT:C183182
label: Hearing Aid
evidence:
- reference: PMID:39157884
reference_title: Auditory Phenotype of a Novel Missense Variant in the CEACAM16 Gene in a Large Russian Family With Autosomal Dominant Nonsyndromic Hearing Loss.
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: Her daughters were fitted with hearing aids after 30 years of age, and her son after 20 years of age (Table 1).
explanation: Observed clinical management in the Russian family, without measured efficacy.
- reference: url:https://pmc.ncbi.nlm.nih.gov/articles/PMC4662267/?report=xml
reference_title: A Novel de novo Mutation in CEACAM16 Associated with Postlingual Hearing Impairment - PMC
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: He was treated with hearing aids at the age of 11 and attends a normal school.
explanation: Hearing-aid use in the de novo case.
- reference: url:https://www.ncbi.nlm.nih.gov/sites/books/NBK1434/?report=reader
reference_title: Genetic Hearing Loss Overview - GeneReviews® - NCBI Bookshelf
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: REVIEW_SYNTHESIS
directness: DIRECT
snippet: Habilitation for hearing loss includes improved access to sound through hearing aids or cochlear implants and, when desired, exposure to and teaching of American Sign Language.
explanation: General genetic-hearing-loss management guidance.
- name: Cochlear Implantation
description: Cochlear implantation may be considered under standard audiologic candidacy criteria when appropriately fitted hearing aids provide insufficient benefit. The reviewed DFNA4B reports do not provide implantation outcomes.
therapeutic_modality: SURGERY
treatment_term:
preferred_term: cochlear device implantation
term:
id: NCIT:C15329
label: Surgical Procedure
qualifiers:
- predicate:
preferred_term: medical device
term:
id: NCIT:C16830
label: Medical Device
value:
preferred_term: cochlear implant
term:
id: NCIT:C157820
label: Cochlear Implant
evidence:
- reference: url:https://www.ncbi.nlm.nih.gov/sites/books/NBK1434/?report=reader
reference_title: Genetic Hearing Loss Overview - GeneReviews® - NCBI Bookshelf
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: REVIEW_SYNTHESIS
directness: DIRECT
snippet: Habilitation for hearing loss includes improved access to sound through hearing aids or cochlear implants and, when desired, exposure to and teaching of American Sign Language.
explanation: General rehabilitation guidance, not DFNA4B-specific outcome evidence.
- name: Genetic Counselling
description: Counseling distinguishes a 50% chance of transmitting a causal heterozygous variant to each child from sibling recurrence in an apparently de novo family. Parental testing, possible mosaicism and variant interpretation inform recurrence counseling. Serial audiometry is appropriate because age of onset and progression vary.
therapeutic_modality: OTHER
treatment_term:
preferred_term: Genetic Counseling
term:
id: NCIT:C15240
label: Genetic Counseling
evidence:
- reference: url:https://www.ncbi.nlm.nih.gov/sites/books/NBK1434/?report=reader
reference_title: Genetic Hearing Loss Overview - GeneReviews® - NCBI Bookshelf
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: REVIEW_SYNTHESIS
directness: DIRECT
snippet: Individuals with autosomal dominant hearing loss have a 50% chance of transmitting the pathogenic variant to each child.
explanation: General dominant-inheritance counseling; the risk refers to offspring, including offspring of a proband with a de novo variant.
- reference: url:https://www.ncbi.nlm.nih.gov/sites/books/NBK1434/?report=reader
reference_title: Genetic Hearing Loss Overview - GeneReviews® - NCBI Bookshelf
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: REVIEW_SYNTHESIS
directness: DIRECT
snippet: Regular follow up is recommended for all individuals with genetic hearing loss
explanation: General surveillance guidance applied to a disorder with variable onset and progression.
- name: Hearing Conservation
description: Avoid repeated excessive noise exposure as part of general hearing conservation. Increased noise vulnerability is not established in DFNA4B, and the BALB/c Ceacam16-null noise challenge did not show increased injury. Reported subjective noise sensitivity does not establish acoustic-trauma susceptibility.
therapeutic_modality: BEHAVIORAL
treatment_term:
preferred_term: preventive behavioral intervention
term:
id: NCIT:C15747
label: Supportive Care
evidence:
- reference: url:https://www.ncbi.nlm.nih.gov/sites/books/NBK1434/?report=reader
reference_title: Genetic Hearing Loss Overview - GeneReviews® - NCBI Bookshelf
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: REVIEW_SYNTHESIS
directness: DIRECT
snippet: persons with documented hearing loss should be counseled appropriately and repeated overexposure to loud noises should be avoided.
explanation: General preventive guidance, not a CEACAM16-specific treatment effect.
- reference: url:https://pmc.ncbi.nlm.nih.gov/articles/PMC3381124/?pdf=1
reference_title: Loss of Mammal-specific Tectorial Membrane Component Carcinoembryonic Antigen Cell Adhesion Molecule 16 (CEACAM16) Leads to Hearing Impairment at Low and High Frequencies - PMC
supports: SUPPORT
evidence_source: MODEL_ORGANISM
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: indicating that vulnerability to cochlear trauma is not increased by loss of the CEACAM16 protein in the cochlea.
explanation: Direct CEACAM16-null evidence limits an inference previously borrowed from another tectorial-membrane gene.
diagnosis:
- name: Audiometry with family history and molecular confirmation
description: Audiometry establishes bilateral sensorineural loss and its frequency pattern, with serial testing to assess progression. Hearing-loss panel or exome testing and segregation analysis can identify a pathogenic or likely pathogenic CEACAM16 variant in the appropriate inheritance context. A heterozygous variant of uncertain significance alone does not confirm DFNA4B. High-frequency onset and a dominant pedigree are helpful but not mandatory, as demonstrated by the flat-audiogram de novo case. Age-related and acquired hearing loss can create phenocopies.
evidence:
- reference: PMID:25589040
reference_title: Exome sequencing identifies a novel CEACAM16 mutation associated with autosomal dominant nonsyndromic hearing loss DFNA4B in a Chinese family.
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: probands were analyzed using exome capture followed by next-generation sequencing
explanation: Establishes exome sequencing as the route to molecular diagnosis in this disorder.
quote_role: PRIMARY_RESULT
directness: DIRECT
- reference: PMID:26648831
reference_title: A Novel de novo Mutation in CEACAM16 Associated with Postlingual Hearing Impairment.
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: we used the TruSight One panel to investigate a parent-child trio without familial history of hearing loss and one affected child
explanation: Documents trio panel sequencing finding a dominant allele where no family history existed, which is why absence of a pedigree does not exclude the diagnosis.
quote_role: PRIMARY_RESULT
directness: DIRECT
- reference: url:https://www.ncbi.nlm.nih.gov/sites/books/NBK1434/?report=reader
reference_title: Genetic Hearing Loss Overview - GeneReviews® - NCBI Bookshelf
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: REVIEW_SYNTHESIS
directness: DIRECT
snippet: The identification of variant(s) of uncertain significance cannot be used to confirm or rule out the diagnosis.
explanation: General molecular-diagnostic interpretation rule; a variant alone is insufficient.
external_assertions:
- name: OMIM deafness, autosomal dominant 4B record
source: OMIM
assertion_type: disease_record
external_id: OMIM:614614
url: https://omim.org/entry/614614
description: OMIM phenotype record for DFNA4B, taken from MONDO's own cross-reference for MONDO:0013823. Recorded here rather than under `mappings:` because the schema's DiseaseMappings container has only a mondo_mappings slot with no OMIM mapping slot.
classifications:
harrisons_chapter:
- classification_value: DISORDER_OF_EAR
evidence:
- reference: PMID:39157884
reference_title: Auditory Phenotype of a Novel Missense Variant in the CEACAM16 Gene in a Large Russian Family With Autosomal Dominant Nonsyndromic Hearing Loss.
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: Eleven family members suffer from hearing impairment, which starts with tinnitus and threshold increase at high frequencies, since the age of 5-20 years.
explanation: The family report identifies an auditory disorder with high-frequency hearing impairment, supporting placement among disorders of the ear.
quote_role: PRIMARY_RESULT
directness: DIRECT
discussions:
- discussion_id: ceacam16_opposite_secretion_phenotypes
kind: OPEN_QUESTION
prompt: How do opposite secretion phenotypes for p.Gly169Arg and p.Arg255Gly lead to dominant hearing loss?
attaches_to:
- pathophysiology#Altered CEACAM16 Secretion
rationale: Transfected-cell studies show reduced secretion for p.Gly169Arg and increased intracellular/extracellular protein for p.Arg255Gly. They did not directly measure mutant/wild-type matrix incorporation or dominant interference. Simple haploinsufficiency is not supported by hearing-normal human heterozygotes in recessive families, but this does not select a unique alternative mechanism.
notes: Resolving this needs the mutant proteins assayed for incorporation into and disruption of a tectorin matrix, not only for secretion efficiency from transfected cells.
- discussion_id: ceacam16_no_dominant_knockin_model
kind: HUMAN_MODEL_MISMATCH
prompt: Which matrix and mechanical abnormalities of null mice are shared by human dominant missense carriers?
attaches_to:
- pathophysiology#Destabilized Tectorial Membrane Matrix Assembly
- animal_models#Ceacam16-null mutant mouse
rationale: Null studies establish important roles in matrix organization and mechanics. Their inheritance, age course and frequency distributions differ from human DFNA4B, and the two null lines differ from one another. No dominant-allele knock-in was identified in the reviewed literature.
notes: Direct measurements in an allele-matched model are needed before presenting null-mouse morphology as established human histopathology.
notes: DFNA4B remains within Mendelian review scope; noise is a possible modifier or symptom, not an environmental disease etiology. No population prevalence estimate is established by the small family series; the number of reported alleles is not a prevalence measure. The p.Thr140Pro glycosylation experiment used a mouse p.Thr142Pro construct, as summarized by the 2024 report; direct full text of the 2011 paper was inaccessible during this review. Glycosylation changes for p.Thr140Ala and p.Thr140Ile are proposed, not experimentally demonstrated by those family reports. ClinicalTrials.gov queries for CEACAM16 and DFNA4B returned no studies on 2026-09-23. Trial absence is limited to those queries. The research report's incorrect phenotype identifiers were not reused.
mechanistic_hypotheses:
- hypothesis_group_id: dfna4b_missense_to_matrix
description: Allele-specific CEACAM16 biochemical changes may disrupt tectorial-membrane organization and mechanics, altering cochlear amplification. The middle and distal steps are chiefly supported by null-mouse and isolated-tissue studies; a common dominant-negative mechanism and the tissue consequences of each human missense allele remain unresolved.
References & Deep Research
References
17Deep Research
1Deep research results are used as seeds for research; they do not undergo the same validation as the main records and may contain errors. How we use deep research.
Evaluations and curation notes (1)
Create: Autosomal Dominant Nonsyndromic Hearing Loss 4B · 2026-09-14T21:29:01Z · View source
Created DFNA4B (CEACAM16) from the primary literature plus an OpenScientist deep-research report. Five-node pathograph from a heterozygous missense monomer through altered secretion and destabilized tectorial-membrane matrix assembly to destabilized cochlear amplification. Curated as a tectorial membrane matrix disease rather than a hair-cell disease. The entry keeps the dominant missense mechanism and the allelic recessive loss-of-function disease apart, because a missense monomer entering the matrix and a null allele removing the protein are different perturbations rather than dosage grades. Two discussions record the open problems: one dominant allele reduces secretion while another increases it, and the only animal model is a null so the dominant-negative step has no animal support; the model's link to the initiating node is curated FAILS_TO_RECAPITULATE with an INVALIDATING BOUNDARY_OMISSION divergence. Four of the seven HP identifiers the deep-research report suggested resolve to unrelated terms (HP:0000421 is Epistaxis, HP:0008542 is Low-frequency hearing impairment, HP:0011476 is Profound sensorineural hearing impairment, HP:0008625 is Severe sensorineural hearing impairment) and the report's own term validation flagged only the last; every binding here was resolved independently. just preflight-dr reports a rival-gene warning for TECTB at 50% of CEACAM16 mentions; this was checked and is a false positive for Named Entity Confusion purposes, since TECTB is CEACAM16's binding partner rather than a second disease entity, and the report's TECTB-specific noise-susceptibility material was deliberately not carried into the entry. Validated with just validate-disorders (31/31 snippets), just validate-terms, check-entity-refs, check-causal-targets, check-duplicate-keys, check-qualifier-terms. During curation I twice assumed a cached reference had no abstract body and wrote placeholder title snippets; on checking, both records had full abstracts, and the placeholders were replaced with real quoted findings.
1. Disease Information
Overview. DFNA4B is a form of hereditary, nonsyndromic (hearing loss occurring in isolation, without associated systemic features), sensorineural hearing impairment inherited in an autosomal dominant pattern. It is one of the allelic disorders caused by variants in CEACAM16 and corresponds to the "B" subtype at the DFNA4 locus. The disease is defined at the disease/gene aggregate level (OMIM, ClinVar, published pedigrees) rather than from individual EHR records; the evidence base is a set of multigenerational families and de novo cases characterized by clinical audiometry and molecular genetics.
Key identifiers.
| Resource | Identifier |
|---|---|
| MONDO | MONDO:0013823 |
| OMIM (phenotype) | #614614 (DFNA4B) |
| Gene | CEACAM16 (HGNC), OMIM *614591 |
| Cytogenetic locus | 19q13.32 (DFNA4 region) |
| Inheritance | Autosomal dominant |
| MeSH (parent concept) | Hearing Loss, Sensorineural; Deafness |
| ICD-10 | H90.5 (unspecified sensorineural hearing loss) — no DFNA4B-specific code |
| ICD-11 | AB50–AB52 range (sensorineural hearing impairment) — no specific code |
Synonyms / alternative names. DFNA4B; deafness, autosomal dominant 4B; nonsyndromic hearing loss DFNA4 (CEACAM16-related dominant form). The recessive allelic disorder is designated DFNB (autosomal recessive nonsyndromic hearing loss, CEACAM16-related; see Section 4).
Information source. Aggregated disease-level resources (OMIM, ClinVar, gene-disease literature) and published family/case reports — not individual patient EHR data.
2. Etiology
Primary cause — genetic. DFNA4B is a monogenic disorder caused by heterozygous variants in CEACAM16. It is not attributable to environmental, infectious, or acquired causes; the etiology is entirely germline genetic. Reported causal variants are predominantly missense changes in conserved regions of the protein (see Section 4).
Genetic risk factors. The disease-determining factor is the presence of a single heterozygous pathogenic CEACAM16 variant. Documented dominant variants include: - p.Thr140Ile — large Russian family PMID: 39157884 - p.Gly169Arg (c.505G>A) — Chinese family PMID: 25589040 - p.Arg255Gly (c.763A>G) — Chinese family PMID: 35292975 - p.Leu365Arg — de novo case PMID: 26648831
Environmental risk factors / modifiers. No specific environmental risk factors are established for DFNA4B itself. However, because the TM is a shared structure whose integrity buffers the cochlea against environmental insults, exposures known to accelerate sensorineural hearing loss generally — noise exposure, aging, ototoxic drugs (aminoglycosides, platinum agents) — are biologically plausible aggravators of an already-compromised TM. Supporting this concept, mice heterozygous for a TM-gene (TECTB) missense variant show increased susceptibility to noise-induced hearing loss despite normal baseline thresholds PMID: 42702881 — evidence (from a related TM gene) that subclinical TM defects sensitize the cochlea to environmental damage. This is an inferred, not directly demonstrated, gene–environment interaction for CEACAM16.
Protective factors. No genetic or environmental protective factors have been specifically characterized for DFNA4B. Avoidance of noise and ototoxins is a reasonable, if unproven, protective strategy.
3. Phenotypes
DFNA4B is a "pure" auditory phenotype. The dominant clinical features and their characteristics are summarized below.
| Phenotype | HPO term | Onset | Severity | Progression | Frequency in affected |
|---|---|---|---|---|---|
| Sensorineural hearing loss | HP:0000407 | Late-onset (~5–30 y) | Mild→severe with age | Slowly progressive | ~100% (defining) |
| High-frequency hearing loss | HP:0000421 / HP:0008542 | Earliest affected range | Initial deficit | Progressive, later involving mid/low frequencies | Characteristic initial pattern |
| Progressive hearing impairment | HP:0001730 | — | — | Progressive | Universal |
| Bilateral hearing loss | HP:0008625 (bilateral SNHL) | — | Symmetric | — | Universal |
| Tinnitus | HP:0000360 | Often the presenting symptom | Variable | — | Common early feature |
| Postlingual onset | HP:0011476 (postlingual sensorineural HL) | After speech acquisition | — | — | Universal |
Detail. The auditory phenotype is postlingual and typically begins in childhood or adolescence with tinnitus and elevation of high-frequency thresholds, then progresses with age to involve middle and lower frequencies, ultimately resembling age-related hearing loss. In a large five-generation Russian family (11 affected individuals), onset occurred between ages 5 and 20 years, "start[ing] with tinnitus and threshold increase at high frequencies" PMID: 39157884. A de novo case matched the described DFNA4B onset and severity PMID: 26648831, and Chinese families likewise showed late-onset, progressive loss PMID: 25589040, PMID: 35292975.
Quality-of-life impact. As a progressive, lifelong sensorineural hearing loss, DFNA4B carries the well-documented QoL burdens of adult-onset deafness: impaired speech comprehension (especially in noise), communication difficulty, social withdrawal, and — in later life — associations with cognitive decline and depression common to presbycusis-like hearing loss. Disease-specific QoL instruments have not been applied to DFNA4B cohorts; generic tools (SF-36, EQ-5D, HHIE) would apply. No mortality or systemic morbidity is associated.
4. Genetic / Molecular Information
Causal gene. CEACAM16 (carcinoembryonic antigen-related cell adhesion molecule 16), OMIM 614591, chromosome 19q13.32, within the DFNA4 linkage region. The gene encodes a secreted glycoprotein* — confirmed by immunofluorescence and Western blot PMID: 25589040 — that is a structural component of the tectorial membrane. The protein contains immunoglobulin-like (Ig) domains (including N-terminal variable/N-type and constant/A-type domains) characteristic of the CEACAM family.
Pathogenic variants (dominant, DFNA4B).
| Variant (protein) | cDNA | Type | Family / origin | Functional consequence | PMID |
|---|---|---|---|---|---|
| p.Thr140Ile | — | Missense | Russian, 5-gen (11 affected) | Dominant | 39157884 |
| p.Gly169Arg | c.505G>A | Missense | Chinese, 5-gen | Reduced mutant secretion efficiency | 25589040 |
| p.Arg255Gly | c.763A>G | Missense | Chinese, 4-gen | Increased secretion of mutant protein | 35292975 |
| p.Leu365Arg | — | Missense | De novo | Dominant, matched DFNA4B phenotype | 26648831 |
Dominant variants are missense and cluster within the conserved Ig-like domains. Functional studies in transfected HEK293T cells reveal two contrasting biochemical consequences: p.Gly169Arg reduces secretion efficiency ("the secretion efficacy of the mutant CEACAM16 is much lower than that of the wild type") PMID: 25589040, whereas p.Arg255Gly increases intracellular and extracellular mutant protein levels PMID: 35292975. Both are interpreted as deleterious, consistent with a dominant-negative / altered-secretion mechanism in which the abnormal monomer poisons assembly of the multimeric TM matrix rather than simply reducing gene dosage.
Variant classification. Reported dominant variants are classified pathogenic/likely pathogenic by ACMG/AMP criteria on the basis of co-segregation in multigenerational families, absence in matched controls (e.g., not present in 200 ancestry-matched controls for p.Gly169Arg), a de novo occurrence (p.Leu365Arg), and supportive functional data.
Allelic recessive form (DFNB). Biallelic loss-of-function variants in CEACAM16 cause autosomal recessive nonsyndromic hearing loss — mechanistically and phenotypically distinct from dominant DFNA4B: - Homozygous splice-altering variants c.37G>T and c.662-1G>C (Iranian families) PMID: 29703829 - Homozygous nonsense c.436C>T (p.Arg146Ter) PMID: 30514912
The authors of the recessive reports concluded that "loss-of-function variants in CEACAM16 cause autosomal recessive hearing loss in humans" PMID: 30514912. Thus CEACAM16 exhibits dual inheritance: dominant missense (poison-monomer) vs. recessive LoF (protein absence).
Allele frequency. Dominant pathogenic variants are private/rare, absent from population controls and gnomAD at appreciable frequency.
Somatic vs germline. Entirely germline; no somatic involvement.
Modifier genes. No formally established modifier genes. Given that CEACAM16 functions within a shared TM protein network, functional variants in interacting partners (TECTA, TECTB, OTOG, OTOGL, OTOA) are plausible modifiers of TM integrity and hearing thresholds, but this is inferential.
Epigenetics / chromosomal abnormalities. No disease-specific DNA-methylation, histone, or large-scale chromosomal abnormalities are reported for DFNA4B. It is a point-mutation disorder.
{{figure:ceacam16_variant_map.png|caption=Schematic map of reported dominant (missense) versus recessive (loss-of-function/splice/nonsense) CEACAM16 variants across the protein's immunoglobulin-fold domain architecture. Dominant DFNA4B variants cluster within conserved Ig domains and act via dominant-negative/altered-secretion mechanisms, whereas biallelic LoF variants truncate the protein and cause the recessive allelic form.}}
5. Environmental Information
DFNA4B is a purely genetic disorder; no environmental, lifestyle, or infectious agent is a necessary cause. However, environmental cochlear stressors are plausible disease aggravators rather than initiators:
- Environmental / occupational: Noise exposure and ototoxic agents (aminoglycoside and platinum-based drugs) may accelerate hearing decline in an already TM-compromised cochlea. Direct evidence in CEACAM16 patients is lacking; the concept is supported by the related TM gene TECTB, where heterozygous mice showed increased noise-induced-hearing-loss susceptibility PMID: 42702881.
- Lifestyle: No specific dietary, smoking, or alcohol associations established for DFNA4B.
- Infectious agents: None. Not applicable.
6. Mechanism / Pathophysiology
Ordered causal chain (initiating lesion → clinical manifestation)
- A heterozygous missense variant in CEACAM16 (e.g., p.Gly169Arg, p.Arg255Gly, p.Thr140Ile, p.Leu365Arg) alters a conserved Ig-domain residue → produces a structurally abnormal CEACAM16 monomer (demonstrated by segregation + in vitro expression).
- The mutant monomer has altered secretion — reduced (p.Gly169Arg) or aberrantly increased/retained (p.Arg255Gly) → impairs correct incorporation of CEACAM16 into the tectorial-membrane matrix (inferred from in vitro secretion assays; PMID: 25589040, PMID: 35292975).
- Because CEACAM16 normally bridges α-tectorin (TECTA) and β-tectorin (TECTB), defective CEACAM16 → destabilizes TECTA–TECTB cross-links and the multimeric TM network, acting as a poison monomer (dominant-negative) (interaction demonstrated in mouse; PMID: 21368133, PMID: 25080593).
- Disrupted cross-linking → failure to form/maintain the striated-sheet matrix and Hensen's stripe, reduced TECTB levels (demonstrated in Ceacam16-null mice; PMID: 25080593).
- Compromised TM structure → abnormal coupling between the TM and the tips of the tallest OHC stereocilia, and with age, accelerated TM degradation/detachment (PMID: 31249509).
- Impaired TM–stereocilia coupling → degraded cochlear amplification and frequency tuning; the active process becomes unstable → elevated/abnormal otoacoustic emissions, including spontaneous OAEs as a micromechanical-instability biomarker (PMID: 25080593, PMID: 26691158, PMID: 34332206).
- Progressive loss of amplification, initially in the high-frequency (basal) cochlea → clinical high-frequency, slowly progressive, bilateral sensorineural hearing loss (the DFNA4B phenotype).
CEACAM16 missense (Ig domain)
│  altered folding/secretion
▼
Poison monomer incorporated into TM matrix  ──(dominant-negative)
│  destabilizes TECTA–TECTB cross-links
▼
↓TECTB, no striated-sheet matrix, loss of Hensen's stripe
│
▼
TM–OHC stereocilia coupling impaired  →  accelerated TM degradation w/ age
│
▼
Cochlear amplifier unstable → abnormal/spontaneous OAEs
│
▼
High-frequency → broadening, slowly progressive bilateral SNHL
Branch (recessive): Biallelic LoF → total absence of CEACAM16 → TM matrix underdevelopment → recessive progressive hearing loss (PMID: 30514912, PMID: 29703829).
Detail by category
- Molecular / structural pathway. The core defect is in extracellular-matrix assembly of the tectorial membrane, not in a classic intracellular signaling cascade (Wnt/MAPK/mTOR are not implicated). CEACAM16 colocalizes and co-immunoprecipitates with α-tectorin PMID: 21368133 and additionally interacts with β-tectorin, "indicating that it may stabilize interactions between TECTA and TECTB" PMID: 25080593.
- Protein dysfunction. Misfolding/altered secretion of an Ig-domain glycoprotein; dominant-negative incorporation of aberrant monomer into a multimeric matrix. GO terms: extracellular matrix structural constituent (GO:0005201).
- Cellular processes. Non-cell-autonomous: the primary lesion is in a secreted matrix, secondarily impairing OHC mechanotransduction/amplification. Relevant GO biological processes: sensory perception of sound (GO:0007605), inner ear morphogenesis (GO:0042472), extracellular matrix organization (GO:0030198).
- Tissue-damage mechanism. Age-accelerated degradation and detachment of the TM (PMID: 31249509); loss of Hensen's stripe correlating with SOAE predominance >15 kHz PMID: 25080593.
- Biochemical / biophysical abnormality. The TM is a passive mechanical element that shapes cochlear tuning; its disruption alters the coupled OHC feedback loop and destabilizes the active process, producing spontaneous otoacoustic emissions in ~70% of Ceacam16-null homozygotes (vs <3% of wild-type) PMID: 25080593.
- Immune involvement / metabolic changes / epigenetics. None implicated. Not applicable.
- Cell types (CL) and anatomy (UBERON): outer hair cell (CL:0000601), inner hair cell (CL:0000589); tectorial membrane (UBERON:0002233), organ of Corti (UBERON:0002227), cochlea (UBERON:0001844).
7. Anatomical Structures Affected
- Organ level: The inner ear / cochlea is the sole affected organ (UBERON:0001846 internal ear; UBERON:0001844 cochlea). No secondary organ involvement; no other body system is affected — consistent with the nonsyndromic designation.
- Body system: Auditory/vestibulocochlear component of the sensory/nervous system (peripheral auditory system). Vestibular function is spared.
- Tissue/cell level: The primary lesion is in the acellular tectorial membrane (UBERON:0002233), an extracellular matrix. Secondarily affected are the sensory outer hair cells (CL:0000601) whose tallest stereocilia are coupled to the TM, and by extension the inner hair cells (CL:0000589) that transmit the encoded signal. The organ of Corti (UBERON:0002227) is the functional epithelium.
- Subcellular level: CEACAM16 is a secreted protein trafficked through the ER/Golgi secretory pathway (GO:0005783 endoplasmic reticulum, GO:0005796 Golgi lumen for biosynthesis) and deposited into the extracellular matrix / tectorial membrane (GO:0031012). Stereociliar tips (GO:0032420 stereocilium) are the coupling site.
- Localization / lateralization: Bilateral and symmetric; the disease progresses along the tonotopic (base→apex) axis, initially affecting the basal (high-frequency) cochlea.
8. Temporal Development
- Onset: Postlingual and late-onset, typically emerging in childhood/adolescence and continuing into adulthood (reported onset ranges ~5–30 years; age 5–20 years in the large Russian pedigree) PMID: 39157884. Onset pattern is insidious/chronic, not acute.
- Progression: Slowly progressive over decades. It begins at high frequencies with tinnitus, then broadens to mid/low frequencies, ultimately resembling presbycusis. Progression rate is slow but effectively lifelong.
- Disease course: Chronic, progressive, non-remitting. There are no episodic or relapsing-remitting features and no spontaneous remission.
- Critical periods / windows for intervention: No pharmacologic window is defined. The relevant intervention window is functional — timely hearing-aid fitting as thresholds rise, and cochlear implantation when hearing loss becomes severe/profound.
9. Inheritance and Population
- Inheritance pattern: Autosomal dominant (dominant DFNA4B, missense). The allelic recessive form is autosomal recessive (biallelic LoF).
- Penetrance: High, consistent with strong co-segregation across multigenerational pedigrees; likely age-dependent given the progressive, late-onset nature (a young carrier may be pre-symptomatic). Formal penetrance estimates are not published.
- Expressivity: Variable in age of onset and rate of progression across and within families.
- De novo mutation: Documented (p.Leu365Arg), so absence of family history does not exclude the diagnosis PMID: 26648831.
- Genetic anticipation / germline mosaicism / founder effects / consanguinity: No anticipation (not a repeat-expansion disorder). Consanguinity is relevant to the recessive allelic form (e.g., Iranian families with homozygous variants PMID: 29703829), not to dominant DFNA4B. No established founder effect for the dominant form.
- Carrier frequency: Not established; dominant variants are private/rare.
- Epidemiology: DFNA4B is a rare cause of hereditary hearing loss; precise prevalence/incidence figures are not established. It is one of ~30+ ADNSHL genes and accounts for a small fraction of dominant nonsyndromic hearing loss PMID: 25589040. Reported dominant families are of Russian and Chinese ancestry, but this reflects ascertainment rather than a demonstrated ethnic predilection.
- Sex ratio: No sex bias expected or reported (autosomal). Male:female ≈ 1:1.
- Geographic distribution: Reported worldwide in individual families; no endemic clustering.
10. Diagnostics
- Audiological testing (core): Pure-tone audiometry reveals bilateral, symmetric, high-frequency-predominant sensorineural hearing loss; otoacoustic emissions (OAE) are notable — TM defects produce abnormal and spontaneous otoacoustic emissions (SOAEs), a potential noninvasive biomarker of TM/cochlear-amplifier instability PMID: 25080593, PMID: 34332206. Auditory brainstem response (ABR) confirms sensorineural pattern and threshold.
- Genetic testing (definitive): Molecular confirmation via CEACAM16 sequencing. Recommended approaches:
- Comprehensive hearing-loss gene panels / targeted NGS (which include CEACAM16 alongside TECTA, TECTB, OTOG, OTOGL, GJB2, etc.) — the practical first-line test PMID: 35292975, PMID: 41897287.
- Whole-exome sequencing (WES) — used to identify CEACAM16 variants in several families, often combined with linkage analysis PMID: 25589040.
- Single-gene testing / Sanger for cascade testing of a known familial variant.
- Linkage analysis historically mapped the locus to 19q13 in extended pedigrees PMID: 25589040.
- Chromosomal microarray, karyotyping, FISH, mtDNA, and repeat-expansion testing are not indicated (point-mutation, autosomal, non-repeat disorder).
- Imaging: No characteristic CT/MRI finding; imaging is used to exclude other causes and for cochlear-implant planning.
- Clinical criteria / differential diagnosis: Diagnosis rests on the combination of a compatible progressive high-frequency SNHL phenotype, dominant family history (or de novo finding), and a pathogenic CEACAM16 variant. Differential diagnosis includes other TM-related dominant hearing losses — especially DFNA8/12 (TECTA) and the newly described TECTB-related dominant hearing loss PMID: 42702881, PMID: 40832383 — as well as OTOG/OTOGL forms and, importantly, age-related hearing loss (presbycusis), which DFNA4B closely mimics. Genetic testing distinguishes these.
- Screening: No population newborn screening for DFNA4B specifically (it is late-onset). Cascade genetic screening of at-risk relatives once a familial variant is identified is appropriate.
11. Outcome / Prognosis
- Survival / mortality: DFNA4B is not life-limiting. Life expectancy is normal; there is no disease-specific mortality.
- Morbidity / disability: The morbidity is functional — progressive hearing disability affecting communication, education, employment, and social participation, with the downstream QoL and (in older adults) cognitive/psychosocial burdens of untreated hearing loss.
- Disease course: Chronic, slowly progressive, lifelong. Without amplification, high-frequency deficits broaden; with hearing aids and, when needed, cochlear implants, functional hearing can be substantially restored.
- Recovery potential: No spontaneous recovery; the structural TM defect is not self-repairing. Functional outcomes with rehabilitation are generally good.
- Prognostic factors: Specific variant, family-specific rate of progression, age at onset, and cumulative environmental exposures (noise/ototoxins) likely modulate the trajectory. Spontaneous/abnormal OAEs are a candidate biomarker of cochlear-amplifier status PMID: 34332206 but are not yet a validated clinical prognostic tool.
12. Treatment
There is no CEACAM16-specific or disease-modifying therapy. Management is rehabilitative and supportive:
- Hearing aids (amplification) for mild–moderate–severe loss (NCIT: Hearing Aid). First-line as thresholds rise.
- Cochlear implantation for severe-to-profound loss not adequately aided (NCIT: Cochlear Implant / Cochlear Implantation).
- Aural rehabilitation / speech-language therapy, assistive listening devices, and communication strategies.
- Genetic counseling for affected individuals and at-risk relatives (autosomal dominant, 50% transmission risk; account for possible de novo and for the distinct recessive allelic form).
- Avoidance of additional cochlear insults — noise protection and avoidance of ototoxic drugs where possible (supportive, not proven disease-modifying).
Pharmacotherapy / pharmacogenomics: None specific. Advanced therapeutics (gene/cell/RNA-based): Not available for DFNA4B. Current cochlear AAV gene therapy has clinical proof-of-concept only for recessive, hair-cell-intrinsic genes: AAV1-hOTOF partially restored hearing in children with DFNB9 — "AAV1-hOTOF gene therapy is safe and efficacious as a novel treatment for children with autosomal recessive deafness 9" PMID: 38280389 — and the field is expanding PMID: 39520052. Because CEACAM16 is a secreted extracellular-matrix protein and DFNA4B is dominant (dominant-negative), simple gene addition is unlikely to suffice; allele-specific silencing or gene editing would be required. The recessive LoF CEACAM16 form is conceptually more tractable for AAV-based gene replacement. No relevant clinical trials (NCT) target CEACAM16 at this time.
13. Prevention
- Primary prevention: Not possible for a germline dominant disorder. Reproductive options (preimplantation genetic testing, prenatal diagnosis) can prevent transmission where a familial variant is known and families elect to use them.
- Secondary prevention: Cascade genetic testing and periodic audiometric monitoring of at-risk/pre-symptomatic carriers enable early amplification. Because onset is late, standard newborn hearing screening will not detect pre-symptomatic carriers.
- Tertiary prevention: Prevent complications of untreated hearing loss (communication failure, social isolation, cognitive decline) through timely amplification/implantation and rehabilitation; minimize additional damage from noise and ototoxins.
- Genetic counseling: Central preventive tool — risk assessment, family planning, and clarification of the dominant vs. recessive allelic distinction.
- Immunization / public-health / environmental interventions: Not applicable (non-infectious, non-environmental etiology).
14. Other Species / Natural Disease
- Taxonomy / orthologs: Ceacam16 is conserved in mouse (Mus musculus, NCBI Taxon 10090) and other mammals; the mouse gene is the principal experimental ortholog. The TM and its protein constituents (TECTA, TECTB, CEACAM16, OTOG, OTOGL) are evolutionarily conserved across mammals, and the disease mechanism is conserved.
- Natural disease in other species: No well-characterized naturally occurring CEACAM16 deafness is documented in companion animals or wildlife (OMIA does not list an established natural CEACAM16 hearing-loss phenotype); the animal evidence is from engineered laboratory models (Section 15).
- Comparative biology: Mouse Ceacam16 models reproduce the human TM pathology, demonstrating strong cross-species conservation of the mechanism.
- Zoonotic potential: Not applicable (genetic disease).
15. Model Organisms
- Principal model — Ceacam16-null (knockout) mouse. This is the best-characterized model and strongly validates the human mechanism. Key findings:
- In the absence of CEACAM16, "TECTB levels are reduced, a clearly defined striated-sheet matrix does not develop, and Hensen's stripe … is absent" PMID: 25080593.
- CEACAM16 "is also shown to interact with TECTB, indicating that it may stabilize interactions between TECTA and TECTB" PMID: 25080593.
- Young null mice have largely normal ABR thresholds and DPOAEs but show enlarged stimulus-frequency and transiently-evoked emissions and spontaneous otoacoustic emissions in ~70% of homozygotes (vs <3% of wild-type); SOAE predominance >15 kHz correlates with loss of Hensen's stripe PMID: 25080593.
- The null shows accelerated age-related degradation/detachment of the TM PMID: 31249509.
- Related TM-mutant models that inform DFNA4B biology: Otoa (otoancorin)-deficient and Tecta mutant mice, in which SOAEs and TM detachment illuminate the general principle that TM defects destabilize the cochlear amplifier PMID: 26691158, PMID: 33340968; and a Tectb-C225Y knock-in showing TM disruption and noise susceptibility PMID: 42702881.
- Phenotype recapitulation: Good for the structural and micromechanical phenotype (TM matrix failure, loss of Hensen's stripe, abnormal OAEs, age-accelerated TM degradation). The null models the recessive (loss-of-function) biology directly; a dominant knock-in carrying a human missense variant (e.g., p.Gly169Arg) would be required to model the dominant-negative DFNA4B mechanism specifically — this is a current gap.
- Model limitations: Frequency ranges and cochlear tonotopy differ between mouse and human; young null mice retain near-normal thresholds, so the model captures early micromechanical instability better than end-stage human hearing loss; no dominant knock-in has been reported.
- Resources: MGI (mouse), and standard inner-ear physiology assays (ABR, DPOAE/SFOAE/SOAE, TM histology).
Mechanistic Model / Interpretation
DFNA4B is fundamentally a tectorial-membrane matrix disease. The unifying interpretation across human genetics and mouse physiology is that CEACAM16 is a molecular "staple" that cross-links the two tectorins (TECTA and TECTB) to build the TM's striated-sheet matrix and Hensen's stripe. Dominant missense variants introduce a defective monomer that poisons matrix assembly (dominant-negative / altered secretion), while biallelic LoF variants simply remove the staple (recessive). Either route weakens the TM's ability to couple to OHC stereocilia and to buffer the active cochlear amplifier. The physiological signature — abnormal and spontaneous otoacoustic emissions — reflects a destabilized but still-active amplifier, and with age the TM progressively degrades, producing the slowly progressive, high-frequency-first sensorineural hearing loss seen clinically.
| Feature | Dominant DFNA4B | Recessive (allelic) |
|---|---|---|
| Variant type | Missense (Ig domains) | LoF: nonsense, splice |
| Mechanism | Dominant-negative / altered secretion (poison monomer) | Protein absence |
| Inheritance | Autosomal dominant | Autosomal recessive |
| Onset | Late-onset, postlingual | Postlingual, progressive |
| Gene-therapy tractability | Hard (needs allele-specific silencing/editing) | More tractable (gene replacement) |
Evidence Base
| PMID | Contribution |
|---|---|
| 21368133 | Foundational: CEACAM16 maps to DFNA4, localizes to tallest OHC stereocilia tips and TM, colocalizes/co-IPs with α-tectorin. |
| 25080593 | Ceacam16-null mouse: reduced TECTB, no striated-sheet matrix, absent Hensen's stripe, SOAEs in 70%; CEACAM16 bridges TECTA–TECTB. |
| 25589040 | p.Gly169Arg dominant family; CEACAM16 is secreted; mutant has reduced secretion (deleterious). |
| 35292975 | p.Arg255Gly dominant family; mutant shows increased secretion — expands mechanism spectrum. |
| 39157884 | Large Russian family (p.Thr140Ile); defines late-onset (5–20 y), tinnitus + high-frequency onset, progression. |
| 26648831 | De novo p.Leu365Arg; confirms DFNA4B designation and postlingual phenotype. |
| 30514912 | Recessive LoF (p.Arg146Ter) — establishes distinct recessive mechanism. |
| 29703829 | Recessive splice variants (c.37G>T, c.662-1G>C). |
| 31249509 | Accelerated age-related TM degradation in Ceacam16-null. |
| 26691158, 34332206, 33340968 | SOAEs as biomarkers of TM defects across mouse mutants. |
| 42702881 | Related TM gene TECTB; noise susceptibility of heterozygous carriers (differential dx + GxE concept). |
| 38280389, 39520052 | Cochlear AAV gene therapy proof-of-concept — for recessive, hair-cell-intrinsic genes, contextualizing why DFNA4B remains molecularly untreated. |
Evidence-type mix: human clinical/genetic (family and de novo reports), in vitro functional (HEK293T secretion assays), and model-organism physiology (mouse Ceacam16 knockout). Mechanistic claims about TECTA/TECTB bridging and TM structure derive primarily from mouse data; human data establish gene–disease causation and the clinical phenotype.
Limitations and Knowledge Gaps
- Rarity and small samples. DFNA4B is defined by a handful of families/individuals; prevalence, penetrance, and expressivity are not quantitatively established.
- No dominant knock-in model. The mouse evidence is chiefly from the null (recessive-like) model; a knock-in of a human dominant missense variant is needed to test the dominant-negative mechanism directly.
- Divergent in vitro secretion phenotypes (reduced for p.Gly169Arg vs increased for p.Arg255Gly) suggest more than one biochemical route to dominance; the unifying mechanism at the matrix level is inferred, not fully resolved.
- Human TM tissue is inaccessible, so structural confirmation in patients (loss of Hensen's stripe, striated-sheet failure) relies on animal analogy.
- No natural-history study with standardized longitudinal audiometry; progression rates are described qualitatively.
- No disease-specific QoL, biomarker validation (e.g., clinical SOAE), or therapeutic development exists for DFNA4B.
Proposed Follow-up Experiments / Actions
- Generate a dominant Ceacam16 knock-in mouse (e.g., p.Gly169Arg or p.Arg255Gly) to directly test the dominant-negative mechanism and characterize TM structure, OAEs, and age-dependent hearing decline.
- Structural biology of CEACAM16 Ig domains (cryo-EM/crystallography or AlphaFold-guided modeling) to map how dominant variants disrupt TECTA/TECTB binding interfaces — informing allele-specific therapeutic design.
- Develop allele-specific silencing (ASO/siRNA) or CRISPR editing targeting the mutant allele, exploiting the recessive tolerance of CEACAM16 haploinsufficiency (biallelic LoF is required for recessive disease, implying heterozygous knockdown may be tolerated).
- Explore AAV gene replacement for the recessive LoF form as the more tractable near-term target, leveraging IHC/OHC-tropic capsids emerging from OTOF programs.
- Establish a prospective natural-history registry with serial audiometry and OAE testing to quantify progression, penetrance, and evaluate SOAEs as a clinical biomarker.
- Cascade genetic testing and counseling protocols for identified families, including reproductive-option discussion.
Report compiled from 28 reviewed publications and 5 confirmed findings. Evidence sources: human clinical/genetic, in vitro functional, and mouse model-organism studies.
Artifacts
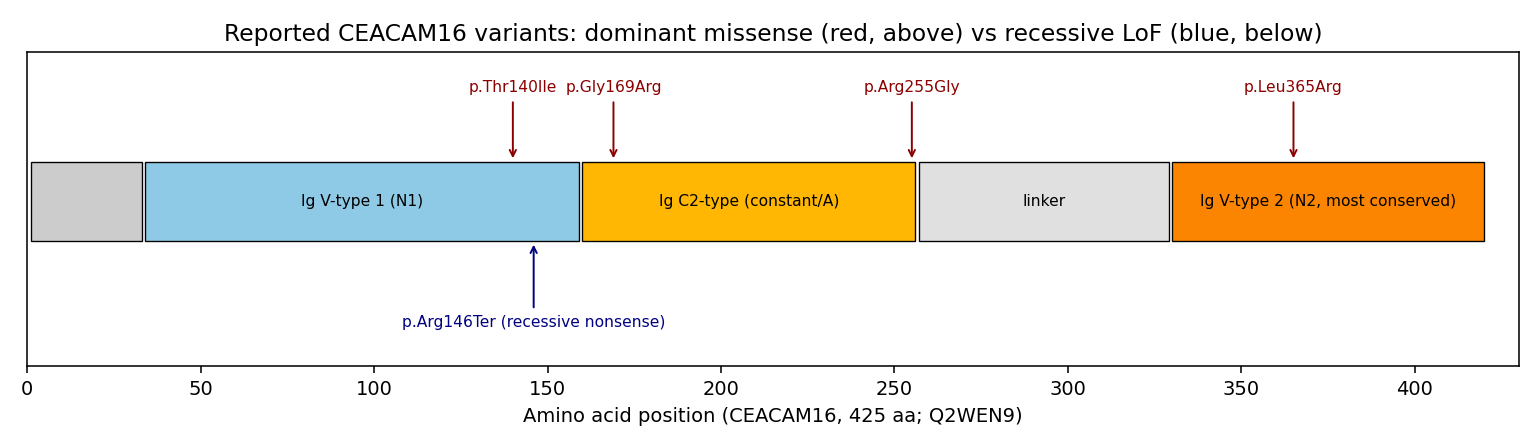
Reference Validation
Checked with linkml-reference-validator 0.2.1.
| Outcome | Count |
|---|---|
| References checked | 17 |
| Resolved | 17 |
| Unresolved (possible confabulation) | 0 |
| Unverifiable | 0 |
| Quoted claims checked | 6 |
| Quoted claims found in source | 4 |
| Quoted claims not found in source | 2 |
| References weighed for topical relevance | 17 |
| On topic | 15 |
| Off topic | 0 |
Quotes not found in the cited source
Searched the abstract, any retrieved full text, and the title. A quote drawn from a part of the paper that was not retrieved will appear here too, so check before treating one as invented:
1 of these was searched against an abstract alone, with no full text retrieved - marked abstract only below. Where full text can be fetched, re-running with it will settle them; where the source publishes only a summary to PubMed, as GeneReviews chapters do, it will not, and the quote has to be checked by hand against the chapter itself.
PMID:39157884: "start[ing] with tinnitus and threshold increase at high frequencies"- closest text in source: "Eleven family members suffer from hearing impairment, which starts with tinnitus and threshold increase at high frequencies, since the age of 5-20 years"
PMID:25080593(abstract only): "TECTB levels are reduced, a clearly defined striated-sheet matrix does not develop, and Hensen's stripe … is absent"- closest text in source: "In the absence of CEACAM16, TECTB levels are reduced, a clearly defined striated-sheet matrix does not develop, and Hensen's stripe, a prominent feature in the basal two-thirds of the TM in WT mice, is absent"
Term Validation
Checked with linkml-term-validator 0.4.5, through the ols: adapter.
| Outcome | Count |
|---|---|
| Terms checked | 22 |
| Resolved | 22 |
| Unresolved (possible confabulation) | 0 |
| Obsolete | 0 |
| Unverifiable | 0 |
| Terms whose name was checked | 11 |
| Terms named correctly | 3 |
| Terms named as a different term | 2 |
| Terms whose name is worth a second look | 6 |
Terms the report names something else
These identifiers resolve, so nothing about them looks wrong, and the ontology calls them something unrelated to what the report calls them. That usually means the identifier is not the one the sentence needs:
MONDO:0013823(2 mentions) - the report calls it "MONDO"; MONDO calls it autosomal dominant nonsyndromic hearing loss 4BHP:0008625(1 mention) - the report calls it "bilateral SNHL"; HP calls it Severe sensorineural hearing impairment
Terms whose name is worth a second look
The report's name for these is recognisably related to the term's own name without being one of them. A loose paraphrase reads the same way as a citation of the wrong sibling term - and so does a related synonym, which the ontology records precisely because it names something adjacent rather than the same thing - so these are listed rather than judged:
HP:0011476(1 mention) - the report calls it "postlingual sensorineural HL"; HP calls it Profound sensorineural hearing impairment, and lists "Profound sensorineural hearing loss" among its other namesCL:0000601(2 mentions) - the report calls it "outer hair cells"; CL calls it cochlear outer hair cellCL:0000589(2 mentions) - the report calls it "inner hair cells"; CL calls it cochlear inner hair cell, and lists "inner hair cell" among its other namesUBERON:0002233(2 mentions) - the report calls it "acellular tectorial membrane"; UBERON calls it tectorial membrane of cochlea, and lists "tectorial membrane" among its other namesUBERON:0002227(2 mentions) - the report calls it "organ of Corti"; UBERON calls it spiral organ of cochlea, and lists "organ of Corti" among its other namesGO:0031012(1 mention) - the report calls it "extracellular matrix / tectorial membrane"; GO calls it extracellular matrix