Acheiropodia
Acheiropodia is a very rare autosomal recessive congenital limb reduction disorder in which all four limbs show a stereotyped bilateral terminal transverse deficiency: the distal humerus and the distal tibial diaphysis are truncated, the radius, ulna and fibula are absent, and the hands and feet do not form. No other organ system is involved and heterozygous carriers are unaffected. Nearly all reported families are Brazilian and carry a homozygous deletion of about 12 kb that removes exon 4 of LMBR1 while leaving the intron-5 ZRS enhancer of SHH intact. The deleted interval has no enhancer activity of its own; it carries three CTCF sites that anchor the long-range contact between the ZRS and the SHH promoter, and in patient cells that contact is lost. The disease is therefore modelled as a loss-of-regulatory-contact disorder in which limb-bud SHH expression is presumed to fail, the opposite direction of travel from the gain-of-ectopic-expression ZRS malformations curated in ZRS-Related Limb Malformation.
Ask OpenScientist
Ask a research question about Acheiropodia. OpenScientist will conduct autonomous deep research using the Disorder Mechanisms Knowledge Base and PubMed literature (typically 10-30 minutes).
Do not include personal health information in your question. Questions and results are cached in your browser's local storage.
Inheritance
1Show evidence (2 references)
Pathophysiology
4Show evidence (2 references)
Show evidence (2 references)
Show evidence (1 reference)
Show evidence (2 references)
Pathograph
Phenotypes
7Limbs 6
Show evidence (2 references)
Show evidence (2 references)
Show evidence (2 references)
Show evidence (2 references)
Show evidence (1 reference)
Show evidence (1 reference)
Other 1
Show evidence (1 reference)
Genetic Associations
1Show evidence (2 references)
Variants (2)
Show evidence (2 references)
Show evidence (1 reference)
Diagnosis
2Show evidence (1 reference)
Show evidence (2 references)
Prevalence
1Show evidence (2 references)
Animal Models
2Show evidence (1 reference)
Show evidence (1 reference)
Show evidence (1 reference)
Source YAML
click to showname: Acheiropodia
creation_date: '2026-10-08T15:37:22Z'
description: >-
Acheiropodia is a very rare autosomal recessive congenital limb reduction
disorder in which all four limbs show a stereotyped bilateral terminal
transverse deficiency: the distal humerus and the distal tibial diaphysis are
truncated, the radius, ulna and fibula are absent, and the hands and feet do
not form. No other organ system is involved and heterozygous carriers are
unaffected. Nearly all reported families are Brazilian and carry a homozygous
deletion of about 12 kb that removes exon 4 of LMBR1 while leaving the
intron-5 ZRS enhancer of SHH intact. The deleted interval has no enhancer
activity of its own; it carries three CTCF sites that anchor the long-range
contact between the ZRS and the SHH promoter, and in patient cells that
contact is lost. The disease is therefore modelled as a
loss-of-regulatory-contact disorder in which limb-bud SHH expression is presumed to fail, the
opposite direction of travel from the gain-of-ectopic-expression ZRS
malformations curated in ZRS-Related Limb Malformation.
category: Mendelian
parents:
- Congenital Disorder
- Limb Malformation
synonyms:
- acheiropody
- ACHP
- handless and footless families of Brazil
disease_term:
preferred_term: acheiropodia
term:
id: MONDO:0008700
label: acheiropody
prevalence:
- population: Brazil
measure_type: BIRTH_PREVALENCE
prevalence_class: BAND_1_9_PER_1000000
rate_per_100000: 0.4
rate_denominator: LIVE_BIRTHS
notes: >-
Estimated at about 1 in 250,000 births in Brazil (Freire-Maia et al. 1975,
as cited by Ianakiev et al.). Outside Brazil only isolated sibships have
been reported (Puerto Rico, Argentina).
evidence:
- reference: PMID:11090342
reference_title: "Acheiropodia is caused by a genomic deletion in C7orf2, the human orthologue of the Lmbr1 gene."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: BACKGROUND
snippet: >-
The incidence of acheiropodia in Brazil has been estimated to be ~1/250,000
births
explanation: >-
States the Brazilian birth-incidence estimate, cited by the gene
identification paper from an earlier epidemiological study.
- reference: PMID:26294167
reference_title: "[Acheiropodia: first case report in Argentina]."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: >-
These patients are the first reported cases of acheiropodia syndrome in
Argentina.
explanation: >-
Documents that cases outside Brazil are exceptional, supporting the
geographically restricted, founder-allele epidemiology.
inheritance:
- name: Autosomal Recessive
inheritance_term:
preferred_term: Autosomal recessive inheritance
term:
id: HP:0000007
label: Autosomal recessive inheritance
description: >-
Affected individuals are homozygous for the deletion, most are offspring of
consanguineous unions, and heterozygous parents are phenotypically normal.
evidence:
- reference: PMID:11090342
reference_title: "Acheiropodia is caused by a genomic deletion in C7orf2, the human orthologue of the Lmbr1 gene."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: >-
Acheiropodia is an autosomal recessive developmental disorder presenting
with bilateral congenital amputations of the upper and lower extremities
and aplasia of the hands and feet.
explanation: >-
Establishes the autosomal recessive mode of inheritance in the
gene-identification study of five Brazilian families.
- reference: PMID:2363435
reference_title: "Acheiropodia: report on four new Brazilian patients."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: >-
This autosomal recessive condition involves all 4 limbs with a
well-defined pattern of defects.
explanation: >-
An independent Brazilian case series describing the condition as
autosomal recessive.
genetic:
- name: LMBR1
gene_term:
preferred_term: LMBR1
term:
id: hgnc:13243
label: LMBR1
affected_regions:
- name: LMBR1 exon 4 CTCF scaffold region
chromosomal_region: 7q36
within_gene:
preferred_term: LMBR1
term:
id: hgnc:13243
label: LMBR1
description: >-
The roughly 12 kb interval around LMBR1 exon 4 removed by the Brazilian
acheiropodia allele. It carries three CTCF sites (LSC3-5) in convergent
orientation to SHH and showed no limb enhancer activity in a transgenic
assay. The ZRS enhancer in LMBR1 intron 5 lies outside the deletion and
is intact.
association: >-
Homozygous deletions within LMBR1 cause acheiropodia. The common Brazilian
allele removes LMBR1 exon 4 and the CTCF sites that tether the ZRS to the
SHH promoter; SHH, about 1 Mb away, is the presumed regulatory target.
relationship_type: CAUSATIVE
variant_origin: GERMLINE
features: >-
LMBR1 is bound here because the causal deletions physically remove LMBR1
sequence and because OMIM and MONDO assign acheiropodia to LMBR1. The
functional evidence does not support loss of the LMBR1 protein as the
mechanism: Ushiki et al. note that a mouse deletion of Lmbr1 exons 1-3 has
no limb phenotype and attribute the disease to loss of the CTCF-mediated
ZRS-SHH promoter contact. The exon-4-skipped transcript and premature stop
codon reported by Ianakiev et al. are therefore recorded as a consequence of
the deletion, not as the disease mechanism.
evidence:
- reference: PMID:11090342
reference_title: "Acheiropodia is caused by a genomic deletion in C7orf2, the human orthologue of the Lmbr1 gene."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: >-
Analysis of C7orf2, the human orthologue of the mouse Lmbr1 gene,
identified a deletion in all five families, thus identifying a common
acheiropodia mutation.
explanation: >-
Identifies a shared homozygous LMBR1 (C7orf2) deletion in five Brazilian
acheiropodia families.
- reference: PMID:10780921
reference_title: "A minimalist approach to gene mapping: locating the gene for acheiropodia, by homozygosity analysis."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: >-
In this region, a maximum two-point LOD score of 3.81 (4.2 with multipoint
analysis) was achieved, and a homozygous haplotype spanning a region of
11.7 cM was seen in all affected in this pedigree.
explanation: >-
Homozygosity mapping and linkage place the acheiropodia locus at 7q36,
the region containing LMBR1 and SHH.
variants:
- name: Homozygous 12 kb deletion encompassing LMBR1 exon 4
description: >-
The common Brazilian founder allele, found homozygous in all five families
studied by Ianakiev et al. Whole-genome sequencing of one proband refined
it to a 12 kb deletion surrounding LMBR1 exon 4. At the RNA level it
produces an LMBR1 transcript lacking exon 4 with a downstream premature
stop codon. It removes three CTCF sites that tether the ZRS to the SHH
promoter; the ZRS itself is not deleted and carried no rare variant in
the sequenced proband.
gene:
preferred_term: LMBR1
term:
id: hgnc:13243
label: LMBR1
regulatory_target_gene:
preferred_term: SHH
term:
id: hgnc:10848
label: SHH
variant_type: deletion
genomic_contexts:
- coding sequence
- intron
affected_regions:
- name: LMBR1 exon 4 CTCF scaffold region
chromosomal_region: 7q36
within_gene:
preferred_term: LMBR1
term:
id: hgnc:13243
label: LMBR1
description: >-
The deleted interval containing CTCF sites LSC3-5; it lies within
LMBR1, centromeric to the intact ZRS enhancer.
clinical_significance: PATHOGENIC
functional_effects:
- function: CTCF-mediated contact between the ZRS enhancer and the SHH promoter
description: >-
In patient lymphoblastoid cells the SHH promoter no longer contacts the
ZRS and instead contacts a CTCF site centromeric to it. Reduced
limb-bud SHH expression is inferred from this and from mouse ZRS and
Shh knockouts; limb-bud SHH expression has not been measured for the
human allele. In patient lymphoblastoid cells SHH mRNA was low overall
and lower than in wild-type cells.
type: loss of enhancer-promoter contact
affected_developmental_stage: embryonic limb development
regulatory_mechanism: >-
Deletion of three convergent CTCF sites that act as a scaffold
bringing the ZRS to the SHH promoter.
evidence:
- reference: PMID:33863876
reference_title: "Deletion of CTCF sites in the SHH locus alters enhancer-promoter interactions and leads to acheiropodia."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: >-
Using whole-genome sequencing, we fine-mapped the acheiropodia-associated
region to 12 kb and show that it does not function as an enhancer.
explanation: >-
Defines the deletion boundaries in a patient genome and establishes that
the deleted interval is not itself an enhancer.
- reference: PMID:11090342
reference_title: "Acheiropodia is caused by a genomic deletion in C7orf2, the human orthologue of the Lmbr1 gene."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: >-
It leads to the production of a C7orf2 transcript lacking exon 4 and
introduces a premature stop codon downstream of exon 3.
explanation: >-
Reports the transcript-level consequence of the deletion for LMBR1.
- name: Homozygous 102 kb deletion at 7q36.3 spanning LMBR1
description: >-
A nullizygous 102 kb deletion spanning LMBR1, found by molecular
karyotyping in a Saudi patient with acheiropodia. Ushiki et al. state that
homozygous ZRS deletion in humans causes acheiropodia; the abstract of the
original report does not give the breakpoints, so whether this allele
removes the ZRS itself is recorded from that secondary statement only.
gene:
preferred_term: LMBR1
term:
id: hgnc:13243
label: LMBR1
regulatory_target_gene:
preferred_term: SHH
term:
id: hgnc:10848
label: SHH
variant_type: deletion
clinical_significance: LIKELY_PATHOGENIC
evidence:
- reference: PMID:26749485
reference_title: "Novel copy number variants and major limb reduction malformation: Report of three cases."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: >-
a third patient with acheiropodia who had a nullizygous deletion of 102 kb
on 7q36.3 spanning LMBR1
explanation: >-
A second, larger homozygous deletion at the LMBR1 locus in a
non-Brazilian patient with acheiropodia.
pathophysiology:
- name: LMBR1 Exon 4 Region Deletion
biological_scale: MOLECULAR
role: trigger
description: >-
Homozygous deletion of the interval surrounding LMBR1 exon 4 (about 12 kb in
the Brazilian founder allele). The deletion removes three CTCF-bound sites
(LSC3-5) in convergent orientation to SHH, which lies about 1 Mb away and is
itself intact. LMBR1 is the host of the deleted sequence, not the proposed
effector.
genetic_context:
allele_type: homozygous ~12 kb deletion encompassing LMBR1 exon 4
variant_type: deletion
genomic_contexts:
- coding sequence
- intron
affected_regions:
- name: LMBR1 exon 4 CTCF scaffold region
chromosomal_region: 7q36
within_gene:
preferred_term: LMBR1
term:
id: hgnc:13243
label: LMBR1
description: >-
Deleted CTCF sites LSC3-5; the intron-5 ZRS enhancer lies outside the
deletion.
zygosity: HOMOZYGOUS
variant_origin: GERMLINE
genes:
- preferred_term: LMBR1
term:
id: hgnc:13243
label: LMBR1
evidence:
- reference: PMID:11090342
reference_title: "Acheiropodia is caused by a genomic deletion in C7orf2, the human orthologue of the Lmbr1 gene."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: >-
Analysis of C7orf2, the human orthologue of the mouse Lmbr1 gene,
identified a deletion in all five families, thus identifying a common
acheiropodia mutation.
explanation: >-
Establishes the shared homozygous deletion at LMBR1 as the initiating
lesion.
- reference: PMID:33863876
reference_title: "Deletion of CTCF sites in the SHH locus alters enhancer-promoter interactions and leads to acheiropodia."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: >-
Using whole-genome sequencing, we fine-mapped the acheiropodia-associated
region to 12 kb and show that it does not function as an enhancer.
explanation: >-
Refines the deletion to 12 kb and shows it is not an enhancer, pointing to
a structural rather than enhancer-loss mechanism.
downstream:
- target: Loss of ZRS-SHH Promoter Chromatin Contact
causal_link_type: DIRECT
description: >-
Removing the CTCF scaffold sites abolishes the ZRS-SHH promoter contact in
patient cells.
evidence:
- reference: PMID:33863876
reference_title: "Deletion of CTCF sites in the SHH locus alters enhancer-promoter interactions and leads to acheiropodia."
supports: SUPPORT
evidence_source: IN_VITRO
snippet: >-
CTCF and RAD21 ChIP-seq together with 4C-seq and DNA FISH identify three
CTCF sites within the acheiropodia-deleted region that mediate the
interaction between the ZRS and the SHH promoter.
explanation: >-
Chromatin assays in patient and control lymphoblastoid cells show that
the deleted CTCF sites mediate the ZRS-SHH promoter interaction.
- name: Loss of ZRS-SHH Promoter Chromatin Contact
biological_scale: MOLECULAR
description: >-
In cells homozygous for the deletion, the SHH promoter no longer interacts
with the ZRS and instead shows increased interaction with a CTCF site (LSC1)
centromeric to the ZRS. This was measured in lymphoblastoid cells, not in
embryonic limb mesenchyme.
biological_processes:
- preferred_term: CTCF-mediated ZRS-SHH promoter looping
term:
id: GO:0140588
label: chromatin looping
modifier: ABNORMAL
genes:
- preferred_term: SHH
term:
id: hgnc:10848
label: SHH
evidence:
- reference: PMID:33863876
reference_title: "Deletion of CTCF sites in the SHH locus alters enhancer-promoter interactions and leads to acheiropodia."
supports: SUPPORT
evidence_source: IN_VITRO
snippet: >-
we did not observe interactions with the ZRS and instead saw increased
interactions between the SHH promoter and LSC1
explanation: >-
4C-seq from the SHH promoter in proband cells shows loss of ZRS contact
and a substituted contact with LSC1.
- reference: PMID:33863876
reference_title: "Deletion of CTCF sites in the SHH locus alters enhancer-promoter interactions and leads to acheiropodia."
supports: SUPPORT
evidence_source: IN_VITRO
snippet: >-
This interaction is substituted with other CTCF sites centromeric to the
ZRS in the disease state.
explanation: >-
Summarizes the rewired contact in the disease state.
downstream:
- target: Loss of Limb-Bud SHH Expression
causal_link_type: UNKNOWN
description: >-
Loss of enhancer-promoter contact is proposed to prevent ZRS-driven SHH
transcription in the limb bud. This step is inferred. SHH expression in
the embryonic limb bud has not been measured for the human allele; the
only human measurement is in EBV-transformed lymphoblastoid cells, where
SHH mRNA was low overall and lower in proband than wild-type cells. The
orthologous mouse deletion leaves Shh expression unchanged because mouse
CTCF architecture differs. Mouse data cut both ways: deleting CTCF sites
around the ZRS reduced limb Shh by about half with no limb phenotype,
whereas an inversion that disrupted ZRS-Shh contact abolished ZPA Shh
and truncated the limb. A partial contact defect may therefore not be
enough, and how completely the human deletion silences limb SHH is
unknown.
evidence:
- reference: PMID:33863876
reference_title: "Deletion of CTCF sites in the SHH locus alters enhancer-promoter interactions and leads to acheiropodia."
supports: SUPPORT
directness: INDIRECT
evidence_source: IN_VITRO
snippet: >-
We also checked the mRNA expression of SHH in the wild-type and mutant
cells, observing overall low expression levels that were significantly
higher in wild-type versus proband cells
explanation: >-
Reduced SHH expression in proband cells is consistent with the lost
contact lowering SHH output. It is indirect because these are
EBV-transformed lymphoblastoid cells, not limb-bud mesenchyme, and SHH
is barely expressed in them.
- reference: PMID:33863876
reference_title: "Deletion of CTCF sites in the SHH locus alters enhancer-promoter interactions and leads to acheiropodia."
supports: REFUTE
directness: INDIRECT
evidence_source: MODEL_ORGANISM
quote_role: BACKGROUND
snippet: >-
Although these triple deletions led to a 52% reduction of Shh expression
in E10.5 limb buds, no limb abnormalities were observed.
explanation: >-
Summarizes an earlier mouse study (Paliou et al.) in which deleting CTCF
sites around the ZRS weakened ZRS-Shh interaction and halved limb Shh
without truncating the limb. This argues that losing CTCF-mediated
contact does not by itself abolish limb SHH, at least in mouse.
- reference: PMID:33863876
reference_title: "Deletion of CTCF sites in the SHH locus alters enhancer-promoter interactions and leads to acheiropodia."
supports: SUPPORT
directness: INDIRECT
evidence_source: MODEL_ORGANISM
quote_role: BACKGROUND
snippet: >-
This led to a complete loss of Shh expression in the ZPA and a limb
truncation phenotype, similar to the ZRS homozygous knockout.
explanation: >-
Summarizes an earlier mouse study (Symmons et al.) in which a 450 kb
inversion disrupting ZRS-Shh interaction abolished ZPA Shh and
truncated the limb, showing that losing the contact can be enough to
cause an acheiropodia-like phenotype.
- reference: PMID:33863876
reference_title: "Deletion of CTCF sites in the SHH locus alters enhancer-promoter interactions and leads to acheiropodia."
supports: SUPPORT
directness: INDIRECT
evidence_source: OTHER
snippet: >-
We reasoned that the likely cause of the acheiropodia in this proband is
altered SHH expression during limb development.
explanation: >-
The authors' inference linking the lost contact to altered limb SHH
expression, made from the chromatin data and prior knockout models
rather than from a limb expression measurement.
- reference: PMID:40604280
reference_title: "Range extender mediates long-distance enhancer activity."
supports: SUPPORT
directness: INDIRECT
evidence_source: MODEL_ORGANISM
snippet: >-
maintains limb activity at short range, but loses its long-range
activity, resulting in severe limb reduction in knock-in mice.
explanation: >-
In mouse, a ZRS with mutated homeodomain motifs that keeps its enhancer activity but cannot act over
long range still produces severe limb reduction. This shows that
failure of the long-distance ZRS-SHH communication, rather than loss of
the enhancer itself, is sufficient for limb reduction; it is indirect
because the mouse lesion alters ZRS motifs, not the CTCF sites deleted
in acheiropodia.
- name: Loss of Limb-Bud SHH Expression
biological_scale: MOLECULAR
mechanism_confidence: PROVISIONAL
regulatory_category: mLOE
conforms_to: limb_digit_patterning_serial_homology#Limb Patterning Signal Perturbation
description: >-
Limb-specific loss of SHH expression from the zone of polarizing activity,
with SHH expression elsewhere unaffected (hence modular loss of
expression). Direct support comes from mouse: deleting the ZRS abolishes
limb-bud Shh expression and truncates the limb distal to the
stylopod-zeugopod junction. Unlike the patterning conformers of the module,
the consequence here is failure of distal outgrowth rather than altered
digit number or identity.
genes:
- preferred_term: SHH
term:
id: hgnc:10848
label: SHH
molecular_functions:
- preferred_term: SHH ligand binding to its patched receptor
term:
id: GO:0005113
label: patched binding
modifier: DECREASED
biological_processes:
- preferred_term: limb-bud SHH signaling
term:
id: GO:0007224
label: smoothened signaling pathway
modifier: DECREASED
locations:
- preferred_term: limb bud
term:
id: UBERON:0004347
label: limb bud
evidence:
- reference: PMID:15677727
reference_title: "Elimination of a long-range cis-regulatory module causes complete loss of limb-specific Shh expression and truncation of the mouse limb."
supports: SUPPORT
directness: INDIRECT
evidence_source: MODEL_ORGANISM
snippet: >-
deletion of this conserved intronic sequence by targeted mutation in the
mouse results in a complete loss of Shh expression in the limb bud
explanation: >-
In mouse, removing the ZRS abolishes limb-bud Shh expression; indirect for
the human 12 kb allele, which spares the ZRS.
downstream:
- target: Failure of Distal Limb Skeletal Element Formation
causal_link_type: DIRECT
evidence:
- reference: PMID:15677727
reference_title: "Elimination of a long-range cis-regulatory module causes complete loss of limb-specific Shh expression and truncation of the mouse limb."
supports: SUPPORT
evidence_source: MODEL_ORGANISM
snippet: >-
complete loss of Shh expression in the limb bud and degeneration of
skeletal elements distal to the stylopod/zygopod junction
explanation: >-
Loss of limb Shh in the ZRS-deleted mouse produces distal limb
truncation at the same level as in acheiropodia.
- name: Failure of Distal Limb Skeletal Element Formation
biological_scale: TISSUE
description: >-
Skeletal elements distal to the stylopod-zeugopod junction fail to form or
degenerate, leaving a truncated distal humerus and tibia and no radius,
ulna, fibula, hand or foot. All four limbs are affected symmetrically.
biological_processes:
- preferred_term: embryonic limb morphogenesis
term:
id: GO:0030326
label: embryonic limb morphogenesis
modifier: ABNORMAL
evidence:
- reference: PMID:15677727
reference_title: "Elimination of a long-range cis-regulatory module causes complete loss of limb-specific Shh expression and truncation of the mouse limb."
supports: SUPPORT
evidence_source: MODEL_ORGANISM
snippet: >-
This sequence contains a major limb-specific Shh enhancer that is
necessary for distal limb development.
explanation: >-
Establishes the requirement of ZRS-driven Shh for distal limb
development in mouse.
- reference: PMID:11090342
reference_title: "Acheiropodia is caused by a genomic deletion in C7orf2, the human orthologue of the Lmbr1 gene."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: >-
Acheiropodia is an autosomal recessive developmental disorder presenting
with bilateral congenital amputations of the upper and lower extremities
and aplasia of the hands and feet.
explanation: >-
The human phenotype is a bilateral distal limb truncation.
downstream:
- target: Four-Limb Terminal Transverse Deficiency
- target: Absent Forearms and Hands
- target: Absent Feet
- target: Fibular Aplasia
- target: Distal Humeral Truncation
- target: Distal Tibial Truncation
phenotypes:
- name: Four-Limb Terminal Transverse Deficiency
category: Musculoskeletal
frequency: VERY_FREQUENT
description: >-
Bilateral congenital amputation-like terminal deficiency of both upper and
both lower limbs.
phenotype_term:
preferred_term: Terminal transverse deficiency of all four limbs
term:
id: HP:6000818
label: Transverse terminal limb defect
evidence:
- reference: PMID:11090342
reference_title: "Acheiropodia is caused by a genomic deletion in C7orf2, the human orthologue of the Lmbr1 gene."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: >-
Acheiropodia is an autosomal recessive developmental disorder presenting
with bilateral congenital amputations of the upper and lower extremities
and aplasia of the hands and feet.
explanation: >-
Defines the condition as bilateral terminal deficiency of upper and lower
limbs.
- reference: PMID:33863876
reference_title: "Deletion of CTCF sites in the SHH locus alters enhancer-promoter interactions and leads to acheiropodia."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: >-
The proband has terminal transverse hemimelia of the four limbs with
truncation of both hands and feet
explanation: >-
Describes the molecularly characterized proband as having terminal
transverse deficiency of all four limbs.
- name: Absent Forearms and Hands
category: Musculoskeletal
frequency: VERY_FREQUENT
description: >-
The radius, ulna, carpals, metacarpals and phalanges are absent bilaterally.
Some affected individuals carry a single rudimentary digit on the upper limb
stump.
phenotype_term:
preferred_term: Absent forearms and hands
term:
id: HP:6000948
label: Hemimelia of forearm
evidence:
- reference: PMID:11090342
reference_title: "Acheiropodia is caused by a genomic deletion in C7orf2, the human orthologue of the Lmbr1 gene."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: >-
and aplasia of the radius, ulna, fibula and of the carpal, metacarpal,
tarsal, metatarsal, and phalangeal bones
explanation: >-
The defining pattern includes absence of the forearm bones and of all
hand bones.
- reference: PMID:26294167
reference_title: "[Acheiropodia: first case report in Argentina]."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: >-
Both twins had limb reduction defect including hands, forearms, feet and
fibulae.
explanation: >-
Independent report of absent hands and forearms in an Argentinian sibship.
- name: Absent Feet
category: Musculoskeletal
frequency: VERY_FREQUENT
phenotype_term:
preferred_term: Absent feet
term:
id: HP:0011301
label: Absent foot
evidence:
- reference: PMID:11090342
reference_title: "Acheiropodia is caused by a genomic deletion in C7orf2, the human orthologue of the Lmbr1 gene."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: >-
The affected individuals have no feet, hands, or forearms.
explanation: >-
Absence of the feet in affected members of a Brazilian pedigree.
- reference: PMID:26294167
reference_title: "[Acheiropodia: first case report in Argentina]."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: >-
Both twins had limb reduction defect including hands, forearms, feet and
fibulae.
explanation: >-
Absent feet in an independent sibship.
- name: Fibular Aplasia
category: Musculoskeletal
frequency: VERY_FREQUENT
phenotype_term:
preferred_term: Fibular aplasia
term:
id: HP:0002990
label: Fibular aplasia
evidence:
- reference: PMID:11090342
reference_title: "Acheiropodia is caused by a genomic deletion in C7orf2, the human orthologue of the Lmbr1 gene."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: >-
and aplasia of the radius, ulna, fibula and of the carpal, metacarpal,
tarsal, metatarsal, and phalangeal bones
explanation: >-
Fibular aplasia is part of the defining skeletal pattern.
- reference: PMID:26294167
reference_title: "[Acheiropodia: first case report in Argentina]."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: >-
Both twins had limb reduction defect including hands, forearms, feet and
fibulae.
explanation: >-
Fibular absence in an independent sibship.
- name: Distal Humeral Truncation
category: Musculoskeletal
frequency: VERY_FREQUENT
description: >-
The distal humeral epiphysis is absent, so the upper limb ends in a
truncated humerus.
phenotype_term:
preferred_term: Truncated distal humeri
term:
id: HP:0005025
label: Hypoplastic distal humeri
evidence:
- reference: PMID:11090342
reference_title: "Acheiropodia is caused by a genomic deletion in C7orf2, the human orthologue of the Lmbr1 gene."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: >-
consisting of a complete amputation of the distal epiphysis of the
humerus
explanation: >-
Absence of the distal humeral epiphysis is part of the defining pattern.
- name: Distal Tibial Truncation
category: Musculoskeletal
frequency: VERY_FREQUENT
description: >-
The tibia is truncated in its distal diaphysis, leaving a short lower-leg
stump.
phenotype_term:
preferred_term: Truncated distal tibiae
term:
id: HP:0005772
label: Aplasia/Hypoplasia of the tibia
notes: >-
The proximal tibia is present and the distal diaphysis is missing, a partial
absence. HPO has no term for distal tibial aplasia or truncation (ols:hp
search l~tibia, 2026-10-12): Absent tibia (HP:0009556) overstates it,
Proximal tibial hypoplasia (HP:0006379) names the wrong end, and Short tibia
(HP:0005736) implies shortening only. Aplasia/Hypoplasia of the tibia covers
partial absence without asserting which.
evidence:
- reference: PMID:11090342
reference_title: "Acheiropodia is caused by a genomic deletion in C7orf2, the human orthologue of the Lmbr1 gene."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: >-
amputation of the distal part of the tibial diaphysis
explanation: >-
Truncation of the distal tibial diaphysis is part of the defining pattern.
- name: Bohomoletz Bone
category: Musculoskeletal
frequency: OCCASIONAL
description: >-
A small ectopic bone at the distal end of the humeral stump, present in some
but not all patients.
notes: >-
Left unbound. HPO searches l~Bohomoletz, l~ectopic bone, l~accessory bone
and l~supernumerary bone (ols:hp, 2026-10-08) returned no term for an
ectopic bone at a limb stump; the hits were carpal, tarsal, metacarpal and
axial supernumerary-bone terms that do not describe this finding.
evidence:
- reference: PMID:2363435
reference_title: "Acheiropodia: report on four new Brazilian patients."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: >-
Two of the patients described here had a small bone fragment in the upper
stumps (Bohomoletz bone), an uncommon finding in acheiropodia.
explanation: >-
Reports the Bohomoletz bone and that it is uncommon.
diagnosis:
- name: Skeletal radiography of the limb stumps
description: >-
Plain radiographs define which skeletal elements are truncated or absent and
identify residual ossicles such as the Bohomoletz bone, which distinguishes
the acheiropodia pattern from other transverse limb deficiencies.
diagnosis_term:
preferred_term: radiography of the limbs
term:
id: NCIT:C38101
label: X-Ray Imaging
evidence:
- reference: PMID:11090342
reference_title: "Acheiropodia is caused by a genomic deletion in C7orf2, the human orthologue of the Lmbr1 gene."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: >-
small fingerlike appendages, bilaterally, which, on x-ray have been
recognized as the Bohomoletz bone.
explanation: >-
Radiographs were used to identify residual stump structures as the
Bohomoletz bone in an affected proband.
- name: Molecular detection of the LMBR1 exon 4 deletion
description: >-
Confirmation by detecting the homozygous deletion around LMBR1 exon 4, by a
targeted genomic or transcript assay for the known founder allele or by
whole-genome sequencing that can resolve structural variants. Coding-only
exome analysis may miss a deletion whose effect is regulatory.
diagnosis_term:
preferred_term: genetic testing
term:
id: NCIT:C15709
label: Genetic Testing
evidence:
- reference: PMID:11090342
reference_title: "Acheiropodia is caused by a genomic deletion in C7orf2, the human orthologue of the Lmbr1 gene."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: >-
The deletion was identified at both the genomic-DNA and mRNA level.
explanation: >-
The founder deletion is detectable in genomic DNA and as an
exon-4-skipped transcript.
- reference: PMID:33863876
reference_title: "Deletion of CTCF sites in the SHH locus alters enhancer-promoter interactions and leads to acheiropodia."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: >-
Using whole-genome sequencing, we fine-mapped the acheiropodia-associated
region to 12 kb and show that it does not function as an enhancer.
explanation: >-
Whole-genome sequencing resolved the deletion breakpoints in a proband.
animal_models:
- name: Mouse ZRS targeted deletion
species: Mus musculus
genotype: Homozygous targeted deletion of the ZRS in Lmbr1 intron 5
publication: PMID:15677727
description: >-
Targeted deletion of the conserved intronic limb enhancer abolishes limb-bud
Shh expression and truncates the limbs distal to the stylopod-zeugopod
junction, phenocopying acheiropodia.
modeled_mechanisms:
- target: Loss of Limb-Bud SHH Expression
relationship: RECAPITULATES
fidelity: MODERATE
model_scale: MOLECULAR
description: >-
Removing the enhancer abolishes limb-bud Shh expression, the presumed
consequence of the human deletion.
limitations: >-
The mouse allele deletes the enhancer itself, whereas the Brazilian human
allele spares the ZRS and removes CTCF scaffold sites; it models the
expression endpoint, not the contact-loss route.
evidence:
- reference: PMID:15677727
reference_title: "Elimination of a long-range cis-regulatory module causes complete loss of limb-specific Shh expression and truncation of the mouse limb."
supports: SUPPORT
evidence_source: MODEL_ORGANISM
snippet: >-
deletion of this conserved intronic sequence by targeted mutation in the
mouse results in a complete loss of Shh expression in the limb bud
explanation: >-
Directly measures loss of limb-bud Shh in the model.
- target: Failure of Distal Limb Skeletal Element Formation
relationship: RECAPITULATES
fidelity: HIGH
model_scale: TISSUE
description: >-
Distal skeletal elements degenerate at the same proximodistal level as in
acheiropodia.
evidence:
- reference: PMID:15677727
reference_title: "Elimination of a long-range cis-regulatory module causes complete loss of limb-specific Shh expression and truncation of the mouse limb."
supports: SUPPORT
evidence_source: MODEL_ORGANISM
snippet: >-
complete loss of Shh expression in the limb bud and degeneration of
skeletal elements distal to the stylopod/zygopod junction
explanation: >-
The skeletal outcome matches the human truncation pattern.
- name: Mouse orthologous 12 kb acheiropodia-region knockout
species: Mus musculus
genotype: Homozygous deletion of the mouse sequence orthologous to the human 12 kb acheiropodia deletion
publication: PMID:33863876
description: >-
Knockout of the orthologous 12 kb region has no limb phenotype and no change
in limb Shh expression, attributed to differences in CTCF site number and
orientation between the mouse and human loci.
modeled_mechanisms:
- target: Loss of ZRS-SHH Promoter Chromatin Contact
relationship: FAILS_TO_RECAPITULATE
fidelity: LOW
description: >-
The orthologous deletion does not reproduce the human disease, so the
mouse cannot be used to test the contact-loss mechanism.
limitations: >-
The mouse orthologous region carries a different set of CTCF motifs and
shows poor conservation with the human interval.
divergences:
- divergence_type: SPECIES_MISMATCH
materiality: INVALIDATING
description: >-
Mouse has one CTCF peak in the orthologous interval where human has
three convergent sites (LSC3-5), so the deletion does not remove the
scaffold the human allele removes.
evidence:
- reference: PMID:33863876
reference_title: "Deletion of CTCF sites in the SHH locus alters enhancer-promoter interactions and leads to acheiropodia."
supports: SUPPORT
evidence_source: MODEL_ORGANISM
snippet: >-
Mouse knockouts of the orthologous 12 kb sequence have no apparent
abnormalities, showcasing the challenges in modelling CTCF alterations in
animal models due to inherent motif differences between species.
explanation: >-
Reports the negative mouse result and its species-difference
explanation.
references:
- reference: PMID:11090342
title: Acheiropodia is caused by a genomic deletion in C7orf2, the human orthologue of the Lmbr1 gene.
- reference: PMID:33863876
title: Deletion of CTCF sites in the SHH locus alters enhancer-promoter interactions and leads to acheiropodia.
notes: >-
Acheiropodia is curated separately from ZRS-Related Limb Malformation. Both
map to the LMBR1-SHH locus, but the ZRS spectrum arises from heterozygous
gain-of-ectopic-expression alleles producing polydactyly, while acheiropodia
arises from homozygous deletions that remove regulatory contact and produce
limb reduction. No treatment is curated: no source describing management
beyond case descriptions was found in a PubMed search for acheiropodia
(32 records, 2026-10-08). The orthopaedic case report PMID:7929505 has no
PubMed abstract, so it could not be quoted.
The falcon deep-research report (research/Acheiropodia-deep-research-falcon.md)
agreed with the curated phenotype pattern and causal chain and added one
mechanistic source used here (PMID:40604280). It also describes general
prosthetic and rehabilitation care, but with no acheiropodia-specific
outcome source, so no treatment is curated from it.
References & Deep Research
References
2Deep Research
1Deep research results are used as seeds for research; they do not undergo the same validation as the main records and may contain errors. How we use deep research.
Evaluations and curation notes (2)
Record notes
Acheiropodia is curated separately from ZRS-Related Limb Malformation. Both map to the LMBR1-SHH locus, but the ZRS spectrum arises from heterozygous gain-of-ectopic-expression alleles producing polydactyly, while acheiropodia arises from homozygous deletions that remove regulatory contact and produce limb reduction. No treatment is curated: no source describing management beyond case descriptions was found in a PubMed search for acheiropodia (32 records, 2026-10-08). The orthopaedic case report PMID:7929505 has no PubMed abstract, so it could not be quoted. The falcon deep-research report (research/Acheiropodia-deep-research-falcon.md) agreed with the curated phenotype pattern and causal chain and added one mechanistic source used here (PMID:40604280). It also describes general prosthetic and rehabilitation care, but with no acheiropodia-specific outcome source, so no treatment is curated from it.
Create: Acheiropodia · 2026-10-08T15:43:12Z · View source
New entry for acheiropodia (MONDO:0008700), curated from the stub queue. Lump/split: kept as a standalone Disease rather than a subtype of ZRS-Related Limb Malformation, which explicitly leaves acheiropodia out of scope because it is a homozygous loss-of-contact limb-reduction disorder rather than a heterozygous ZRS gain-of-ectopic-expression polydactyly. The entry was first curated directly from PubMed (all 32 records for acheiropody/acheiropodia were screened by title), then reconciled against a falcon deep-research report (research/Acheiropodia-deep-research-falcon.md; 7/7 references resolved, the one term-validation flag is a parsing artifact that labels MONDO:0008700 'if available'). The report agreed with the curated phenotype pattern and causal chain; it added one source used here, Bower et al. 2025 (PMID:40604280, mouse ZRS homeodomain-motif knock-in loses long-range activity and causes limb reduction), cited as indirect support for the contact-loss to SHH-loss edge. Its other leads were not used: the cohesin study (Kane et al. 2022) had no cached abstract, the ETV2 study concerns Shh initiation in general, and its supportive-care section had no acheiropodia-specific source. just preflight-dr could not be run because it requires the local MONDO build (about 1.3 GB), which is absent and was not downloaded because disk space is limited; the report was checked by hand to be about acheiropodia/LMBR1/SHH throughout. LMBR1 lands on the deletion node, whose lesion is deleted DNA (CTCF scaffold sites) rather than loss of an LMBR1 protein activity, so no molecular function is true of that node; it was grandfathered in tests/gene_activity_grounding_baseline.txt, and the SHH node carries GO:0005113 patched binding. Key sources: Ianakiev et al. 2001 (PMID:11090342, common LMBR1 exon 4 deletion in five Brazilian families, phenotype pattern, inheritance, birth incidence), Ushiki et al. 2021 (PMID:33863876, 12 kb fine-mapping, loss of CTCF-mediated ZRS-SHH promoter contact in patient cells, negative orthologous mouse knockout), Sagai et al. 2005 (PMID:15677727, mouse ZRS deletion abolishes limb Shh and truncates distal limbs), Escamilla et al. 2000 (PMID:10780921, 7q36 linkage), Shamseldin et al. 2016 (PMID:26749485, 102 kb homozygous deletion), Fett-Conte and Richieri-Costa 1990 (PMID:2363435, Bohomoletz bone) and Ormazabal et al. 2015 (PMID:26294167, Argentinian twins). Three older case reports (PMID:9788736, PMID:6660253, PMID:7929505) have no abstract and were not cited. The limb-bud SHH loss node is PROVISIONAL because it has not been measured for the human allele and is supported directly only by the mouse ZRS deletion. Bohomoletz bone is left without an HPO binding after searching ols:hp. No treatment was curated because no management source with a quotable abstract was found. No GeneReviews chapter exists (just check-genereviews). Validation: just validate, validate-terms, count-verified-snippets (all snippets verified), the offline gates, and validate-disorders.
Question: You are an expert researcher providing comprehensive, well-cited information.
Provide detailed information focusing on: 1. Key concepts and definitions with current understanding 2. Recent developments and latest research (prioritize 2023-2024 sources) 3. Current applications and real-world implementations 4. Expert opinions and analysis from authoritative sources 5. Relevant statistics and data from recent studies
Format as a comprehensive research report with proper citations. Include URLs and publication dates where available. Always prioritize recent, authoritative sources and provide specific citations for all major claims.
Disease Characteristics Research Template
Target Disease
- Disease Name: Acheiropodia
- MONDO ID: MONDO:0008700 (if available)
- Category: Mendelian
Research Objectives
Please provide a comprehensive research report on Acheiropodia covering all of the disease characteristics listed below. This report will be used to populate a disease knowledge base entry. Be thorough and cite primary literature (PMID preferred) for all claims.
For each section, suggested databases/resources are listed. These are the first places you should search for information on each topic.
1. Disease Information
Search first: OMIM, Orphanet, ICD-10/ICD-11, MeSH, PubMed
- What is the disease? Provide a concise overview.
- What are the key identifiers? (OMIM, Orphanet, ICD-10/ICD-11, MeSH, Mondo)
- What are the common synonyms and alternative names?
- Is the information derived from individual patients (e.g., EHR) or aggregated disease-level resources?
2. Etiology
- Disease Causal Factors: What are the primary causes? (genetic, environmental, infectious, mechanistic)
- Risk Factors:
Search first: PubMed, Cochrane Library, UpToDate, clinical guidelines, ClinVar, ClinGen, GWAS Catalog, PheGenI, CTD, CDC, WHO, epidemiological databases
- Genetic risk factors (causal variants, susceptibility loci, modifier genes)
- Environmental risk factors (toxins, lifestyle, occupational exposures, age, sex, family history)
- Protective Factors:
Search first: PubMed, Cochrane Library, clinical trial databases, GWAS Catalog, gnomAD, WHO, CDC, nutrition databases
- Genetic protective factors (protective variants, modifier alleles)
- Environmental protective factors (diet, lifestyle, exposures that reduce risk)
- Gene-Environment Interactions: How do genetic and environmental factors interact to influence disease?
Search first: CTD, PubMed, PheGenI, GxE databases
3. Phenotypes
Search first: HPO (Human Phenotype Ontology), OMIM, Orphanet, PubMed, clinicaltrials.gov, MedDRA, SNOMED CT, DECIPHER, LOINC
For each phenotype, provide: - Phenotype type: symptoms, clinical signs, physical manifestations, behavioral changes, or laboratory abnormalities
For symptoms/signs: HPO, OMIM, Orphanet, PubMed For behavioral changes: HPO, DSM, RDoC (Research Domain Criteria), PubMed For laboratory abnormalities: LOINC, SNOMED CT, LabTests Online, PubMed - Phenotype characteristics: Search first: OMIM, Orphanet, HPO, PubMed - Age of symptom onset (neonatal, childhood, adult-onset, late-onset) - Symptom severity (mild, moderate, severe, variable) - Symptom progression (stable, progressive, episodic, fluctuating) - Frequency among affected individuals (percentage or qualitative) - Quality of life impact: Effects on daily functioning and well-being (per-phenotype when possible) Search first: EQ-5D database, SF-36, WHO QOL databases, PubMed - Suggest HPO (Human Phenotype Ontology) terms for each phenotype
4. Genetic/Molecular Information
- Causal Genes: Gene mutations or chromosomal abnormalities responsible for disease (gene symbols, OMIM IDs)
Search first: OMIM, ClinVar, HGMD, Ensembl, NCBI Gene
- Pathogenic Variants:
- Affected genes (gene symbols, HGNC IDs) > Search first: OMIM, NCBI Gene, Ensembl, HGNC, UniProt, GeneCards
- Variant classification (pathogenic, likely pathogenic, VUS per ACMG/AMP guidelines) > Search first: ClinVar, ClinGen, ACMG/AMP guidelines, VarSome
- Variant type/class (missense, frameshift, nonsense, splice-site, structural)
- Allele frequency in population databases > Search first: gnomAD, 1000 Genomes, ExAC, TOPMed, dbSNP
- Somatic vs germline origin > Search first: COSMIC (somatic), ClinVar, ICGC, TCGA
- Functional consequences (loss of function, gain of function, dominant negative)
- Modifier Genes: Genes that modify disease severity or expression
- Epigenetic Information: DNA methylation, histone modifications, chromatin changes affecting disease
Search first: ENCODE, Roadmap Epigenomics, MethBase, DiseaseMeth
- Chromosomal Abnormalities: Large-scale genetic changes (aneuploidy, translocations, inversions)
Search first: DECIPHER, ClinVar, ECARUCA, UCSC Genome Browser
5. Environmental Information
- Environmental Factors: Non-genetic contributing factors (toxins, radiation, pollution, occupational exposure)
Search first: CTD (Comparative Toxicogenomics Database), TOXNET, PubMed, EPA databases
- Lifestyle Factors: Behavioral factors (smoking, diet, exercise, alcohol consumption)
Search first: CDC databases, WHO, PubMed, NHANES
- Infectious Agents: If applicable, pathogens causing or triggering disease (bacteria, viruses, fungi, parasites)
Search first: NCBI Taxonomy, ViPR, BV-BRC, MicrobeDB, GIDEON
6. Mechanism / Pathophysiology
Present this section as an ordered causal chain first, then the detail below. Open with a numbered sequence of mechanistic steps running from the initiating lesion (mutation, exposure, infection) to the clinical manifestation, one step per line, each naming what it causes next. State the causal verb explicitly ("leads to", "results in") and say where a step is inferred rather than demonstrated. Where the mechanism branches, show the branch. The categories below are a checklist of what to cover within those steps, not the organizing structure — a step may draw on several of them, and a category may contribute to several steps.
- Molecular Pathways: Specific signaling cascades or biochemical pathways involved (Wnt, MAPK, mTOR, PI3K-AKT, etc.)
Search first: KEGG, Reactome, WikiPathways, PathBank, BioCyc
- Cellular Processes: Cell-level mechanisms (apoptosis, autophagy, cell cycle dysregulation, inflammation, etc.)
Search first: Gene Ontology (GO), Reactome, KEGG, PubMed
- Protein Dysfunction: How protein structure or function is altered (misfolding, aggregation, loss of function, gain of function)
Search first: UniProt, PDB (Protein Data Bank), InterPro, Pfam, AlphaFold
- Metabolic Changes: Alterations in metabolic processes (energy metabolism, lipid metabolism, amino acid metabolism)
Search first: KEGG, BioCyc, HMDB (Human Metabolome Database), BRENDA
- Immune System Involvement: Role of immune response (autoimmunity, immunodeficiency, chronic inflammation)
Search first: ImmPort, Immunome Database, IEDB, Gene Ontology
- Tissue Damage Mechanisms: How tissues/ are injured (oxidative stress, ischemia, fibrosis, necrosis)
Search first: PubMed, Gene Ontology, Reactome
- Biochemical Abnormalities: Specific molecular defects (enzyme deficiencies, receptor dysfunction, ion channel defects)
Search first: BRENDA, UniProt, KEGG, OMIM, PubMed
- Epigenetic Changes: DNA methylation, histone modifications affecting gene expression in disease
Search first: ENCODE, Roadmap Epigenomics, MethBase, DiseaseMeth
- Molecular Profiling (if available):
- Transcriptomics/gene expression changes > Search first: GEO (Gene Expression Omnibus), ArrayExpress, GTEx, Human Cell Atlas, SRA
- Proteomics findings > Search first: PRIDE, ProteomeXchange, Human Protein Atlas, STRING, BioGRID
- Metabolomics signatures > Search first: MetaboLights, Metabolomics Workbench, HMDB, METLIN
- Lipidomics alterations > Search first: LIPID MAPS, SwissLipids, LipidHome, Metabolomics Workbench
- Genomic structural features > Search first: UCSC Genome Browser, Ensembl, NCBI, dbVar, DGV
- Advanced Technologies (if applicable):
- Single-cell analysis findings (cell-type specific mechanisms, cellular heterogeneity) > Search first: Human Cell Atlas, Single Cell Portal, GEO, CELLxGENE
- Spatial transcriptomics findings > Search first: GEO, Spatial Research, Vizgen, 10x Genomics data
- Multi-omics integration results > Search first: TCGA, ICGC, cBioPortal, LinkedOmics, PubMed
- Functional genomics screens (CRISPR, RNAi) > Search first: DepMap, GenomeRNAi, PubMed, BioGRID ORCS
For each mechanism, describe: - The causal chain from initial trigger to clinical manifestation - Which mechanisms are upstream vs downstream - What cell types and biological processes are involved - Suggest GO terms for biological processes and CL terms for cell types
7. Anatomical Structures Affected
- Organ Level:
- Primary organs directly affected
- Secondary organ involvement (complications, secondary effects)
- Body systems involved (cardiovascular, nervous, digestive, respiratory, endocrine, etc.)
Search first: Uberon, FMA (Foundational Model of Anatomy), OMIM, HPO, ICD-11, MeSH, SNOMED CT
- Tissue and Cell Level:
- Specific tissue types affected (epithelial, connective, muscle, nervous)
- Specific cell populations targeted (with Cell Ontology terms)
Search first: Uberon, Human Protein Atlas, Cell Ontology, Human Cell Atlas, CellMarker, PanglaoDB
- Subcellular Level:
- Cellular compartments involved (mitochondria, nucleus, ER, lysosomes) (with GO Cellular Component terms)
Search first: Gene Ontology (Cellular Component), UniProt, Human Protein Atlas
- Localization:
- Specific anatomical sites (with UBERON terms) > Search first: FMA, Uberon, NeuroNames (for brain), SNOMED CT
- Lateralization (unilateral, bilateral, asymmetric) > Search first: HPO, clinical literature, imaging databases
8. Temporal Development
- Onset:
- Typical age of onset (congenital, pediatric, adult, geriatric)
- Onset pattern (acute, subacute, chronic, insidious)
Search first: OMIM, Orphanet, HPO, PubMed
- Progression:
- Disease stages (early, intermediate, advanced, end-stage) > Search first: Cancer Staging Manual (AJCC), WHO classifications, PubMed
- Progression rate (rapid, slow, variable)
- Disease course pattern (episodic, relapsing-remitting, progressive, stable)
- Disease duration (self-limited, chronic lifelong)
Search first: Disease registries, longitudinal cohort databases, natural history studies, PubMed, Orphanet, OMIM
- Patterns:
- Remission patterns (spontaneous, treatment-induced) > Search first: Clinical trial databases, disease registries, PubMed
- Critical periods (time windows of vulnerability or opportunity for intervention) > Search first: PubMed, developmental biology databases, clinical guidelines
9. Inheritance and Population
- Epidemiology:
- Prevalence (cases per 100,000 at given time)
- Incidence (new cases per 100,000 per year)
Search first: Orphanet, CDC, WHO, GBD (Global Burden of Disease), national registries, SEER, disease registries
- For Genetic Etiology:
- Inheritance pattern (AD, AR, X-linked, mitochondrial, multifactorial, polygenic) > Search first: OMIM, Orphanet, ClinVar, GTR (Genetic Testing Registry)
- Penetrance (complete, incomplete, age-dependent) > Search first: ClinVar, OMIM, PubMed, ClinGen
- Expressivity (variable, consistent) > Search first: OMIM, ClinVar, PubMed
- Genetic anticipation (increasing severity in successive generations) > Search first: OMIM, PubMed (especially for repeat expansion disorders)
- Germline mosaicism > Search first: ClinVar, OMIM, genetic counseling literature, PubMed
- Founder effects (population-specific mutations) > Search first: gnomAD, population genetics databases, PubMed
- Consanguinity role > Search first: OMIM, population studies, genetic counseling resources
- Carrier frequency > Search first: gnomAD, carrier screening databases, GeneReviews, GTR
- Population Demographics:
- Affected populations (ethnic or demographic groups with higher prevalence) > Search first: gnomAD, 1000 Genomes, PAGE Study, PubMed, population registries
- Geographic distribution (endemic areas, regional variation) > Search first: WHO, CDC, GBD, Orphanet, geographic epidemiology databases
- Geographic distribution of specific variants
- Sex ratio (male:female) > Search first: Disease registries, OMIM, PubMed, epidemiological databases
- Age distribution of affected individuals > Search first: CDC, disease registries, SEER, Orphanet
10. Diagnostics
- Clinical Tests:
- Laboratory tests (blood, urine, tissue chemistry, specific enzyme assays) > Search first: LOINC, LabTests Online, PubMed
- Biomarkers (proteins, metabolites, genetic markers, circulating biomarkers) > Search first: FDA Biomarker List, BEST (Biomarkers, EndpointS, and other Tools), PubMed
- Imaging studies (X-ray, CT, MRI, PET, ultrasound) > Search first: RadLex, DICOM, Radiopaedia, imaging databases
- Functional tests (pulmonary function, cardiac stress tests) > Search first: LOINC, clinical guidelines, PubMed
- Electrophysiology (EEG, EMG, ECG, nerve conduction studies) > Search first: LOINC, clinical neurophysiology databases, PubMed
- Biopsy findings (histopathology, immunohistochemistry) > Search first: SNOMED CT, College of American Pathologists resources, PubMed
- Pathology findings (microscopic examination) > Search first: SNOMED CT, Digital Pathology databases, PubMed
- Genetic Testing:
Search first: GTR (Genetic Testing Registry), GeneReviews, ClinGen
- Overview of recommended genetic testing approach
- Whole genome sequencing (WGS) utility > Search first: GTR, ClinVar, GEL (Genomics England), gnomAD
- Whole exome sequencing (WES) utility > Search first: GTR, ClinVar, OMIM, GeneMatcher
- Gene panels (which panels, which genes) > Search first: GTR, ClinVar, laboratory-specific databases
- Single gene testing > Search first: GTR, ClinVar, OMIM, GeneReviews
- Chromosomal microarray (CMA) > Search first: DECIPHER, ClinVar, dbVar, ECARUCA
- Karyotyping > Search first: Chromosome Abnormality Database, ClinVar, cytogenetics resources
- FISH > Search first: ClinVar, cytogenetics databases, PubMed
- Mitochondrial DNA testing > Search first: MITOMAP, MSeqDR, ClinVar, GTR
- Repeat expansion testing > Search first: GTR, ClinVar, repeat expansion databases, PubMed
- Omics-Based Diagnostics (if applicable):
- RNA sequencing / transcriptomics > Search first: GEO, ArrayExpress, GTEx, RNA-seq databases
- Proteomics > Search first: PRIDE, ProteomeXchange, FDA Biomarker database
- Metabolomics > Search first: MetaboLights, Metabolomics Workbench, HMDB
- Epigenomics > Search first: GEO, ENCODE, Roadmap Epigenomics, MethBase
- Liquid biopsy > Search first: COSMIC, ClinVar, liquid biopsy databases, PubMed
- Clinical Criteria:
- Standardized diagnostic criteria (DSM, ICD, society guidelines) > Search first: DSM-5, ICD-11, clinical society guidelines, UpToDate
- Differential diagnosis (other conditions to rule out, with distinguishing features) > Search first: DynaMed, UpToDate, clinical decision support systems
- Screening:
- Screening methods for asymptomatic individuals (newborn screening, carrier screening, cascade screening) > Search first: ACMG recommendations, CDC newborn screening, GTR
11. Outcome/Prognosis
- Survival and Mortality:
- Survival rate (5-year, 10-year, overall) > Search first: SEER, cancer registries, disease-specific registries, PubMed
- Life expectancy (with and without treatment if applicable) > Search first: Orphanet, disease registries, actuarial databases, PubMed
- Mortality rate > Search first: CDC, WHO, GBD, national mortality databases
- Disease-specific mortality (deaths directly attributable to disease) > Search first: Disease registries, CDC Wonder, GBD, PubMed
- Morbidity and Function:
- Morbidity (disease-related disability and health impacts) > Search first: GBD, WHO, disability databases, PubMed
- Disability outcomes (long-term functional impairments) > Search first: ICF (International Classification of Functioning), disability registries
- Quality of life measures (EQ-5D, SF-36, PROMIS, disease-specific tools) > Search first: EQ-5D database, SF-36, PROMIS, PubMed
- Disease Course:
- Complications (secondary problems: infections, organ failure, etc.) > Search first: ICD codes, disease registries, clinical databases, PubMed
- Recovery potential (likelihood and extent of recovery, with vs without treatment) > Search first: Natural history studies, rehabilitation databases, PubMed
- Prediction:
- Prognostic factors (age, disease severity, biomarkers, treatment response) > Search first: Prognostic models databases, clinical calculators, PubMed
- Prognostic biomarkers (molecular markers predicting disease course) > Search first: FDA Biomarker database, PubMed, cancer prognostic databases
12. Treatment
- Pharmacotherapy:
- Pharmacological treatments (drug names, drug classes, mechanisms of action) > Search first: DrugBank, RxNorm, ATC classification, DailyMed, FDA databases
- Pharmacogenomics (how genetic variants affect drug metabolism, efficacy, toxicity) > Search first: PharmGKB, CPIC (Clinical Pharmacogenetics), FDA Table of PGx Biomarkers
- Advanced Therapeutics:
- Gene therapy (viral vectors, CRISPR, gene replacement, gene editing) > Search first: ClinicalTrials.gov, FDA gene therapy database, ASGCT resources
- Cell therapy (stem cell transplant, CAR-T, cellular therapeutics) > Search first: ClinicalTrials.gov, FDA cell therapy database, FACT standards
- RNA-based therapies (ASOs, siRNA, mRNA therapies) > Search first: ClinicalTrials.gov, FDA approvals, PubMed
- Targeted therapies (treatments directed at specific molecular targets) > Search first: My Cancer Genome, OncoKB, ClinicalTrials.gov, FDA approvals
- Immunotherapies (checkpoint inhibitors, monoclonal antibodies) > Search first: Cancer Immunotherapy Database, FDA approvals, ClinicalTrials.gov
- Surgical and Interventional:
- Surgical interventions (types of surgery, timing, outcomes) > Search first: CPT codes, surgical registries, clinical guidelines, PubMed
- Supportive and Rehabilitative:
- Supportive care (symptom management, pain control, nutrition) > Search first: Clinical guidelines, Cochrane Library, PubMed
- Rehabilitation (physical therapy, occupational therapy, speech therapy) > Search first: Rehabilitation medicine databases, clinical guidelines, PubMed
- Experimental:
- Experimental treatments in clinical trials (with NCT identifiers if available) > Search first: ClinicalTrials.gov, EU Clinical Trials Register, WHO ICTRP
- Treatment Outcomes:
- Treatment response rates > Search first: Clinical trial databases, FDA reviews, systematic reviews, PubMed
- Side effects and adverse events > Search first: FDA Adverse Event Reporting System (FAERS), MedWatch, PubMed
- Treatment Strategy:
- Treatment algorithms (clinical pathways, decision trees) > Search first: Clinical practice guidelines, NCCN Guidelines, UpToDate
- Combination therapies > Search first: ClinicalTrials.gov, treatment guidelines, PubMed
- Personalized medicine approaches (genotype-guided treatment) > Search first: My Cancer Genome, CIViC, PharmGKB, precision medicine databases
For each treatment, suggest NCIT (NCI Thesaurus) clinical-intervention terms where applicable.
13. Prevention
- Prevention Levels:
- Primary prevention (preventing disease occurrence: vaccination, risk factor modification) > Search first: CDC, WHO, USPSTF recommendations, Cochrane Library
- Secondary prevention (early detection and treatment: screening programs, early intervention) > Search first: USPSTF, CDC screening guidelines, WHO
- Tertiary prevention (preventing complications in those with disease) > Search first: Clinical guidelines, disease management protocols, PubMed
- Immunization: Vaccine strategies (if applicable)
Search first: CDC vaccine schedules, WHO immunization, FDA vaccine database
- Screening and Early Detection:
- Screening programs (population-based: newborn screening, cancer screening) > Search first: CDC screening programs, USPSTF, cancer screening databases
- Genetic screening (carrier screening, preimplantation genetic diagnosis, prenatal testing) > Search first: ACMG recommendations, ACOG guidelines, GTR
- Risk stratification (identifying high-risk individuals for targeted prevention) > Search first: Risk prediction models, clinical calculators, PubMed
- Behavioral Interventions: Lifestyle modifications to reduce risk
Search first: CDC, WHO, behavioral intervention databases, Cochrane Library
- Counseling: Genetic counseling (risk assessment, family planning guidance)
Search first: NSGC resources, ACMG guidelines, GeneReviews
- Public Health:
- Public health interventions (sanitation, vector control, health education) > Search first: CDC, WHO, public health databases, PubMed
- Environmental interventions (reducing environmental risk factors) > Search first: EPA databases, WHO environmental health, PubMed
- Prophylaxis: Preventive medications or procedures
Search first: Clinical guidelines, FDA approvals, PubMed
14. Other Species / Natural Disease
- Taxonomy: Species affected (with NCBI Taxon identifiers)
Search first: NCBI Taxonomy
- Breed: Specific breeds affected (with VBO identifiers if applicable)
Search first: VBO (Vertebrate Breed Ontology)
- Gene: Orthologous genes in other species (with NCBI Gene IDs)
Search first: NCBI Gene
- Natural Disease:
- Naturally occurring disease in other species (companion animals, wildlife) > Search first: OMIA (Online Mendelian Inheritance in Animals), VetCompass, PubMed
- Veterinary relevance and importance in animal health > Search first: OMIA, veterinary databases, PubMed
- Comparative Biology:
- Comparative pathology (similarities and differences across species) > Search first: OMIA, comparative pathology databases, PubMed
- Evolutionary conservation of disease mechanisms > Search first: HomoloGene, OrthoMCL, Alliance of Genome Resources
- Transmission (if applicable):
- Zoonotic potential > Search first: CDC zoonotic diseases, WHO zoonoses, GIDEON
- Cross-species susceptibility > Search first: NCBI Taxonomy, veterinary databases, PubMed
15. Model Organisms
- Model Types:
- Model organism type (mammalian, invertebrate, cellular, in vitro) > Search first: Alliance of Genome Resources, model organism databases
- Specific model systems (mouse, rat, zebrafish, Drosophila, C. elegans, yeast, cell lines, organoids, iPSCs) > Search first: MGI, RGD, ZFIN, FlyBase, WormBase, SGD, ATCC, Cellosaurus
- Induced models (drug treatment, surgical intervention, environmental manipulation) > Search first: MGI, model organism databases, PubMed
- Genetic Models:
- Types available (knockout, knock-in, transgenic, conditional, humanized) > Search first: MGI, IMPC, KOMP, EuMMCR, IMSR
- Model Characteristics:
- Phenotype recapitulation (how well model reproduces human disease features) > Search first: Model organism databases, comparative studies, PubMed
- Model limitations (aspects of human disease not captured) > Search first: Model organism databases, PubMed, review articles
- Applications:
- Research applications (what aspects of disease can be studied) > Search first: Model organism databases, PubMed
- Resources:
- Model databases > Search first: MGI, RGD, ZFIN, FlyBase, WormBase, IMSR, EMMA, MMRRC
Citation Requirements
- Cite primary literature (PMID preferred) for all mechanistic and clinical claims
- Prioritize recent reviews and landmark papers
- Include direct quotes from abstracts where possible to support key statements
- Distinguish evidence source types: human clinical, model organism, in vitro, computational
Output Format
Structure your response as a comprehensive narrative organized by the sections above. For each section, provide: - Factual content with specific details (numbers, percentages, gene names, variant nomenclature) - Ontology term suggestions (HPO, GO, CL, UBERON, CHEBI, NCIT, MONDO) where applicable - Evidence citations with PMIDs - Direct quotes from abstracts to support key claims - Clear indication when information is not available or not applicable for this disease
This report will be used to populate a disease knowledge base entry with: - Pathophysiology descriptions with causal chains - Gene/protein annotations (HGNC, GO terms) - Phenotype associations (HP terms) with frequencies - Cell type involvement (CL terms) - Anatomical locations (UBERON terms) - Chemical entities (CHEBI terms) - Treatment annotations (NCIT terms) - Evidence items with PMIDs and exact abstract quotes - Epidemiology, prognosis, diagnostic, and prevention information - Animal model descriptions with phenotype recapitulation details
Acheiropodia: disease-characteristics research report
Scope and evidence standard. Acheiropodia is an exceptionally rare congenital limb-development disorder. The principal disease-specific evidence consists of a five-family Brazilian genetic study published in 2001 and a 2021 study combining a patient genome, patient-derived cells, and mouse experiments. Consequently, many requested knowledge-base fields cannot be populated with reliable disease-specific measurements. Evidence from other SHH-locus disorders or animal models is identified separately below. The information here is aggregated from publications and a disease–target resource, not extracted from a patient's electronic health record. (ianakiev2001acheiropodiaiscaused pages 1-3, ushiki2021deletionofctcf pages 1-2, OpenTargets Search: Acheiropodia)
The following evidence summary separates observations from interpretations and identifies the principal quantitative findings.
| Topic | Validated human observation or numeric detail | Evidence type and important limitation | Citation IDs |
|---|---|---|---|
| Disease identifier | Acheiropodia; OMIM/MIM 200500 | Explicitly reported in primary human literature; no unverified Orphanet or HPO identifiers included | (ianakiev2001acheiropodiaiscaused pages 1-3) |
| Epidemiology | Historical incidence estimate in Brazil: approximately 1 per 250,000 births | Historical estimate cited in 2001; not a modern prevalence or incidence measurement; global rates are unknown | (ianakiev2001acheiropodiaiscaused pages 1-3) |
| Core phenotype | Congenital, severe, bilateral transverse truncation of all four limbs, with absent hands, feet, and forearms; defects include distal humeral and tibial truncation and absence of the radius, ulna, fibula, and distal hand and foot bones | Human clinical observations from Brazilian families; published cohorts are extremely small | (ianakiev2001acheiropodiaiscaused pages 1-3, ianakiev2001acheiropodiaiscaused media 964b69bc) |
| Phenotypic variation | Some affected individuals had an implanted residual digit, small finger-like appendages, or an ectopic Bohomoletz bone near the distal humerus | Human case-series evidence; frequencies were not quantified | (ianakiev2001acheiropodiaiscaused pages 1-3) |
| Inheritance | Autosomal recessive; affected individuals were homozygous, whereas heterozygous parents were clinically unaffected; consanguinity was common in the original families | Human pedigree evidence; penetrance is supported within reported families but has not been estimated in a population cohort | (ianakiev2001acheiropodiaiscaused pages 1-3, ushiki2021deletionofctcf pages 1-2) |
| Founder evidence | The same mutation and an approximately 1.3-cM shared haplotype occurred in five unrelated Brazilian families, supporting inheritance from a common ancestor | Linkage and haplotype evidence from five families; mutation-age estimates were imprecise | (ianakiev2001acheiropodiaiscaused pages 4-7) |
| Initial deletion definition, 2001 | PCR initially delimited a 4–6-kb genomic deletion spanning C7orf2/LMBR1 exon 4 | Primary human molecular study; boundaries were approximate because contemporary WGS was unavailable | (ianakiev2001acheiropodiaiscaused pages 4-7) |
| Refined deletion definition, 2021 | WGS resolved a homozygous 12,041-bp deletion at GRCh38 chr7:156,816,030–156,828,070, corresponding to GRCh37/hg19 chr7:156,608,724–156,620,764; a two-base CA insertion occurred at the breakpoint | Human WGS with PCR and Sanger validation in a proband and carrier parents; evidence concerns the recurrent Brazilian lesion rather than every possible acheiropodia allele | (ushiki2021deletionofctcf pages 1-2, ushiki2021deletionofctcf pages 10-10) |
| Original molecular interpretation | Exon 4 loss produced an exon 3–exon 5 junction, frameshift, and premature stop codon in exon 6; originally interpreted as a C7orf2/LMBR1 null allele | Human RNA evidence; this coding-loss interpretation was later supplemented by chromatin-architecture evidence | (ianakiev2001acheiropodiaiscaused pages 4-7) |
| Revised regulatory interpretation | The 12-kb interval contains three CTCF sites, LSC3–LSC5, with RAD21 occupancy; deletion alters SHH-locus contacts and removes detectable ZRS–SHH-promoter interaction in proband cells | Patient-derived lymphoblastoid-cell ChIP-seq, 4C-seq, and DNA-FISH evidence; lymphoblasts are not embryonic limb-bud cells | (ushiki2021deletionofctcf pages 4-5) |
| ZRS status | The limb enhancer ZRS remains physically intact outside the deleted interval; no rare causal ZRS variant was identified, and the deleted sequence lacked consistent limb-enhancer activity | Human sequencing plus mouse transgenic reporter evidence; supports disrupted enhancer–promoter communication rather than deletion of ZRS | (ushiki2021deletionofctcf pages 2-4, ushiki2021deletionofctcf pages 1-2) |
| SHH expression | SHH expression was low in lymphoblastoid cells but significantly lower in proband cells than in wild-type cells | Human cell-line evidence; no fold-change was available, and embryonic limb expression was not measured directly | (ushiki2021deletionofctcf pages 4-5) |
| Mouse ortholog deletion | Homozygous deletion of the orthologous interval caused no apparent limb phenotype and no significant E11.5 autopod Shh-expression difference, P = 0.7796 | Engineered mouse model; the negative result highlights species-specific CTCF architecture and limits direct phenotypic recapitulation | (ushiki2021deletionofctcf pages 4-5, ushiki2021deletionofctcf pages 7-9) |
Table: Compact evidence table separating verified human findings from model-organism results and noting major limitations. It reconciles the original exon-loss interpretation with the later CTCF-mediated SHH regulatory mechanism.
1. Disease information and identifiers
Acheiropodia, also indexed as acheiropody, is an autosomal-recessive disorder characterized by severe, usually bilateral congenital truncation of upper and lower limbs with absent hands and feet. The original investigators emphasized that the reported phenotype appeared confined to the extremities; this should not be converted into a claim that extracranial or systemic findings have been systematically excluded in a large cohort. Their abstract states: “This severely handicapping condition appears to affect only the extremities, with no other systemic manifestations reported.” (ianakiev2001acheiropodiaiscaused pages 1-3, OpenTargets Search: Acheiropodia)
Verified identifiers: MONDO:0008700, displayed as “acheiropody” in Open Targets; OMIM/MIM 200500, explicitly given in the clinical genetics literature. The implicated chromosomal region is 7q36. For disease indexing, https://mondo.monarchinitiative.org/ and https://omim.org/entry/200500 are useful entry points; the MONDO–LMBR1 association was directly recovered from Open Targets. A disease-specific Orphanet number, MeSH descriptor, and ICD-10/ICD-11 code were not independently verified in the sources examined. Generic congenital-limb-deficiency codes should not be represented as unique identifiers for acheiropodia. “Terminal transverse hemimelia” describes its limb presentation, not necessarily an exact database synonym. (OpenTargets Search: Acheiropodia, ianakiev2001acheiropodiaiscaused pages 1-3, ushiki2021deletionofctcf pages 1-2)
2. Etiology, risk, protection, and gene–environment interaction
The established causal association is biallelic, germline deletion of a region of LMBR1—called C7orf2 in the original report—that also contains regulatory architecture governing distant SHH expression. The original families shared a deletion and a linked haplotype; unaffected heterozygous parents establish the recessive pattern in those pedigrees. Consanguinity increases the probability that relatives inherit two copies of the same rare ancestral allele, but is neither a biological prerequisite nor an environmental cause. There is no demonstrated infectious, toxic, dietary, behavioral, occupational, sex-specific, or age-related cause of this particular genetic syndrome, and no validated protective allele or exposure. Potential effects of embryonic exposures on limb formation in general must not be misclassified as evidence for this genotype-specific condition. Disease-specific gene–environment interaction and modifier-gene studies were not identified. (ianakiev2001acheiropodiaiscaused pages 1-3, ianakiev2001acheiropodiaiscaused pages 4-7, ushiki2021deletionofctcf pages 1-2)
3. Phenotypes and functional effects
The following are congenital physical signs, rather than acquired amputations or laboratory abnormalities. In the small published series, bilateral upper- and lower-limb involvement, major transverse truncation, and absent hands and feet define the reported syndrome. The investigators describe distal humeral and tibial truncation; aplasia of the radius, ulna, fibula, and distal hand and foot skeleton; and absent forearms in some descriptions. Some patients instead retain a single implanted digit, finger-like appendages, or an ectopic distal-humeral ossicle termed the Bohomoletz bone. A published clinical photograph and radiographs illustrate the bilateral defects and skeletal configuration. The original abstract describes “bilateral congenital amputations of the upper and lower extremities and aplasia of the hands and feet.” (ianakiev2001acheiropodiaiscaused pages 1-3, ianakiev2001acheiropodiaiscaused media 964b69bc)
Suggested phenotype annotations, pending confirmation against the live Human Phenotype Ontology: bilateral limb reduction/terminal transverse limb defect; absent hand; absent foot; absent forearm; absent radius; absent ulna; absent fibula; shortened humerus; shortened tibia; abnormal finger morphology or residual digit where documented. Apply terms per patient, rather than assigning every anatomical finding to every affected person; verified HP identifiers and population-level phenotype percentages were unavailable from the consulted primary reports. Onset is prenatal, the anatomical defect is apparent at birth, and the skeletal absence is permanent rather than an episodic or progressively degenerative process. Severe limitations to grasping, mobility, and self-care are plausible consequences, but phenotype-specific EQ-5D, SF-36, PROMIS, pain, educational, and employment data have not been established for this disorder. (ianakiev2001acheiropodiaiscaused pages 1-3, ianakiev2001acheiropodiaiscaused media 964b69bc, ushiki2021deletionofctcf pages 1-2)
4. Genetics and molecular lesion
The disease-associated gene locus is LMBR1, formerly C7orf2; SHH is the distant developmental target of its limb regulatory landscape, not a separately demonstrated pathogenic SHH coding variant in the Brazilian families. Open Targets identifies LMBR1 as the associated target under MONDO:0008700. The 2001 study used PCR and RNA analysis to estimate a 4–6-kb deletion encompassing LMBR1 exon 4 in affected members of five Brazilian families. Exon 4 skipping joined exon 3 to exon 5, shifted the reading frame, and introduced a premature stop in exon 6. The authors therefore initially interpreted the allele as a likely LMBR1 null. This remains a documented transcript consequence, but is not sufficient by itself to establish that loss of LMBR1 protein, rather than altered SHH regulation, produces the limb phenotype. (OpenTargets Search: Acheiropodia, ianakiev2001acheiropodiaiscaused pages 4-7)
Whole-genome sequencing in 2021 refined the associated deletion to 12,041 bp, GRCh38/hg38 chr7:156,816,030–156,828,070; the paper also reports GRCh37/hg19 chr7:156,608,724–156,620,764. A two-base CA insertion occurs at the breakpoint; PCR/Sanger sequencing confirmed the junction. The proband was homozygous and both unaffected parents heterozygous. The approximately 4–6-kb early PCR estimate and the approximately 12-kb subsequent genome-sequencing definition should not be entered as distinct established disease alleles without independent breakpoint evidence. Importantly, the ZRS limb enhancer lies outside the deleted interval and was retained; the investigators did not find a rare explanatory sequence variant in ZRS. A standardized HGVS expression, rsID, confirmed ClinVar accession/classification, and allele frequency in gnomAD or other population datasets were not verified. The deletion is best described here as a disease-associated pathogenic structural deletion supported by segregation and functional evidence, not assigned an invented database classification. (ushiki2021deletionofctcf pages 1-2, ushiki2021deletionofctcf pages 10-10, ushiki2021deletionofctcf pages 2-4)
The predicted LMBR1 product was described historically as a 490-amino-acid multipass membrane protein, but the stronger current disease mechanism concerns the deleted cis-regulatory architecture. Three CTCF-bound sites within the interval are lost, affecting SHH enhancer–promoter organization; do not annotate CTCF or RAD21 as independently mutated causal genes. Disease-specific modifier genes, altered DNA-methylation patterns, epigenetic biomarkers, non-founder pathogenic alleles, and additional pathogenic chromosomal abnormalities have not been validated in the examined studies. (ianakiev2001acheiropodiaiscaused pages 4-7, ushiki2021deletionofctcf pages 4-5, ushiki2021deletionofctcf pages 7-9)
5. Environmental and infectious information
No environmental exposure, lifestyle practice, or pathogen has been demonstrated to cause or trigger genetically defined acheiropodia. There is no reported zoonotic or infectious transmission. The relevant non-genetic context is reproductive relatedness, which changes the probability of homozygosity for a segregating founder deletion rather than changing limb development through an exposure mechanism. Environmental prevention claims should therefore not be inferred from data on unrelated causes of congenital limb reduction. (ianakiev2001acheiropodiaiscaused pages 1-3, ianakiev2001acheiropodiaiscaused pages 4-7)
6. Mechanism and pathophysiology
Ordered causal chain; “inferred” identifies a step not directly demonstrated in affected human embryonic limbs:
- Inherited homozygosity for the LMBR1-region deletion leads to loss of exon 4 and three adjacent SHH-locus CTCF-binding sites; the ZRS enhancer itself remains present. The exon-loss and CTCF-site losses are experimentally documented. (ianakiev2001acheiropodiaiscaused pages 4-7, ushiki2021deletionofctcf pages 1-2)
- Loss of those CTCF sites leads to altered CTCF/RAD21 occupancy and altered SHH-locus chromatin contacts in patient-derived lymphoblastoid cells, resulting in reduced detectable ZRS–SHH-promoter contact and alternative contacts with other CTCF sites. (ushiki2021deletionofctcf pages 4-5, ushiki2021deletionofctcf pages 5-7)
- Altered long-range contacts likely lead to insufficiently robust SHH transcription in embryonic posterior limb-bud mesenchyme; this limb-specific step is inferred, because the human patient assays used lymphoblastoid cells, not affected embryonic limb buds. One reported cell-line comparison found low SHH expression in both groups but significantly higher expression in controls than proband cells; no reliable patient-limb expression magnitude is available. (ushiki2021deletionofctcf pages 4-5, ushiki2021deletionofctcf pages 1-2)
- Inferred reduction in localized SHH signaling leads to disturbed embryonic limb outgrowth and skeletal patterning, resulting in congenital bilateral distal upper- and lower-limb truncation and loss of hands and feet. Mouse experiments manipulating the ZRS/SHH pathway support biological plausibility, but they are not direct observation of this final sequence in human embryos. (bower2024conservedcisactingrangeextender pages 4-8, koyanonakagawa2022etv2regulatesenhancer pages 1-2, ianakiev2001acheiropodiaiscaused pages 1-3)
- Parallel branch: exon 4 loss leads to a frameshifted LMBR1 transcript and predicted premature termination; whether this independently contributes to the patient's limb phenotype, rather than accompanying the CTCF-site deletion, remains unresolved. A second proposed architectural possibility involving a changed domain boundary was not definitively excluded. (ianakiev2001acheiropodiaiscaused pages 4-7, ushiki2021deletionofctcf pages 9-10)
Thus, the upstream lesion is a structural variant, the intermediate biochemical processes are CTCF/cohesin-associated chromosome organization and long-range enhancer control, and the downstream tissue process is impaired embryonic limb formation. This is not established as an enzyme deficiency, metabolic disease, immune disease, chronic inflammation, or postnatal oxidative/ischemic injury. Appropriate GO-process suggestions, without asserting verified accession numbers, include regulation of transcription by RNA polymerase II, chromatin organization, embryonic limb morphogenesis, and Hedgehog signaling; appropriate cellular-component annotations include the nucleus/chromatin for CTCF, RAD21, and regulatory DNA. Posterior limb-bud mesenchymal cells are the biologically relevant proposed target cell population; a precise Cell Ontology ID requires independent validation. These annotations describe the proposed pathway, not direct single-cell profiling of human affected limbs. (ushiki2021deletionofctcf pages 4-5, koyanonakagawa2022etv2regulatesenhancer pages 1-2, kane2022cohesinisrequired pages 1-3)
Orthogonal experimental support and important restraint. Patient-derived cell experiments used CTCF/RAD21 ChIP-seq, chromosome-conformation capture/4C-seq, and DNA-FISH; a mouse transgenic enhancer assay failed to show consistent limb-enhancer activity in the deleted human interval. In 2022, a separate mouse-cell perturbation study found that cohesin, but not CTCF, was required for activation by distant SHH enhancers in its specific experimental assay: this establishes context dependence, not a refutation of the disease-associated human CTCF-site findings. A separate 2022 mouse study demonstrated ETV2-dependent ZRS opening and initiation of Shh transcription; it did not show that ETV2 is mutated in acheiropodia. A 2024 bioRxiv preprint, subsequently published in 2025, found that conserved motifs within ZRS support long-range activity and that disrupting them produces severe mouse limb reduction; those are comparative mechanistic findings, not new 2023–2024 human acheiropodia cases. The 2021 disease abstract states: “Using whole-genome sequencing, we fine-mapped the acheiropodia-associated region to 12 kb and show that it does not function as an enhancer.” (ushiki2021deletionofctcf pages 2-4, kane2022cohesinisrequired pages 1-3, koyanonakagawa2022etv2regulatesenhancer pages 1-2, bower2024conservedcisactingrangeextender pages 4-8, ushiki2021deletionofctcf pages 1-2)
No disease-specific human-embryo transcriptomic, proteomic, metabolomic, lipidomic, single-cell, spatial-transcriptomic, or integrated multi-omic signature was established in the available sources. The human cell-line chromatin profiling and WGS are research assays, not validated clinical molecular-profiling biomarkers. (ushiki2021deletionofctcf pages 1-2, ushiki2021deletionofctcf pages 4-5)
7. Affected anatomy and localization
The primary anatomical structures are the bilateral upper and lower limbs, especially distal upper-arm/forearm and lower-leg segments, hands, feet, and their developing skeleton. Reported defects include absent or truncated radius, ulna, fibula, distal humerus and tibia, with absent distal hand and foot bones; variable rudimentary digits or a distal-humeral Bohomoletz bone may be present. The apparent bilateral pattern is characteristic, although accessory appendages may be unilateral or bilateral. The musculoskeletal system is directly affected; systemic organ involvement is not a recognized feature of the described families. The relevant developmental tissue is limb-bud mesenchyme and nascent cartilage/bone; no established disease-specific lesion of muscle, nerve, vessels, or epithelia is reported. Suggested UBERON labels are upper limb, lower limb, forearm, hand, foot, humerus, radius, ulna, tibia, and fibula; exact UBERON and CL accession numbers should be resolved against ontology releases before ingestion. At the mechanistic subcellular level, the demonstrable regulatory interactions involve nuclear chromatin, not an established mitochondrial, lysosomal, or ER defect. (ianakiev2001acheiropodiaiscaused pages 1-3, ianakiev2001acheiropodiaiscaused media 964b69bc, ushiki2021deletionofctcf pages 4-5)
8. Temporal development
The initiating disruption acts during embryonic limb development; anatomical manifestations are congenital and visible at birth. The limb difference remains lifelong and is not described as episodic, relapsing, remitting, or progressively destructive. The embryonic limb-bud patterning period is the biologically important vulnerability window; mouse developmental data demonstrate that ZRS accessibility and Shh initiation are timed events, but a validated gestational-week threshold for human acheiropodia or a prenatal rescue window is unavailable. No formal clinical stage system, longitudinal progression rate, spontaneous remission, or postnatal anatomical recovery has been established. (ianakiev2001acheiropodiaiscaused pages 1-3, koyanonakagawa2022etv2regulatesenhancer pages 1-2, ushiki2021deletionofctcf pages 1-2)
9. Inheritance and population epidemiology
Inheritance is autosomal recessive. One affected individual in each of five Brazilian families shared the deletion; several families involved consanguinity. All five shared an approximately 1.3-cM haplotype, consistent with a founder allele. A mutation age estimated using combined historical families was approximately 20 generations, with a broad 95% interval of 4–60 generations; this is a population-genetic inference, not a dated historical event. An older report cited by the 2001 authors described two affected siblings in Puerto Rico; most previously reported cases were Brazilian. Approximately one affected birth per 250,000 births in Brazil is an historical estimate quoted by the original authors, not a current global prevalence estimate and not one affected person per 250,000 living people. A corresponding crude rate would be about 0.4 per 100,000 births, subject to the original estimate's substantial uncertainty. Current incidence, worldwide prevalence, carrier frequency, sex ratio, and variant frequencies by ancestry are unknown from these data. (ianakiev2001acheiropodiaiscaused pages 1-3, ianakiev2001acheiropodiaiscaused pages 4-7)
Heterozygotes in the reported pedigrees were clinically unaffected; the observed affected subjects had severe manifestations, with some variation in residual digits and ectopic bone. Formal population penetrance and expressivity estimates, anticipation, germline mosaicism, and reproducible modifier alleles have not been quantified. For a couple both heterozygous for the same fully recessive causal allele, the Mendelian probability of a biallelic child is 25% per pregnancy; that number is a segregation calculation, not an observed incidence in the study families. (ianakiev2001acheiropodiaiscaused pages 1-3, ushiki2021deletionofctcf pages 1-2)
10. Diagnosis and differential diagnosis
Clinical assessment: congenital bilateral four-limb transverse reduction with absent hands/feet prompts examination of residual limb anatomy, family history, and plain radiographs to characterize missing or truncated bones and any residual ossicle. Photographs and radiographs in the original publication document this approach. Prenatal anatomical ultrasound may reasonably detect major limb deficiencies, but an acheiropodia-specific screening sensitivity, diagnostic ultrasound protocol, and confirmatory imaging accuracy were not reported. There is no validated disease-specific blood chemistry, protein/metabolite biomarker, enzymatic assay, biopsy morphology, electrophysiological signature, or functional cardiopulmonary test. (ianakiev2001acheiropodiaiscaused pages 1-3, ianakiev2001acheiropodiaiscaused media 964b69bc)
Molecular confirmation: when a familial breakpoint is known, a targeted deletion/breakpoint assay with parental segregation testing is the most direct molecular approach. Otherwise, use a clinically validated assay capable of detecting structural variants spanning LMBR1 exon 4 and adjacent noncoding CTCF sites; genome sequencing with structural-variant analysis is specifically supported by the 2021 discovery. A limb-malformation panel is useful only if it has appropriate copy-number/structural-variant and noncoding-region coverage. Exome sequencing focused on coding single-nucleotide variants may miss or incompletely characterize this lesion; standard CMA resolution also requires verification against the approximately 12-kb event. Karyotype and FISH are not established first-line confirmatory tests for this small founder deletion; mitochondrial, repeat-expansion, liquid-biopsy, methylome, proteome, and metabolome tests have no established role. No disease-specific standardized diagnostic criteria, clinical genetic-testing sensitivity, or professional diagnostic algorithm was identified. (ushiki2021deletionofctcf pages 1-2, ushiki2021deletionofctcf pages 10-10, ianakiev2001acheiropodiaiscaused pages 4-7)
Differential diagnosis: other congenital transverse limb deficiencies and other genetic or non-genetic causes of severe limb reduction. Bilateral symmetry, the specific distal humeral/tibial and radial/ulnar/fibular pattern, affected siblings/consanguinity, and confirmation of biallelic LMBR1-region deletion help distinguish the documented syndrome. Preaxial polydactyly and triphalangeal thumb can also involve SHH/ZRS regulatory variation, but have materially different digit phenotypes and generally should not be collapsed into acheiropodia. Diagnostic comparison with specific competing syndromes requires patient-specific imaging and genetics. (ianakiev2001acheiropodiaiscaused pages 1-3, vandermeer2011cis‐regulatorymutationsare pages 6-7, ushiki2021deletionofctcf pages 2-4)
11. Outcome and prognosis
The principal established morbidity is persistent, severe limb-related disability. Congenital limb truncation does not biologically regrow; improvement in activities may nevertheless be possible through individualized adaptations and rehabilitation. No disease-specific prospective natural-history cohort supports a numerical survival rate, lifespan estimate, mortality rate, complication frequency, or validated prognostic biomarker. Descriptions of apparently isolated limb involvement do not prove normal life expectancy; similarly, the documented residual-digit variability does not establish a predictive genotype–phenotype scale. Quantitative patient-reported quality-of-life results for acheiropodia were not found. (ianakiev2001acheiropodiaiscaused pages 1-3, ianakiev2001acheiropodiaiscaused media 964b69bc)
12. Treatment and clinical implementation
No pharmacotherapy, molecularly targeted drug, approved gene therapy, RNA treatment, cell therapy, immunotherapy, or established intervention restores the absent limbs; disease-specific response rates and adverse-event estimates are unavailable. Management is therefore individualized supportive care: multidisciplinary assessment of limb function, prosthetic and assistive-device options, occupational/physical rehabilitation, mobility and self-care adaptations, and psychosocial support. These are reasonable applications of general congenital limb-deficiency practice, not interventions whose effectiveness was measured in the identified acheiropodia case series. Surgical procedures, if considered for an individual residual limb or device interface, cannot be assigned an acheiropodia-specific recommended operation, timing, or success rate from the available evidence. ClinicalTrials.gov searches did not yield a substantiated disease-specific interventional trial or NCT efficacy dataset. (ianakiev2001acheiropodiaiscaused pages 1-3, ushiki2021deletionofctcf pages 1-2)
Suggested intervention concepts for later NCIT mapping: prosthetic device, occupational therapy, physical therapy, rehabilitation, and genetic counseling. Exact NCIT codes and disease-specific approved treatment annotations were not verified; no drug or ChEBI therapeutic entity should be entered on the basis of this evidence. (ianakiev2001acheiropodiaiscaused pages 1-3, ushiki2021deletionofctcf pages 1-2)
13. Prevention, screening, and counseling
There is no demonstrated vaccine, environmental avoidance measure, diet, supplement, or prophylactic medication that prevents a child who has inherited the causal biallelic deletion from developing the syndrome. The evidence-based actionable context is reproductive risk assessment: offer genetic counseling and targeted carrier/cascade testing after confirming a familial variant, discuss the 25% per-pregnancy recessive risk for two carriers, and consider prenatal molecular diagnosis or preimplantation genetic testing for a known familial allele according to access, preferences, and local practice. These are general Mendelian reproductive options, not proven population-level acheiropodia screening programs or evidence that treatment during gestation rescues the phenotype. Early recognition after birth can facilitate function-oriented services but is not primary prevention of the developmental lesion. Routine population newborn screening and universal carrier screening specifically for acheiropodia were not documented. (ianakiev2001acheiropodiaiscaused pages 1-3, ushiki2021deletionofctcf pages 1-2)
14. Natural disease in other species
The clinically defined human founder-deletion syndrome has not been demonstrated as the same naturally occurring disorder in another species in the materials reviewed. SHH/ZRS-related limb abnormalities occur in other animals—including naturally occurring ZRS-related polydactyly in cats and a chicken model with altered ZRS-associated limb patterning—but these are comparative limb-development phenotypes, not animal acheiropodia diagnoses. Mouse Lmbr1 and Shh are orthologous components of the human regulatory landscape; taxonomically, the principal experimentally investigated species are human (Homo sapiens, NCBI Taxon 9606) and laboratory mouse (Mus musculus, NCBI Taxon 10090). Breed-specific VBO entries, verified orthologue NCBI Gene IDs, veterinary incidence, and zoonotic transmission are unavailable or inapplicable; the condition is inherited, not contagious. (ianakiev2001acheiropodiaiscaused pages 4-7, ushiki2021deletionofctcf pages 7-9, bower2024conservedcisactingrangeextender pages 4-8)
15. Model organisms and model limitations
Closest engineered model: a homozygous mouse deletion of the region orthologous to the human approximately 12-kb interval was produced by genome editing. Contrary to a straightforward disease-model prediction, the mice had no apparent limb defects and no significant E11.5 autopod Shh expression difference; the reported comparison was P = 0.7796. The mouse edit also had different boundaries from the exact human deletion, and the human and mouse CTCF motif distribution/occupancy differ. Therefore, this is a useful negative mechanistic model, not a phenotype-recapitulating acheiropodia mouse. The 2021 abstract explicitly states: “Mouse knockouts of the orthologous 12 kb sequence have no apparent abnormalities.” (ushiki2021deletionofctcf pages 4-5, ushiki2021deletionofctcf pages 7-9, ushiki2021deletionofctcf pages 1-2)
Related, non-equivalent models: mouse ZRS disruption produces severe limb reduction by preventing limb-specific Shh activation; ETV2 perturbation affects ZRS accessibility and Shh initiation; mouse-cell cohesin degradation affects distant-enhancer action. A 2024 mouse enhancer-range preprint and its later peer-reviewed publication further explore motifs required for long-distance ZRS activity. These systems test pathway components, but a ZRS knockout, ETV2 edit, or cohesin perturbation is not the patient's intact-ZRS, LMBR1-region founder deletion. No validated patient-derived limb-bud organoid, human iPSC developmental model, or single-cell acheiropodia atlas was established by the retrieved studies. The 2021 engineered-deletion study is the primary source for an MGI/IMSR-style model annotation; exact stock or allele registry IDs were not verified. (bower2024conservedcisactingrangeextender pages 4-8, koyanonakagawa2022etv2regulatesenhancer pages 1-2, kane2022cohesinisrequired pages 1-3, ushiki2021deletionofctcf pages 7-9, ushiki2021deletionofctcf pages 2-4)
Source record and research currency
- Ianakiev P et al. “Acheiropodia Is Caused by a Genomic Deletion in C7orf2, the Human Orthologue of the Lmbr1 Gene.” American Journal of Human Genetics 68:38–45, electronically published 22 November 2000, journal issue January 2001. https://doi.org/10.1086/316955. Primary human pedigrees, anatomy, and transcript analysis. (ianakiev2001acheiropodiaiscaused pages 1-3, ianakiev2001acheiropodiaiscaused pages 4-7)
- Ushiki A et al. “Deletion of CTCF sites in the SHH locus alters enhancer–promoter interactions and leads to acheiropodia.” Nature Communications 12:2282, April 2021. https://doi.org/10.1038/s41467-021-22470-z. Primary human genome/patient-cell and engineered-mouse study. (ushiki2021deletionofctcf pages 1-2, ushiki2021deletionofctcf pages 4-5)
- Koyano-Nakagawa N et al. “Etv2 regulates enhancer chromatin status to initiate Shh expression in the limb bud.” Nature Communications 13, July 2022. https://doi.org/10.1038/s41467-022-31848-6. Related mouse developmental mechanism, not an acheiropodia cohort. (koyanonakagawa2022etv2regulatesenhancer pages 1-2)
- Kane L et al. “Cohesin is required for long-range enhancer action at the Shh locus.” Nature Structural & Molecular Biology 29:891–897, September 2022. https://doi.org/10.1038/s41594-022-00821-8. Related experimental chromatin mechanism. (kane2022cohesinisrequired pages 1-3)
- Bower G et al. “Conserved Cis-Acting Range Extender Element Mediates Extreme Long-Range Enhancer Activity in Mammals,” May 2024 preprint: https://doi.org/10.1101/2024.05.26.595809; later published as “Range extender mediates long-distance enhancer activity,” Nature, July 2025, https://doi.org/10.1038/s41586-025-09221-6. Related mouse mechanism; no additional human acheiropodia patients. (bower2024conservedcisactingrangeextender pages 4-8)
Bibliographic limitation: PMID numbers for these papers were not independently retrieved and are deliberately not supplied rather than guessed. Likewise, current Orphanet, HPO, GO, CL, UBERON, HGNC, ChEBI, and NCIT accession numbers require direct vocabulary validation before knowledge-base import. No verified 2023–2024 study updating the human clinical natural history or treatment of acheiropodia was identified; recent advances above principally concern the relevant regulatory biology. (ushiki2021deletionofctcf pages 1-2, ianakiev2001acheiropodiaiscaused pages 1-3, bower2024conservedcisactingrangeextender pages 4-8)
References
-
(ianakiev2001acheiropodiaiscaused pages 1-3): P. Ianakiev, M.J. van Baren, M.J. Daly, S.P.A. Toledo, M.G. Cavalcanti, J. Correa Neto, E. Lemos Silveira, A. Freire-Maia, P. Heutink, M.W. Kilpatrick, and P. Tsipouras. Acheiropodia is caused by a genomic deletion in c7orf2, the human orthologue of the lmbr1 gene. American journal of human genetics, 68 1:38-45, Jan 2001. URL: https://doi.org/10.1086/316955, doi:10.1086/316955. This article has 126 citations and is from a highest quality peer-reviewed journal.
-
(ushiki2021deletionofctcf pages 1-2): Aki Ushiki, Yichi Zhang, Chenling Xiong, Jingjing Zhao, Ilias Georgakopoulos-Soares, Lauren Kane, Kirsty Jamieson, Michael J. Bamshad, Deborah A. Nickerson, Yin Shen, Laura A. Lettice, Elizabeth Lemos Silveira-Lucas, Florence Petit, and Nadav Ahituv. Deletion of ctcf sites in the shh locus alters enhancer–promoter interactions and leads to acheiropodia. Nature Communications, Apr 2021. URL: https://doi.org/10.1038/s41467-021-22470-z, doi:10.1038/s41467-021-22470-z. This article has 74 citations and is from a highest quality peer-reviewed journal.
-
(OpenTargets Search: Acheiropodia): Open Targets Query (Acheiropodia, 1 results). Buniello, A. et al. (2025). Open Targets Platform: facilitating therapeutic hypotheses building in drug discovery. Nucleic Acids Research.
-
(ianakiev2001acheiropodiaiscaused media 964b69bc): P. Ianakiev, M.J. van Baren, M.J. Daly, S.P.A. Toledo, M.G. Cavalcanti, J. Correa Neto, E. Lemos Silveira, A. Freire-Maia, P. Heutink, M.W. Kilpatrick, and P. Tsipouras. Acheiropodia is caused by a genomic deletion in c7orf2, the human orthologue of the lmbr1 gene. American journal of human genetics, 68 1:38-45, Jan 2001. URL: https://doi.org/10.1086/316955, doi:10.1086/316955. This article has 126 citations and is from a highest quality peer-reviewed journal.
-
(ianakiev2001acheiropodiaiscaused pages 4-7): P. Ianakiev, M.J. van Baren, M.J. Daly, S.P.A. Toledo, M.G. Cavalcanti, J. Correa Neto, E. Lemos Silveira, A. Freire-Maia, P. Heutink, M.W. Kilpatrick, and P. Tsipouras. Acheiropodia is caused by a genomic deletion in c7orf2, the human orthologue of the lmbr1 gene. American journal of human genetics, 68 1:38-45, Jan 2001. URL: https://doi.org/10.1086/316955, doi:10.1086/316955. This article has 126 citations and is from a highest quality peer-reviewed journal.
-
(ushiki2021deletionofctcf pages 10-10): Aki Ushiki, Yichi Zhang, Chenling Xiong, Jingjing Zhao, Ilias Georgakopoulos-Soares, Lauren Kane, Kirsty Jamieson, Michael J. Bamshad, Deborah A. Nickerson, Yin Shen, Laura A. Lettice, Elizabeth Lemos Silveira-Lucas, Florence Petit, and Nadav Ahituv. Deletion of ctcf sites in the shh locus alters enhancer–promoter interactions and leads to acheiropodia. Nature Communications, Apr 2021. URL: https://doi.org/10.1038/s41467-021-22470-z, doi:10.1038/s41467-021-22470-z. This article has 74 citations and is from a highest quality peer-reviewed journal.
-
(ushiki2021deletionofctcf pages 4-5): Aki Ushiki, Yichi Zhang, Chenling Xiong, Jingjing Zhao, Ilias Georgakopoulos-Soares, Lauren Kane, Kirsty Jamieson, Michael J. Bamshad, Deborah A. Nickerson, Yin Shen, Laura A. Lettice, Elizabeth Lemos Silveira-Lucas, Florence Petit, and Nadav Ahituv. Deletion of ctcf sites in the shh locus alters enhancer–promoter interactions and leads to acheiropodia. Nature Communications, Apr 2021. URL: https://doi.org/10.1038/s41467-021-22470-z, doi:10.1038/s41467-021-22470-z. This article has 74 citations and is from a highest quality peer-reviewed journal.
-
(ushiki2021deletionofctcf pages 2-4): Aki Ushiki, Yichi Zhang, Chenling Xiong, Jingjing Zhao, Ilias Georgakopoulos-Soares, Lauren Kane, Kirsty Jamieson, Michael J. Bamshad, Deborah A. Nickerson, Yin Shen, Laura A. Lettice, Elizabeth Lemos Silveira-Lucas, Florence Petit, and Nadav Ahituv. Deletion of ctcf sites in the shh locus alters enhancer–promoter interactions and leads to acheiropodia. Nature Communications, Apr 2021. URL: https://doi.org/10.1038/s41467-021-22470-z, doi:10.1038/s41467-021-22470-z. This article has 74 citations and is from a highest quality peer-reviewed journal.
-
(ushiki2021deletionofctcf pages 7-9): Aki Ushiki, Yichi Zhang, Chenling Xiong, Jingjing Zhao, Ilias Georgakopoulos-Soares, Lauren Kane, Kirsty Jamieson, Michael J. Bamshad, Deborah A. Nickerson, Yin Shen, Laura A. Lettice, Elizabeth Lemos Silveira-Lucas, Florence Petit, and Nadav Ahituv. Deletion of ctcf sites in the shh locus alters enhancer–promoter interactions and leads to acheiropodia. Nature Communications, Apr 2021. URL: https://doi.org/10.1038/s41467-021-22470-z, doi:10.1038/s41467-021-22470-z. This article has 74 citations and is from a highest quality peer-reviewed journal.
-
(ushiki2021deletionofctcf pages 5-7): Aki Ushiki, Yichi Zhang, Chenling Xiong, Jingjing Zhao, Ilias Georgakopoulos-Soares, Lauren Kane, Kirsty Jamieson, Michael J. Bamshad, Deborah A. Nickerson, Yin Shen, Laura A. Lettice, Elizabeth Lemos Silveira-Lucas, Florence Petit, and Nadav Ahituv. Deletion of ctcf sites in the shh locus alters enhancer–promoter interactions and leads to acheiropodia. Nature Communications, Apr 2021. URL: https://doi.org/10.1038/s41467-021-22470-z, doi:10.1038/s41467-021-22470-z. This article has 74 citations and is from a highest quality peer-reviewed journal.
-
(bower2024conservedcisactingrangeextender pages 4-8): Grace Bower, Ethan W. Hollingsworth, Sandra Jacinto, Benjamin Clock, Kaitlyn Cao, Mandy Liu, Adam Dziulko, Ana Alcaina-Caro, Qianlan Xu, Dorota Skowronska-Krawczyk, Javier Lopez-Rios, Diane E. Dickel, Anaïs F. Bardet, Len A. Pennacchio, Axel Visel, and Evgeny Z. Kvon. Conservedcis-acting range extender element mediates extreme long-range enhancer activity in mammals. BioRxiv, May 2024. URL: https://doi.org/10.1101/2024.05.26.595809, doi:10.1101/2024.05.26.595809. This article has 25 citations.
-
(koyanonakagawa2022etv2regulatesenhancer pages 1-2): Naoko Koyano-Nakagawa, Wuming Gong, Satyabrata Das, Joshua W. M. Theisen, Tran B. Swanholm, Daniel Van Ly, Nikita Dsouza, Bhairab N. Singh, Hiroko Kawakami, Samantha Young, Katherine Q. Chen, Yasuhiko Kawakami, and Daniel J. Garry. Etv2 regulates enhancer chromatin status to initiate shh expression in the limb bud. Nature Communications, Jul 2022. URL: https://doi.org/10.1038/s41467-022-31848-6, doi:10.1038/s41467-022-31848-6. This article has 22 citations and is from a highest quality peer-reviewed journal.
-
(ushiki2021deletionofctcf pages 9-10): Aki Ushiki, Yichi Zhang, Chenling Xiong, Jingjing Zhao, Ilias Georgakopoulos-Soares, Lauren Kane, Kirsty Jamieson, Michael J. Bamshad, Deborah A. Nickerson, Yin Shen, Laura A. Lettice, Elizabeth Lemos Silveira-Lucas, Florence Petit, and Nadav Ahituv. Deletion of ctcf sites in the shh locus alters enhancer–promoter interactions and leads to acheiropodia. Nature Communications, Apr 2021. URL: https://doi.org/10.1038/s41467-021-22470-z, doi:10.1038/s41467-021-22470-z. This article has 74 citations and is from a highest quality peer-reviewed journal.
-
(kane2022cohesinisrequired pages 1-3): Lauren Kane, Iain Williamson, Ilya M. Flyamer, Yatendra Kumar, Robert E. Hill, Laura A. Lettice, and Wendy A. Bickmore. Cohesin is required for long-range enhancer action at the shh locus. Nature Structural & Molecular Biology, 29:891-897, Sep 2022. URL: https://doi.org/10.1038/s41594-022-00821-8, doi:10.1038/s41594-022-00821-8. This article has 192 citations and is from a highest quality peer-reviewed journal.
-
(vandermeer2011cis‐regulatorymutationsare pages 6-7): Julia E. VanderMeer and Nadav Ahituv. Cis‐regulatory mutations are a genetic cause of human limb malformations. Developmental Dynamics, 240:920-930, May 2011. URL: https://doi.org/10.1002/dvdy.22535, doi:10.1002/dvdy.22535. This article has 68 citations and is from a peer-reviewed journal.
Artifacts
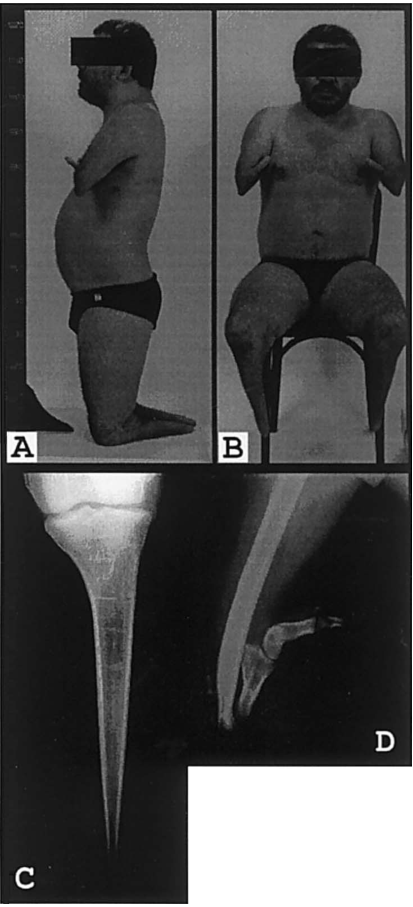
Reference Validation
Checked with linkml-reference-validator 0.3.0.
| Outcome | Count |
|---|---|
| References checked | 7 |
| Resolved | 7 |
| Unresolved (possible confabulation) | 0 |
| Unverifiable | 0 |
| References weighed for topical relevance | 7 |
| On topic | 1 |
| Off topic | 0 |
All extracted references resolved successfully.
Term Validation
Checked with linkml-term-validator 0.4.5, through the ols: adapter.
| Outcome | Count |
|---|---|
| Terms checked | 1 |
| Resolved | 1 |
| Unresolved (possible confabulation) | 0 |
| Obsolete | 0 |
| Unverifiable | 0 |
| Terms whose name was checked | 1 |
| Terms named correctly | 0 |
| Terms named as a different term | 1 |
Terms the report names something else
These identifiers resolve, so nothing about them looks wrong, and the ontology calls them something unrelated to what the report calls them. That usually means the identifier is not the one the sentence needs:
MONDO:0008700(3 mentions) - the report calls it "if available"; MONDO calls it acheiropody