ATP6V0C-Related Epilepsy
An epilepsy and neurodevelopmental disorder caused by heterozygous, often de novo, variants in ATP6V0C, encoding the c subunit of the membrane V0 domain of the vacuolar H+-ATPase. Its clinical spectrum includes familial febrile seizures or epilepsy with febrile seizures plus and normal development, developmental delay and intellectual disability with or without epilepsy, and Dravet-like developmental and epileptic encephalopathy persisting into adulthood. Seizures often begin in infancy or early childhood and can be fever-sensitive. Callosal or cerebellar vermian abnormalities and delayed myelination occur in some patients; brain MRI can also be normal. Yeast and invertebrate experiments support impaired V-ATPase function, but the intervening mechanisms producing the human neurological phenotype remain incompletely resolved. ATP6V0C is a proposed contributor to overlapping multigene 16p13.3 deletion phenotypes; these deletions do not isolate its effect.
Ask OpenScientist
Ask a research question about ATP6V0C-Related Epilepsy. OpenScientist will conduct autonomous deep research using the Disorder Mechanisms Knowledge Base and PubMed literature (typically 10-30 minutes).
Do not include personal health information in your question. Questions and results are cached in your browser's local storage.
Inheritance
1Show evidence (3 references)
Subtypes
3Show evidence (1 reference)
Show evidence (1 reference)
Show evidence (2 references)
Mechanistic Hypotheses
2Show evidence (1 reference)
Show evidence (1 reference)
Discussions and Knowledge Gaps
1Show evidence (1 reference)
Pathophysiology
7Show evidence (4 references)
Show evidence (1 reference)
Show evidence (2 references)
Show evidence (1 reference)
Show evidence (1 reference)
Show evidence (1 reference)
Show evidence (1 reference)
Pathograph
Phenotypes
28Cardiovascular 3
Show evidence (1 reference)
Show evidence (1 reference)
Show evidence (1 reference)
Head and Neck 1
Show evidence (1 reference)
Musculoskeletal 1
Show evidence (1 reference)
Nervous System 23
Show evidence (3 references)
Context-specific annotations (1)
Show evidence (1 reference)
Show evidence (2 references)
Context-specific annotations (1)
Show evidence (1 reference)
Show evidence (1 reference)
Context-specific annotations (1)
Show evidence (1 reference)
Show evidence (1 reference)
Context-specific annotations (1)
Show evidence (1 reference)
Show evidence (1 reference)
Context-specific annotations (1)
Show evidence (1 reference)
Show evidence (2 references)
Context-specific annotations (1)
Show evidence (1 reference)
Show evidence (2 references)
Show evidence (3 references)
Context-specific annotations (1)
Show evidence (1 reference)
Show evidence (1 reference)
Context-specific annotations (1)
Show evidence (1 reference)
Show evidence (1 reference)
Context-specific annotations (1)
Show evidence (1 reference)
Show evidence (1 reference)
Context-specific annotations (1)
Show evidence (1 reference)
Show evidence (1 reference)
Show evidence (2 references)
Show evidence (1 reference)
Show evidence (2 references)
Show evidence (1 reference)
Show evidence (1 reference)
Show evidence (2 references)
Show evidence (1 reference)
Show evidence (1 reference)
Show evidence (2 references)
Show evidence (1 reference)
Show evidence (2 references)
Genetic Associations
1Show evidence (4 references)
Medical Actions
1Show evidence (4 references)
Diagnosis
1Show evidence (2 references)
Prevalence
1Differential Diagnoses
3Conditions with similar clinical presentations that must be differentiated from ATP6V0C-Related Epilepsy:
- A pathogenic ATP6V0C variant with no SCN1A variant favors this disorder.
Show evidence (2 references)
- Segregating ATP6V0C variants were reported in two FS/EFS+ families with normal development; clinical overlap alone does not identify the causal gene.
Show evidence (1 reference)
- Microcephaly is common in the deletion.
- Detected by chromosomal microarray.
Show evidence (2 references)
Experimental Models
1Show evidence (2 references)
Animal Models
2Show evidence (3 references)
Show evidence (4 references)
Source YAML
click to showname: ATP6V0C-Related Epilepsy
creation_date: "2026-10-01T00:00:00Z"
description: >-
An epilepsy and neurodevelopmental disorder caused by heterozygous, often de novo, variants in ATP6V0C, encoding the c subunit of the membrane V0 domain of the vacuolar H+-ATPase. Its clinical spectrum includes familial febrile seizures or epilepsy with febrile seizures plus and normal development, developmental delay and intellectual disability with or without epilepsy, and Dravet-like developmental and epileptic encephalopathy persisting into adulthood. Seizures often begin in infancy or early childhood and can be fever-sensitive. Callosal or cerebellar vermian abnormalities and delayed myelination occur in some patients; brain MRI can also be normal. Yeast and invertebrate experiments support impaired V-ATPase function, but the intervening mechanisms producing the human neurological phenotype remain incompletely resolved. ATP6V0C is a proposed contributor to overlapping multigene 16p13.3 deletion phenotypes; these deletions do not isolate its effect.
category: Mendelian
parents:
- Epilepsy
- Neurodevelopmental Disorder
synonyms:
- ATP6V0C-related neurodevelopmental disorder
- ATP6V0C deficiency
disease_term:
preferred_term: epilepsy, early-onset, 3, with or without developmental delay
term:
id: MONDO:0958196
label: epilepsy, early-onset, 3, with or without developmental delay
inheritance:
- name: Autosomal Dominant
inheritance_term:
preferred_term: Autosomal dominant inheritance
term:
id: HP:0000006
label: Autosomal dominant inheritance
description: >-
Heterozygous variants act dominantly. Most arise de novo, with somatic
mosaicism in a few probands; in milder fever-associated families the variant
cosegregates with affected relatives.
evidence:
- reference: PMID:36074901
reference_title: "ATP6V0C variants impair V-ATPase function causing a neurodevelopmental disorder often associated with epilepsy."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "Second, in 24 patients where biologic parent DNA was available, the variants were found to have occurred de novo."
explanation: >-
De novo occurrence of heterozygous variants in every tested trio of the
founding cohort supports dominant action.
- reference: PMID:35600075
reference_title: "ATP6V0C Is Associated With Febrile Seizures and Epilepsy With Febrile Seizures Plus."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "We identified two heterozygous ATP6V0C mutations that co-segregated with the disease in two unrelated families with six individuals affected."
explanation: >-
Cosegregation of heterozygous variants with febrile seizures in two
pedigrees shows familial dominant transmission at the mild end.
- reference: PMID:36074901
reference_title: "ATP6V0C variants impair V-ATPase function causing a neurodevelopmental disorder often associated with epilepsy."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "Patients 4, 9, 15 and 19 were each found to be mosaic for their identified ATP6V0C variant."
explanation: >-
Four of 27 founding-cohort patients carry their variant in mosaic form.
mechanistic_hypotheses:
- hypothesis_group_id: vesicle_loading_deficit
hypothesis_label: >-
Reduced V-ATPase pumping underfills synaptic vesicles and lowers transmitter
release
status: EMERGING
description: >-
Patient-equivalent variants impair vacuolar acidification in yeast. Applying the established dependence of synaptic vesicle loading on the proton electrochemical gradient suggests a route through vesicle underfilling and altered neurotransmission. The ATP6V0C clinical and functional studies cited here do not directly measure vesicle transmitter content in patient neurons or establish preferential failure of inhibitory transmission.
evidence:
- reference: PMID:29652249
reference_title: "Clathrin coat controls synaptic vesicle acidification by blocking vacuolar ATPase activity."
supports: SUPPORT
evidence_source: IN_VITRO
directness: INDIRECT
snippet: "Newly-formed synaptic vesicles (SVs) are rapidly acidified by vacuolar adenosine triphosphatases (vATPases), generating a proton electrochemical gradient that drives neurotransmitter loading."
explanation: >-
States the general dependency of vesicle loading on V-ATPase pumping that
the hypothesis assumes.
quote_role: BACKGROUND
- hypothesis_group_id: v0c_fusion_facilitation
hypothesis_label: >-
Loss of a proton-pump-independent c-subunit role in vesicle fusion
status: EMERGING
description: >-
Acute c-subunit inactivation inhibits transmission in CA3 pyramidal neurons, supporting an acidification-independent contribution to neurotransmission. Impaired fusion is a proposed route for patient variants; the cited experiment does not test those variants or demonstrate a disease-specific fusion defect.
evidence:
- reference: PMID:30155790
reference_title: "Chromophore-Assisted Light Inactivation of the V-ATPase V0c Subunit Inhibits Neurotransmitter Release Downstream of Synaptic Vesicle Acidification."
supports: SUPPORT
evidence_source: IN_VITRO
directness: INDIRECT
snippet: "The c-subunit interacts with the v-SNARE VAMP2 and facilitates neurotransmission."
explanation: >-
Provides the general V0c/VAMP2 interaction underlying the hypothesis, not a patient-variant result.
quote_role: BACKGROUND
pathophysiology:
- name: ATP6V0C Heterozygous Variant
conforms_to: "synaptic_vesicle_cycle#Synaptic Vesicle Cycle Protein Deficiency"
description: >-
Heterozygous ATP6V0C variants are the initiating genetic lesion; both de novo and familial variants occur. Selected patient-equivalent missense variants impair V-ATPase function in yeast. Structural modelling proposes interference with c-ring rotation, but the yeast null-complementation experiment does not establish dominant-negative action in human heterozygotes. Reduced total leukocyte RNA in one stop-loss carrier suggests a dosage contribution; mutant RNA persists, and the balance between haploinsufficiency and altered-protein effects remains unresolved.
notes: >-
Four founding-cohort patients were mosaic. Their seizure histories varied, including early-onset epilepsy, late onset, unconfirmed seizure-like episodes and no reported seizures; mosaicism does not guarantee a mild presentation.
biological_scale: MOLECULAR
role: trigger
genes:
- preferred_term: ATP6V0C
term:
id: hgnc:855
label: ATP6V0C
genetic_context:
variant_origin: GERMLINE
zygosity: HETEROZYGOUS
molecular_functions:
- preferred_term: V-ATPase proton pumping activity
term:
id: GO:0046961
label: proton-transporting ATPase activity, rotational mechanism
modifier: DECREASED
cell_types:
- preferred_term: neuron
term:
id: CL:0000540
label: neuron
evidence:
- reference: PMID:36074901
reference_title: "ATP6V0C variants impair V-ATPase function causing a neurodevelopmental disorder often associated with epilepsy."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "We describe heterozygous point variants in ATP6V0C, encoding the c-subunit in the membrane bound integral domain of the vacuolar H+-ATPase, in 27 patients with neurodevelopmental abnormalities with or without epilepsy."
explanation: >-
Founding cohort establishing heterozygous ATP6V0C variants as the lesion.
- reference: PMID:33190975
reference_title: "Novel de novo mutation substantiates ATP6V0C as a gene causing epilepsy with intellectual disability."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "real-time PCR demonstrated that the patient's ATP6V0C RNA level was approximately half of that in her parents, suggesting haploinsufficiency as a pathomechanism."
explanation: >-
Approximately halved total leukocyte RNA in one stop-loss carrier suggests haploinsufficiency, without establishing the mechanism of every stop-loss or truncating allele.
- reference: PMID:36074901
reference_title: "ATP6V0C variants impair V-ATPase function causing a neurodevelopmental disorder often associated with epilepsy."
supports: SUPPORT
evidence_source: COMPUTATIONAL
directness: INDIRECT
snippet: "Therefore, we speculate that missense variants and those predicted to escape NMD act via a dominant negative mechanism, while nonsense variants and microdeletions containing ATP6V0C act via haploinsufficiency as demonstrated by our Drosophila data"
explanation: >-
The authors explicitly speculate about dominant-negative effects in missense and NMD-escaping variants; structural modelling is not an experimental demonstration of dominant-negative action.
- reference: PMID:33190975
reference_title: Novel de novo mutation substantiates ATP6V0C as a gene causing epilepsy with intellectual disability.
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: Using RNA from leukocytes, RT-PCR followed by Sanger sequencing showed the existence of the mutant RNA
explanation: Mutant transcript persistence limits a simple assumption of complete nonsense-mediated decay.
downstream:
- target: Reduced V-ATPase Proton Pumping
causal_link_type: DIRECT
description: >-
Patient-equivalent substitutions in yeast VMA3 reduce acidification in a vma3-null complementation assay, with variant-dependent residual activity.
evidence:
- reference: PMID:36074901
reference_title: "ATP6V0C variants impair V-ATPase function causing a neurodevelopmental disorder often associated with epilepsy."
supports: SUPPORT
evidence_source: IN_VITRO
snippet: "When the uptake of LysoSensor was measured, we saw that nine variants were associated with little to no fluorescence, indicating significant reduction or loss of V-ATPase activity."
explanation: >-
Nine of twelve tested patient-equivalent variants gave little or no LysoSensor fluorescence. Two retained intermediate signal and F137L-equivalent signal was comparable to wild type; fluorescence is an acidification surrogate.
- target: Impaired V0c-Dependent Facilitation of Vesicle Fusion
causal_link_type: DIRECT
hypothesis_groups:
- v0c_fusion_facilitation
description: >-
Hypothesized extrapolation from acute c-subunit inactivation in neurons; the cited experiment did not test ATP6V0C patient alleles.
evidence:
- reference: PMID:30155790
reference_title: Chromophore-Assisted Light Inactivation of the V-ATPase V0c Subunit Inhibits Neurotransmitter Release Downstream of Synaptic Vesicle Acidification.
supports: SUPPORT
evidence_source: IN_VITRO
quote_role: PRIMARY_RESULT
directness: INDIRECT
snippet: We show that V-ATPase c-subunit V0c is a key element in modulating neurotransmission and that its specific inactivation rapidly inhibited neurotransmission.
explanation: Acute inactivation supports a possible route, not a demonstrated effect of a patient allele.
- target: Global Developmental Delay
description: Genetic evidence links ATP6V0C variants to the neurodevelopmental phenotype; the intervening cellular pathway and any contribution from seizures remain unresolved.
causal_link_type: INDIRECT_UNKNOWN_INTERMEDIATES
evidence:
- reference: PMID:36074901
reference_title: ATP6V0C variants impair V-ATPase function causing a neurodevelopmental disorder often associated with epilepsy.
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: Collectively, these data show that ATP6V0C variants cause a human syndrome of developmental delay, intellectual disability and epilepsy.
explanation: Supports gene-to-phenotype causality without assuming that seizures cause the developmental impairment.
- target: Intellectual Disability
description: Genetic evidence links ATP6V0C variants to the neurodevelopmental phenotype; the intervening cellular pathway and any contribution from seizures remain unresolved.
causal_link_type: INDIRECT_UNKNOWN_INTERMEDIATES
evidence:
- reference: PMID:36074901
reference_title: ATP6V0C variants impair V-ATPase function causing a neurodevelopmental disorder often associated with epilepsy.
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: Collectively, these data show that ATP6V0C variants cause a human syndrome of developmental delay, intellectual disability and epilepsy.
explanation: Supports gene-to-phenotype causality without assuming that seizures cause the developmental impairment.
- target: Delayed Speech and Language Development
description: Language impairment is part of the genetically associated neurodevelopmental phenotype; no separate cellular pathway has been established.
causal_link_type: INDIRECT_UNKNOWN_INTERMEDIATES
evidence:
- reference: PMID:40085430
reference_title: Variants in ATP6V0C are associated with Dravet-like developmental and epileptic encephalopathy.
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: INDIRECT
snippet: At age 31 years, he had autism spectrum disorder and severe ID, speaking only 1- to 2-word sentences.
explanation: Observed in an adult with a de novo ATP6V0C variant; the intermediate mechanism is unresolved.
- target: Aplasia or Hypoplasia of the Corpus Callosum
description: Observed within the ATP6V0C-associated neurodevelopmental phenotype; the intervening developmental or cellular mechanism is not established.
causal_link_type: INDIRECT_UNKNOWN_INTERMEDIATES
evidence:
- reference: PMID:36074901
reference_title: "ATP6V0C variants impair V-ATPase function causing a neurodevelopmental disorder often associated with epilepsy."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "Five patients with MRIs show hypoplasia or agenesis of the corpus callosum."
explanation: >-
Corpus callosum agenesis or hypoplasia in 5 of 21 imaged patients (24%).
directness: INDIRECT
- target: Aplasia or Hypoplasia of the Cerebellar Vermis
description: Observed within the ATP6V0C-associated neurodevelopmental phenotype; the intervening developmental or cellular mechanism is not established.
causal_link_type: INDIRECT_UNKNOWN_INTERMEDIATES
evidence:
- reference: PMID:36074901
reference_title: ATP6V0C variants impair V-ATPase function causing a neurodevelopmental disorder often associated with epilepsy.
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: INDIRECT
snippet: Common findings in MRIs included agenesis/hypoplasia of the corpus callosum (Supplementary Fig. 2) or cerebellar vermis (Patients 5, 6, 18, 20 and 26), and delayed myelination (Patients 6, 18, 22 and 24).
explanation: Supports vermian malformation without assigning the combined patient list entirely to that finding.
- target: Delayed Myelination
description: Observed within the ATP6V0C-associated neurodevelopmental phenotype; the intervening developmental or cellular mechanism is not established.
causal_link_type: INDIRECT_UNKNOWN_INTERMEDIATES
evidence:
- reference: PMID:36074901
reference_title: "ATP6V0C variants impair V-ATPase function causing a neurodevelopmental disorder often associated with epilepsy."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "and delayed myelination (Patients 6, 18, 22 and 24)"
explanation: >-
Delayed myelination in 4 of 21 imaged patients (19%).
directness: INDIRECT
- target: Autism
description: Observed within the ATP6V0C-associated neurodevelopmental phenotype; the intervening developmental or cellular mechanism is not established.
causal_link_type: INDIRECT_UNKNOWN_INTERMEDIATES
evidence:
- reference: PMID:40085430
reference_title: Variants in ATP6V0C are associated with Dravet-like developmental and epileptic encephalopathy.
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: INDIRECT
snippet: At age 31 years, he had autism spectrum disorder and severe ID, speaking only 1- to 2-word sentences.
explanation: Patient 1 adult phenotype.
- target: Absent Speech
description: Observed within the genetically associated neurodevelopmental phenotype; no separate cellular intermediate is established.
causal_link_type: INDIRECT_UNKNOWN_INTERMEDIATES
evidence:
- reference: url:https://pmc.ncbi.nlm.nih.gov/articles/PMC10319782/?report=reader
reference_title: ATP6V0C variants impair V-ATPase function causing a neurodevelopmental disorder often associated with epilepsy - PMC
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: INDIRECT
snippet: Developmental delay ... p.A95P ... Yes, motor and non-verbal
explanation: 'Table 1 developmental-delay column, patient 10 (p.A95P): non-verbal status. Ellipses omit intervening table rows and columns.'
- target: Developmental Regression
description: Observed within the genetically associated neurodevelopmental phenotype; no separate cellular intermediate is established.
causal_link_type: INDIRECT_UNKNOWN_INTERMEDIATES
evidence:
- reference: url:https://pmc.ncbi.nlm.nih.gov/articles/PMC10319782/?report=reader
reference_title: ATP6V0C variants impair V-ATPase function causing a neurodevelopmental disorder often associated with epilepsy - PMC
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: INDIRECT
snippet: Developmental delay ... p.A95T ... Yes, regression to non-verbal
explanation: 'Table 1 developmental-delay column, patient 9 (p.A95T): regression to non-verbal status. Ellipses omit intervening table rows and columns.'
- name: Reduced V-ATPase Proton Pumping
description: >-
V-ATPase dysfunction is supported by reduced vacuolar acidification and calcium-sensitive growth in yeast expressing patient-equivalent VMA3 variants. These readouts are indirect measures of pump function; they do not directly measure proton flux in human neurons. Effects vary by allele and assay condition.
biological_scale: MOLECULAR
biological_processes:
- preferred_term: proton transmembrane transport
term:
id: GO:1902600
label: proton transmembrane transport
modifier: DECREASED
- preferred_term: vacuolar acidification
term:
id: GO:0007035
label: vacuolar acidification
modifier: DECREASED
evidence:
- reference: PMID:36074901
reference_title: "ATP6V0C variants impair V-ATPase function causing a neurodevelopmental disorder often associated with epilepsy."
supports: SUPPORT
evidence_source: IN_VITRO
snippet: "Consistent with decreased vacuolar H+-ATPase activity, functional analyses conducted in Saccharomyces cerevisiae revealed reduced LysoSensor fluorescence and reduced growth in media containing varying concentrations of CaCl2."
explanation: >-
Yeast complementation assays show reduced V-ATPase function for patient
variants.
downstream:
- target: Impaired Synaptic Vesicle Acidification
causal_link_type: DIRECT
hypothesis_groups:
- vesicle_loading_deficit
description: >-
Proposed transfer of the yeast pump defect to neuronal vesicles, based on the established function of V-ATPase in synaptic vesicle acidification.
mechanism_confidence: PROVISIONAL
- name: Impaired Synaptic Vesicle Acidification
description: >-
Reduced V-ATPase function could diminish the proton electrochemical gradient across synaptic vesicle membranes. This is a disease-specific inference from yeast assays and general vesicle physiology, not a patient-neuron measurement.
biological_scale: CELLULAR
cell_types:
- preferred_term: neuron
term:
id: CL:0000540
label: neuron
biological_processes:
- preferred_term: synaptic vesicle lumen acidification
term:
id: GO:0097401
label: synaptic vesicle lumen acidification
modifier: DECREASED
evidence:
- reference: PMID:30155790
reference_title: "Chromophore-Assisted Light Inactivation of the V-ATPase V0c Subunit Inhibits Neurotransmitter Release Downstream of Synaptic Vesicle Acidification."
supports: SUPPORT
evidence_source: IN_VITRO
quote_role: BACKGROUND
directness: INDIRECT
snippet: "Active acidification of synaptic vesicles, triggered by the V-ATPase, is necessary for neurotransmitter storage."
explanation: >-
States the dependency of transmitter storage on V-ATPase acidification.
- reference: PMID:35600075
reference_title: "ATP6V0C Is Associated With Febrile Seizures and Epilepsy With Febrile Seizures Plus."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: BACKGROUND
snippet: "As a component of V0 domain of V-ATPase, ATP6V0C forms the proteolipid c-ring of the V-ATPase and plays a pivotal role in synaptic vesicle proton gradient generation and regulation of intra and extracellular pH value"
explanation: >-
Places ATP6V0C specifically in synaptic vesicle proton-gradient generation.
downstream:
- target: Reduced Neurotransmitter Loading
description: The vesicular proton electrochemical gradient drives neurotransmitter uptake; diminished loading in ATP6V0C-related disease is inferred.
causal_link_type: DIRECT
hypothesis_groups:
- vesicle_loading_deficit
evidence:
- reference: PMID:29652249
reference_title: Clathrin coat controls synaptic vesicle acidification by blocking vacuolar ATPase activity.
supports: SUPPORT
evidence_source: IN_VITRO
quote_role: BACKGROUND
directness: INDIRECT
snippet: Newly-formed synaptic vesicles (SVs) are rapidly acidified by vacuolar adenosine triphosphatases (vATPases), generating a proton electrochemical gradient that drives neurotransmitter loading.
explanation: General dependence of loading on acidification.
mechanism_confidence: HYPOTHETICAL
- name: Reduced Neurotransmitter Loading
description: A reduced vesicular proton gradient could reduce neurotransmitter filling. The identity of the most affected transmitter system and vesicle cargo levels in patient neurons were not established by the cited ATP6V0C studies.
biological_scale: CELLULAR
mechanism_confidence: HYPOTHETICAL
evidence:
- reference: PMID:29652249
reference_title: Clathrin coat controls synaptic vesicle acidification by blocking vacuolar ATPase activity.
supports: SUPPORT
evidence_source: IN_VITRO
quote_role: BACKGROUND
directness: INDIRECT
snippet: Newly-formed synaptic vesicles (SVs) are rapidly acidified by vacuolar adenosine triphosphatases (vATPases), generating a proton electrochemical gradient that drives neurotransmitter loading.
explanation: Background physiology supports this proposed downstream consequence.
biological_processes:
- preferred_term: neurotransmitter loading into synaptic vesicle
term:
id: GO:0098700
label: neurotransmitter loading into synaptic vesicle
modifier: DECREASED
downstream:
- target: Reduced Neurotransmitter Release
description: Vesicle underfilling could reduce transmitter output per fusion event; this route remains hypothetical for ATP6V0C patient variants.
causal_link_type: DIRECT
hypothesis_groups:
- vesicle_loading_deficit
- name: Impaired V0c-Dependent Facilitation of Vesicle Fusion
conforms_to: "synaptic_vesicle_cycle#Impaired Calcium-Triggered SNARE-Mediated Vesicle Fusion"
description: >-
The membrane V0 sector interacts with SNARE machinery, and acute light inactivation of V0c in CA3 pyramidal neurons inhibits transmission. Impaired facilitation of fusion is one proposed disease mechanism, distinct from vesicle acidification; the cited neuronal experiment did not test patient variants.
biological_scale: CELLULAR
cell_types:
- preferred_term: neuron
term:
id: CL:0000540
label: neuron
evidence:
- reference: PMID:30155790
reference_title: "Chromophore-Assisted Light Inactivation of the V-ATPase V0c Subunit Inhibits Neurotransmitter Release Downstream of Synaptic Vesicle Acidification."
supports: SUPPORT
evidence_source: IN_VITRO
snippet: "We show that V-ATPase c-subunit V0c is a key element in modulating neurotransmission and that its specific inactivation rapidly inhibited neurotransmission."
explanation: >-
Direct inactivation of V0c in CA3 pyramidal neurons inhibits transmission.
directness: INDIRECT
downstream:
- target: Reduced Neurotransmitter Release
causal_link_type: DIRECT
hypothesis_groups:
- v0c_fusion_facilitation
mechanism_confidence: HYPOTHETICAL
- name: Reduced Neurotransmitter Release
conforms_to: "synaptic_vesicle_cycle#Neurotransmitter Release Failure and Synaptic Transmission Deficit"
description: >-
Reduced vesicle filling or impaired V0c-dependent facilitation of release could diminish transmitter output. Acute V0c inactivation inhibits neuronal transmission, but neither that experiment nor worm aldicarb hypersensitivity identifies a selective inhibitory-transmitter deficit in ATP6V0C-related disease.
biological_scale: CELLULAR
biological_processes:
- preferred_term: chemical synaptic transmission
term:
id: GO:0007268
label: chemical synaptic transmission
modifier: DECREASED
evidence:
- reference: PMID:30155790
reference_title: "Chromophore-Assisted Light Inactivation of the V-ATPase V0c Subunit Inhibits Neurotransmitter Release Downstream of Synaptic Vesicle Acidification."
supports: SUPPORT
evidence_source: IN_VITRO
directness: INDIRECT
snippet: "We show that V-ATPase c-subunit V0c is a key element in modulating neurotransmission and that its specific inactivation rapidly inhibited neurotransmission."
explanation: >-
Loss of c-subunit function reduces transmission in neurons; the patient
alleles themselves were not tested.
downstream:
- target: Neuronal Hyperexcitability and Seizure Susceptibility
causal_link_type: INDIRECT_UNKNOWN_INTERMEDIATES
hypothesis_groups:
- vesicle_loading_deficit
- v0c_fusion_facilitation
description: >-
A proposed transition from altered synaptic output to seizure susceptibility. The ATP6V0C studies do not establish the intervening network mechanism or preferential loss of inhibition.
mechanism_confidence: HYPOTHETICAL
- name: Neuronal Hyperexcitability and Seizure Susceptibility
conforms_to: "epilepsy_excitation_inhibition_imbalance#Neuronal Hyperexcitability and Hypersynchrony"
description: >-
Pan-neuronal knockdown of the fly orthologue prolongs electroshock-induced seizure-like behavior, supporting a link between reduced c-subunit function and seizure susceptibility. The model does not directly measure human network hyperexcitability or explain fever sensitivity.
biological_scale: TISSUE
locations:
- preferred_term: brain
term:
id: UBERON:0000955
label: brain
evidence:
- reference: PMID:36074901
reference_title: "ATP6V0C variants impair V-ATPase function causing a neurodevelopmental disorder often associated with epilepsy."
supports: SUPPORT
evidence_source: MODEL_ORGANISM
snippet: "Knockdown of ATP6V0C in Drosophila resulted in increased duration of seizure-like behaviour"
explanation: >-
Neuronal loss of the orthologue increases seizure susceptibility in vivo.
downstream:
- target: Seizures
causal_link_type: DIRECT
evidence:
- reference: PMID:36074901
reference_title: "ATP6V0C variants impair V-ATPase function causing a neurodevelopmental disorder often associated with epilepsy."
supports: SUPPORT
evidence_source: MODEL_ORGANISM
snippet: "These results are consistent with the hypothesis that haploinsufficiency of ATP6V0C contributes to seizures."
explanation: >-
The Drosophila knockdown result connects reduced ATP6V0C dosage to
seizures.
- target: Febrile Seizures
causal_link_type: DIRECT
- target: Bilateral Tonic-Clonic Seizures
causal_link_type: DIRECT
- target: Focal-Onset Seizures
causal_link_type: DIRECT
- target: Atonic Seizures
causal_link_type: DIRECT
- target: Myoclonic Seizures
causal_link_type: DIRECT
- target: Multifocal EEG Abnormalities
causal_link_type: DIRECT
- target: Status Epilepticus
description: Clinical manifestation of the seizure-susceptibility phenotype; specific circuitry is unresolved.
causal_link_type: INDIRECT_UNKNOWN_INTERMEDIATES
- target: Atypical Absence Seizures
description: Clinical manifestation of the seizure-susceptibility phenotype; specific circuitry is unresolved.
causal_link_type: INDIRECT_UNKNOWN_INTERMEDIATES
- target: Infantile Spasms
description: Another seizure manifestation documented in the ATP6V0C cohort; the specific neuronal circuitry is unknown.
causal_link_type: INDIRECT_UNKNOWN_INTERMEDIATES
evidence:
- reference: url:https://pmc.ncbi.nlm.nih.gov/articles/PMC10319782/?report=reader
reference_title: ATP6V0C variants impair V-ATPase function causing a neurodevelopmental disorder often associated with epilepsy - PMC
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: Infantile spasms, GTCS, At, Myo
explanation: Table 1, patient 6, documents infantile spasms together with generalized tonic-clonic, atonic and myoclonic seizures.
mechanism_confidence: PROVISIONAL
phenotypes:
- name: Seizures
category: Neurological
description: >-
Seizures begin in early childhood; in the founding cohort the mean onset age
was about two years and most began before 24 months. A few individuals,
including some mosaic carriers, have no seizures.
phenotype_term:
preferred_term: Seizure
term:
id: HP:0001250
label: Seizure
evidence:
- reference: PMID:36074901
reference_title: "ATP6V0C variants impair V-ATPase function causing a neurodevelopmental disorder often associated with epilepsy."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "The mean age of seizure onset was 24.6 ± 8.0 months, with 14 of 18 patients for whom this information was available having onset before 24 months."
explanation: >-
Quantifies early-childhood seizure onset in the founding cohort.
- reference: PMID:37161035
reference_title: "ATP6V0C gene variants were identified in individuals with epilepsy, with or without developmental delay."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "Whole exome sequencing was performed in three unrelated patients with early-onset epilepsy, with or without developmental delay and intellectual disability."
explanation: >-
An independent series of early-onset epilepsy with de novo ATP6V0C variants.
- reference: PMID:36074901
reference_title: "ATP6V0C variants impair V-ATPase function causing a neurodevelopmental disorder often associated with epilepsy."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "Patient 19 (p.G142D) has not reported any seizure or seizure-like episodes."
explanation: >-
A mosaic carrier without seizures shows that seizures are not universal.
phenotype_contexts:
- population: 'Mattison 2023 founding cohort: patients with seizure-onset information (n=18)'
frequency: 14/18
onset:
max_age_years: 2.0
notes: Onset strictly before 24 months in this subset; the upper bound is exclusive.
notes: Frequency refers to seizures beginning before age two among those with known onset, not overall seizure frequency or seizures assessed at age two.
evidence:
- reference: PMID:36074901
reference_title: "ATP6V0C variants impair V-ATPase function causing a neurodevelopmental disorder often associated with epilepsy."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "The mean age of seizure onset was 24.6 ± 8.0 months, with 14 of 18 patients for whom this information was available having onset before 24 months."
explanation: >-
Quantifies early-childhood seizure onset in the founding cohort.
- name: Febrile Seizures
category: Neurological
description: >-
Infantile febrile seizures occur in mild familial presentations and in severe Dravet-like cases. All six affected relatives in the two-family study had onset at 7–8 months. A later study selected patients for fever sensitivity, so its diagnostic yield is not a disease-wide phenotype frequency.
phenotype_term:
preferred_term: Febrile seizure
term:
id: HP:0002373
label: Febrile seizure (within the age range of 3 months to 6 years)
evidence:
- reference: PMID:35600075
reference_title: "ATP6V0C Is Associated With Febrile Seizures and Epilepsy With Febrile Seizures Plus."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "All the six affected individuals suffered from their first FS at the age of 7-8 months."
explanation: >-
Febrile seizures in infancy in every affected member of two families.
- reference: PMID:42526150
reference_title: "Epilepsy with fever-sensitivity in patients with ATP6V0C pathogenic variants."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "Notably, ATP6V0C variants accounted for 2/16 (12.5%) of all pathogenic/likely pathogenic findings and were identified in 3.8% of tested fever-sensitive epilepsy patients, ranking among the most frequently implicated genes in this cohort."
explanation: >-
A selected diagnostic cohort identified ATP6V0C variants in two of 53 fever-sensitive epilepsy patients (2/16 positive genetic findings); this does not estimate population prevalence or the fraction of ATP6V0C patients with fever sensitivity.
phenotype_contexts:
- population: 'Tian 2022: affected members of two FS/EFS+ families (n=6)'
frequency: 6/6
onset:
min_age_years: 0.5833333333333334
max_age_years: 0.6666666666666666
notes: First febrile seizure at 7–8 months; age bounds converted from months.
notes: Affected relatives were selected for study; this does not estimate penetrance among all variant carriers.
evidence:
- reference: PMID:35600075
reference_title: "ATP6V0C Is Associated With Febrile Seizures and Epilepsy With Febrile Seizures Plus."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "All the six affected individuals suffered from their first FS at the age of 7-8 months."
explanation: >-
Febrile seizures in infancy in every affected member of two families.
- name: Bilateral Tonic-Clonic Seizures
category: Neurological
phenotype_term:
preferred_term: Generalized tonic-clonic seizure
term:
id: HP:0002069
label: Bilateral tonic-clonic seizure
evidence:
- reference: PMID:36074901
reference_title: "ATP6V0C variants impair V-ATPase function causing a neurodevelopmental disorder often associated with epilepsy."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "On the basis of clinical information from 19 patients, the most common seizure types observed were generalized tonic-clonic (12/19), focal (7/19), atonic (6/19) and myoclonic (5/19)."
explanation: >-
Generalized tonic-clonic seizures in 12 of 19 patients (63%).
description: Reported in 12 of 19 founding-cohort patients with seizure-type information; seizure categories overlap and these counts are not population penetrance estimates.
phenotype_contexts:
- population: 'Mattison 2023 founding cohort: patients with seizure-type data (n=19)'
frequency: 12/19
notes: Selected clinical cohort; this count is not a population penetrance estimate.
evidence:
- reference: PMID:36074901
reference_title: "ATP6V0C variants impair V-ATPase function causing a neurodevelopmental disorder often associated with epilepsy."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "On the basis of clinical information from 19 patients, the most common seizure types observed were generalized tonic-clonic (12/19), focal (7/19), atonic (6/19) and myoclonic (5/19)."
explanation: >-
Generalized tonic-clonic seizures in 12 of 19 patients (63%).
- name: Focal-Onset Seizures
category: Neurological
phenotype_term:
preferred_term: Focal-onset seizure
term:
id: HP:0007359
label: Focal-onset seizure
evidence:
- reference: PMID:36074901
reference_title: "ATP6V0C variants impair V-ATPase function causing a neurodevelopmental disorder often associated with epilepsy."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "On the basis of clinical information from 19 patients, the most common seizure types observed were generalized tonic-clonic (12/19), focal (7/19), atonic (6/19) and myoclonic (5/19)."
explanation: >-
Focal seizures in 7 of 19 patients (37%).
description: Reported in 7 of 19 founding-cohort patients with seizure-type information; seizure categories overlap and these counts are not population penetrance estimates.
phenotype_contexts:
- population: 'Mattison 2023 founding cohort: patients with seizure-type data (n=19)'
frequency: 7/19
notes: Selected clinical cohort; this count is not a population penetrance estimate.
evidence:
- reference: PMID:36074901
reference_title: "ATP6V0C variants impair V-ATPase function causing a neurodevelopmental disorder often associated with epilepsy."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "On the basis of clinical information from 19 patients, the most common seizure types observed were generalized tonic-clonic (12/19), focal (7/19), atonic (6/19) and myoclonic (5/19)."
explanation: >-
Focal seizures in 7 of 19 patients (37%).
- name: Atonic Seizures
category: Neurological
phenotype_term:
preferred_term: Atonic seizure
term:
id: HP:0010819
label: Atonic seizure
evidence:
- reference: PMID:36074901
reference_title: "ATP6V0C variants impair V-ATPase function causing a neurodevelopmental disorder often associated with epilepsy."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "On the basis of clinical information from 19 patients, the most common seizure types observed were generalized tonic-clonic (12/19), focal (7/19), atonic (6/19) and myoclonic (5/19)."
explanation: >-
Atonic seizures in 6 of 19 patients (32%).
description: Reported in 6 of 19 founding-cohort patients with seizure-type information; seizure categories overlap and these counts are not population penetrance estimates.
phenotype_contexts:
- population: 'Mattison 2023 founding cohort: patients with seizure-type data (n=19)'
frequency: 6/19
notes: Selected clinical cohort; this count is not a population penetrance estimate.
evidence:
- reference: PMID:36074901
reference_title: "ATP6V0C variants impair V-ATPase function causing a neurodevelopmental disorder often associated with epilepsy."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "On the basis of clinical information from 19 patients, the most common seizure types observed were generalized tonic-clonic (12/19), focal (7/19), atonic (6/19) and myoclonic (5/19)."
explanation: >-
Atonic seizures in 6 of 19 patients (32%).
- name: Myoclonic Seizures
category: Neurological
phenotype_term:
preferred_term: Myoclonic seizure
term:
id: HP:0032794
label: Myoclonic seizure
evidence:
- reference: PMID:36074901
reference_title: "ATP6V0C variants impair V-ATPase function causing a neurodevelopmental disorder often associated with epilepsy."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "On the basis of clinical information from 19 patients, the most common seizure types observed were generalized tonic-clonic (12/19), focal (7/19), atonic (6/19) and myoclonic (5/19)."
explanation: >-
Myoclonic seizures in 5 of 19 patients (26%).
- reference: PMID:35600075
reference_title: ATP6V0C Is Associated With Febrile Seizures and Epilepsy With Febrile Seizures Plus.
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: Ictal video EEG recordings showed generalized polyspike-slow waves associated with myoclonic seizures
explanation: Myoclonic seizures were documented in the mildly affected family 1 proband; this quote does not assert a medication-specific response.
description: Reported in 5 of 19 founding-cohort patients with seizure-type information; seizure categories overlap and these counts are not population penetrance estimates. They also occurred in the family 1 proband with normal development in the two-family study.
phenotype_contexts:
- population: 'Mattison 2023 founding cohort: patients with seizure-type data (n=19)'
frequency: 5/19
notes: Selected clinical cohort; this count is not a population penetrance estimate.
evidence:
- reference: PMID:36074901
reference_title: "ATP6V0C variants impair V-ATPase function causing a neurodevelopmental disorder often associated with epilepsy."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "On the basis of clinical information from 19 patients, the most common seizure types observed were generalized tonic-clonic (12/19), focal (7/19), atonic (6/19) and myoclonic (5/19)."
explanation: >-
Myoclonic seizures in 5 of 19 patients (26%).
- name: Multifocal EEG Abnormalities
category: Neurological
description: >-
Multifocal EEG abnormalities were reported in the fever-sensitive series; the adult Dravet-like cases had multiple independent spike foci. Initially normal EEG recordings do not exclude the diagnosis.
phenotype_term:
preferred_term: Multifocal epileptiform discharges
term:
id: HP:0010841
label: Multifocal epileptiform discharges
evidence:
- reference: PMID:42526150
reference_title: "Epilepsy with fever-sensitivity in patients with ATP6V0C pathogenic variants."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "The associated phenotype appears characterized by early febrile-triggered motor seizures, initially normal EEG followed by multifocal abnormalities, normal MRI and subsequent onset of neurodevelopmental impairment."
explanation: >-
Describes the evolution from a normal to a multifocally abnormal EEG.
- reference: PMID:40085430
reference_title: Variants in ATP6V0C are associated with Dravet-like developmental and epileptic encephalopathy.
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: The EEG features are also in keeping with DS, with generalized and multifocal-onset seizures.
explanation: The adult report and its table document multifocal epileptiform activity alongside generalized discharges.
- name: Global Developmental Delay
category: Neurological
description: >-
Developmental delay was reported in 21 of 23 founding-cohort patients with available information, but not in the six affected members of the mild familial series. Two adult Dravet-like patients had a plateau around age four without regression.
phenotype_term:
preferred_term: Global developmental delay
term:
id: HP:0001263
label: Global developmental delay
evidence:
- reference: PMID:36074901
reference_title: "ATP6V0C variants impair V-ATPase function causing a neurodevelopmental disorder often associated with epilepsy."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "Development delay was seen in 21/23 patients."
explanation: >-
Developmental delay in 21 of 23 patients (91%).
- reference: PMID:35600075
reference_title: "ATP6V0C Is Associated With Febrile Seizures and Epilepsy With Febrile Seizures Plus."
supports: REFUTE
evidence_source: HUMAN_CLINICAL
snippet: "They all displayed favorable outcomes without intellectual or developmental abnormalities, although afebrile seizures or frequent seizures occurred."
explanation: >-
Developmental delay is not universal: affected members of two
fever-associated families developed normally.
- reference: PMID:40085430
reference_title: Variants in ATP6V0C are associated with Dravet-like developmental and epileptic encephalopathy.
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: Normal early development, followed by development delay and plateau around age 4 years, without regression.
explanation: A developmental plateau, rather than loss of acquired abilities, characterized these two severe cases.
phenotype_contexts:
- population: 'Mattison 2023 founding cohort: patients with developmental information (n=23)'
frequency: 21/23
notes: Selected clinical cohort; this count is not a population penetrance estimate.
evidence:
- reference: PMID:36074901
reference_title: "ATP6V0C variants impair V-ATPase function causing a neurodevelopmental disorder often associated with epilepsy."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "Development delay was seen in 21/23 patients."
explanation: >-
Developmental delay in 21 of 23 patients (91%).
- name: Intellectual Disability
category: Neurological
description: >-
Mild to severe intellectual disability was reported in all 16 founding-cohort patients old enough for assessment and with available information. The mild familial cases had normal development; the two adult Dravet-like cases had severe intellectual disability.
phenotype_term:
preferred_term: Intellectual disability
term:
id: HP:0001249
label: Intellectual disability
evidence:
- reference: PMID:36074901
reference_title: "ATP6V0C variants impair V-ATPase function causing a neurodevelopmental disorder often associated with epilepsy."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "Intellectual disability, ranging from mild to severe, was seen in 16/16 patients who were old enough for a formal diagnosis and for whom this information was available."
explanation: >-
Intellectual disability in all 16 assessable patients.
phenotype_contexts:
- population: 'Mattison 2023 founding cohort: patients old enough for formal diagnosis and with available information (n=16)'
frequency: 16/16
notes: Selected clinical cohort; this count is not a population penetrance estimate.
evidence:
- reference: PMID:36074901
reference_title: "ATP6V0C variants impair V-ATPase function causing a neurodevelopmental disorder often associated with epilepsy."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "Intellectual disability, ranging from mild to severe, was seen in 16/16 patients who were old enough for a formal diagnosis and for whom this information was available."
explanation: >-
Intellectual disability in all 16 assessable patients.
- name: Aplasia or Hypoplasia of the Corpus Callosum
category: Neurological
phenotype_term:
preferred_term: Agenesis or hypoplasia of the corpus callosum
term:
id: HP:0007370
label: Aplasia/Hypoplasia of the corpus callosum
evidence:
- reference: PMID:36074901
reference_title: "ATP6V0C variants impair V-ATPase function causing a neurodevelopmental disorder often associated with epilepsy."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "Five patients with MRIs show hypoplasia or agenesis of the corpus callosum."
explanation: >-
Corpus callosum agenesis or hypoplasia in 5 of 21 imaged patients (24%).
description: Five founding-cohort patients had callosal agenesis or hypoplasia; 21 patients underwent MRI. Normal MRI was reported in both adult Dravet-like cases.
phenotype_contexts:
- population: 'Mattison 2023 founding cohort: patients who underwent brain MRI (n=21)'
frequency: 5/21
notes: Selected clinical cohort; this count is not a population penetrance estimate.
evidence:
- reference: PMID:36074901
reference_title: "ATP6V0C variants impair V-ATPase function causing a neurodevelopmental disorder often associated with epilepsy."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "Five patients with MRIs show hypoplasia or agenesis of the corpus callosum."
explanation: >-
Corpus callosum agenesis or hypoplasia in 5 of 21 imaged patients (24%).
- name: Delayed Myelination
category: Neurological
phenotype_term:
preferred_term: Delayed CNS myelination
term:
id: HP:0002188
label: Delayed CNS myelination
evidence:
- reference: PMID:36074901
reference_title: "ATP6V0C variants impair V-ATPase function causing a neurodevelopmental disorder often associated with epilepsy."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "and delayed myelination (Patients 6, 18, 22 and 24)"
explanation: >-
Delayed myelination in 4 of 21 imaged patients (19%).
description: Delayed myelination was reported in four founding-cohort patients; 21 underwent MRI. This is a variable imaging finding.
phenotype_contexts:
- population: 'Mattison 2023 founding cohort: patients who underwent brain MRI (n=21)'
frequency: 4/21
notes: Selected clinical cohort; this count is not a population penetrance estimate.
evidence:
- reference: PMID:36074901
reference_title: "ATP6V0C variants impair V-ATPase function causing a neurodevelopmental disorder often associated with epilepsy."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "and delayed myelination (Patients 6, 18, 22 and 24)"
explanation: >-
Delayed myelination in 4 of 21 imaged patients (19%).
- name: Aplasia or Hypoplasia of the Cerebellar Vermis
category: Neurological
description: Agenesis or hypoplasia of the cerebellar vermis was among the founding-cohort MRI findings. The combined callosal/vermian narrative does not give a separate vermian denominator.
phenotype_term:
preferred_term: Aplasia/Hypoplasia of the cerebellar vermis
term:
id: HP:0006817
label: Aplasia/Hypoplasia of the cerebellar vermis
evidence:
- reference: PMID:36074901
reference_title: ATP6V0C variants impair V-ATPase function causing a neurodevelopmental disorder often associated with epilepsy.
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: Common findings in MRIs included agenesis/hypoplasia of the corpus callosum (Supplementary Fig. 2) or cerebellar vermis (Patients 5, 6, 18, 20 and 26), and delayed myelination (Patients 6, 18, 22 and 24).
explanation: Supports vermian malformation without assigning the combined patient list entirely to that finding.
- name: Dental Enamel Defects
category: Dental
description: Dental enamel defects were reported in patients 5 and 24 of the founding cohort; patient 24 lacked enamel.
phenotype_term:
preferred_term: Abnormal dental enamel morphology
term:
id: HP:0000682
label: Abnormal dental enamel morphology
evidence:
- reference: PMID:36074901
reference_title: ATP6V0C variants impair V-ATPase function causing a neurodevelopmental disorder often associated with epilepsy.
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: Patients 5 and 24 showed dental enamel defects, with Patient 24 lacking dental enamel.
explanation: Two reported dental findings; the study does not establish a general population frequency.
- name: Pulmonary Valve Stenosis
category: Cardiovascular
description: Reported in founding-cohort patient 3.
phenotype_term:
preferred_term: Valvular pulmonary stenosis
term:
id: HP:0034350
label: Valvular pulmonary stenosis
evidence:
- reference: PMID:36074901
reference_title: ATP6V0C variants impair V-ATPase function causing a neurodevelopmental disorder often associated with epilepsy.
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: 'Four patients were reported to have cardiac abnormalities: Patient 3 had pulmonary valve stenosis, Patient 6 had a thickened left ventricular wall, Patient 7 had a heart murmur and Patient 13 exhibited several cardiac defects including hypertrophic cardiomyopathy, mitral valve prolapse and mild to moderate mitral valve regurgitation.'
explanation: Patient-level cardiac observations rather than a uniform ATP6V0C cardiac syndrome.
- name: Left Ventricular Wall Thickening
category: Cardiovascular
description: Reported in founding-cohort patient 6; this observation alone does not establish primary hypertrophic cardiomyopathy.
phenotype_term:
preferred_term: Left ventricular hypertrophy
term:
id: HP:0001712
label: Left ventricular hypertrophy
evidence:
- reference: PMID:36074901
reference_title: ATP6V0C variants impair V-ATPase function causing a neurodevelopmental disorder often associated with epilepsy.
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: 'Four patients were reported to have cardiac abnormalities: Patient 3 had pulmonary valve stenosis, Patient 6 had a thickened left ventricular wall, Patient 7 had a heart murmur and Patient 13 exhibited several cardiac defects including hypertrophic cardiomyopathy, mitral valve prolapse and mild to moderate mitral valve regurgitation.'
explanation: Patient-level cardiac observations rather than a uniform ATP6V0C cardiac syndrome.
- name: Heart Murmur
category: Cardiovascular
description: Reported in patient 7, who also had a de novo 20q11.22–11.23 deletion. Attribution to ATP6V0C alone is uncertain.
phenotype_term:
preferred_term: Heart murmur
term:
id: HP:0030148
label: Heart murmur
evidence:
- reference: PMID:36074901
reference_title: ATP6V0C variants impair V-ATPase function causing a neurodevelopmental disorder often associated with epilepsy.
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: 'Four patients were reported to have cardiac abnormalities: Patient 3 had pulmonary valve stenosis, Patient 6 had a thickened left ventricular wall, Patient 7 had a heart murmur and Patient 13 exhibited several cardiac defects including hypertrophic cardiomyopathy, mitral valve prolapse and mild to moderate mitral valve regurgitation.'
explanation: Patient-level cardiac observations rather than a uniform ATP6V0C cardiac syndrome.
- name: Status Epilepticus
category: Neurological
description: Both adult Dravet-like cases had episodes of status epilepticus in their seizure histories.
phenotype_term:
preferred_term: Status epilepticus
term:
id: HP:0002133
label: Status epilepticus
evidence:
- reference: PMID:40085430
reference_title: Variants in ATP6V0C are associated with Dravet-like developmental and epileptic encephalopathy.
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: He had several episodes of status epilepticus (often initially associated with fever) requiring induced coma.
explanation: Patient 1 experienced severe prolonged seizures.
- reference: PMID:40085430
reference_title: Variants in ATP6V0C are associated with Dravet-like developmental and epileptic encephalopathy.
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: P2 had clusters of BTC seizures 2–3 days a week, with a frequency of 7–30 seizures per cluster, sometimes leading to status epilepticus (SE).
explanation: Patient 2 also had clusters progressing to status epilepticus.
- name: Atypical Absence Seizures
category: Neurological
description: Reported in the two adult Dravet-like cases, alongside other generalized and focal seizure types.
phenotype_term:
preferred_term: Atypical absence seizure
term:
id: HP:0007270
label: Atypical absence seizure
evidence:
- reference: PMID:40085430
reference_title: Variants in ATP6V0C are associated with Dravet-like developmental and epileptic encephalopathy.
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: Other seizure types recorded over the years included myoclonic, focal motor onset with impaired awareness (later associated with falls), focal tonic, and atypical absence seizures.
explanation: Patient 2 narrative documents atypical absence seizures.
- name: Autism
category: Neurological
description: Both adult Dravet-like cases had autism spectrum disorder and severe intellectual disability.
phenotype_term:
preferred_term: Autism
term:
id: HP:0000717
label: Autism
evidence:
- reference: PMID:40085430
reference_title: Variants in ATP6V0C are associated with Dravet-like developmental and epileptic encephalopathy.
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: At age 31 years, he had autism spectrum disorder and severe ID, speaking only 1- to 2-word sentences.
explanation: Patient 1 adult phenotype.
- reference: PMID:40085430
reference_title: Variants in ATP6V0C are associated with Dravet-like developmental and epileptic encephalopathy.
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: At age 27 years, P2 had autism spectrum disorder (ASD) and severe ID.
explanation: Patient 2 adult phenotype.
- name: Crouch Gait
category: Neurological
description: Crouch gait occurred in the adult Dravet-like cases. In patient 1 it was accompanied by truncal hypotonia and difficulty on stairs.
phenotype_term:
preferred_term: Crouch gait
term:
id: HP:0025682
label: Crouch gait
evidence:
- reference: PMID:40085430
reference_title: Variants in ATP6V0C are associated with Dravet-like developmental and epileptic encephalopathy.
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: Truncal hypotonia was documented at age 15 years in addition to crouch gait with everted feet.
explanation: Longitudinal motor findings in patient 1.
- name: Parkinsonism
category: Neurological
description: Parkinsonism was reported in both adult Dravet-like cases treated with antipsychotics. Its attribution to the disease versus medication remains unresolved.
phenotype_term:
preferred_term: Parkinsonism
term:
id: HP:0001300
label: Parkinsonism
evidence:
- reference: PMID:40085430
reference_title: Variants in ATP6V0C are associated with Dravet-like developmental and epileptic encephalopathy.
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: It is unclear if the parkinsonism observed in both patients is part of the natural history of this condition or if it is a side effect of antipsychotic treatment.
explanation: The authors explicitly identify confounding; this is an observed association, not established ATP6V0C-mediated neurodegeneration.
- name: Delayed Speech and Language Development
category: Neurological
description: Language was among the domains most affected in the adult Dravet-like cases; patient 1 used only one- to two-word sentences at age 31.
phenotype_term:
preferred_term: Delayed speech and language development
term:
id: HP:0000750
label: Delayed speech and language development
evidence:
- reference: PMID:40085430
reference_title: Variants in ATP6V0C are associated with Dravet-like developmental and epileptic encephalopathy.
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: At age 31 years, he had autism spectrum disorder and severe ID, speaking only 1- to 2-word sentences.
explanation: Severely limited adult language in patient 1.
- reference: PMID:40085430
reference_title: Variants in ATP6V0C are associated with Dravet-like developmental and epileptic encephalopathy.
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: P2 also had a slowing in developmental milestones and plateaued at the age of 4 years, with language and fine motor skills being most affected.
explanation: Patient 2 had developmental language impairment without reported regression.
- name: Axial Hypotonia
category: Neurological
description: Truncal hypotonia was documented at age 15 in patient 1 of the adult Dravet-like report.
phenotype_term:
preferred_term: Axial hypotonia
term:
id: HP:0008936
label: Axial hypotonia
evidence:
- reference: PMID:40085430
reference_title: Variants in ATP6V0C are associated with Dravet-like developmental and epileptic encephalopathy.
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: Truncal hypotonia was documented at age 15 years in addition to crouch gait with everted feet.
explanation: Case-series observation; no disease-wide frequency is established.
- name: Ataxia
category: Neurological
description: Ataxia is listed among the motor manifestations in the two-adult Dravet-like report; the narrative does not establish a population frequency or a specific cerebellar mechanism.
phenotype_term:
preferred_term: Ataxia
term:
id: HP:0001251
label: Ataxia
evidence:
- reference: PMID:40085430
reference_title: Variants in ATP6V0C are associated with Dravet-like developmental and epileptic encephalopathy.
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: Motor manifestations with hypotonia during childhood, later occurrence of crouch gait, ataxia, and parkinsonism as an adult.
explanation: Case-series observation; no disease-wide frequency is established.
- name: Aggressive Behavior
category: Neurological
description: Behavioral difficulties including aggression requiring antipsychotic treatment were reported in both adult Dravet-like cases.
phenotype_term:
preferred_term: Aggressive behavior
term:
id: HP:0000718
label: Aggressive behavior
evidence:
- reference: PMID:40085430
reference_title: Variants in ATP6V0C are associated with Dravet-like developmental and epileptic encephalopathy.
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: P2 also had behavioral problems starting from childhood, including aggression requiring antipsychotic drugs.
explanation: Case-series observation; no disease-wide frequency is established.
- name: Infantile Spasms
category: Neurological
description: Infantile spasms were recorded for founding-cohort patients 6 and 7, and infantile flexor spasms for patient 24. Patient 7 also had a 20q11.22–11.23 deletion, limiting single-gene attribution.
phenotype_term:
preferred_term: Infantile spasms
term:
id: HP:0012469
label: Infantile spasms
evidence:
- reference: url:https://pmc.ncbi.nlm.nih.gov/articles/PMC10319782/?report=reader
reference_title: ATP6V0C variants impair V-ATPase function causing a neurodevelopmental disorder often associated with epilepsy - PMC
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: Infantile spasms, GTCS, At, Myo
explanation: Table 1, patient 6, documents infantile spasms together with generalized tonic-clonic, atonic and myoclonic seizures.
- reference: url:https://pmc.ncbi.nlm.nih.gov/articles/PMC10319782/?report=reader
reference_title: ATP6V0C variants impair V-ATPase function causing a neurodevelopmental disorder often associated with epilepsy - PMC
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: Infantile flexor spasms, T (w/asymmetrical limb stiffening)
explanation: Table 1, patient 24, documents infantile flexor spasms and tonic seizures.
- name: Absent Speech
category: Neurological
description: Non-verbal developmental status was recorded in founding-cohort patients 6, 10, 16 and 19; patient 9 was described as regressing to non-verbal status. This differs from the one- to two-word speech retained by an adult in the separate Dravet-like report.
phenotype_term:
preferred_term: Absent speech
term:
id: HP:0001344
label: Absent speech
evidence:
- reference: url:https://pmc.ncbi.nlm.nih.gov/articles/PMC10319782/?report=reader
reference_title: ATP6V0C variants impair V-ATPase function causing a neurodevelopmental disorder often associated with epilepsy - PMC
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: Developmental delay ... p.A95P ... Yes, motor and non-verbal
explanation: 'Table 1 developmental-delay column, patient 10 (p.A95P): non-verbal status. Ellipses omit intervening table rows and columns.'
- name: Developmental Regression
category: Neurological
description: 'Regression was reported in founding-cohort patients 2, 9 and 27, including regression to non-verbal status in patient 9. Patient 14 was described as having regression in adulthood. Regression is not universal: the two separately reported adult Dravet-like cases had developmental plateau without regression.'
phenotype_term:
preferred_term: Developmental regression
term:
id: HP:0002376
label: Developmental regression
evidence:
- reference: url:https://pmc.ncbi.nlm.nih.gov/articles/PMC10319782/?report=reader
reference_title: ATP6V0C variants impair V-ATPase function causing a neurodevelopmental disorder often associated with epilepsy - PMC
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: Developmental delay ... p.A95T ... Yes, regression to non-verbal
explanation: 'Table 1 developmental-delay column, patient 9 (p.A95T): regression to non-verbal status. Ellipses omit intervening table rows and columns.'
- reference: PMID:40085430
reference_title: Variants in ATP6V0C are associated with Dravet-like developmental and epileptic encephalopathy.
supports: REFUTE
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: Normal early development, followed by development delay and plateau around age 4 years, without regression.
explanation: Neither adult Dravet-like case had reported regression; this does not negate regression in other presentations.
genetic:
- name: ATP6V0C
gene_term:
preferred_term: ATP6V0C
term:
id: hgnc:855
label: ATP6V0C
association: Causative
relationship_type: CAUSATIVE
presence: Positive
variant_origin: GERMLINE
inheritance:
- name: Autosomal Dominant
inheritance_term:
preferred_term: Autosomal dominant inheritance
term:
id: HP:0000006
label: Autosomal dominant inheritance
notes: >-
Missense variants predominate in the founding 27-patient cohort (22 patients); four frameshifts and one stop-loss were also reported. Independent cases include an in-frame deletion and a splice-region indel. The founding series includes three previously published patients, including the stop-loss RNA case, so the reports should not be summed as independent cohorts. Multigene deletions support a possible dosage contribution but do not isolate ATP6V0C. Patient 13 also had biallelic LZTR1 variants and hypertrophic cardiomyopathy/mitral valve abnormalities; these cardiac findings cannot be confidently attributed to ATP6V0C.
evidence:
- reference: PMID:36074901
reference_title: "ATP6V0C variants impair V-ATPase function causing a neurodevelopmental disorder often associated with epilepsy."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "In summary, we report 27 patients with heterozygous ATP6V0C variants who presented with developmental delay, early onset epilepsy and intellectual disability."
explanation: >-
Founding cohort for the gene-disease relationship.
- reference: PMID:37161035
reference_title: "ATP6V0C gene variants were identified in individuals with epilepsy, with or without developmental delay."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "We identified de novo heterozygous variants (p.Arg119Trp, p.Val99_Ser102del, c.260_263 + 11delinsGCCCA) in the ATP6V0C gene, which encodes a subunit of vacuolar ATPase."
explanation: >-
Independent replication with missense, in-frame deletion and splice variants.
- reference: PMID:33090716
reference_title: "Haploinsufficiency of ATP6V0C possibly underlies 16p13.3 deletions that cause microcephaly, seizures, and neurodevelopmental disorder."
supports: SUPPORT
evidence_source: COMPUTATIONAL
snippet: "Analysis of loss of function constraint metrics, transcript-aware evaluation of the population variants, GeVIR scores, analysis of reported pathogenic point variants, detailed review of the known functions of gene products and their animal models showed that the haploinsufficiency of ATP6V0C likely underlies the phenotype of this condition."
explanation: >-
In-silico prioritization proposes ATP6V0C as a contributor to the deletion phenotype; the observation does not isolate single-gene causality.
- reference: PMID:36074901
reference_title: ATP6V0C variants impair V-ATPase function causing a neurodevelopmental disorder often associated with epilepsy.
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: Patient 13 has biallelic variants in LZTR1, which has been associated with Noonan-like syndrome
explanation: A second genetic diagnosis limits attribution of this patient's cardiac findings.
differential_diagnoses:
- name: Dravet syndrome
disease_term:
preferred_term: Dravet syndrome
term:
id: MONDO:0100135
label: Dravet syndrome
description: >-
The severe end of ATP6V0C-related epilepsy can meet clinical criteria for
Dravet syndrome. The distinction matters for treatment, because SCN1A-directed
therapies do not apply.
distinguishing_features:
- A pathogenic ATP6V0C variant with no SCN1A variant favors this disorder.
evidence:
- reference: PMID:40085430
reference_title: "Variants in ATP6V0C are associated with Dravet-like developmental and epileptic encephalopathy."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "We propose that abnormal ATP6V0C function can, at the severe end of the clinical spectrum, be associated with Dravet-like phenotype."
explanation: >-
Two adults with clinically classic Dravet syndrome carried de novo ATP6V0C
variants.
- reference: PMID:40085430
reference_title: "Variants in ATP6V0C are associated with Dravet-like developmental and epileptic encephalopathy."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "This is relevant, as these patients would not qualify for disease-modifying antisense nucleotide or gene therapies targeting SCN1A."
explanation: >-
States the treatment consequence of distinguishing the two disorders.
- name: Generalized epilepsy with febrile seizures plus
disease_term:
preferred_term: generalized epilepsy with febrile seizures plus
term:
id: MONDO:0018214
label: generalized epilepsy with febrile seizures plus
description: >-
Familial ATP6V0C variants produce febrile seizures and epilepsy with febrile
seizures plus with normal development, overlapping the other GEFS+ genes.
distinguishing_features:
- Segregating ATP6V0C variants were reported in two FS/EFS+ families with normal development; clinical overlap alone does not identify the causal gene.
evidence:
- reference: PMID:35600075
reference_title: "ATP6V0C Is Associated With Febrile Seizures and Epilepsy With Febrile Seizures Plus."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "This study suggests that ATP6V0C is potentially a candidate pathogenic gene of FS and EFS+."
explanation: >-
Places ATP6V0C in the febrile-seizure and EFS+ differential.
- name: 16p13.3 microdeletion including TBC1D24, ATP6V0C and PDPK1
description: >-
Overlapping 16p13.3 deletions involving ATP6V0C, TBC1D24, PDPK1 and other genes cause a contiguous-gene syndrome. All eight individuals in the original deletion series had developmental delay, intellectual disability and seizures, and six had microcephaly. Chromosomal microarray defines the deletion. ATP6V0C has been proposed as a major contributor, but the multigene context precludes assigning all features to it alone.
distinguishing_features:
- Microcephaly is common in the deletion.
- Detected by chromosomal microarray.
evidence:
- reference: PMID:30245510
reference_title: "A new microdeletion syndrome involving TBC1D24, ATP6V0C, and PDPK1 causes epilepsy, microcephaly, and developmental delay."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "We propose that 16p13.3 microdeletions resulting in simultaneous haploinsufficiencies of TBC1D24, ATP6V0C, and PDPK1 cause a novel rare contiguous gene deletion syndrome of microcephaly, developmental delay, intellectual disability, and epilepsy."
explanation: >-
Defines the contiguous-deletion syndrome that includes ATP6V0C.
- reference: PMID:39273013
reference_title: "V-ATPase Dysfunction in the Brain: Genetic Insights and Therapeutic Opportunities."
supports: SUPPORT
evidence_source: OTHER
quote_role: REVIEW_SYNTHESIS
snippet: "despite the presence of the epilepsy gene TBC1D24 in the genomic deletion, ATP6V0C haploinsufficiency was proposed as the primary contributor to the clinical features"
explanation: >-
Review synthesis describes a proposed contributor, not proof that the other deleted genes have no effect.
animal_models:
- name: Drosophila pan-neuronal Vha16-3 knockdown
species: Fruit fly (Drosophila melanogaster)
genotype: elaV-GAL4 > Vha16-3 (CG32090) RNAi
publication: PMID:36074901
description: >-
Pan-neuronal Vha16-3 RNAi prolonged electroshock recovery in third-instar larvae. Pretreatment with levetiracetam, topiramate, lamotrigine or valproate shortened recovery versus vehicle; phenytoin did not significantly change it at the same 2 mM food concentration. These experiments do not establish comparative clinical efficacy.
modeled_mechanisms:
- target: Neuronal Hyperexcitability and Seizure Susceptibility
relationship: RECAPITULATES
fidelity: MODERATE
model_scale: ORGANISM
description: >-
Reduced neuronal dosage of the orthologue produces increased seizure
susceptibility in vivo.
limitations: >-
Knockdown models haploinsufficiency only, not the proposed dominant-negative missense effect, and larval electroshock recovery is a proxy for human seizures. Drug effects were measured after larval pretreatment, not in patients.
evidence:
- reference: PMID:36074901
reference_title: "ATP6V0C variants impair V-ATPase function causing a neurodevelopmental disorder often associated with epilepsy."
supports: SUPPORT
evidence_source: MODEL_ORGANISM
snippet: "Knockdown of the Drosophila orthologue of ATP6V0C increases seizure duration."
explanation: >-
Neuronal knockdown increases seizure duration.
- reference: PMID:36074901
reference_title: "ATP6V0C variants impair V-ATPase function causing a neurodevelopmental disorder often associated with epilepsy."
supports: SUPPORT
evidence_source: MODEL_ORGANISM
snippet: "Lamotrigine (LAM) and valproate (VAL) were also effective, but not phenytoin (PHY)."
explanation: >-
Drug responses in the knockdown larvae, including the lack of a
phenytoin effect.
- reference: PMID:36074901
reference_title: ATP6V0C variants impair V-ATPase function causing a neurodevelopmental disorder often associated with epilepsy.
supports: SUPPORT
evidence_source: MODEL_ORGANISM
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: Phenytoin, at the same concentration (2 mM in fly food) did not significantly alter recovery time.
explanation: A negative result at this experimental concentration is not a human contraindication.
- name: Caenorhabditis elegans homozygous patient-equivalent knock-ins
species: Nematode (Caenorhabditis elegans)
genotype: 'CRISPR–Cas9 knock-ins: vha-3 G69A (human G63A), vha-2 F143L (human F137L), vha-3 L156F (human L150F); homozygous'
publication: PMID:36074901
description: >-
Homozygous ortholog knock-ins reduced body size and lifespan and increased age-associated paralysis. Day-one movement in physiological M9 was normal; osmotic stress exposed variant-dependent motor abnormalities. A fourth A95T-equivalent homozygous line was sterile and excluded from subsequent assays. Aldicarb hypersensitivity supports altered synaptic signaling without resolving whether acetylcholine or GABA signaling is primarily affected.
modeled_mechanisms:
- target: ATP6V0C Heterozygous Variant
relationship: PARTIALLY_RECAPITULATES
fidelity: LOW
model_scale: ORGANISM
description: >-
Patient-equivalent substitutions impair organismal function in homozygous worm orthologues.
limitations: >-
The human disorder is heterozygous; these are homozygous ortholog knock-ins. Growth, lifespan, motility and aldicarb paralysis are indirect readouts, not measurements of human epilepsy, neuronal transmitter cargo or selective inhibitory release.
evidence:
- reference: PMID:36074901
reference_title: "ATP6V0C variants impair V-ATPase function causing a neurodevelopmental disorder often associated with epilepsy."
supports: SUPPORT
evidence_source: MODEL_ORGANISM
snippet: "the expression of selected patient variants in Caenorhabditis elegans led to reduced growth, motor dysfunction and reduced lifespan"
explanation: >-
In vivo expression of patient variants is deleterious.
- reference: PMID:36074901
reference_title: ATP6V0C variants impair V-ATPase function causing a neurodevelopmental disorder often associated with epilepsy.
supports: SUPPORT
evidence_source: MODEL_ORGANISM
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: Mutations were knocked-in via CRISPR–Cas9, and homozygous worms were studied.
explanation: Methods establish the knock-in design and zygosity.
- reference: PMID:36074901
reference_title: ATP6V0C variants impair V-ATPase function causing a neurodevelopmental disorder often associated with epilepsy.
supports: SUPPORT
evidence_source: MODEL_ORGANISM
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: When tested in liquid physiological M9 over a 4-h period, movement of Day 1 young adult worms with each mutation was comparable to N2 worms
explanation: Baseline movement was preserved despite abnormalities under other conditions.
- reference: PMID:36074901
reference_title: ATP6V0C variants impair V-ATPase function causing a neurodevelopmental disorder often associated with epilepsy.
supports: SUPPORT
evidence_source: MODEL_ORGANISM
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: Worms expressing each patient variant showed greater paralysis in presence of aldicarb, compared to N2 worms
explanation: Aldicarb hypersensitivity is a synaptic-function readout with multiple possible mechanistic explanations.
treatments:
- name: Individualized Antiseizure Pharmacotherapy
description: >-
Treatment evidence is limited to individual cases and small series. One mild familial proband became seizure-free on valproate; another became seizure-free after lamotrigine was added to valproate and nitrazepam. In the adult Dravet-like report, patient 1 improved after switching carbamazepine/phenytoin to valproate, while patient 2 still had monthly seizures on valproate, levetiracetam and lamotrigine and had worsened during a zonisamide trial. These observations do not establish a preferred regimen or general drug contraindications.
therapeutic_modality: SMALL_MOLECULE
treatment_term:
preferred_term: Pharmacotherapy
term:
id: NCIT:C15986
label: Pharmacotherapy
therapeutic_agent:
- preferred_term: valproic acid
term:
id: CHEBI:39867
label: valproic acid
- preferred_term: lamotrigine
term:
id: CHEBI:6367
label: lamotrigine
- preferred_term: levetiracetam
term:
id: CHEBI:6437
label: levetiracetam
- preferred_term: nitrazepam
term:
id: CHEBI:7581
label: nitrazepam
evidence:
- reference: PMID:35600075
reference_title: "ATP6V0C Is Associated With Febrile Seizures and Epilepsy With Febrile Seizures Plus."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "The patient became seizure-free with VPA of 23 mg/kg/day."
explanation: >-
Valproate monotherapy controlled seizures in a familial case.
- reference: PMID:40085430
reference_title: Variants in ATP6V0C are associated with Dravet-like developmental and epileptic encephalopathy.
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: His seizure frequency improved after switching from those two drugs to valproate.
explanation: Patient 1 improved after switching from carbamazepine and phenytoin; this single observation is not a comparative trial.
- reference: PMID:40085430
reference_title: Variants in ATP6V0C are associated with Dravet-like developmental and epileptic encephalopathy.
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: Currently he is taking valproate, levetiracetam, and lamotrigine. A trial of zonisamide led to worsening seizure frequency.
explanation: Patient 2 remained on combination treatment; worsening on zonisamide was an individual observation.
- reference: PMID:35600075
reference_title: ATP6V0C Is Associated With Febrile Seizures and Epilepsy With Febrile Seizures Plus.
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: At the age of 3 years and 8 months, lamotrigine (LTG) was added with a decrease of NZP, and the patient became seizure-free with VPA of 24.2 mg/kg/day, NZP of 0.3 mg/kg/day, and LTG of 1.2 mg/kg/day.
explanation: The case narrative documents seizure freedom during valproate/nitrazepam/lamotrigine combination therapy, not lamotrigine monotherapy.
target_mechanisms:
- target: Seizures
treatment_effect: INHIBITS
description: Symptomatic seizure reduction or seizure freedom was observed in individual patients; responses varied.
evidence:
- reference: PMID:35600075
reference_title: "ATP6V0C Is Associated With Febrile Seizures and Epilepsy With Febrile Seizures Plus."
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
snippet: "The patient became seizure-free with VPA of 23 mg/kg/day."
explanation: >-
Valproate monotherapy controlled seizures in a familial case.
has_subtypes:
- name: Familial febrile seizures and epilepsy with febrile seizures plus
description: 'A reported mild presentation, rather than a separate molecular subtype: six affected members of two families had infantile febrile seizures and normal development; the two probands also developed afebrile seizures.'
evidence:
- reference: PMID:35600075
reference_title: ATP6V0C Is Associated With Febrile Seizures and Epilepsy With Febrile Seizures Plus.
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: They all displayed favorable outcomes without intellectual or developmental abnormalities, although afebrile seizures or frequent seizures occurred.
explanation: Normal development in the two reported families contrasts with the neurodevelopmental presentations.
- name: Neurodevelopmental disorder with or without epilepsy
description: The founding 27-patient series included developmental delay and intellectual disability, variable MRI findings and frequent early-onset epilepsy. Some individuals, including mosaic carriers, had no reported seizures. These are clinical presentations within a spectrum, not established allele-specific categories.
evidence:
- reference: PMID:36074901
reference_title: ATP6V0C variants impair V-ATPase function causing a neurodevelopmental disorder often associated with epilepsy.
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: We describe heterozygous point variants in ATP6V0C, encoding the c-subunit in the membrane bound integral domain of the vacuolar H+-ATPase, in 27 patients with neurodevelopmental abnormalities with or without epilepsy.
explanation: Defines the founding neurodevelopmental spectrum.
- name: Dravet-like developmental and epileptic encephalopathy
description: Two adult men had infantile febrile seizures followed by multiple afebrile seizure types and status epilepticus, severe intellectual disability, autism and motor abnormalities. Development plateaued around age four without reported regression.
evidence:
- reference: PMID:40085430
reference_title: Variants in ATP6V0C are associated with Dravet-like developmental and epileptic encephalopathy.
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: The identification of P/LP ATP6V0C variants in two unrelated adult patients with typical DS-like phenotype expands the clinical spectrum of ATP6V0C-associated epilepsy to include DS-like DEE.
explanation: Two adult cases establish a severe presentation without implying that all ATP6V0C-related epilepsy is Dravet-like.
- reference: PMID:40085430
reference_title: Variants in ATP6V0C are associated with Dravet-like developmental and epileptic encephalopathy.
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: Normal early development, followed by development delay and plateau around age 4 years, without regression.
explanation: The longitudinal report distinguishes plateau from regression.
diagnosis:
- name: Molecular diagnosis and segregation analysis
description: Trio exome sequencing identified the causal or likely causal ATP6V0C variants in the two adult Dravet-like cases; chromosomal microarray excluded copy-number alternatives. Familial presentations were investigated by exome sequencing and segregation testing. Interpretation integrates variant evidence and phenotype rather than diagnosing from fever sensitivity or a VUS alone.
evidence:
- reference: PMID:40085430
reference_title: Variants in ATP6V0C are associated with Dravet-like developmental and epileptic encephalopathy.
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: The two patients reported below had their ATP6V0C variants identified through trio exome sequencing
explanation: Documents the molecular diagnostic route in the adult cases.
- reference: PMID:35600075
reference_title: ATP6V0C Is Associated With Febrile Seizures and Epilepsy With Febrile Seizures Plus.
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: Whole blood samples were collected from the probands, their parents, and available family members to ascertain whether the genetic variants were co-segregation.
explanation: Family studies establish segregation of candidate variants.
experimental_models:
- name: Yeast VMA3 null-complementation assay
experimental_model_type: OTHER
description: Twelve patient-equivalent missense variants were introduced into yeast VMA3 and expressed in vma3-null cells. Nine gave little or no LysoSensor fluorescence, G63A and L150F retained intermediate fluorescence, and F137L was comparable to wild type in that assay. Calcium-sensitive growth provided an additional functional readout, including impairment of F137L at higher calcium concentrations.
publication: PMID:36074901
modeled_mechanisms:
- target: Reduced V-ATPase Proton Pumping
description: Ortholog complementation probes residual V-ATPase function through vacuolar acidification and calcium-sensitive growth.
evidence:
- reference: PMID:36074901
reference_title: ATP6V0C variants impair V-ATPase function causing a neurodevelopmental disorder often associated with epilepsy.
supports: SUPPORT
evidence_source: IN_VITRO
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: When the uptake of LysoSensor was measured, we saw that nine variants were associated with little to no fluorescence, indicating significant reduction or loss of V-ATPase activity.
explanation: Variant-dependent acidification impairment in the yeast assay.
- reference: PMID:36074901
reference_title: ATP6V0C variants impair V-ATPase function causing a neurodevelopmental disorder often associated with epilepsy.
supports: SUPPORT
evidence_source: IN_VITRO
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: One patient variant (p.F137L) showed levels of fluorescence intensity that were comparable to wild-type rescue.
explanation: The fluorescence readout was not abnormal for every patient-equivalent variant.
relationship: PARTIALLY_RECAPITULATES
model_scale: CELLULAR
limitations: Yeast vacuoles are not human synaptic vesicles. Null complementation does not reproduce heterozygous wild-type/mutant coexpression or establish dominant-negative action; LysoSensor is an indirect readout of pump function.
fidelity: MODERATE
references:
- reference: PMID:36074901
title: "ATP6V0C variants impair V-ATPase function causing a neurodevelopmental disorder often associated with epilepsy."
- reference: PMID:35600075
title: "ATP6V0C Is Associated With Febrile Seizures and Epilepsy With Febrile Seizures Plus."
- reference: PMID:29652249
title: "Clathrin coat controls synaptic vesicle acidification by blocking vacuolar ATPase activity."
- reference: PMID:30155790
title: "Chromophore-Assisted Light Inactivation of the V-ATPase V0c Subunit Inhibits Neurotransmitter Release Downstream of Synaptic Vesicle Acidification."
- reference: PMID:33190975
title: "Novel de novo mutation substantiates ATP6V0C as a gene causing epilepsy with intellectual disability."
- reference: PMID:37161035
title: "ATP6V0C gene variants were identified in individuals with epilepsy, with or without developmental delay."
- reference: PMID:42526150
title: "Epilepsy with fever-sensitivity in patients with ATP6V0C pathogenic variants."
- reference: PMID:40085430
title: Variants in ATP6V0C are associated with Dravet-like developmental and epileptic encephalopathy.
- reference: PMID:33090716
title: "Haploinsufficiency of ATP6V0C possibly underlies 16p13.3 deletions that cause microcephaly, seizures, and neurodevelopmental disorder."
- reference: PMID:30245510
title: "A new microdeletion syndrome involving TBC1D24, ATP6V0C, and PDPK1 causes epilepsy, microcephaly, and developmental delay."
- reference: PMID:39273013
title: "V-ATPase Dysfunction in the Brain: Genetic Insights and Therapeutic Opportunities."
- reference: url:https://pmc.ncbi.nlm.nih.gov/articles/PMC10319782/?report=reader
title: ATP6V0C variants impair V-ATPase function causing a neurodevelopmental disorder often associated with epilepsy - PMC
discussions:
- discussion_id: atp6v0c_cellular_routes
prompt: Which cellular defects connect ATP6V0C dysfunction to neurodevelopmental impairment and epilepsy?
kind: KNOWLEDGE_GAP
status: OPEN
attaches_to:
- pathophysiology#Reduced V-ATPase Proton Pumping
rationale: The founding study proposes combined effects on synaptic signaling, trans-Golgi trafficking and lysosomal/autophagic function. Evidence from other V-ATPase genes motivates these possibilities but does not establish the corresponding pathway defects in ATP6V0C patient neurons. Worm aldicarb sensitivity cannot by itself distinguish GABA from acetylcholine dysfunction. Direct cellular measurements would be needed to resolve the relative contributions.
evidence:
- reference: PMID:36074901
reference_title: ATP6V0C variants impair V-ATPase function causing a neurodevelopmental disorder often associated with epilepsy.
supports: SUPPORT
evidence_source: OTHER
quote_role: BACKGROUND
directness: INDIRECT
snippet: We hypothesize that the epilepsy and other neurodevelopmental phenotypes seen in patients with pathogenic V-ATPase variants may be due to a combinatorial effect of impaired synaptic signalling, trafficking and sorting of various membrane bound proteins, and defects along the endomembrane system including the lysosomal/autophagy degradation pathway.
explanation: The authors explicitly frame these cellular routes as a hypothesis.
prevalence:
- population: 'ATP6V0C-related neurodevelopmental disorder: ascertainment context of the Mattison 2023 report'
measure_type: UNKNOWN
notes: The founding report raised under-ascertainment because ATP6V0C was absent from commercial epilepsy/intellectual-disability panels at that time. This is a historical testing caveat, not a claim about current panel content or a quantitative prevalence estimate.
evidence:
- reference: PMID:36074901
reference_title: ATP6V0C variants impair V-ATPase function causing a neurodevelopmental disorder often associated with epilepsy.
supports: SUPPORT
evidence_source: HUMAN_CLINICAL
quote_role: PRIMARY_RESULT
directness: DIRECT
snippet: The prevalence of neurodevelopmental disorders, including epilepsy, resulting from variants in ATP6V0C is probably underestimated as this gene, to the best of our knowledge, is not currently included on commercially available epilepsy or intellectual disability gene panels.
explanation: Study-era ascertainment limitation; no numerical population prevalence follows from it.
References & Deep Research
References
12Deep Research
1Deep research results are used as seeds for research; they do not undergo the same validation as the main records and may contain errors. How we use deep research.
Evaluations and curation notes (1)
Create: ATP6V0C-Related Epilepsy · 2026-10-01T05:41:53Z · View source
New entry for MONDO:0958196 (early-onset epilepsy 3, ATP6V0C). Built from the primary literature: the 27-patient founding cohort (PMID:36074901, full text), the familial FS/EFS+ report (PMID:35600075, full text), independent case series (PMID:37161035, PMID:33190975), the Dravet-like severe end (PMID:40085430), fever-sensitive epilepsy (PMID:42526150), the 16p13.3 deletion papers (PMID:30245510, PMID:33090716), and V-ATPase synaptic biology (PMID:29652249, PMID:30155790, PMID:20839327). Falcon deep research (research/ATP6V0C-Related_Epilepsy-deep-research-falcon.md) agreed with this source set; its reference block set needs_review because of one ellipsis-shortened quote of the PMID:36074901 abstract (the entry quotes the full sentence) and a template placeholder in place of the MONDO label, and its one additional source, the 2024 V-ATPase review (PMID:39273013), is cited for one synthesis claim. Steps downstream of reduced pump activity (vesicle loading, release, excitability) are not measured for patient alleles and are carried in two EMERGING hypothesis groups. Developmental delay and intellectual disability are deliberately not wired into the pathograph. just preflight-dr was not run: it requires a local mondo.db, which this environment does not have. just check-genereviews --online raised a ValueError in bookshelf.py rank() on this entry; the offline check reports NO_CHAPTER for GeneReviews and StatPearls, matching a PubMed genereviews[book] search. A fresh-context review pass found missing reference titles, an over-specific congenital heart binding, an unsourced mosaicism claim and treatment wording; all were fixed before commit. Validation: just validate, validate-terms, count-verified-snippets (50/50) and the offline gates pass.
Question: You are an expert researcher providing comprehensive, well-cited information.
Provide detailed information focusing on: 1. Key concepts and definitions with current understanding 2. Recent developments and latest research (prioritize 2023-2024 sources) 3. Current applications and real-world implementations 4. Expert opinions and analysis from authoritative sources 5. Relevant statistics and data from recent studies
Format as a comprehensive research report with proper citations. Include URLs and publication dates where available. Always prioritize recent, authoritative sources and provide specific citations for all major claims.
Disease Characteristics Research Template
Target Disease
- Disease Name: ATP6V0C-Related Epilepsy
- MONDO ID: MONDO:0958196 (if available)
- Category: Mendelian
Research Objectives
Please provide a comprehensive research report on ATP6V0C-Related Epilepsy covering all of the disease characteristics listed below. This report will be used to populate a disease knowledge base entry. Be thorough and cite primary literature (PMID preferred) for all claims.
For each section, suggested databases/resources are listed. These are the first places you should search for information on each topic.
1. Disease Information
Search first: OMIM, Orphanet, ICD-10/ICD-11, MeSH, PubMed
- What is the disease? Provide a concise overview.
- What are the key identifiers? (OMIM, Orphanet, ICD-10/ICD-11, MeSH, Mondo)
- What are the common synonyms and alternative names?
- Is the information derived from individual patients (e.g., EHR) or aggregated disease-level resources?
2. Etiology
- Disease Causal Factors: What are the primary causes? (genetic, environmental, infectious, mechanistic)
- Risk Factors:
Search first: PubMed, Cochrane Library, UpToDate, clinical guidelines, ClinVar, ClinGen, GWAS Catalog, PheGenI, CTD, CDC, WHO, epidemiological databases
- Genetic risk factors (causal variants, susceptibility loci, modifier genes)
- Environmental risk factors (toxins, lifestyle, occupational exposures, age, sex, family history)
- Protective Factors:
Search first: PubMed, Cochrane Library, clinical trial databases, GWAS Catalog, gnomAD, WHO, CDC, nutrition databases
- Genetic protective factors (protective variants, modifier alleles)
- Environmental protective factors (diet, lifestyle, exposures that reduce risk)
- Gene-Environment Interactions: How do genetic and environmental factors interact to influence disease?
Search first: CTD, PubMed, PheGenI, GxE databases
3. Phenotypes
Search first: HPO (Human Phenotype Ontology), OMIM, Orphanet, PubMed, clinicaltrials.gov, MedDRA, SNOMED CT, DECIPHER, LOINC
For each phenotype, provide: - Phenotype type: symptoms, clinical signs, physical manifestations, behavioral changes, or laboratory abnormalities
For symptoms/signs: HPO, OMIM, Orphanet, PubMed For behavioral changes: HPO, DSM, RDoC (Research Domain Criteria), PubMed For laboratory abnormalities: LOINC, SNOMED CT, LabTests Online, PubMed - Phenotype characteristics: Search first: OMIM, Orphanet, HPO, PubMed - Age of symptom onset (neonatal, childhood, adult-onset, late-onset) - Symptom severity (mild, moderate, severe, variable) - Symptom progression (stable, progressive, episodic, fluctuating) - Frequency among affected individuals (percentage or qualitative) - Quality of life impact: Effects on daily functioning and well-being (per-phenotype when possible) Search first: EQ-5D database, SF-36, WHO QOL databases, PubMed - Suggest HPO (Human Phenotype Ontology) terms for each phenotype
4. Genetic/Molecular Information
- Causal Genes: Gene mutations or chromosomal abnormalities responsible for disease (gene symbols, OMIM IDs)
Search first: OMIM, ClinVar, HGMD, Ensembl, NCBI Gene
- Pathogenic Variants:
- Affected genes (gene symbols, HGNC IDs) > Search first: OMIM, NCBI Gene, Ensembl, HGNC, UniProt, GeneCards
- Variant classification (pathogenic, likely pathogenic, VUS per ACMG/AMP guidelines) > Search first: ClinVar, ClinGen, ACMG/AMP guidelines, VarSome
- Variant type/class (missense, frameshift, nonsense, splice-site, structural)
- Allele frequency in population databases > Search first: gnomAD, 1000 Genomes, ExAC, TOPMed, dbSNP
- Somatic vs germline origin > Search first: COSMIC (somatic), ClinVar, ICGC, TCGA
- Functional consequences (loss of function, gain of function, dominant negative)
- Modifier Genes: Genes that modify disease severity or expression
- Epigenetic Information: DNA methylation, histone modifications, chromatin changes affecting disease
Search first: ENCODE, Roadmap Epigenomics, MethBase, DiseaseMeth
- Chromosomal Abnormalities: Large-scale genetic changes (aneuploidy, translocations, inversions)
Search first: DECIPHER, ClinVar, ECARUCA, UCSC Genome Browser
5. Environmental Information
- Environmental Factors: Non-genetic contributing factors (toxins, radiation, pollution, occupational exposure)
Search first: CTD (Comparative Toxicogenomics Database), TOXNET, PubMed, EPA databases
- Lifestyle Factors: Behavioral factors (smoking, diet, exercise, alcohol consumption)
Search first: CDC databases, WHO, PubMed, NHANES
- Infectious Agents: If applicable, pathogens causing or triggering disease (bacteria, viruses, fungi, parasites)
Search first: NCBI Taxonomy, ViPR, BV-BRC, MicrobeDB, GIDEON
6. Mechanism / Pathophysiology
Present this section as an ordered causal chain first, then the detail below. Open with a numbered sequence of mechanistic steps running from the initiating lesion (mutation, exposure, infection) to the clinical manifestation, one step per line, each naming what it causes next. State the causal verb explicitly ("leads to", "results in") and say where a step is inferred rather than demonstrated. Where the mechanism branches, show the branch. The categories below are a checklist of what to cover within those steps, not the organizing structure — a step may draw on several of them, and a category may contribute to several steps.
- Molecular Pathways: Specific signaling cascades or biochemical pathways involved (Wnt, MAPK, mTOR, PI3K-AKT, etc.)
Search first: KEGG, Reactome, WikiPathways, PathBank, BioCyc
- Cellular Processes: Cell-level mechanisms (apoptosis, autophagy, cell cycle dysregulation, inflammation, etc.)
Search first: Gene Ontology (GO), Reactome, KEGG, PubMed
- Protein Dysfunction: How protein structure or function is altered (misfolding, aggregation, loss of function, gain of function)
Search first: UniProt, PDB (Protein Data Bank), InterPro, Pfam, AlphaFold
- Metabolic Changes: Alterations in metabolic processes (energy metabolism, lipid metabolism, amino acid metabolism)
Search first: KEGG, BioCyc, HMDB (Human Metabolome Database), BRENDA
- Immune System Involvement: Role of immune response (autoimmunity, immunodeficiency, chronic inflammation)
Search first: ImmPort, Immunome Database, IEDB, Gene Ontology
- Tissue Damage Mechanisms: How tissues/ are injured (oxidative stress, ischemia, fibrosis, necrosis)
Search first: PubMed, Gene Ontology, Reactome
- Biochemical Abnormalities: Specific molecular defects (enzyme deficiencies, receptor dysfunction, ion channel defects)
Search first: BRENDA, UniProt, KEGG, OMIM, PubMed
- Epigenetic Changes: DNA methylation, histone modifications affecting gene expression in disease
Search first: ENCODE, Roadmap Epigenomics, MethBase, DiseaseMeth
- Molecular Profiling (if available):
- Transcriptomics/gene expression changes > Search first: GEO (Gene Expression Omnibus), ArrayExpress, GTEx, Human Cell Atlas, SRA
- Proteomics findings > Search first: PRIDE, ProteomeXchange, Human Protein Atlas, STRING, BioGRID
- Metabolomics signatures > Search first: MetaboLights, Metabolomics Workbench, HMDB, METLIN
- Lipidomics alterations > Search first: LIPID MAPS, SwissLipids, LipidHome, Metabolomics Workbench
- Genomic structural features > Search first: UCSC Genome Browser, Ensembl, NCBI, dbVar, DGV
- Advanced Technologies (if applicable):
- Single-cell analysis findings (cell-type specific mechanisms, cellular heterogeneity) > Search first: Human Cell Atlas, Single Cell Portal, GEO, CELLxGENE
- Spatial transcriptomics findings > Search first: GEO, Spatial Research, Vizgen, 10x Genomics data
- Multi-omics integration results > Search first: TCGA, ICGC, cBioPortal, LinkedOmics, PubMed
- Functional genomics screens (CRISPR, RNAi) > Search first: DepMap, GenomeRNAi, PubMed, BioGRID ORCS
For each mechanism, describe: - The causal chain from initial trigger to clinical manifestation - Which mechanisms are upstream vs downstream - What cell types and biological processes are involved - Suggest GO terms for biological processes and CL terms for cell types
7. Anatomical Structures Affected
- Organ Level:
- Primary organs directly affected
- Secondary organ involvement (complications, secondary effects)
- Body systems involved (cardiovascular, nervous, digestive, respiratory, endocrine, etc.)
Search first: Uberon, FMA (Foundational Model of Anatomy), OMIM, HPO, ICD-11, MeSH, SNOMED CT
- Tissue and Cell Level:
- Specific tissue types affected (epithelial, connective, muscle, nervous)
- Specific cell populations targeted (with Cell Ontology terms)
Search first: Uberon, Human Protein Atlas, Cell Ontology, Human Cell Atlas, CellMarker, PanglaoDB
- Subcellular Level:
- Cellular compartments involved (mitochondria, nucleus, ER, lysosomes) (with GO Cellular Component terms)
Search first: Gene Ontology (Cellular Component), UniProt, Human Protein Atlas
- Localization:
- Specific anatomical sites (with UBERON terms) > Search first: FMA, Uberon, NeuroNames (for brain), SNOMED CT
- Lateralization (unilateral, bilateral, asymmetric) > Search first: HPO, clinical literature, imaging databases
8. Temporal Development
- Onset:
- Typical age of onset (congenital, pediatric, adult, geriatric)
- Onset pattern (acute, subacute, chronic, insidious)
Search first: OMIM, Orphanet, HPO, PubMed
- Progression:
- Disease stages (early, intermediate, advanced, end-stage) > Search first: Cancer Staging Manual (AJCC), WHO classifications, PubMed
- Progression rate (rapid, slow, variable)
- Disease course pattern (episodic, relapsing-remitting, progressive, stable)
- Disease duration (self-limited, chronic lifelong)
Search first: Disease registries, longitudinal cohort databases, natural history studies, PubMed, Orphanet, OMIM
- Patterns:
- Remission patterns (spontaneous, treatment-induced) > Search first: Clinical trial databases, disease registries, PubMed
- Critical periods (time windows of vulnerability or opportunity for intervention) > Search first: PubMed, developmental biology databases, clinical guidelines
9. Inheritance and Population
- Epidemiology:
- Prevalence (cases per 100,000 at given time)
- Incidence (new cases per 100,000 per year)
Search first: Orphanet, CDC, WHO, GBD (Global Burden of Disease), national registries, SEER, disease registries
- For Genetic Etiology:
- Inheritance pattern (AD, AR, X-linked, mitochondrial, multifactorial, polygenic) > Search first: OMIM, Orphanet, ClinVar, GTR (Genetic Testing Registry)
- Penetrance (complete, incomplete, age-dependent) > Search first: ClinVar, OMIM, PubMed, ClinGen
- Expressivity (variable, consistent) > Search first: OMIM, ClinVar, PubMed
- Genetic anticipation (increasing severity in successive generations) > Search first: OMIM, PubMed (especially for repeat expansion disorders)
- Germline mosaicism > Search first: ClinVar, OMIM, genetic counseling literature, PubMed
- Founder effects (population-specific mutations) > Search first: gnomAD, population genetics databases, PubMed
- Consanguinity role > Search first: OMIM, population studies, genetic counseling resources
- Carrier frequency > Search first: gnomAD, carrier screening databases, GeneReviews, GTR
- Population Demographics:
- Affected populations (ethnic or demographic groups with higher prevalence) > Search first: gnomAD, 1000 Genomes, PAGE Study, PubMed, population registries
- Geographic distribution (endemic areas, regional variation) > Search first: WHO, CDC, GBD, Orphanet, geographic epidemiology databases
- Geographic distribution of specific variants
- Sex ratio (male:female) > Search first: Disease registries, OMIM, PubMed, epidemiological databases
- Age distribution of affected individuals > Search first: CDC, disease registries, SEER, Orphanet
10. Diagnostics
- Clinical Tests:
- Laboratory tests (blood, urine, tissue chemistry, specific enzyme assays) > Search first: LOINC, LabTests Online, PubMed
- Biomarkers (proteins, metabolites, genetic markers, circulating biomarkers) > Search first: FDA Biomarker List, BEST (Biomarkers, EndpointS, and other Tools), PubMed
- Imaging studies (X-ray, CT, MRI, PET, ultrasound) > Search first: RadLex, DICOM, Radiopaedia, imaging databases
- Functional tests (pulmonary function, cardiac stress tests) > Search first: LOINC, clinical guidelines, PubMed
- Electrophysiology (EEG, EMG, ECG, nerve conduction studies) > Search first: LOINC, clinical neurophysiology databases, PubMed
- Biopsy findings (histopathology, immunohistochemistry) > Search first: SNOMED CT, College of American Pathologists resources, PubMed
- Pathology findings (microscopic examination) > Search first: SNOMED CT, Digital Pathology databases, PubMed
- Genetic Testing:
Search first: GTR (Genetic Testing Registry), GeneReviews, ClinGen
- Overview of recommended genetic testing approach
- Whole genome sequencing (WGS) utility > Search first: GTR, ClinVar, GEL (Genomics England), gnomAD
- Whole exome sequencing (WES) utility > Search first: GTR, ClinVar, OMIM, GeneMatcher
- Gene panels (which panels, which genes) > Search first: GTR, ClinVar, laboratory-specific databases
- Single gene testing > Search first: GTR, ClinVar, OMIM, GeneReviews
- Chromosomal microarray (CMA) > Search first: DECIPHER, ClinVar, dbVar, ECARUCA
- Karyotyping > Search first: Chromosome Abnormality Database, ClinVar, cytogenetics resources
- FISH > Search first: ClinVar, cytogenetics databases, PubMed
- Mitochondrial DNA testing > Search first: MITOMAP, MSeqDR, ClinVar, GTR
- Repeat expansion testing > Search first: GTR, ClinVar, repeat expansion databases, PubMed
- Omics-Based Diagnostics (if applicable):
- RNA sequencing / transcriptomics > Search first: GEO, ArrayExpress, GTEx, RNA-seq databases
- Proteomics > Search first: PRIDE, ProteomeXchange, FDA Biomarker database
- Metabolomics > Search first: MetaboLights, Metabolomics Workbench, HMDB
- Epigenomics > Search first: GEO, ENCODE, Roadmap Epigenomics, MethBase
- Liquid biopsy > Search first: COSMIC, ClinVar, liquid biopsy databases, PubMed
- Clinical Criteria:
- Standardized diagnostic criteria (DSM, ICD, society guidelines) > Search first: DSM-5, ICD-11, clinical society guidelines, UpToDate
- Differential diagnosis (other conditions to rule out, with distinguishing features) > Search first: DynaMed, UpToDate, clinical decision support systems
- Screening:
- Screening methods for asymptomatic individuals (newborn screening, carrier screening, cascade screening) > Search first: ACMG recommendations, CDC newborn screening, GTR
11. Outcome/Prognosis
- Survival and Mortality:
- Survival rate (5-year, 10-year, overall) > Search first: SEER, cancer registries, disease-specific registries, PubMed
- Life expectancy (with and without treatment if applicable) > Search first: Orphanet, disease registries, actuarial databases, PubMed
- Mortality rate > Search first: CDC, WHO, GBD, national mortality databases
- Disease-specific mortality (deaths directly attributable to disease) > Search first: Disease registries, CDC Wonder, GBD, PubMed
- Morbidity and Function:
- Morbidity (disease-related disability and health impacts) > Search first: GBD, WHO, disability databases, PubMed
- Disability outcomes (long-term functional impairments) > Search first: ICF (International Classification of Functioning), disability registries
- Quality of life measures (EQ-5D, SF-36, PROMIS, disease-specific tools) > Search first: EQ-5D database, SF-36, PROMIS, PubMed
- Disease Course:
- Complications (secondary problems: infections, organ failure, etc.) > Search first: ICD codes, disease registries, clinical databases, PubMed
- Recovery potential (likelihood and extent of recovery, with vs without treatment) > Search first: Natural history studies, rehabilitation databases, PubMed
- Prediction:
- Prognostic factors (age, disease severity, biomarkers, treatment response) > Search first: Prognostic models databases, clinical calculators, PubMed
- Prognostic biomarkers (molecular markers predicting disease course) > Search first: FDA Biomarker database, PubMed, cancer prognostic databases
12. Treatment
- Pharmacotherapy:
- Pharmacological treatments (drug names, drug classes, mechanisms of action) > Search first: DrugBank, RxNorm, ATC classification, DailyMed, FDA databases
- Pharmacogenomics (how genetic variants affect drug metabolism, efficacy, toxicity) > Search first: PharmGKB, CPIC (Clinical Pharmacogenetics), FDA Table of PGx Biomarkers
- Advanced Therapeutics:
- Gene therapy (viral vectors, CRISPR, gene replacement, gene editing) > Search first: ClinicalTrials.gov, FDA gene therapy database, ASGCT resources
- Cell therapy (stem cell transplant, CAR-T, cellular therapeutics) > Search first: ClinicalTrials.gov, FDA cell therapy database, FACT standards
- RNA-based therapies (ASOs, siRNA, mRNA therapies) > Search first: ClinicalTrials.gov, FDA approvals, PubMed
- Targeted therapies (treatments directed at specific molecular targets) > Search first: My Cancer Genome, OncoKB, ClinicalTrials.gov, FDA approvals
- Immunotherapies (checkpoint inhibitors, monoclonal antibodies) > Search first: Cancer Immunotherapy Database, FDA approvals, ClinicalTrials.gov
- Surgical and Interventional:
- Surgical interventions (types of surgery, timing, outcomes) > Search first: CPT codes, surgical registries, clinical guidelines, PubMed
- Supportive and Rehabilitative:
- Supportive care (symptom management, pain control, nutrition) > Search first: Clinical guidelines, Cochrane Library, PubMed
- Rehabilitation (physical therapy, occupational therapy, speech therapy) > Search first: Rehabilitation medicine databases, clinical guidelines, PubMed
- Experimental:
- Experimental treatments in clinical trials (with NCT identifiers if available) > Search first: ClinicalTrials.gov, EU Clinical Trials Register, WHO ICTRP
- Treatment Outcomes:
- Treatment response rates > Search first: Clinical trial databases, FDA reviews, systematic reviews, PubMed
- Side effects and adverse events > Search first: FDA Adverse Event Reporting System (FAERS), MedWatch, PubMed
- Treatment Strategy:
- Treatment algorithms (clinical pathways, decision trees) > Search first: Clinical practice guidelines, NCCN Guidelines, UpToDate
- Combination therapies > Search first: ClinicalTrials.gov, treatment guidelines, PubMed
- Personalized medicine approaches (genotype-guided treatment) > Search first: My Cancer Genome, CIViC, PharmGKB, precision medicine databases
For each treatment, suggest NCIT (NCI Thesaurus) clinical-intervention terms where applicable.
13. Prevention
- Prevention Levels:
- Primary prevention (preventing disease occurrence: vaccination, risk factor modification) > Search first: CDC, WHO, USPSTF recommendations, Cochrane Library
- Secondary prevention (early detection and treatment: screening programs, early intervention) > Search first: USPSTF, CDC screening guidelines, WHO
- Tertiary prevention (preventing complications in those with disease) > Search first: Clinical guidelines, disease management protocols, PubMed
- Immunization: Vaccine strategies (if applicable)
Search first: CDC vaccine schedules, WHO immunization, FDA vaccine database
- Screening and Early Detection:
- Screening programs (population-based: newborn screening, cancer screening) > Search first: CDC screening programs, USPSTF, cancer screening databases
- Genetic screening (carrier screening, preimplantation genetic diagnosis, prenatal testing) > Search first: ACMG recommendations, ACOG guidelines, GTR
- Risk stratification (identifying high-risk individuals for targeted prevention) > Search first: Risk prediction models, clinical calculators, PubMed
- Behavioral Interventions: Lifestyle modifications to reduce risk
Search first: CDC, WHO, behavioral intervention databases, Cochrane Library
- Counseling: Genetic counseling (risk assessment, family planning guidance)
Search first: NSGC resources, ACMG guidelines, GeneReviews
- Public Health:
- Public health interventions (sanitation, vector control, health education) > Search first: CDC, WHO, public health databases, PubMed
- Environmental interventions (reducing environmental risk factors) > Search first: EPA databases, WHO environmental health, PubMed
- Prophylaxis: Preventive medications or procedures
Search first: Clinical guidelines, FDA approvals, PubMed
14. Other Species / Natural Disease
- Taxonomy: Species affected (with NCBI Taxon identifiers)
Search first: NCBI Taxonomy
- Breed: Specific breeds affected (with VBO identifiers if applicable)
Search first: VBO (Vertebrate Breed Ontology)
- Gene: Orthologous genes in other species (with NCBI Gene IDs)
Search first: NCBI Gene
- Natural Disease:
- Naturally occurring disease in other species (companion animals, wildlife) > Search first: OMIA (Online Mendelian Inheritance in Animals), VetCompass, PubMed
- Veterinary relevance and importance in animal health > Search first: OMIA, veterinary databases, PubMed
- Comparative Biology:
- Comparative pathology (similarities and differences across species) > Search first: OMIA, comparative pathology databases, PubMed
- Evolutionary conservation of disease mechanisms > Search first: HomoloGene, OrthoMCL, Alliance of Genome Resources
- Transmission (if applicable):
- Zoonotic potential > Search first: CDC zoonotic diseases, WHO zoonoses, GIDEON
- Cross-species susceptibility > Search first: NCBI Taxonomy, veterinary databases, PubMed
15. Model Organisms
- Model Types:
- Model organism type (mammalian, invertebrate, cellular, in vitro) > Search first: Alliance of Genome Resources, model organism databases
- Specific model systems (mouse, rat, zebrafish, Drosophila, C. elegans, yeast, cell lines, organoids, iPSCs) > Search first: MGI, RGD, ZFIN, FlyBase, WormBase, SGD, ATCC, Cellosaurus
- Induced models (drug treatment, surgical intervention, environmental manipulation) > Search first: MGI, model organism databases, PubMed
- Genetic Models:
- Types available (knockout, knock-in, transgenic, conditional, humanized) > Search first: MGI, IMPC, KOMP, EuMMCR, IMSR
- Model Characteristics:
- Phenotype recapitulation (how well model reproduces human disease features) > Search first: Model organism databases, comparative studies, PubMed
- Model limitations (aspects of human disease not captured) > Search first: Model organism databases, PubMed, review articles
- Applications:
- Research applications (what aspects of disease can be studied) > Search first: Model organism databases, PubMed
- Resources:
- Model databases > Search first: MGI, RGD, ZFIN, FlyBase, WormBase, IMSR, EMMA, MMRRC
Citation Requirements
- Cite primary literature (PMID preferred) for all mechanistic and clinical claims
- Prioritize recent reviews and landmark papers
- Include direct quotes from abstracts where possible to support key statements
- Distinguish evidence source types: human clinical, model organism, in vitro, computational
Output Format
Structure your response as a comprehensive narrative organized by the sections above. For each section, provide: - Factual content with specific details (numbers, percentages, gene names, variant nomenclature) - Ontology term suggestions (HPO, GO, CL, UBERON, CHEBI, NCIT, MONDO) where applicable - Evidence citations with PMIDs - Direct quotes from abstracts to support key claims - Clear indication when information is not available or not applicable for this disease
This report will be used to populate a disease knowledge base entry with: - Pathophysiology descriptions with causal chains - Gene/protein annotations (HGNC, GO terms) - Phenotype associations (HP terms) with frequencies - Cell type involvement (CL terms) - Anatomical locations (UBERON terms) - Chemical entities (CHEBI terms) - Treatment annotations (NCIT terms) - Evidence items with PMIDs and exact abstract quotes - Epidemiology, prognosis, diagnostic, and prevention information - Animal model descriptions with phenotype recapitulation details
ATP6V0C-related epilepsy: disease-characteristics report
Evidence scope. ATP6V0C disease spans febrile seizures with normal development through severe neurodevelopmental disorder with epilepsy. The strongest ATP6V0C-specific evidence is a 2023 series of 27 people with heterozygous point variants, a 2022 report of six affected relatives in two families, and earlier reports of multigene 16p13.3 deletions. These are clinically ascertained groups, not a population registry; their phenotype frequencies must not be interpreted as penetrance or prevalence. The 2024 literature principally synthesizes these findings rather than establishing an ATP6V0C-specific treatment. (mattison2023atp6v0cvariantsimpair pages 1-3, mattison2023atp6v0cvariantsimpair pages 5-6, tian2022atp6v0cisassociated pages 4-6, tian2022atp6v0cisassociated pages 6-7, falace2024vatpasedysfunctionin pages 6-8)
1. Disease information
ATP6V0C-related epilepsy is an autosomal-dominant spectrum caused by disease-associated variation in the gene encoding the c subunit of vacuolar H⁺-ATPase (V-ATPase). Epilepsy is not obligatory: developmental problems without reported seizures and familial febrile-seizure presentations both occur. A published disease designation is early-onset epilepsy 3, with or without developmental delay (EEO3; OMIM 620465); the ATP6V0C gene is OMIM 108745, at 16p13.3, with clinical-report reference transcript NM_001694.4. The requested MONDO:0958196 is retained as a supplied identifier, but its current mapping was not independently verified here. An ATP6V0C-specific Orphanet, MeSH, ICD-10, or ICD-11 code was not established from the retrieved sources; generic epilepsy codes should not be represented as disease-specific identifiers. Synonyms suitable for text search include ATP6V0C-associated early-onset epilepsy, ATP6V0C-related neurodevelopmental disorder with or without epilepsy, and EEO3. (mattison2023atp6v0cvariantsimpair pages 1-3, mattison2023atp6v0cvariantsimpair pages 3-4, falace2024vatpasedysfunctionin pages 3-4, carpentieri2024dominantlyactingvariants pages 1-2)
Provenance: The numerical observations below derive from individual patients described in published clinical series and subsequently aggregated by investigators, not from an identified individual’s electronic health record. Mattison and colleagues assembled deidentified clinical information through clinicians, GeneMatcher, sequencing projects, ClinVar, and prior publications; some cases had appeared elsewhere and must not be counted again as independent discoveries. (mattison2023atp6v0cvariantsimpair pages 4-5, mattison2023atp6v0cvariantsimpair pages 5-6)
Primary abstract, Mattison et al. (2023): “We describe heterozygous point variants in ATP6V0C … in 27 patients with neurodevelopmental abnormalities with or without epilepsy.” [Brain 146:1357–1372; https://doi.org/10.1093/brain/awac330]. (mattison2023atp6v0cvariantsimpair pages 1-3)
2. Etiology and risk or protective factors
Established causal factor: Germline heterozygous ATP6V0C missense, frameshift, and stop-loss variants are associated with the phenotype. In the 2023 series, 22/27 had missense substitutions, 4/27 frameshifts, and 1/27 a stop-loss variant; among 24 with parental DNA, variants arose de novo. Four affected individuals were reported mosaic. Familial transmission, rather than an exclusively de novo mechanism, is documented by the two families with febrile seizures reported in 2022. A deletion encompassing ATP6V0C can also contribute, but neighboring TBC1D24 and PDPK1 complicate assignment of a multigene-deletion phenotype solely to ATP6V0C. (mattison2023atp6v0cvariantsimpair pages 5-6, mattison2023atp6v0cvariantsimpair pages 6-7, tian2022atp6v0cisassociated pages 4-6, tian2022atp6v0cisassociated pages 6-7, tinker2021haploinsufficiencyofatp6v0c pages 1-5)
Environment and interaction: Fever accompanied the first seizures in all six members of the 2022 families, making febrile illness a documented seizure precipitant in those carriers, not a cause of their genetic disorder. In experimental worms, high-salt osmotic stress worsened motor/paralysis phenotypes, but this is not evidence that dietary salt modifies disease in humans. No disease-specific causal association was established for smoking, diet, toxins, occupation, pathogens, sex, or geographic ancestry; no protective allele, modifier gene, preventive diet, or human gene–environment interaction estimate was identified. (tian2022atp6v0cisassociated pages 6-7, mattison2023atp6v0cvariantsimpair pages 11-14)
3. Phenotypes
Clinical findings range from episodic febrile seizures with normal development to persistent epilepsy and substantial cognitive or communication disability. The following denominators belong only to the 2023 point-variant cohort and vary with availability of records. (mattison2023atp6v0cvariantsimpair pages 6-7)
| Feature | Observed finding | Available denominator | Interpretation |
|---|---|---|---|
| Developmental delay | 21/23 (91.3%) | 23/27 | Ascertainment-selected; data missing for 4 individuals |
| Intellectual disability | 16/16 (100%) | 16 age-eligible, assessed individuals | Severity ranged from mild to profound; not evaluable or unavailable for the remainder |
| Seizure onset before 24 months | 14/18 (77.8%) | 18 individuals with onset reported | Early onset was common among individuals with available data |
| Mean seizure-onset age | 24.6 ± 8.0 months | 18 individuals with onset reported | Summary estimate from available observations; not the entire cohort |
| Generalized tonic–clonic seizures | 12/19 (63.2%) | 19 individuals with seizure-type data | Seizure types were non-exclusive |
| Focal seizures | 7/19 (36.8%) | 19 individuals with seizure-type data | Seizure types were non-exclusive |
| Atonic seizures | 6/19 (31.6%) | 19 individuals with seizure-type data | Seizure types were non-exclusive |
| Myoclonic seizures | 5/19 (26.3%) | 19 individuals with seizure-type data | Seizure types were non-exclusive |
| Abnormal brain MRI | 13/21 (61.9%) | 21 individuals with MRI data | Findings included callosal/cerebellar-vermian abnormalities and delayed myelination |
| Cohort-level caveat | 27 heterozygous point-variant cases | — | Frequencies are descriptive of a clinically ascertained series and must not be extrapolated to all ATP6V0C variant carriers (mattison2023atp6v0cvariantsimpair pages 6-7) |
Table: Observed findings in the ascertainment-selected 27-person Mattison ATP6V0C point-variant cohort; denominators vary because clinical data were incomplete. The separate Tian 2022 familial cohort—six individuals, all with febrile-seizure onset at 7–8 months and normal development—is not pooled here (tian2022atp6v0cisassociated pages 6-7).
Seizure types overlap within individuals; the 12/19, 7/19, 6/19, and 5/19 figures therefore cannot be summed. MRI abnormalities included callosal or cerebellar-vermian agenesis/hypoplasia and delayed myelination. Four patients were reported to have cardiac findings—pulmonary valve stenosis, thickened ventricular wall, murmur, or cardiomyopathy/valvular disease—but one with multiple cardiac defects also had biallelic LZTR1 variants, a competing explanation. Dental enamel defects occurred in two reported patients. A separate patient carried a potentially confounding 20q11.22–q11.23 deletion. (mattison2023atp6v0cvariantsimpair pages 6-7, mattison2023atp6v0cvariantsimpair pages 7-8)
The 2022 familial series is clinically different: all 6/6 had first febrile seizures at 7–8 months, 2/6 later had afebrile epilepsy, and 6/6 were described as having normal intellectual and physical development. Some relatives became seizure-free without antiseizure drugs. This contrast supports variable expressivity and cautions against labeling every ATP6V0C-associated seizure as a severe developmental and epileptic encephalopathy. (tian2022atp6v0cisassociated pages 6-7)
Knowledge-base phenotype suggestions—map to current HPO releases before ingestion: seizures/epilepsy, febrile seizures, generalized tonic–clonic seizures, focal seizures, atonic seizures, myoclonic seizures, infantile spasms, global developmental delay, intellectual disability, speech delay or absent speech, developmental regression, corpus-callosum hypoplasia/agenesis, delayed myelination, cerebellar-vermian hypoplasia, and cardiac structural anomaly. HP:0001250 (seizure), HP:0001263 (global developmental delay), HP:0001249 (intellectual disability), and HP:0000252 (microcephaly) are candidate mappings; microcephaly is especially pertinent to the multigene deletion literature and should not be assigned the point-variant cohort’s frequency. Quality-of-life consequences are inferred from seizure burden, delayed walking or speech, and intellectual disability; no ATP6V0C-specific EQ-5D, SF-36, or PROMIS estimates were identified. (mattison2023atp6v0cvariantsimpair pages 6-7, tian2022atp6v0cisassociated pages 6-7, tinker2021haploinsufficiencyofatp6v0c pages 1-5)
4. Genetic and molecular information
The implicated protein is a 155-amino-acid, four-transmembrane V₀ c subunit; its gene lies at 16p13.3. For reproducible HGVS reporting, the 2023 table uses NM_001694.4. Examples include c.188G>C (p.Gly63Ala), c.283G>A (p.Ala95Thr), c.409T>C (p.Phe137Leu), c.412G>C (p.Ala138Pro), c.445G>A (p.Ala149Thr), c.448C>T (p.Leu150Phe), and the frameshift c.134_135delCT [p.(Ser45CysfsTer37)]. The reported stop-loss is c.467A>T [p.(Ter156LeuextTer35)]. The 2022 familial variants were c.64G>A (p.Ala22Thr) and c.361_373del (p.Thr121Profs*7); check transcript/version when importing these records. (mattison2023atp6v0cvariantsimpair pages 3-4, mattison2023atp6v0cvariantsimpair pages 6-7, mattison2023atp6v0cvariantsimpair pages 7-8, tian2022atp6v0cisassociated pages 4-6)
The 2023 authors judged their patient variants likely pathogenic or pathogenic under ACMG/AMP criteria, but this is a study-level statement, not a substitute for checking each current ClinVar assertion. Their variants were absent from gnomAD v2.1.1 at the time; 21 observed missense variants versus 108 expected and zero observed loss-of-function variants versus 4.5 expected indicated population constraint. Those dated database observations are not present-day global allele-frequency estimates, and rare population missense variants cannot automatically be called benign or pathogenic: three tested population variants also affected yeast readouts to differing degrees. The study found missense-variant enrichment in transmembrane domain 4 (P = 0.006). (mattison2023atp6v0cvariantsimpair pages 5-6, mattison2023atp6v0cvariantsimpair pages 4-5, mattison2023atp6v0cvariantsimpair pages 8-11)
Functional classification: Reduced V-ATPase-dependent activity is experimentally supported in yeast for tested patient substitutions; haploinsufficiency is supported as a plausible mechanism by frameshifts/deletions and fly knockdown. A dominant-negative action for some outward-facing missense variants remains a structural hypothesis, not an experimentally established universal mechanism. A previously reported stop-loss transcript escaped nonsense-mediated decay, leaving its precise mechanism unsettled. No ATP6V0C-specific epigenetic signature or validated severity modifier was established. The previously described 16p13.3 deletions are contiguous-gene copy-number variants, not evidence for recurrent ATP6V0C-only aneuploidy or translocation. (mattison2023atp6v0cvariantsimpair pages 7-8, mattison2023atp6v0cvariantsimpair pages 3-4, mattison2023atp6v0cvariantsimpair pages 8-11, tinker2021haploinsufficiencyofatp6v0c pages 1-5)
5. Environmental and infectious information
This is a noninfectious Mendelian disorder; there is no implicated causative microorganism or zoonotic transmission. Fever may trigger seizures in some carriers, but the data do not establish that infection changes genetic penetrance. No ATP6V0C-specific human evidence establishes pollution, radiation, occupational agents, alcohol, smoking, exercise, or dietary factors as causes or protective interventions. (tian2022atp6v0cisassociated pages 4-6, tian2022atp6v0cisassociated pages 6-7)
6. Mechanism and pathophysiology
Ordered causal chain; “inferred” means not directly demonstrated in ATP6V0C patient neurons:
- Heterozygous ATP6V0C variant or ATP6V0C-containing deletion leads to altered amount or structure of the V₀ c subunit; reduced dosage is plausible for truncations/deletions, whereas variant-specific effects differ. (mattison2023atp6v0cvariantsimpair pages 5-6, tinker2021haploinsufficiencyofatp6v0c pages 1-5)
- Altered c subunit leads to disrupted c-ring function or c–a-subunit coupling during ATP-driven rotation; for individual missense variants, the interface change is inferred from structural modeling. The c-ring glutamate Glu139 and a-subunit arginine participate in proton translocation. (mattison2023atp6v0cvariantsimpair pages 3-4, mattison2023atp6v0cvariantsimpair pages 7-8)
- Disrupted V-ATPase function results in reduced acidification capacity: directly supported in yeast by diminished LysoSensor signal and poor growth under calcium challenge, but not demonstrated as a measured organellar-pH change in ATP6V0C patient neurons. (mattison2023atp6v0cvariantsimpair pages 8-11, mattison2023atp6v0cvariantsimpair pages 1-3)
- Branch A—synaptic vesicles: reduced proton gradient is inferred to lead to altered neurotransmitter loading/release and neuronal signaling; variant-bearing worms’ aldicarb hypersensitivity supports altered neuromuscular signaling, not direct measurement of human synaptic-vesicle cargo. (mattison2023atp6v0cvariantsimpair pages 11-14, mattison2023atp6v0cvariantsimpair pages 14-15)
- Branch B—Golgi/endolysosomes: altered organelle acidification is inferred to lead to disturbed membrane-protein sorting, lysosomal enzyme activity, and autophagic degradation. These are biologically plausible V-ATPase pathways, not confirmed ATP6V0C-patient multi-omics signatures; experiments on other V-ATPase genes cannot be relabeled ATP6V0C results. (mattison2023atp6v0cvariantsimpair pages 14-15, falace2024vatpasedysfunctionin pages 6-8, carpentieri2024dominantlyactingvariants pages 1-2)
- Changed neural development/circuit activity results in or is inferred to contribute to developmental impairment and seizures in humans; neuron-specific fly knockdown prolongs electroshock-induced seizure-like behavior, furnishing model-organism, not human circuit-level, support. (mattison2023atp6v0cvariantsimpair pages 1-3, mattison2023atp6v0cvariantsimpair pages 7-8, mattison2023atp6v0cvariantsimpair pages 6-7)
The upstream biochemical lesion is proton-pump dysfunction; the proposed synaptic and endomembrane branches are downstream. No ATP6V0C-specific causal role has been demonstrated for Wnt, MAPK, PI3K–AKT, mTOR, inflammation, ferroptosis, or altered DNA methylation. The related-gene 2024 literature contains lysosomal, autophagic, and even increased-acidification findings for other subunits; those mechanisms must not be assumed to apply unchanged to ATP6V0C. No disease-specific human transcriptomic, proteomic, metabolomic, lipidomic, single-cell, spatial-transcriptomic, or multi-omics profile was identified. GO suggestions: proton transmembrane transport, organelle acidification, lysosome organization, endocytosis, autophagy, and synaptic-vesicle transport; assign exact GO accessions only after release-specific validation. CL suggestions: neuron, excitatory neuron, inhibitory neuron; these reflect plausible vulnerable cell populations, not experimentally demonstrated human-cell selectivity. (mattison2023atp6v0cvariantsimpair pages 14-15, falace2024vatpasedysfunctionin pages 6-8, carpentieri2024dominantlyactingvariants pages 1-2)
Primary abstract, Mattison et al. (2023): “Functional analyses conducted in Saccharomyces cerevisiae revealed reduced LysoSensor fluorescence and reduced growth … Knockdown of ATP6V0C in Drosophila resulted in increased duration of seizure-like behaviour.” https://doi.org/10.1093/brain/awac330. (mattison2023atp6v0cvariantsimpair pages 1-3)
7. Anatomical structures affected
The central nervous system is the predominant clinically affected system: the developing brain and distributed seizure-generating neural circuits, with documented callosal/cerebellar-vermian structural findings in some patients. Cardiac findings and enamel abnormalities have been reported but are not proven invariant primary tissue manifestations. There is no established consistent lateralization. At the cellular level, neurons are strongly implicated by the fly neuron-specific experiment; the affected human excitatory versus inhibitory neuronal populations remain unresolved. Relevant subcellular compartments are the V-ATPase-containing synaptic-vesicle, endosomal, lysosomal, and Golgi membranes, with involvement varying by tissue and complex. Ontology suggestions: UBERON brain, cerebral cortex, corpus callosum, cerebellum, heart; CL neuron; GO cellular-component vacuolar proton-transporting V-type ATPase complex, lysosomal membrane, and synaptic vesicle membrane. These are anatomical/mechanistic annotations, not proof of lesion in every structure. (mattison2023atp6v0cvariantsimpair pages 7-8, mattison2023atp6v0cvariantsimpair pages 3-4, mattison2023atp6v0cvariantsimpair pages 14-15, falace2024vatpasedysfunctionin pages 6-8)
8. Temporal development
The disorder begins in infancy or childhood in the best-characterized cases: 14/18 with available onset information in the 2023 series had seizures before 24 months, although individual reported onset included later childhood. In the distinct 2022 family series, onset was 7–8 months, with some seizure courses ending spontaneously and normal later development. Severe cases in the 2023 table included persistent developmental deficits and occasional regression; the literature does not define a standardized sequence of stages, a uniform progression rate, a critical intervention window, or lifelong seizure probability. Thus, the course can be episodic for seizures and persistent or variable for developmental disability. (mattison2023atp6v0cvariantsimpair pages 6-7, mattison2023atp6v0cvariantsimpair pages 7-8, tian2022atp6v0cisassociated pages 6-7)
9. Inheritance and population
Inheritance is autosomal dominant, including de novo and transmitted heterozygous variants; mosaic affected individuals are documented. The 2022 families demonstrate relatively mild expression, whereas the 2023 series shows substantial variation, so complete penetrance, age-specific penetrance, sex ratio, genetic anticipation, founder effects, carrier frequency, incidence, and prevalence cannot be estimated from these ascertainment-selected reports. A 2022 study found its two variants in 2/64 proband alleles from 32 selected FS/EFS+ families and 0/280,788 comparator alleles in its cited aggregated gnomAD dataset; this is a study-specific enrichment comparison, not a population incidence or allele frequency for the disease. No ethnic or geographic concentration has been established. Germline mosaicism in an unaffected parent is a counseling possibility in dominant disease, but was not established as a measured recurrence rate in these sources. (mattison2023atp6v0cvariantsimpair pages 5-6, tian2022atp6v0cisassociated pages 4-6, tian2022atp6v0cisassociated pages 6-7, falace2024vatpasedysfunctionin pages 3-4)
10. Diagnostics
Practical evaluation, extrapolated from the described presentations: characterize seizure events and development; obtain EEG/video-EEG when clinically indicated, brain MRI when warranted by epilepsy or developmental findings, and cardiac assessment when signs or structural concerns are present. Familial probands had generalized approximately 2–4-Hz or 2.5–3.5-Hz abnormalities, whereas a child with a multigene deletion and febrile seizures had an unrevealing 24-hour EEG; therefore neither an EEG pattern nor MRI abnormality is required for a molecular diagnosis. Routine blood, CSF, and metabolic tests were normal in a reported familial proband and are not established ATP6V0C biomarkers. Biopsy and a clinical lysosomal-enzyme assay have no validated disease-specific role. (tian2022atp6v0cisassociated pages 6-7, tinker2021haploinsufficiencyofatp6v0c pages 1-5, mattison2023atp6v0cvariantsimpair pages 6-7)
Molecular confirmation: Favor trio exome or genome sequencing, or a validated epilepsy/neurodevelopmental panel confirmed to include ATP6V0C, with segregation and ACMG/AMP variant assessment. Review copy-number calls at 16p13.3 if phenotype or sequencing suggests a deletion, interpreting TBC1D24/PDPK1 co-deletion separately. Targeted familial-variant testing is appropriate once an informative variant is established. The 2023 authors cautioned that ATP6V0C was missing from some commercial panels at that time; present-day panel content must be checked rather than assumed. CMA can detect a sufficiently large deletion but does not replace sequence testing for small variants; karyotype/FISH, mitochondrial sequencing, repeat-expansion tests, liquid biopsy, and diagnostic RNA/protein/metabolomic panels are not established routine ATP6V0C tests. There is no independently validated disease-specific clinical scoring criterion. Differential diagnosis includes SCN1A-related febrile seizure/Dravet presentations, other genetic developmental epilepsies, and ATP6V0A1/ATP6V1A-related disorders; clinical similarity alone does not distinguish their genotypes. (mattison2023atp6v0cvariantsimpair pages 4-5, mattison2023atp6v0cvariantsimpair pages 14-15, tian2022atp6v0cisassociated pages 6-7, falace2024vatpasedysfunctionin pages 3-4, tinker2021haploinsufficiencyofatp6v0c pages 1-5)
11. Outcome and prognosis
Prognosis is heterogeneous. In the familial six-person series, development remained normal and several individuals stopped having seizures, whereas the separately ascertained 2023 series included severe-to-profound intellectual disability, absent speech or regression in some, and recurrent epilepsy. This supports meaningful morbidity and a likely effect on daily independence, but no validated disease-specific quality-of-life score, survival curve, life expectancy, mortality rate, or prognostic biomarker was identified. Mouse or worm lethality is not a human survival estimate. Genotype-specific risk prediction is premature, particularly where multigene deletions or other variants confound attribution. (tian2022atp6v0cisassociated pages 6-7, mattison2023atp6v0cvariantsimpair pages 6-7, mattison2023atp6v0cvariantsimpair pages 7-8, mattison2023atp6v0cvariantsimpair pages 11-14)
12. Treatment and real-world implementation
Current clinical practice is symptomatic, not ATP6V0C-corrective. In the 2022 report, one proband received valproate, nitrazepam, and then lamotrigine and became seizure-free on the combination; the second improved and became seizure-free on valproate. These are individual case outcomes, not disease-wide response rates or evidence that lamotrigine is universally preferable to another drug. Standard individualized epilepsy care, rescue planning, developmental assessment, speech/occupational/physical therapies when indicated, and evaluation of associated findings are reasonable clinical extrapolations rather than tested ATP6V0C protocols. Seizure-directed surgery has no demonstrated ATP6V0C-specific indication; eligibility would depend on ordinary individualized epilepsy evaluation. (tian2022atp6v0cisassociated pages 4-6, tian2022atp6v0cisassociated pages 6-7, mattison2023atp6v0cvariantsimpair pages 7-8)
Preclinical treatment result: In ATP6V0C-ortholog knockdown fly larvae, levetiracetam and topiramate shortened electroshock recovery most clearly; lamotrigine and valproate also had effects, whereas phenytoin did not significantly change recovery at the tested fly dose. This assay does not establish human comparative efficacy or contraindicate phenytoin. The 2024 review discusses other V-ATPase/lysosome-directed ideas, but none is established as an ATP6V0C-specific human therapy. An ATP6V0C-specific interventional trial or NCT identifier was not identified by the clinical-trials search; there is no established gene replacement/editing, ASO, cell therapy, immunotherapy, or genotype-guided pharmacogenomic rule. Suggested NCIT intervention labels for ontology matching: antiepileptic therapy, valproate therapy, lamotrigine therapy, levetiracetam therapy, physical therapy, occupational therapy, and speech therapy; validate exact NCIT codes before entry. (mattison2023atp6v0cvariantsimpair pages 7-8, falace2024vatpasedysfunctionin pages 14-15, falace2024vatpasedysfunctionin pages 12-14)
13. Prevention
There is no known way to prevent occurrence of a pathogenic de novo variant through lifestyle change or immunization. For an established familial variant, genetic counseling and informed reproductive options, including targeted prenatal or preimplantation testing where appropriate, are relevant; recurrence estimates require the family’s actual segregation and mosaicism findings rather than a blanket rate. Early recognition of seizures, individualized treatment, developmental services, and ordinary seizure-safety measures are secondary or tertiary prevention of consequences, not proven prevention of the genetic disease. Avoid claiming a disease-specific newborn screen, vaccine, preventive medication, or public-health environmental intervention: none was identified. (mattison2023atp6v0cvariantsimpair pages 5-6, tian2022atp6v0cisassociated pages 6-7, tinker2021haploinsufficiencyofatp6v0c pages 1-5)
14. Other species and naturally occurring disease
The human disorder is not transmissible and has no zoonotic potential. Conservation of ATP6V0C orthologs supports comparative experiments in mouse (Mus musculus), zebrafish (Danio rerio), fruit fly (Drosophila melanogaster), nematode (Caenorhabditis elegans), and budding yeast (Saccharomyces cerevisiae). These are experimental ortholog systems, not established reports of a naturally occurring ATP6V0C epilepsy in a veterinary breed. No VBO breed entry, veterinary carrier frequency, or naturally affected animal population was established; species-specific NCBI Taxon and ortholog Gene accessions require database validation before structured import. (mattison2023atp6v0cvariantsimpair pages 3-4, mattison2023atp6v0cvariantsimpair pages 4-5, tian2022atp6v0cisassociated pages 6-7)
15. Model organisms and functional resources
- Yeast, S. cerevisiae: VMA3 ortholog variants were expressed in a vma3Δ background. 11/12 tested patient variants reduced LysoSensor fluorescence versus wild-type rescue; 11/12 reduced growth under the reported 5 mM CaCl₂ condition. This is strong evidence for disrupted proton-pump-associated function, but a haploid yeast rescue does not reproduce human heterozygosity or epilepsy. (mattison2023atp6v0cvariantsimpair pages 4-5, mattison2023atp6v0cvariantsimpair pages 8-11)
- Fruit fly, D. melanogaster: pan-neuronal RNAi against Vha16-3/CG32090 prolonged recovery from electroshock-induced seizure-like behavior. This models reduced neuronal gene dosage and a treatment-screening readout, not spontaneous human epilepsy. (mattison2023atp6v0cvariantsimpair pages 4-5, mattison2023atp6v0cvariantsimpair pages 7-8)
- Nematode, C. elegans: CRISPR-generated ortholog substitutions corresponding to p.Gly63Ala, p.Phe137Leu, and p.Leu150Phe caused smaller size, movement/paralysis abnormalities, aldicarb hypersensitivity, and shortened lifespan; salt stress amplified some findings. A fourth modeled substitution corresponding to p.Ala95Thr caused homozygous sterility. These homozygous invertebrate genotypes do not replicate the typical human heterozygous state or directly demonstrate spontaneous seizures. (mattison2023atp6v0cvariantsimpair pages 4-5, mattison2023atp6v0cvariantsimpair pages 11-14)
- Mouse and zebrafish: reported Atp6v0c homozygous knockout mouse lethality and zebrafish ortholog perturbation support essential conserved biology, but were not validated here as variant-specific ATP6V0C human-epilepsy models. No ATP6V0C patient-derived iPSC-neuron model, disease-specific mouse knock-in with measured human-like seizure phenotype, or organoid rescue was established in the retrieved ATP6V0C primary study. MGI, FlyBase, WormBase, SGD, and ZFIN are suitable next resources for release-verified model identifiers, not proof that each hosts a disease-specific model. (tian2022atp6v0cisassociated pages 6-7, mattison2023atp6v0cvariantsimpair pages 4-5, mattison2023atp6v0cvariantsimpair pages 1-3)
Evidence and identifier caution. The principal paper is Mattison et al., Brain 2023;146:1357–1372, DOI 10.1093/brain/awac330; the familial report is Tian et al., Frontiers in Molecular Neuroscience May 2022, DOI 10.3389/fnmol.2022.889534; the contextual review is Falace et al., Cells August 2024, DOI 10.3390/cells13171441. Deletion context comes from Mucha et al., Genetics in Medicine 2019, DOI 10.1038/s41436-018-0290-3, and Tinker et al., American Journal of Medical Genetics A 2021, DOI 10.1002/ajmg.a.61905. PMIDs were not present in the retrievable source metadata and are intentionally not invented. Exact HGNC, NCIT, ChEBI, CL, UBERON, and most HPO/GO accession assignments likewise need authoritative ontology lookup before database insertion; plausible chemical-label suggestions include H⁺, ATP, CaCl₂, valproate, and lamotrigine, without asserting unverified ChEBI codes. (mattison2023atp6v0cvariantsimpair pages 1-3, tian2022atp6v0cisassociated pages 4-6, falace2024vatpasedysfunctionin pages 6-8, tinker2021haploinsufficiencyofatp6v0c pages 1-5)
References
-
(mattison2023atp6v0cvariantsimpair pages 1-3): Kari A Mattison, Gilles Tossing, Fred Mulroe, Callum Simmons, Kameryn M Butler, Alison Schreiber, Adnan Alsadah, Derek E Neilson, Karin Naess, Anna Wedell, Anna Wredenberg, Arthur Sorlin, Emma McCann, George J Burghel, Beatriz Menendez, George E Hoganson, Lorenzo D Botto, Francis M Filloux, Ángel Aledo-Serrano, Antonio Gil-Nagel, Katrina Tatton-Brown, Nienke E Verbeek, Bert van der Zwaag, Kyrieckos A Aleck, Andrew C Fazenbaker, Jorune Balciuniene, Holly A Dubbs, Eric D Marsh, Kathryn Garber, Jakob Ek, Morten Duno, Christina E Hoei-Hansen, Matthew A Deardorff, Gordana Raca, Catherine Quindipan, Michele van Hirtum-Das, Jeroen Breckpot, Trine Bjørg Hammer, Rikke S Møller, Andrea Whitney, Andrew G L Douglas, Mira Kharbanda, Nicola Brunetti-Pierri, Manuela Morleo, Vincenzo Nigro, Halie J May, James X Tao, Emanuela Argilli, Elliot H Sherr, William B Dobyns, Richard A Baines, Jim Warwicker, J Alex Parker, Siddharth Banka, Philippe M Campeau, and Andrew Escayg. atp6v0c variants impair v-atpase function causing a neurodevelopmental disorder often associated with epilepsy. Brain, 146:1357-1372, Sep 2023. URL: https://doi.org/10.1093/brain/awac330, doi:10.1093/brain/awac330. This article has 36 citations and is from a highest quality peer-reviewed journal.
-
(mattison2023atp6v0cvariantsimpair pages 5-6): Kari A Mattison, Gilles Tossing, Fred Mulroe, Callum Simmons, Kameryn M Butler, Alison Schreiber, Adnan Alsadah, Derek E Neilson, Karin Naess, Anna Wedell, Anna Wredenberg, Arthur Sorlin, Emma McCann, George J Burghel, Beatriz Menendez, George E Hoganson, Lorenzo D Botto, Francis M Filloux, Ángel Aledo-Serrano, Antonio Gil-Nagel, Katrina Tatton-Brown, Nienke E Verbeek, Bert van der Zwaag, Kyrieckos A Aleck, Andrew C Fazenbaker, Jorune Balciuniene, Holly A Dubbs, Eric D Marsh, Kathryn Garber, Jakob Ek, Morten Duno, Christina E Hoei-Hansen, Matthew A Deardorff, Gordana Raca, Catherine Quindipan, Michele van Hirtum-Das, Jeroen Breckpot, Trine Bjørg Hammer, Rikke S Møller, Andrea Whitney, Andrew G L Douglas, Mira Kharbanda, Nicola Brunetti-Pierri, Manuela Morleo, Vincenzo Nigro, Halie J May, James X Tao, Emanuela Argilli, Elliot H Sherr, William B Dobyns, Richard A Baines, Jim Warwicker, J Alex Parker, Siddharth Banka, Philippe M Campeau, and Andrew Escayg. atp6v0c variants impair v-atpase function causing a neurodevelopmental disorder often associated with epilepsy. Brain, 146:1357-1372, Sep 2023. URL: https://doi.org/10.1093/brain/awac330, doi:10.1093/brain/awac330. This article has 36 citations and is from a highest quality peer-reviewed journal.
-
(tian2022atp6v0cisassociated pages 4-6): Yang Tian, Qiong-Xiang Zhai, Xiao-Jing Li, Zhen Shi, Chuan-Fang Cheng, Cui-Xia Fan, Bin Tang, Ying Zhang, Yun-Yan He, Wen-Bin Li, Sheng Luo, Chi Hou, Wen-Xiong Chen, Wei-Ping Liao, and Jie Wang. Atp6v0c is associated with febrile seizures and epilepsy with febrile seizures plus. Frontiers in Molecular Neuroscience, May 2022. URL: https://doi.org/10.3389/fnmol.2022.889534, doi:10.3389/fnmol.2022.889534. This article has 28 citations.
-
(tian2022atp6v0cisassociated pages 6-7): Yang Tian, Qiong-Xiang Zhai, Xiao-Jing Li, Zhen Shi, Chuan-Fang Cheng, Cui-Xia Fan, Bin Tang, Ying Zhang, Yun-Yan He, Wen-Bin Li, Sheng Luo, Chi Hou, Wen-Xiong Chen, Wei-Ping Liao, and Jie Wang. Atp6v0c is associated with febrile seizures and epilepsy with febrile seizures plus. Frontiers in Molecular Neuroscience, May 2022. URL: https://doi.org/10.3389/fnmol.2022.889534, doi:10.3389/fnmol.2022.889534. This article has 28 citations.
-
(falace2024vatpasedysfunctionin pages 6-8): Antonio Falace, Greta Volpedo, Marcello Scala, Federico Zara, Pasquale Striano, and Anna Fassio. V-atpase dysfunction in the brain: genetic insights and therapeutic opportunities. Cells, 13:1441, Aug 2024. URL: https://doi.org/10.3390/cells13171441, doi:10.3390/cells13171441. This article has 30 citations.
-
(mattison2023atp6v0cvariantsimpair pages 3-4): Kari A Mattison, Gilles Tossing, Fred Mulroe, Callum Simmons, Kameryn M Butler, Alison Schreiber, Adnan Alsadah, Derek E Neilson, Karin Naess, Anna Wedell, Anna Wredenberg, Arthur Sorlin, Emma McCann, George J Burghel, Beatriz Menendez, George E Hoganson, Lorenzo D Botto, Francis M Filloux, Ángel Aledo-Serrano, Antonio Gil-Nagel, Katrina Tatton-Brown, Nienke E Verbeek, Bert van der Zwaag, Kyrieckos A Aleck, Andrew C Fazenbaker, Jorune Balciuniene, Holly A Dubbs, Eric D Marsh, Kathryn Garber, Jakob Ek, Morten Duno, Christina E Hoei-Hansen, Matthew A Deardorff, Gordana Raca, Catherine Quindipan, Michele van Hirtum-Das, Jeroen Breckpot, Trine Bjørg Hammer, Rikke S Møller, Andrea Whitney, Andrew G L Douglas, Mira Kharbanda, Nicola Brunetti-Pierri, Manuela Morleo, Vincenzo Nigro, Halie J May, James X Tao, Emanuela Argilli, Elliot H Sherr, William B Dobyns, Richard A Baines, Jim Warwicker, J Alex Parker, Siddharth Banka, Philippe M Campeau, and Andrew Escayg. atp6v0c variants impair v-atpase function causing a neurodevelopmental disorder often associated with epilepsy. Brain, 146:1357-1372, Sep 2023. URL: https://doi.org/10.1093/brain/awac330, doi:10.1093/brain/awac330. This article has 36 citations and is from a highest quality peer-reviewed journal.
-
(falace2024vatpasedysfunctionin pages 3-4): Antonio Falace, Greta Volpedo, Marcello Scala, Federico Zara, Pasquale Striano, and Anna Fassio. V-atpase dysfunction in the brain: genetic insights and therapeutic opportunities. Cells, 13:1441, Aug 2024. URL: https://doi.org/10.3390/cells13171441, doi:10.3390/cells13171441. This article has 30 citations.
-
(carpentieri2024dominantlyactingvariants pages 1-2): Giovanna Carpentieri, Serena Cecchetti, Gianfranco Bocchinfuso, Francesca Clementina Radio, Chiara Leoni, Roberta Onesimo, Paolo Calligari, Agostina Pietrantoni, Andrea Ciolfi, Marco Ferilli, Cristina Calderan, Gerarda Cappuccio, Simone Martinelli, Elena Messina, Viviana Caputo, Ulrike Hüffmeier, Cyril Mignot, Stéphane Auvin, Yline Capri, Charles Marques Lourenco, Bianca E. Russell, Ahna Neustad, Nicola Brunetti Pierri, Boris Keren, André Reis, Julie S. Cohen, Alexis Heidlebaugh, Clay Smith, Christian T. Thiel, Leonardo Salviati, Giuseppe Zampino, Philippe M. Campeau, Lorenzo Stella, Marco Tartaglia, and Elisabetta Flex. Dominantly acting variants in atp6v1c1 and atp6v1b2 cause a multisystem phenotypic spectrum by altering lysosomal and/or autophagosome function. Human Genetics and Genomics Advances, 5:100349, Oct 2024. URL: https://doi.org/10.1016/j.xhgg.2024.100349, doi:10.1016/j.xhgg.2024.100349. This article has 14 citations and is from a peer-reviewed journal.
-
(mattison2023atp6v0cvariantsimpair pages 4-5): Kari A Mattison, Gilles Tossing, Fred Mulroe, Callum Simmons, Kameryn M Butler, Alison Schreiber, Adnan Alsadah, Derek E Neilson, Karin Naess, Anna Wedell, Anna Wredenberg, Arthur Sorlin, Emma McCann, George J Burghel, Beatriz Menendez, George E Hoganson, Lorenzo D Botto, Francis M Filloux, Ángel Aledo-Serrano, Antonio Gil-Nagel, Katrina Tatton-Brown, Nienke E Verbeek, Bert van der Zwaag, Kyrieckos A Aleck, Andrew C Fazenbaker, Jorune Balciuniene, Holly A Dubbs, Eric D Marsh, Kathryn Garber, Jakob Ek, Morten Duno, Christina E Hoei-Hansen, Matthew A Deardorff, Gordana Raca, Catherine Quindipan, Michele van Hirtum-Das, Jeroen Breckpot, Trine Bjørg Hammer, Rikke S Møller, Andrea Whitney, Andrew G L Douglas, Mira Kharbanda, Nicola Brunetti-Pierri, Manuela Morleo, Vincenzo Nigro, Halie J May, James X Tao, Emanuela Argilli, Elliot H Sherr, William B Dobyns, Richard A Baines, Jim Warwicker, J Alex Parker, Siddharth Banka, Philippe M Campeau, and Andrew Escayg. atp6v0c variants impair v-atpase function causing a neurodevelopmental disorder often associated with epilepsy. Brain, 146:1357-1372, Sep 2023. URL: https://doi.org/10.1093/brain/awac330, doi:10.1093/brain/awac330. This article has 36 citations and is from a highest quality peer-reviewed journal.
-
(mattison2023atp6v0cvariantsimpair pages 6-7): Kari A Mattison, Gilles Tossing, Fred Mulroe, Callum Simmons, Kameryn M Butler, Alison Schreiber, Adnan Alsadah, Derek E Neilson, Karin Naess, Anna Wedell, Anna Wredenberg, Arthur Sorlin, Emma McCann, George J Burghel, Beatriz Menendez, George E Hoganson, Lorenzo D Botto, Francis M Filloux, Ángel Aledo-Serrano, Antonio Gil-Nagel, Katrina Tatton-Brown, Nienke E Verbeek, Bert van der Zwaag, Kyrieckos A Aleck, Andrew C Fazenbaker, Jorune Balciuniene, Holly A Dubbs, Eric D Marsh, Kathryn Garber, Jakob Ek, Morten Duno, Christina E Hoei-Hansen, Matthew A Deardorff, Gordana Raca, Catherine Quindipan, Michele van Hirtum-Das, Jeroen Breckpot, Trine Bjørg Hammer, Rikke S Møller, Andrea Whitney, Andrew G L Douglas, Mira Kharbanda, Nicola Brunetti-Pierri, Manuela Morleo, Vincenzo Nigro, Halie J May, James X Tao, Emanuela Argilli, Elliot H Sherr, William B Dobyns, Richard A Baines, Jim Warwicker, J Alex Parker, Siddharth Banka, Philippe M Campeau, and Andrew Escayg. atp6v0c variants impair v-atpase function causing a neurodevelopmental disorder often associated with epilepsy. Brain, 146:1357-1372, Sep 2023. URL: https://doi.org/10.1093/brain/awac330, doi:10.1093/brain/awac330. This article has 36 citations and is from a highest quality peer-reviewed journal.
-
(tinker2021haploinsufficiencyofatp6v0c pages 1-5): Rory J Tinker, George J Burghel, Shruti Garg, Maggie Steggall, Sara Cuvertino, and Siddharth Banka. Haploinsufficiency of atp6v0c possibly underlies 16p13.3 deletions that cause microcephaly, seizures, and neurodevelopmental disorder. American Journal of Medical Genetics Part A, 185:196-202, Oct 2021. URL: https://doi.org/10.1002/ajmg.a.61905, doi:10.1002/ajmg.a.61905. This article has 19 citations.
-
(mattison2023atp6v0cvariantsimpair pages 11-14): Kari A Mattison, Gilles Tossing, Fred Mulroe, Callum Simmons, Kameryn M Butler, Alison Schreiber, Adnan Alsadah, Derek E Neilson, Karin Naess, Anna Wedell, Anna Wredenberg, Arthur Sorlin, Emma McCann, George J Burghel, Beatriz Menendez, George E Hoganson, Lorenzo D Botto, Francis M Filloux, Ángel Aledo-Serrano, Antonio Gil-Nagel, Katrina Tatton-Brown, Nienke E Verbeek, Bert van der Zwaag, Kyrieckos A Aleck, Andrew C Fazenbaker, Jorune Balciuniene, Holly A Dubbs, Eric D Marsh, Kathryn Garber, Jakob Ek, Morten Duno, Christina E Hoei-Hansen, Matthew A Deardorff, Gordana Raca, Catherine Quindipan, Michele van Hirtum-Das, Jeroen Breckpot, Trine Bjørg Hammer, Rikke S Møller, Andrea Whitney, Andrew G L Douglas, Mira Kharbanda, Nicola Brunetti-Pierri, Manuela Morleo, Vincenzo Nigro, Halie J May, James X Tao, Emanuela Argilli, Elliot H Sherr, William B Dobyns, Richard A Baines, Jim Warwicker, J Alex Parker, Siddharth Banka, Philippe M Campeau, and Andrew Escayg. atp6v0c variants impair v-atpase function causing a neurodevelopmental disorder often associated with epilepsy. Brain, 146:1357-1372, Sep 2023. URL: https://doi.org/10.1093/brain/awac330, doi:10.1093/brain/awac330. This article has 36 citations and is from a highest quality peer-reviewed journal.
-
(mattison2023atp6v0cvariantsimpair pages 7-8): Kari A Mattison, Gilles Tossing, Fred Mulroe, Callum Simmons, Kameryn M Butler, Alison Schreiber, Adnan Alsadah, Derek E Neilson, Karin Naess, Anna Wedell, Anna Wredenberg, Arthur Sorlin, Emma McCann, George J Burghel, Beatriz Menendez, George E Hoganson, Lorenzo D Botto, Francis M Filloux, Ángel Aledo-Serrano, Antonio Gil-Nagel, Katrina Tatton-Brown, Nienke E Verbeek, Bert van der Zwaag, Kyrieckos A Aleck, Andrew C Fazenbaker, Jorune Balciuniene, Holly A Dubbs, Eric D Marsh, Kathryn Garber, Jakob Ek, Morten Duno, Christina E Hoei-Hansen, Matthew A Deardorff, Gordana Raca, Catherine Quindipan, Michele van Hirtum-Das, Jeroen Breckpot, Trine Bjørg Hammer, Rikke S Møller, Andrea Whitney, Andrew G L Douglas, Mira Kharbanda, Nicola Brunetti-Pierri, Manuela Morleo, Vincenzo Nigro, Halie J May, James X Tao, Emanuela Argilli, Elliot H Sherr, William B Dobyns, Richard A Baines, Jim Warwicker, J Alex Parker, Siddharth Banka, Philippe M Campeau, and Andrew Escayg. atp6v0c variants impair v-atpase function causing a neurodevelopmental disorder often associated with epilepsy. Brain, 146:1357-1372, Sep 2023. URL: https://doi.org/10.1093/brain/awac330, doi:10.1093/brain/awac330. This article has 36 citations and is from a highest quality peer-reviewed journal.
-
(mattison2023atp6v0cvariantsimpair pages 8-11): Kari A Mattison, Gilles Tossing, Fred Mulroe, Callum Simmons, Kameryn M Butler, Alison Schreiber, Adnan Alsadah, Derek E Neilson, Karin Naess, Anna Wedell, Anna Wredenberg, Arthur Sorlin, Emma McCann, George J Burghel, Beatriz Menendez, George E Hoganson, Lorenzo D Botto, Francis M Filloux, Ángel Aledo-Serrano, Antonio Gil-Nagel, Katrina Tatton-Brown, Nienke E Verbeek, Bert van der Zwaag, Kyrieckos A Aleck, Andrew C Fazenbaker, Jorune Balciuniene, Holly A Dubbs, Eric D Marsh, Kathryn Garber, Jakob Ek, Morten Duno, Christina E Hoei-Hansen, Matthew A Deardorff, Gordana Raca, Catherine Quindipan, Michele van Hirtum-Das, Jeroen Breckpot, Trine Bjørg Hammer, Rikke S Møller, Andrea Whitney, Andrew G L Douglas, Mira Kharbanda, Nicola Brunetti-Pierri, Manuela Morleo, Vincenzo Nigro, Halie J May, James X Tao, Emanuela Argilli, Elliot H Sherr, William B Dobyns, Richard A Baines, Jim Warwicker, J Alex Parker, Siddharth Banka, Philippe M Campeau, and Andrew Escayg. atp6v0c variants impair v-atpase function causing a neurodevelopmental disorder often associated with epilepsy. Brain, 146:1357-1372, Sep 2023. URL: https://doi.org/10.1093/brain/awac330, doi:10.1093/brain/awac330. This article has 36 citations and is from a highest quality peer-reviewed journal.
-
(mattison2023atp6v0cvariantsimpair pages 14-15): Kari A Mattison, Gilles Tossing, Fred Mulroe, Callum Simmons, Kameryn M Butler, Alison Schreiber, Adnan Alsadah, Derek E Neilson, Karin Naess, Anna Wedell, Anna Wredenberg, Arthur Sorlin, Emma McCann, George J Burghel, Beatriz Menendez, George E Hoganson, Lorenzo D Botto, Francis M Filloux, Ángel Aledo-Serrano, Antonio Gil-Nagel, Katrina Tatton-Brown, Nienke E Verbeek, Bert van der Zwaag, Kyrieckos A Aleck, Andrew C Fazenbaker, Jorune Balciuniene, Holly A Dubbs, Eric D Marsh, Kathryn Garber, Jakob Ek, Morten Duno, Christina E Hoei-Hansen, Matthew A Deardorff, Gordana Raca, Catherine Quindipan, Michele van Hirtum-Das, Jeroen Breckpot, Trine Bjørg Hammer, Rikke S Møller, Andrea Whitney, Andrew G L Douglas, Mira Kharbanda, Nicola Brunetti-Pierri, Manuela Morleo, Vincenzo Nigro, Halie J May, James X Tao, Emanuela Argilli, Elliot H Sherr, William B Dobyns, Richard A Baines, Jim Warwicker, J Alex Parker, Siddharth Banka, Philippe M Campeau, and Andrew Escayg. atp6v0c variants impair v-atpase function causing a neurodevelopmental disorder often associated with epilepsy. Brain, 146:1357-1372, Sep 2023. URL: https://doi.org/10.1093/brain/awac330, doi:10.1093/brain/awac330. This article has 36 citations and is from a highest quality peer-reviewed journal.
-
(falace2024vatpasedysfunctionin pages 14-15): Antonio Falace, Greta Volpedo, Marcello Scala, Federico Zara, Pasquale Striano, and Anna Fassio. V-atpase dysfunction in the brain: genetic insights and therapeutic opportunities. Cells, 13:1441, Aug 2024. URL: https://doi.org/10.3390/cells13171441, doi:10.3390/cells13171441. This article has 30 citations.
-
(falace2024vatpasedysfunctionin pages 12-14): Antonio Falace, Greta Volpedo, Marcello Scala, Federico Zara, Pasquale Striano, and Anna Fassio. V-atpase dysfunction in the brain: genetic insights and therapeutic opportunities. Cells, 13:1441, Aug 2024. URL: https://doi.org/10.3390/cells13171441, doi:10.3390/cells13171441. This article has 30 citations.
Artifacts
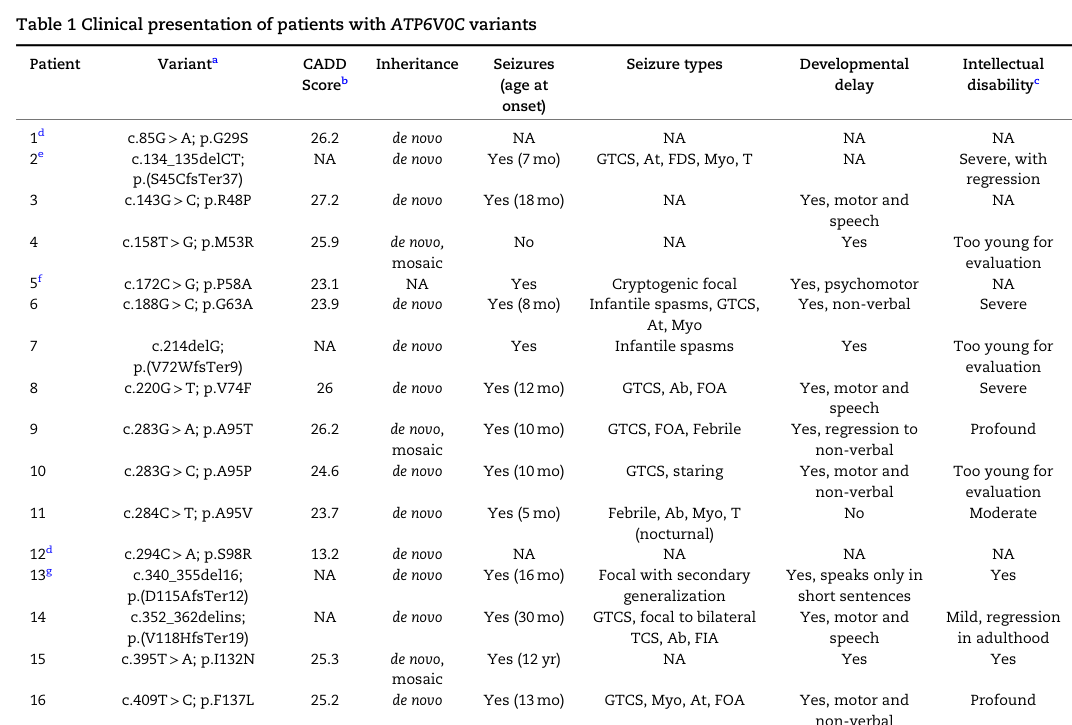
Reference Validation
Checked with linkml-reference-validator 0.3.0rc3.
| Outcome | Count |
|---|---|
| References checked | 5 |
| Resolved | 5 |
| Unresolved (possible confabulation) | 0 |
| Unverifiable | 0 |
| Quoted claims checked | 1 |
| Quoted claims found in source | 0 |
| Quoted claims not found in source | 1 |
| References weighed for topical relevance | 5 |
| On topic | 4 |
| Off topic | 0 |
Quotes not found in the cited source
Searched the abstract, any retrieved full text, and the title. A quote drawn from a part of the paper that was not retrieved will appear here too, so check before treating one as invented:
Every one of these was searched against an abstract alone, with no full text retrieved - marked abstract only below. Where full text can be fetched, re-running with it will settle them; where the source publishes only a summary to PubMed, as GeneReviews chapters do, it will not, and the quote has to be checked by hand against the chapter itself.
DOI:10.1093/brain/awac330(abstract only): "We describe heterozygous point variants in ATP6V0C … in 27 patients with neurodevelopmental abnormalities with or without epilepsy."- closest text in source: "We describe heterozygous point variants in ATP6V0C, encoding the c-subunit in the membrane bound integral domain of the vacuolar H+-ATPase, in 27 patients with neurodevelopmental abnormalities with or without epilepsy"
Term Validation
Checked with linkml-term-validator 0.4.5, through the ols: adapter.
| Outcome | Count |
|---|---|
| Terms checked | 5 |
| Resolved | 5 |
| Unresolved (possible confabulation) | 0 |
| Obsolete | 0 |
| Unverifiable | 0 |
| Terms whose name was checked | 5 |
| Terms named correctly | 4 |
| Terms named as a different term | 1 |
Terms the report names something else
These identifiers resolve, so nothing about them looks wrong, and the ontology calls them something unrelated to what the report calls them. That usually means the identifier is not the one the sentence needs:
MONDO:0958196(2 mentions) - the report calls it "if available"; MONDO calls it epilepsy, early-onset, 3, with or without developmental delay